通过范德瓦尔斯相互作用建模对氧化纳米颗粒的几何依赖性多克索鲁比辛加载的实验和理论评估
Panyada Sripaturad1, Sereysonita Keo2, Anongnat Wongpan2
1Department of Mathematics, Faculty of Science, Mahidol University, Bangkok, 10400, Thailand.
Scientific reports
|January 24, 2026

概括
纳米粒子几何学显著影响药物加载效率. 虽然模型预测球形和板形的结合,但它们与圆柱形相扎,凸显了复杂交互模型的需要.
科学领域:
- 材料科学 材料科学 材料科学
- 纳米技术 纳米技术
- 计算化学计算化学
背景情况:
- 纳米粒子药物递送系统提供有针对性的治疗潜力.
- 了解药物-纳米粒子相互作用对于优化药物递送效率至关重要.
- 纳米粒子几何学是影响药物加载的尚未探索的因素.
研究的目的:
- 调查纳米粒子几何学如何影响多克索鲁比 (DOX) 负载效率.
- 开发和验证用于预测药物-纳米粒子相互作用的分析模型.
- 在复杂的纳米生物系统中评估范德瓦尔斯 (vdW) 模型的局限性.
主要方法:
- 合成和表征球形,板形和圆柱形的纳米粒子.
- 实验测量了每个几何形状的多克索鲁比 (DOX) 负载效率.
- 开发了分析模型来计算vdW相互作用能量,用于理论负载和吸附场景.
主要成果:
- 实验结果显示,球形和板状纳米颗粒的高DOX加载效率.
- 分析模型准确地预测了球形和板状几何体的高结合能,与实验数据相关联.
- 对圆柱形纳米粒子观察到一个差异,预测的结合能量与高实验负载效率不匹配.
结论:
- 纳米粒子几何学在药物加载效率方面发挥着至关重要的作用.
- 基于真空的简单VDW模型不足以完全捕捉复杂的药物-纳米粒子相互作用.
- 结合实验和建模方法,为理解和完善药物输送系统提供了一个框架.
相关概念视频
 01:24
Van der Waals Interactions
71.0K
Atoms and molecules interact with each other through intermolecular forces. These electrostatic forces arise from attractive or repulsive interactions between particles with permanent, partial, or temporary charges. The intermolecular forces between neutral atoms and molecules are ion–dipole, dipole–dipole, and dispersion forces, collectively known as van der Waals forces.
71.0K
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
38.9K
Thus far, the ideal gas law, PV = nRT, has been applied to a variety of different types of problems, ranging from reaction stoichiometry and empirical and molecular formula problems to determining the density and molar mass of a gas. However, the behavior of a gas is often non-ideal, meaning that the observed relationships between its pressure, volume, and temperature are not accurately described by the gas laws.
38.9K
 01:10
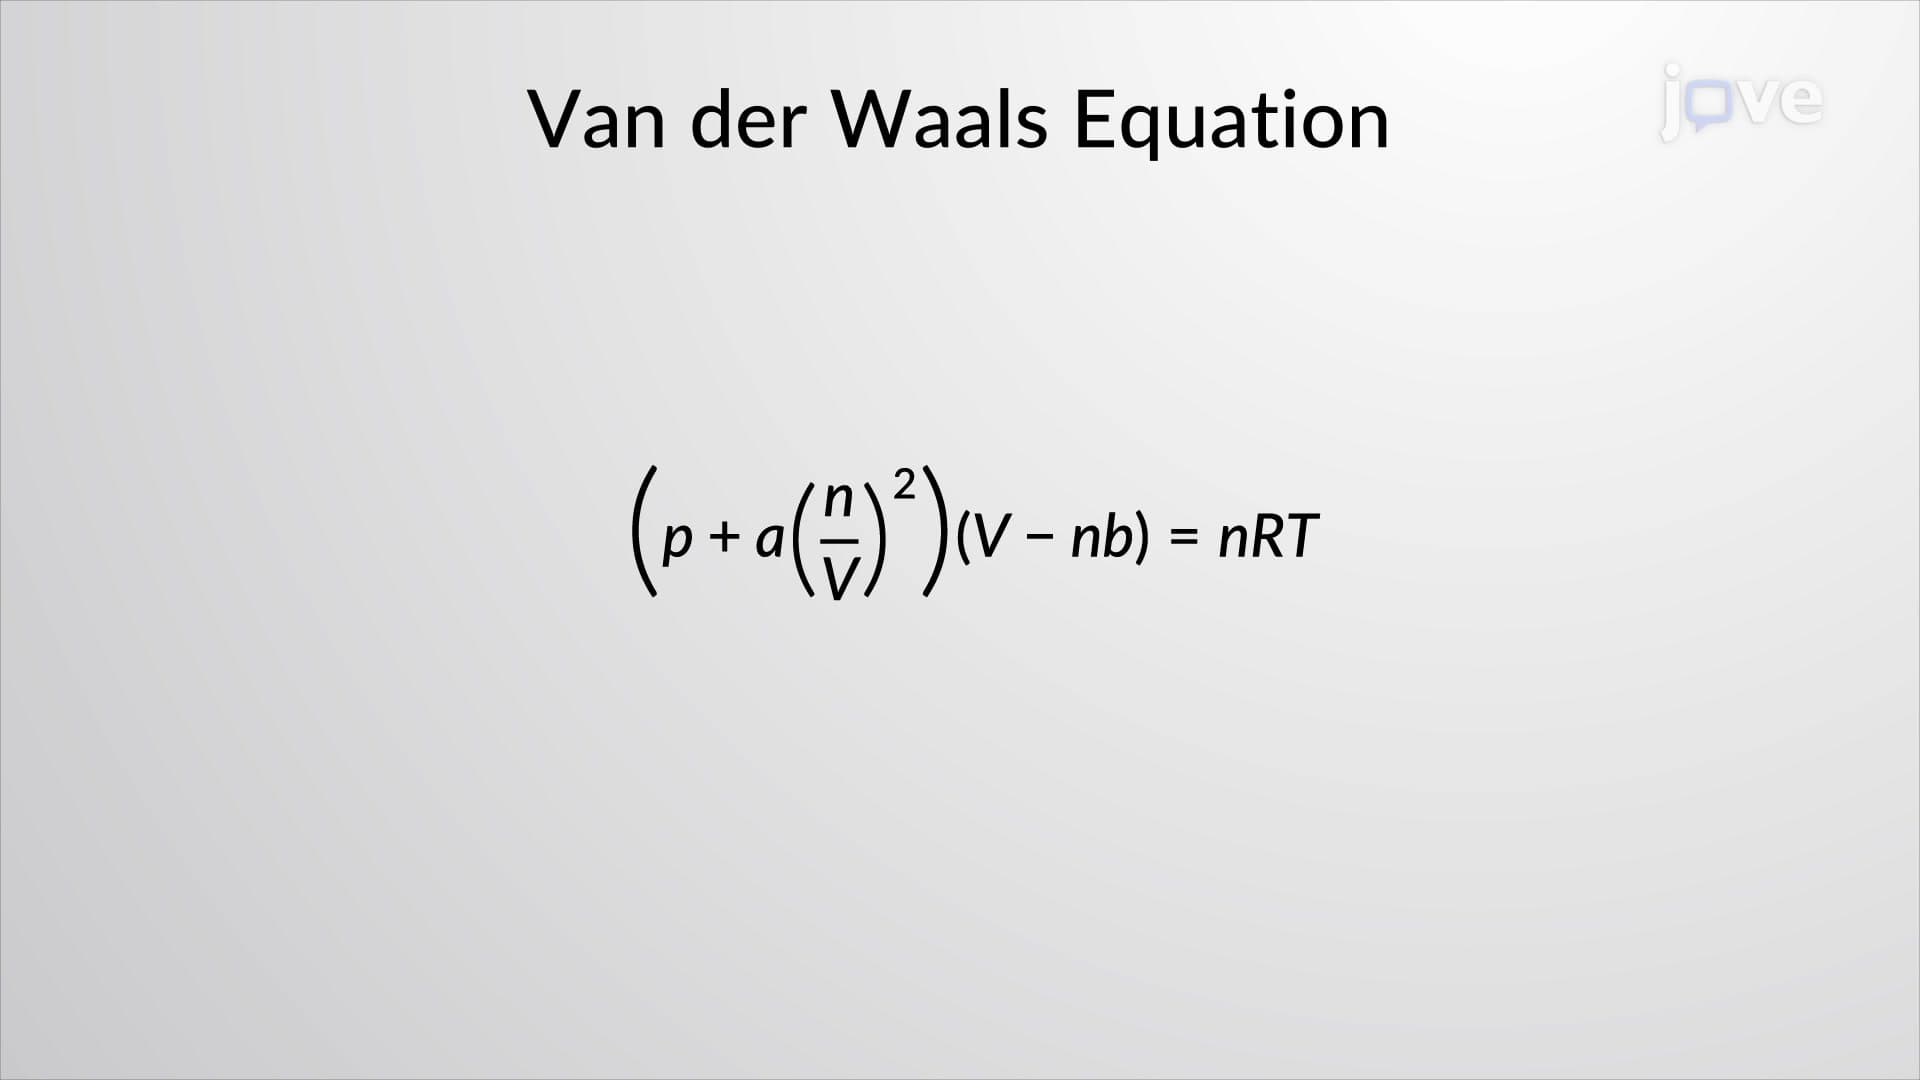
Van der Waals Equation
6.2K
The ideal gas law is an approximation that works well at high temperatures and low pressures. The van der Waals equation of state (named after the Dutch physicist Johannes van der Waals, 1837−1923) improves it by considering two factors.
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
6.2K
 02:57
Coordination Number and Geometry
19.0K
For transition metal complexes, the coordination number determines the geometry around the central metal ion. Table 1 compares coordination numbers to molecular geometry. The most common structures of the complexes in coordination compounds are octahedral, tetrahedral, and square planar.
19.0K
 02:27
Predicting Molecular Geometry
45.6K
VSEPR Theory for Determination of Electron Pair Geometries
45.6K
 02:35
Noncovalent Attractions in Biomolecules
64.3K
Noncovalent attractions are associations within and between molecules that influence the shape and structural stability of complexes. These interactions differ from covalent bonding in that they do not involve sharing of electrons.
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
64.3K