
在固体电解质Li10GeP2S12中模拟四极性NMR动态
Tabea Huss1, Federico Civaia1, Simone S Köcher1,2
1Fritz-Haber Institute of the Max Planck Society, Berlin (DE), Germany.

机器学习加速了固体电解质的分子动力学模拟,例如10硫化 (LGPS). 这种方法准确地预测了离子扩散动态,改进了电池材料分析.
科学领域:
- 固态化学 固态化学
- 材料科学是一种材料科学.
- 计算化学是一种计算化学.
背景情况:
- 四极固态核磁共振 (NMR) 光谱对固体电解质中的离子扩散动态敏感.
- 由于材料复杂性和运动缩小效应,解释NMR数据具有挑战性.
- 原子模拟在计算上昂贵,并且经常使用理想化的模型.
研究的目的:
- 开发机器学习 (ML) 辅助的工作流程,以解决电池材料的固态NMR研究中的实验复杂性.
- 为了实现微秒级分子动力学 (MD) 模拟,并预测离子导体的电场梯度 (EFG) 张量.
- 准确预测NMR可观测物和区分离子电池材料中的离子动力学.
主要方法:
- 用于微秒级MD模拟Li10GeP2S12 (LGPS) 的ML加速.
- 从MD轨迹中使用张量模型进行高效的EFG张量预测.
- 在中计算了温度依赖的7Li NMR四极可观测物和模拟的自旋对齐回声 (SAE) 实验.
主要成果:
- 在四边形LGPS的预测 (24 kHz) 和实验 (23 kHz) 四极合之间取得了很好的一致性.
- 成功预测了7Li NMR四极可观测的温度依赖性,解释了运动缩小.
- 在不同的LGPS晶体结构中,在模拟的SAE数据上使用反拉普拉斯变换提取了离子运动的相关时间.
结论:
- 这种ML辅助的工作流有效地克服了对电池材料复杂固态NMR数据的解释方面的局限性.
- 可以准确预测NMR可观测量和离子动态,从而提高对离子扩散的理解.
- 开发的方法表明,有望在固体电解质中区分谷物间与谷物内离子动态.
更多相关视频
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
相关概念视频
 02:42
Valence Bond Theory
 02:54
Molecular and Ionic Solids
Molecular Solids
Molecular crystalline solids, such as ice, sucrose (table sugar), and iodine, are solids that are composed of neutral molecules as their constituent units. These molecules are held together by weak intermolecular forces such as London dispersion forces, dipole-dipole interactions, or hydrogen bonds, which...
 01:27
Electrostatic Boundary Conditions in Dielectrics
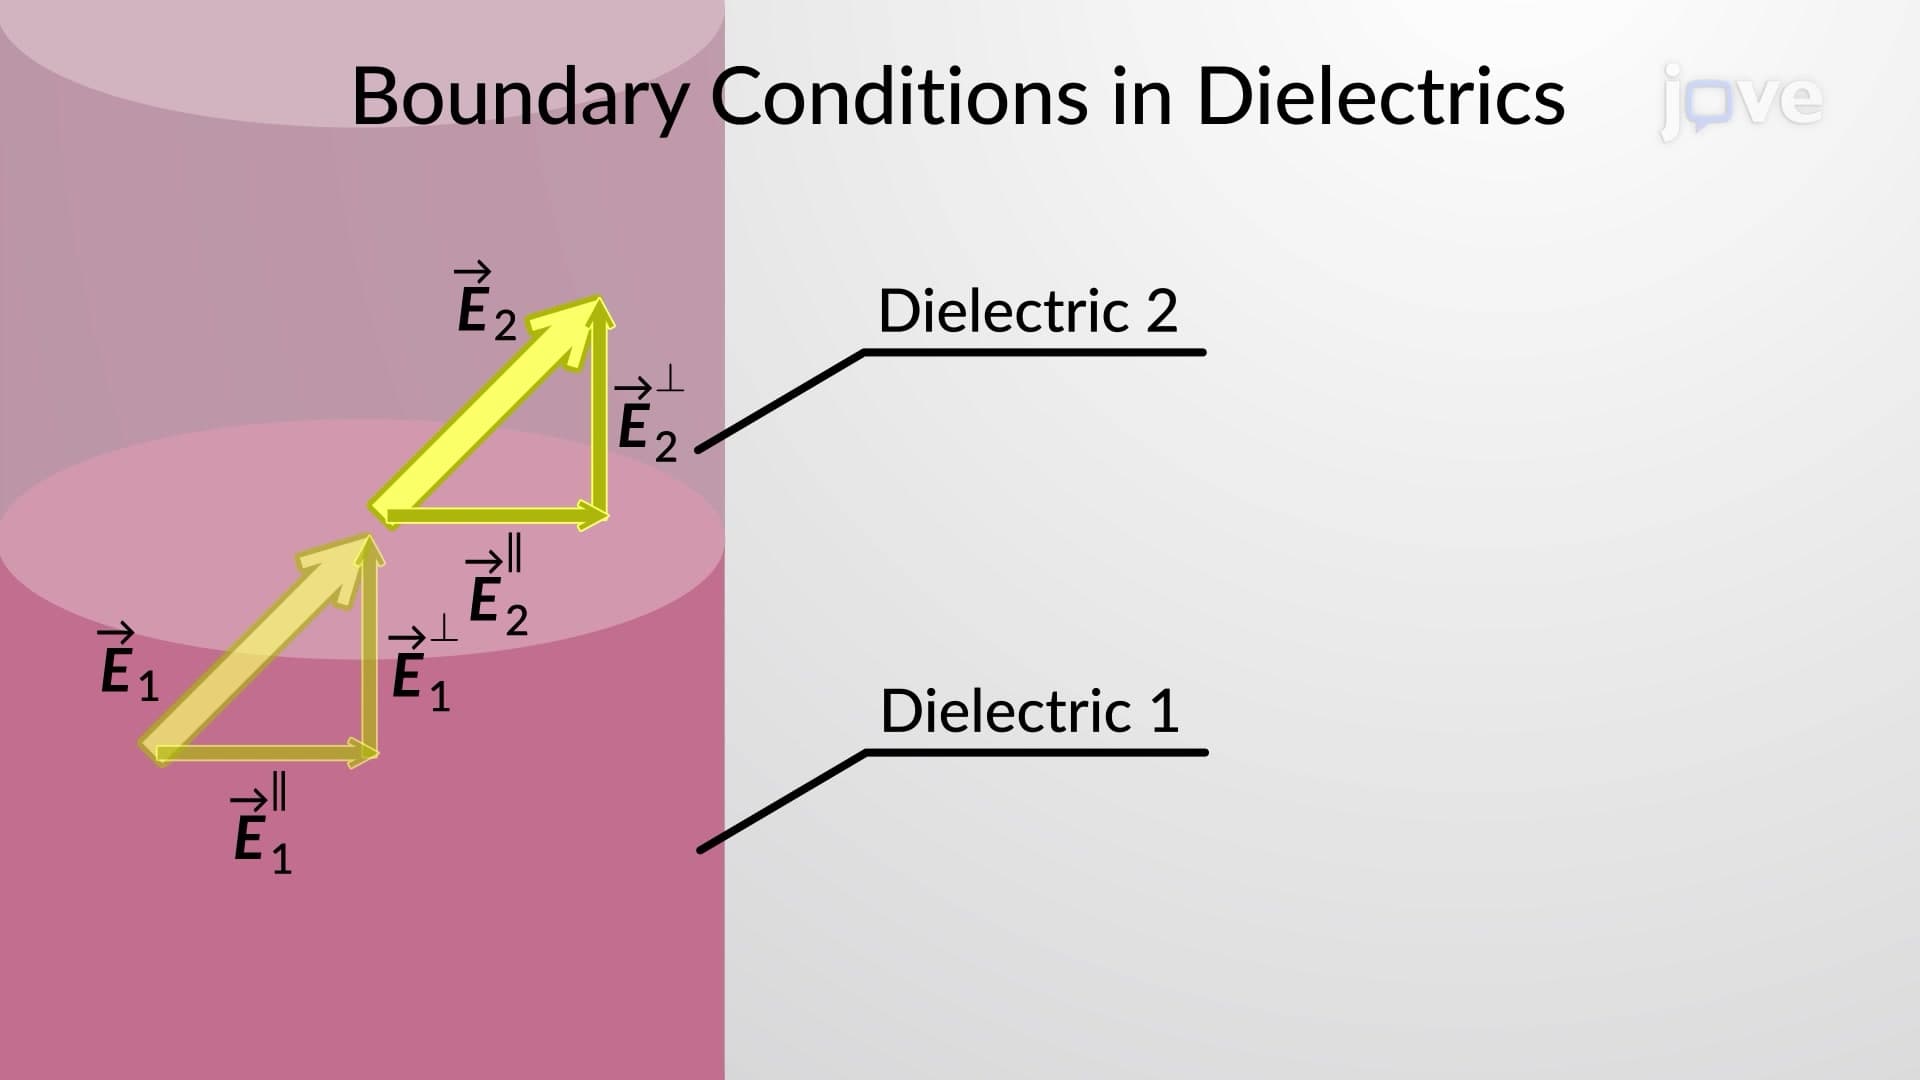
Consider a case where both the mediums across a boundary are two different dielectric materials. Recall that the electric field and electric displacement are proportional and related through the material's permittivity....
 02:54
Trends in Lattice Energy: Ion Size and Charge
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...