
对高度配方的蛋白质与蛋白质相互作用的辅助剂效应的实验评估和xDLVO建模
Kai Ge1, Jiabin Shen1, Yuanhui Ji2
1Key Laboratory of Pollution Exposure and Health Intervention of Zhejiang Province, College of Biology and Environmental Engineering, Zhejiang Shuren University, Hangzhou 310015, P. R. China.
Molecular pharmaceutics
|March 2, 2026

概括
这项研究评估了稳定高度蛋白质配方的辅助剂. 糖,多和氨基酸增强蛋白质排斥力,而盐减少它,帮助配方设计.
科学领域:
- 生物制药开发 生物制药开发
- 物理化学 物理化学
- 蛋白质科学 蛋白质科学
背景情况:
- 由于聚合,设计高度蛋白质配方具有挑战性.
- 辅助剂查对于稳定蛋白质药物至关重要.
- 了解辅助剂-蛋白质相互作用是制剂成功的关键.
研究的目的:
- 系统地评估各种辅助剂对蛋白质相互作用的影响.
- 为了研究蛋白质溶液中的分子间力量的辅助剂介导调制.
- 为生物制药开发中辅助剂查开发一个预测框架.
主要方法:
- 结合实验和建模方法.
- 使用静态光散射测量透式二次病毒系数 (B22).
- 应用扩展的德贾金 - 兰道 - 维维 - 奥弗比克 (xDLVO) 模型.
主要成果:
- 辅助剂的类型和度显著改变了蛋白质相互作用.
- 糖,多和氨基酸通常会增强排斥性相互作用.
- 盐通过静电屏蔽来降低B22,而糖/多醇则通过排斥体积和水竞争来起作用.
结论:
- 一个预测模型 (xDLVO) 准确地捕捉了辅助剂对蛋白质相互作用的影响.
- 这一框架有助于合理选择用于稳定高度蛋白质配方的辅助剂.
- 这项研究为生物制药开发中辅助剂的早期查提供了强大的工具.
相关概念视频
 01:09
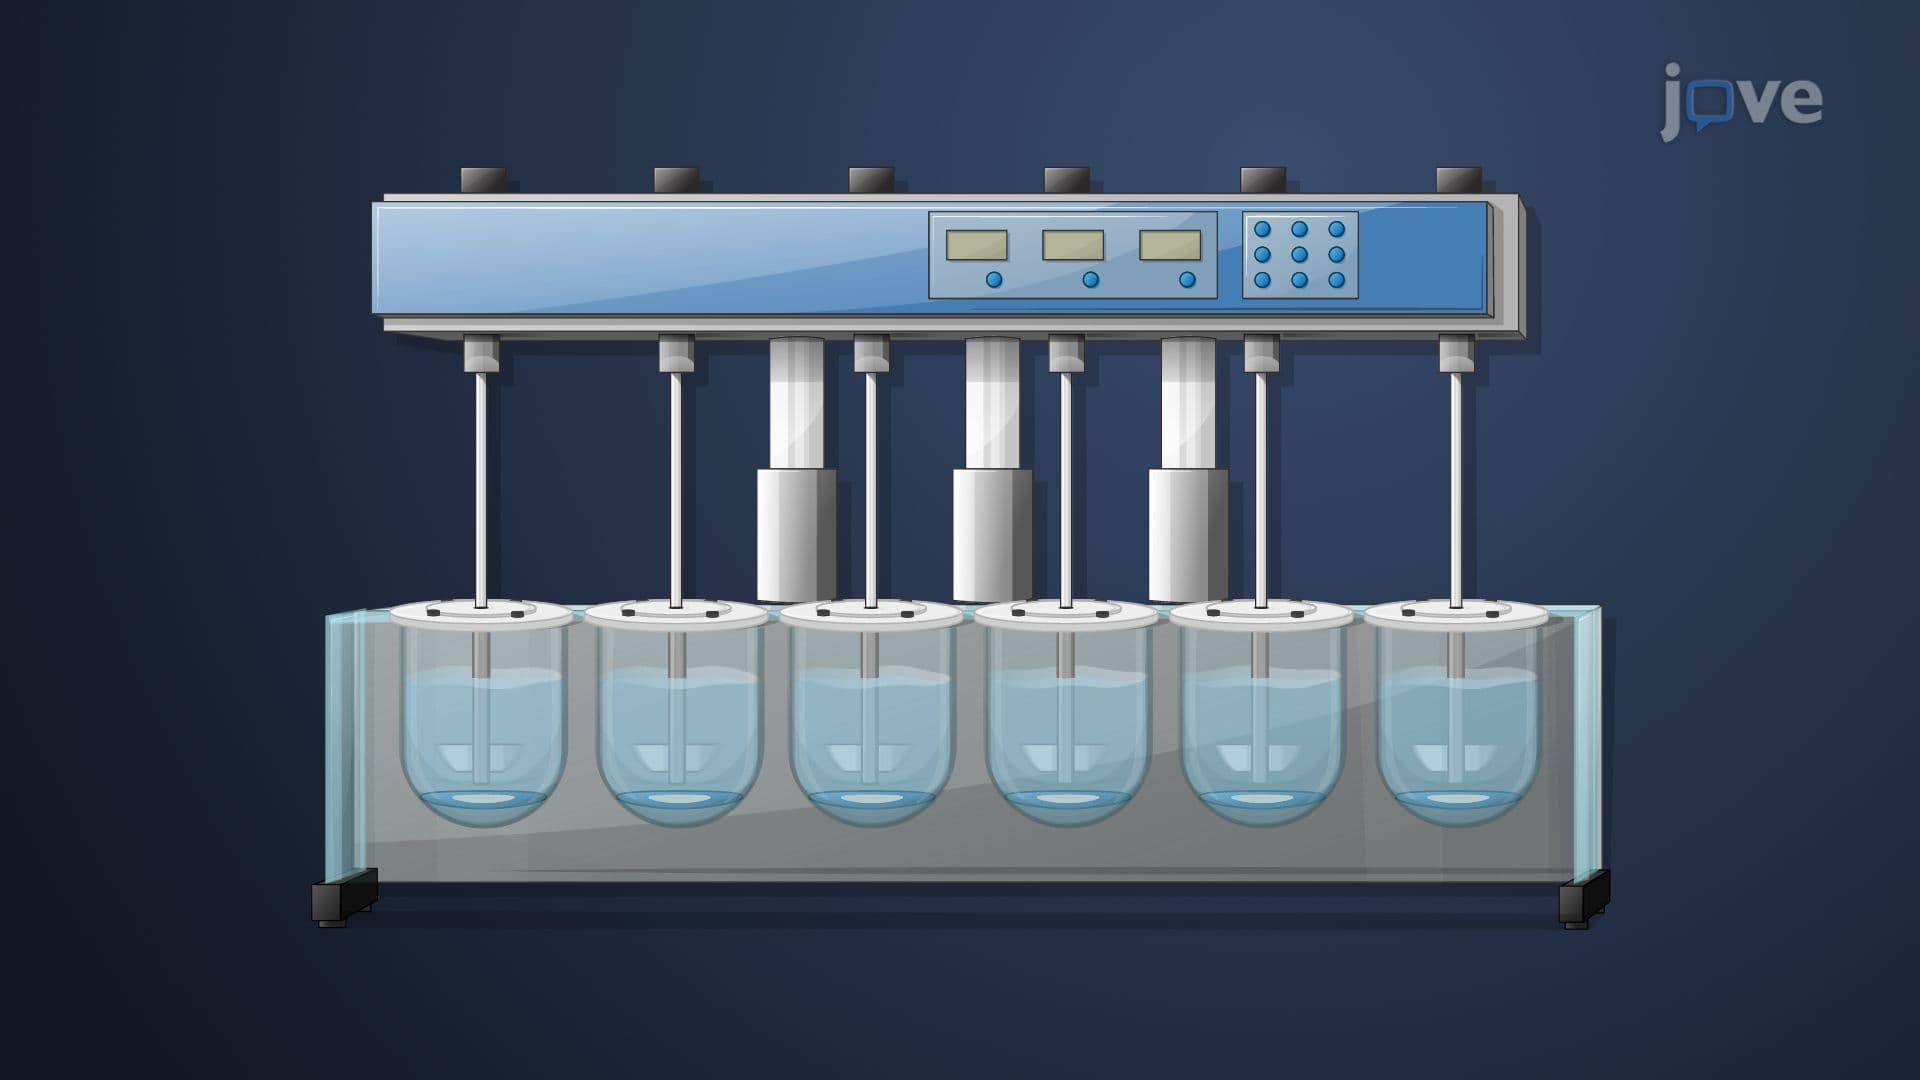
In Vitro Drug Dissolution: Compendial Testing Models II
412
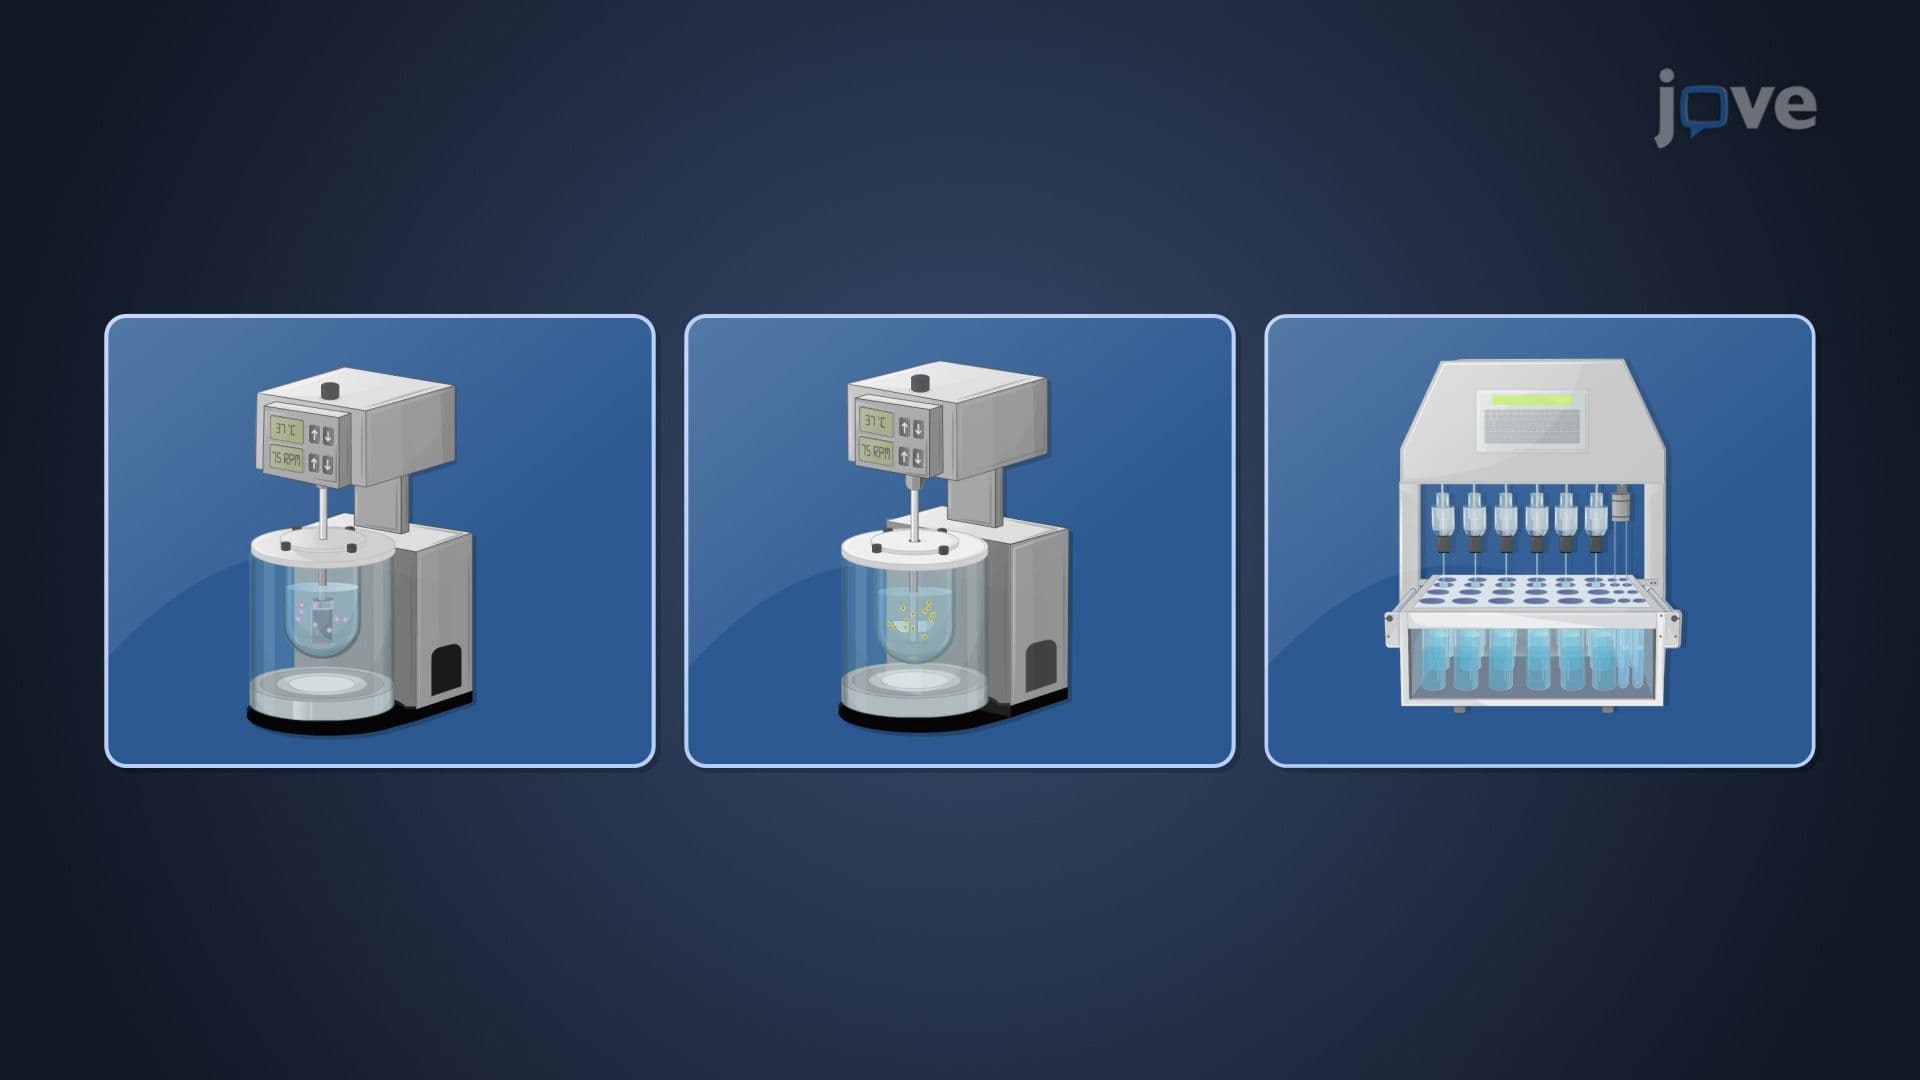
Various dissolution methods are utilized to assess a drug’s dissolution rate, including the flow-through cell, paddle-over-disk, cylinder, and reciprocating disk methods.The flow-through cell apparatus (USP (United States Pharmacopeia) method 4) comprises a reservoir for the dissolution medium and a pump that propels the medium through the cell containing the test sample. This method is crucial for assessing modified-release dosage forms with minimally soluble active ingredients,...
412
 01:13
In Vitro Drug Dissolution: Compendial Testing Models I
385
Compendial dissolution methods are standardized procedures defined by pharmacopeias to evaluate the rate at which a drug dissolves in a specific medium. These methods ensure batch-to-batch consistency, enable quality control, and support the prediction of drug bioavailability. They are critical for both immediate and modified-release drug products.The apparatuses used for dissolution testing differ in their design and mechanical function, but all aim to simulate the physiological environment of...
385
 01:28
Factors Influencing Drug Absorption: Pharmaceutical Parameters
667
Solid dosage forms such as tablets and capsules undergo rigorous manufacturing processes to ensure stability and effectiveness. Their dissolution and absorption properties are influenced significantly by the choice of excipients (inactive ingredients that serve various roles in the formulation), and the methodology applied during production. The manufacturing parameters, such as compression force and granulation techniques, significantly affect dissolution rates. Elevated compression forces...
667
 01:18
Pharmacodynamic Models: Emax Drug–Concentration Effect Model
81
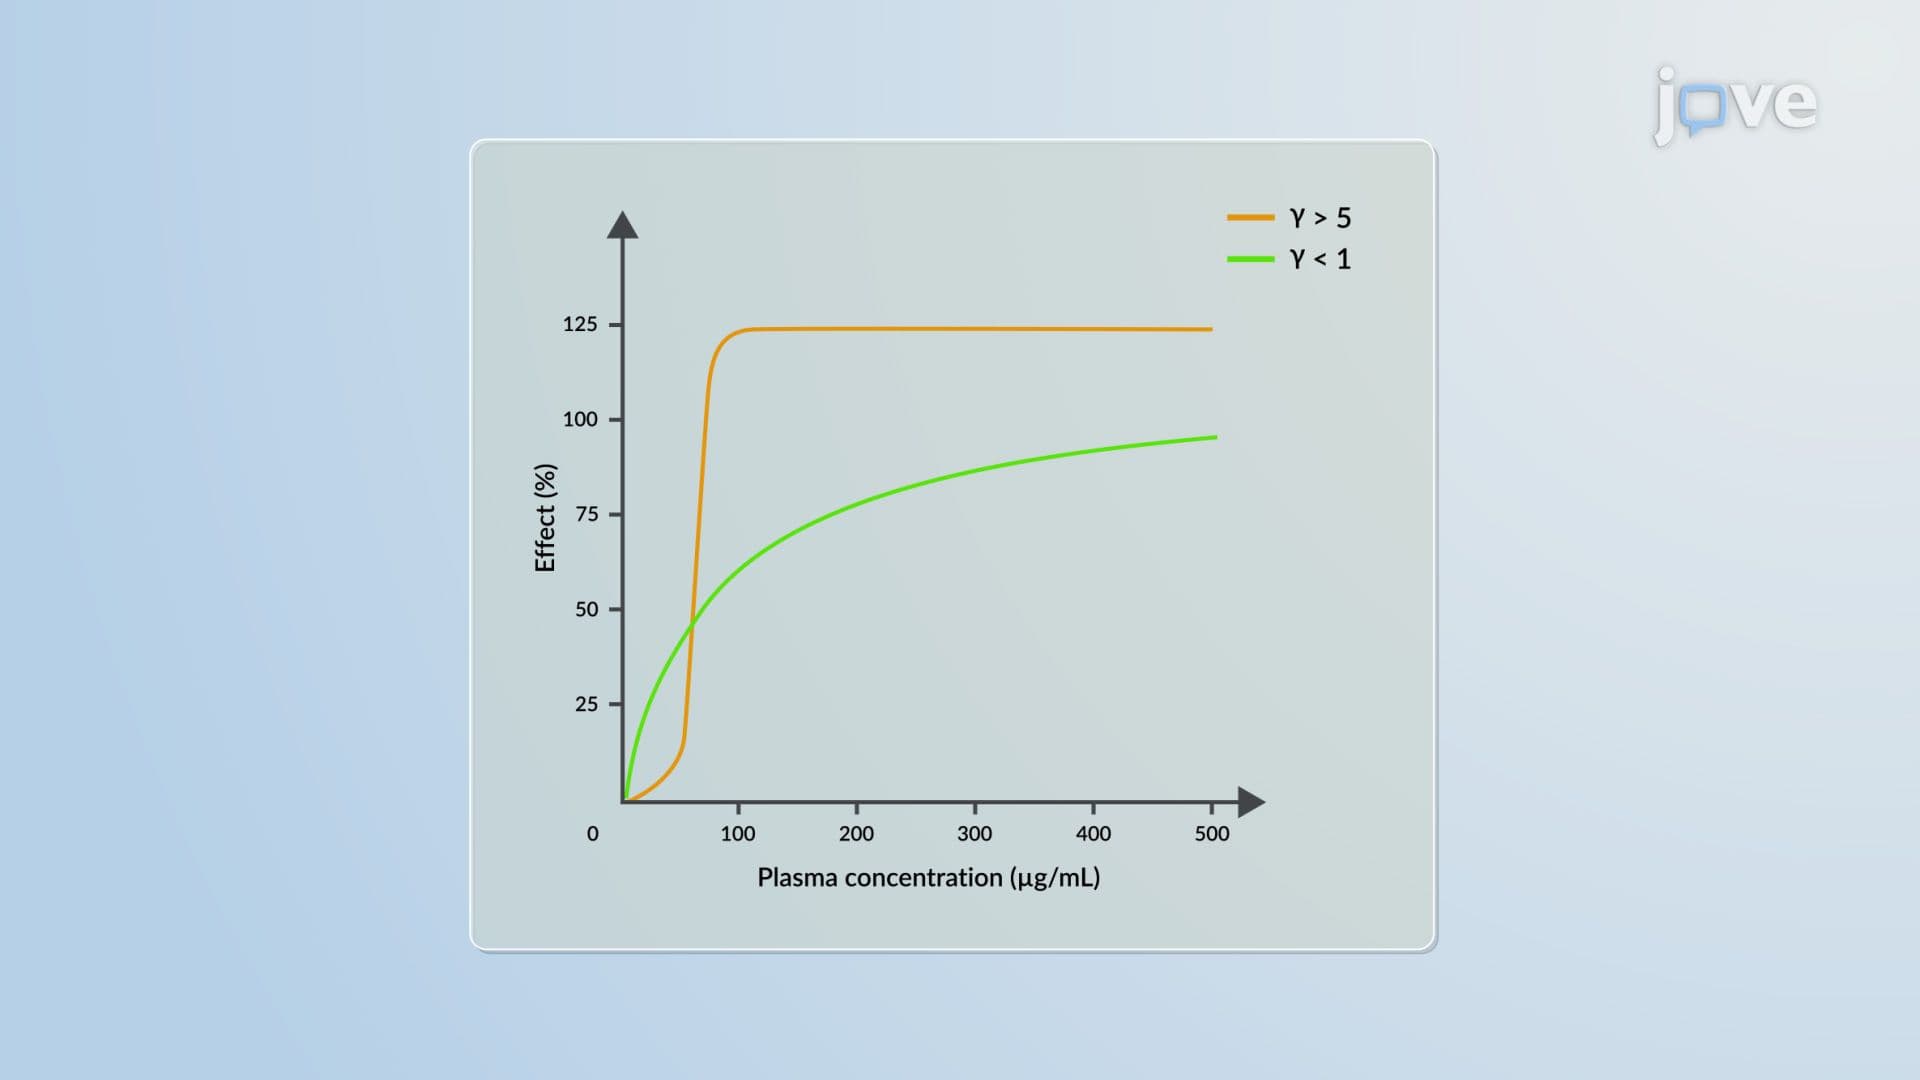
The Emax drug-concentration effect model is central to pharmacodynamics in drug discovery and development. This model is predicated on the receptor occupancy theory, which posits that the effect of a drug is directly related to the number of receptors occupied by the drug and the resultant complex formation.The model describes the reversible interaction between a drug (C) and a receptor (R) to form a drug-receptor complex (RC). The kinetics of this interaction are quantified by an equation that...
81
 01:09
Theories of Dissolution: The Danckwerts' Model and Interfacial Barrier Model
882
Various dissolution theories provide insight into the factors that influence the dissolution rate. Danckwerts' Model suggests that turbulence, rather than a stagnant layer, characterizes the dissolution medium at the solid-liquid interface. In this model, the agitated solvent contains macroscopic packets that move to the interface via eddy currents, facilitating the absorption and delivery of the drug to the bulk solution. The regular replenishment of solvent packets maintains the...
882
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model
41
The linear concentration–effect model, underpinned by the principle that pharmacological effect (E) is directly proportional to plasma drug concentration (C), emerges as a pivotal simplification of the Emax model for conditions where C is significantly less than EC50. This model portrays a linear trajectory of the concentration–effect relationship when drug levels are markedly below the EC50 threshold.Despite its inherent assumption of continuous effect augmentation with increasing...
41