Kearns-Sayre syndrome presenting as 2-oxoadipic aciduria

B A Barshop1, W L Nyhan, R K Naviaux

  • 1Departments of Pediatrics, Medicine and Neurosciences, The Institute of Molecular Genetics, University of California San Diego, California 92093, USA.

Related Concept Videos

Protein Import into the Peroxisomes01:27

Protein Import into the Peroxisomes

Cells contain membrane-bound organelles called peroxisomes that oxidize organic molecules by transferring hydrogen atoms to oxygen, producing hydrogen peroxide. Peroxisomes enzymatically convert the released hydrogen peroxide into water and oxygen.
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
Overview of Protein Metabolism01:21

Overview of Protein Metabolism

Proteins are broken down into amino acids during digestion. Unlike fats and carbohydrates, which are stored for later use, proteins are not. Instead, amino acids are either used to produce ATP through oxidation or contribute to the creation of new proteins for the growth and repair of the body. Any surplus amino acids from the diet are converted into glucose or triglycerides rather than excreted.
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...
Inborn Errors of Metabolism01:20

Inborn Errors of Metabolism

Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...
Diagnosing Acidosis and Alkalosis01:24

Diagnosing Acidosis and Alkalosis

Diagnosing acid-base imbalances involves systematically analyzing arterial blood samples, focusing on three key measurements: pH, bicarbonate (HCO3−) concentration, and carbon dioxide partial pressure (PCO2). This analysis follows a four-step process that helps identify the imbalance's underlying cause and nature.
First, the pH level is assessed to determine whether the blood pH is normal (7.35–7.45), low (acidosis), or high (alkalosis).
Next, the PCO2  and HCO3−  values are examined to...
Diabetic Ketoacidosis l: Introduction01:25

Diabetic Ketoacidosis l: Introduction
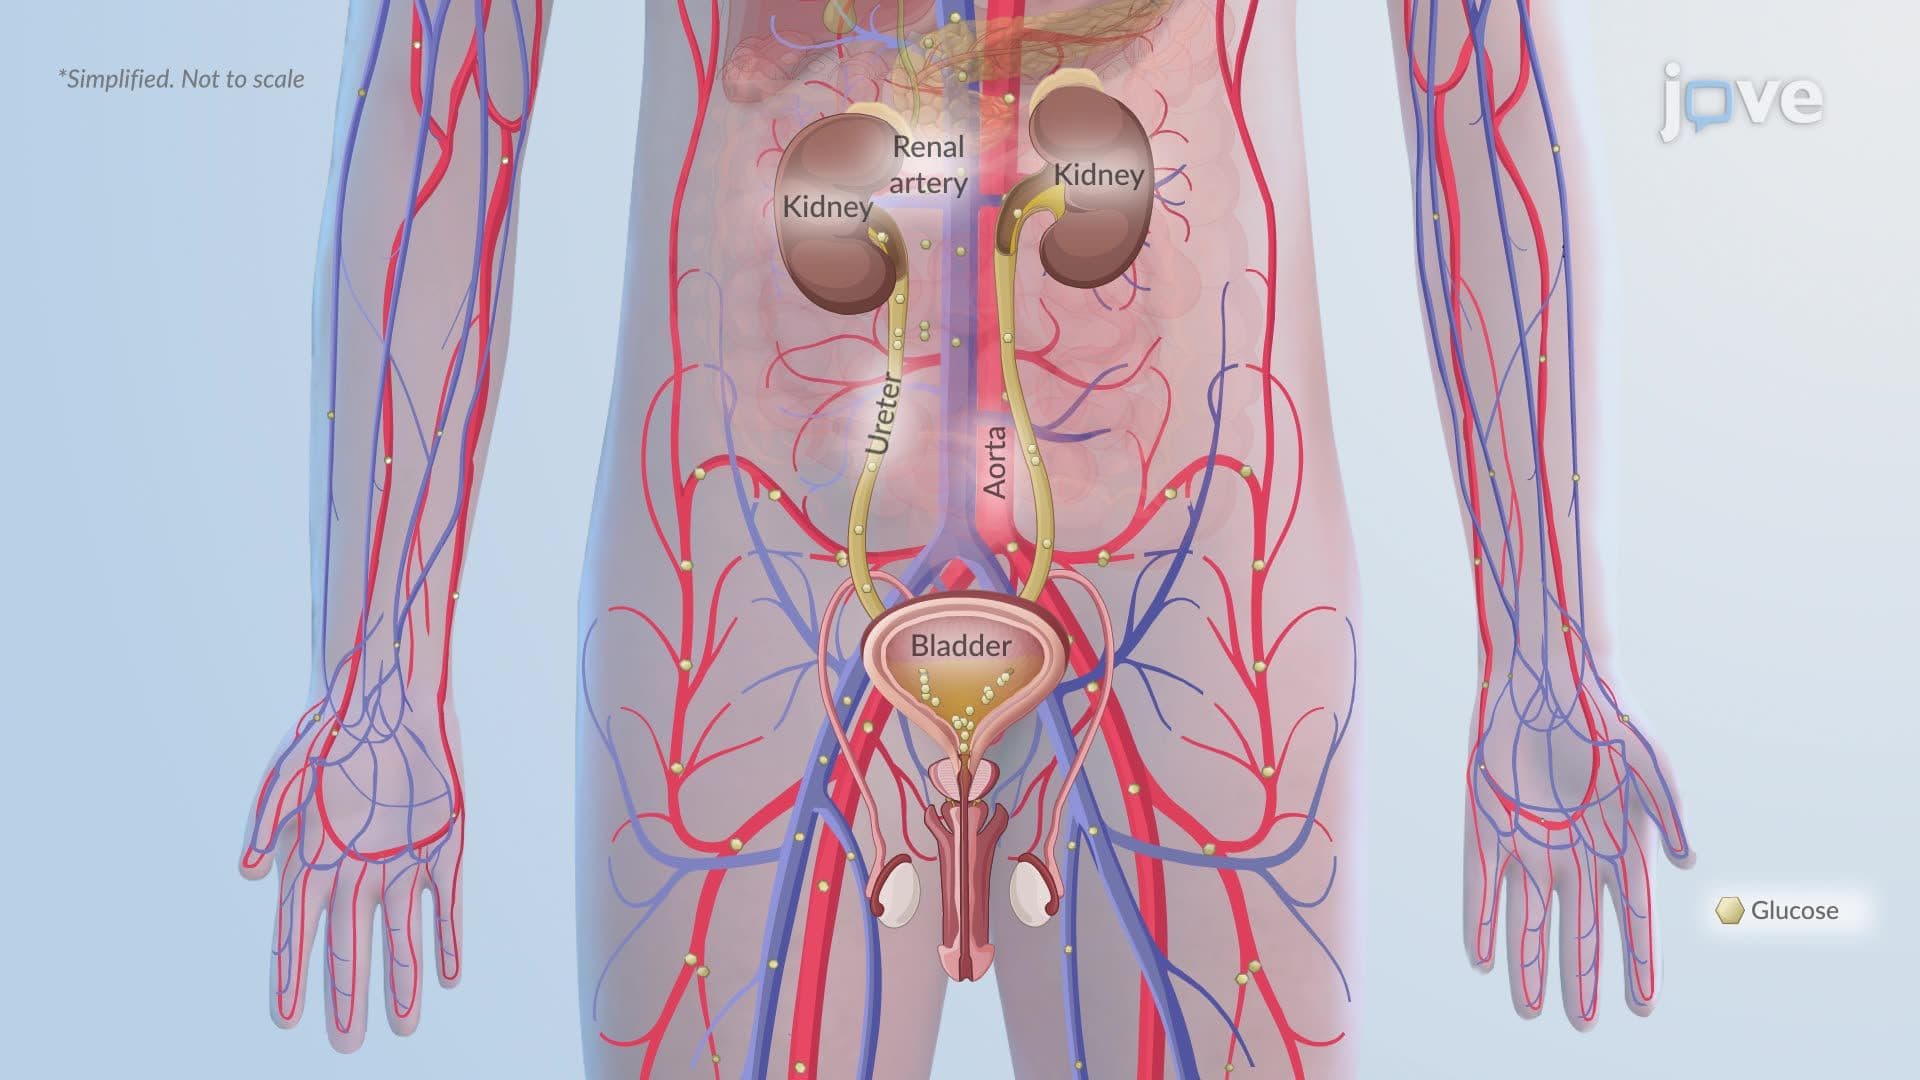
DefinitionDiabetic ketoacidosis (DKA) is an acute, life-threatening complication of diabetes mellitus, characterized by a triad of hyperglycemia (blood glucose >250 mg/dL), ketonemia or ketonuria, and metabolic acidosis (arterial pH <7.30 and serum bicarbonate <18 mEq/L). It results from insulin deficiency combined with elevated levels of counterregulatory hormones—glucagon, catecholamines, cortisol, and growth hormone—leading to increased lipolysis, hepatic ketone production, and...
Diabetic Ketoacidosis ll: Pathophysiology01:22

Diabetic Ketoacidosis ll: Pathophysiology

Diabetic ketoacidosis (DKA) is a metabolic emergency characterized by hyperglycemia, ketonemia, and metabolic acidosis. It results from severe insulin deficiency and an excess of counterregulatory hormones, leading to uncontrolled lipolysis, ketogenesis, and widespread electrolyte and fluid disturbances.Pathophysiology The central event in DKA is a profound loss of insulin action. Without insulin, glucose uptake in insulin-dependent tissues is impaired, while hepatic glucose production...