Related Experiment Video
Updated: Aug 9, 2026


Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
Correlations among amino acid sites in bHLH protein domains: an information theoretic analysis
W R Atchley1, K R Wollenberg, W M Fitch
1Department of Genetics, North Carolina State University, USA.

Abstract:
An information theoretic approach is used to examine the magnitude and origin of associations among amino acid sites in the basic helix-loop-helix (bHLH) family of transcription factors. Entropy and mutual information values are used to summarize the variability and covariability of amino acids comprising the bHLH domain for 242 sequences. When these quantitative measures are integrated with crystal structure data and summarized using helical wheels, they provide important insights into the evolution of three-dimensional structure in these proteins. We show that amino acid sites in the bHLH domain known to pack against each other have very low entropy values, indicating little residue diversity at these contact sites. Noncontact sites, on the other hand, exhibit significantly larger entropy values, as well as statistically significant levels of mutual information or association among sites. High levels of mutual information indicate significant amounts of intercorrelation among amino acid residues at these various sites. Using computer simulations based on a parametric bootstrap procedure, we are able to partition the observed covariation among various amino acid sites into that arising from phylogenetic (common ancestry) and stochastic causes and those resulting from structural and functional constraints. These results show that a significant amount of the observed covariation among amino acid sites is due to structural/functional constraints, over and above the covariation arising from phylogenetic constraints. These quantitative analyses provide a highly integrated evolutionary picture of the multidimensional dynamics of sequence diversity and protein structure.
More Related Videos
 06:50
Computational Prediction of Amino Acid Preferences of Potentially Multispecific Peptide-Binding Domains Involved in Protein-Protein Interactions
Published on: January 26, 2024
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to form...
 02:21
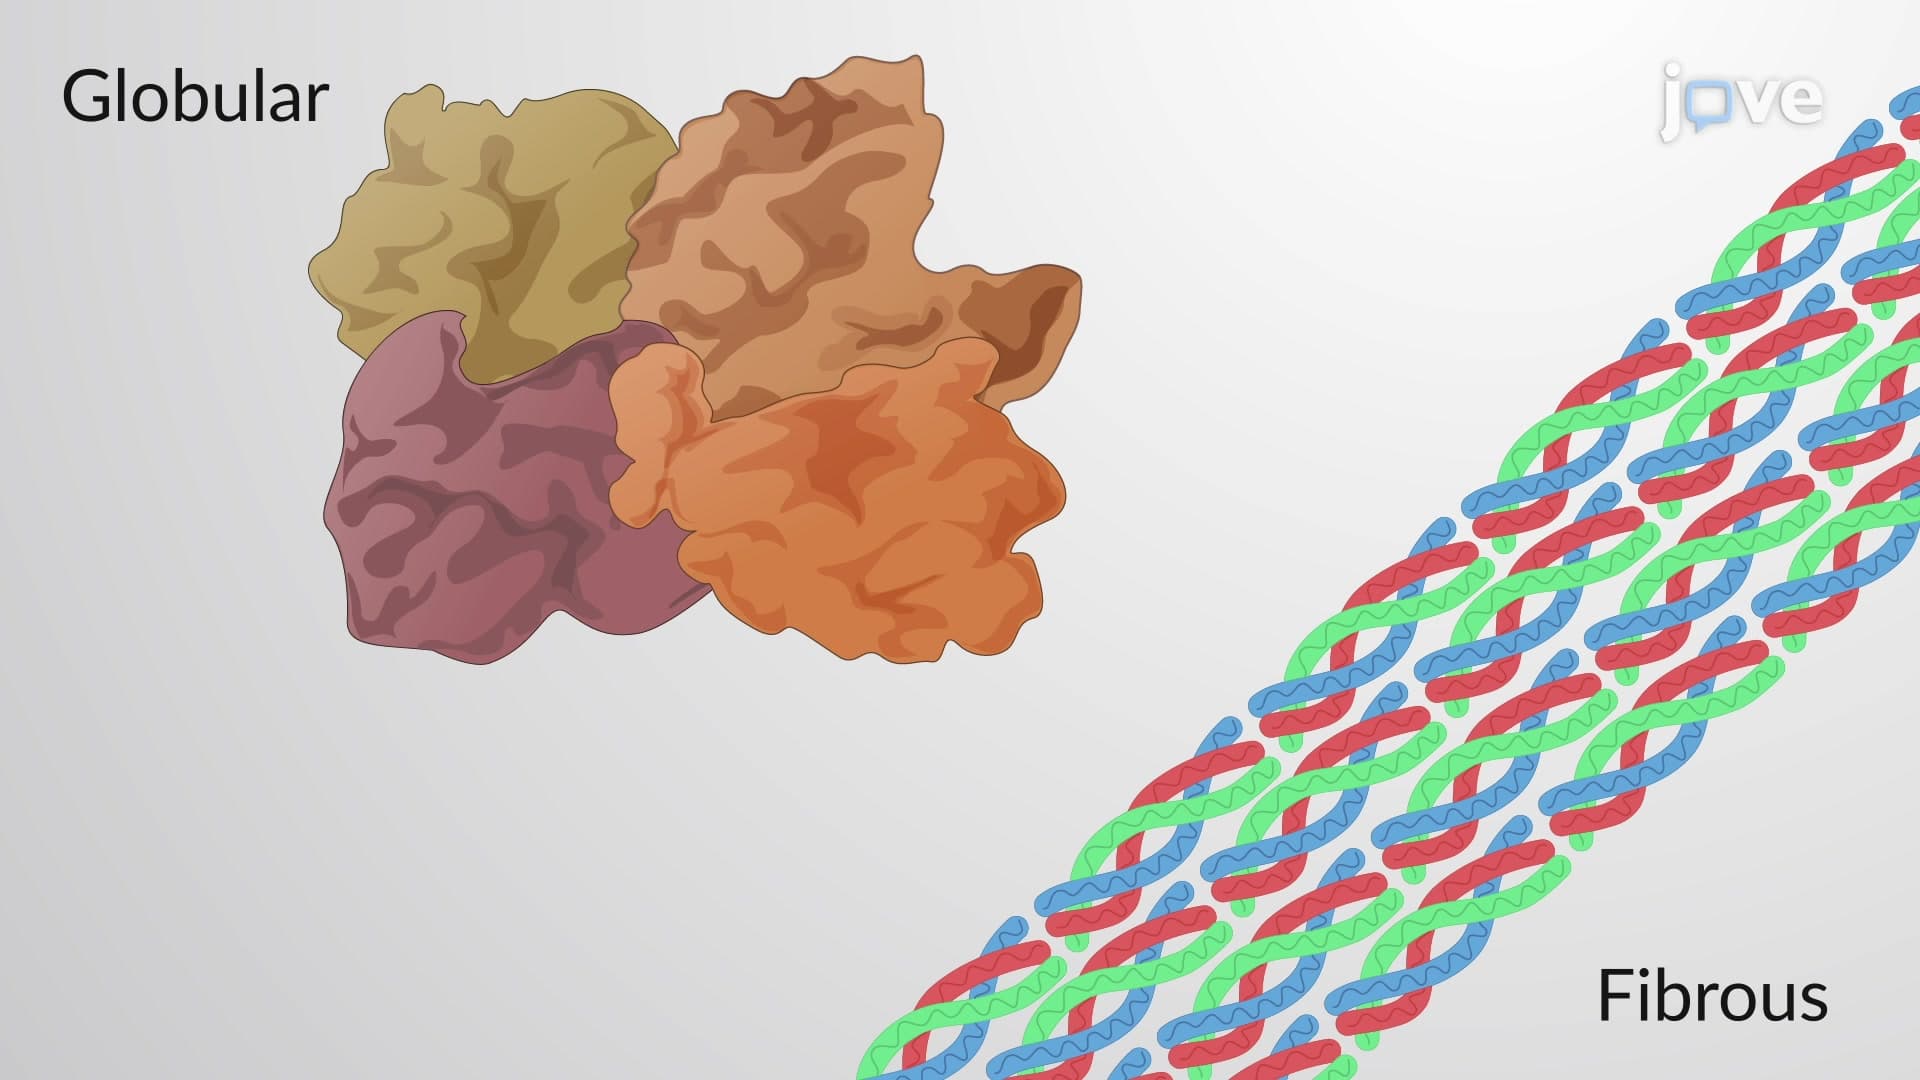
Globular and Fibrous Proteins
Globular proteins are also known as spheroproteins and typically are approximately round in shape. They contain a mix of amino acid types and contain differing sequences in their primary structures. Globular proteins have many different functions, such as enzymes, cellular messengers, and molecular transporters. These roles often require the proteins to be...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 00:49
Ligand Binding and Linkage
01:49Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...