Related Experiment Video
Updated: Oct 16, 2026
Human Liver Microphysiological System for Assessing Drug-Induced Liver Toxicity In Vitro
Published on: January 31, 2022
Nonspecific binding of drugs to human liver microsomes
J A McLure1, J O Miners, D J Birkett
1Department of Clinical Pharmacology, Flinders Medical Centre and Flinders University School of Medicine, Adelaide, Australia.

Aims:
To characterize the nonspecific binding to human liver microsomes of drugs with varying physicochemical characteristics, and to develop a model for the effect of nonspecific binding on the in vitro kinetics of drug metabolism enzymes.
Methods:
The extent of nonspecific binding to human liver microsomes of the acidic drugs caffeine, naproxen, tolbutamide and phenytoin, and of the basic drugs amiodarone, amitriptyline and nortriptyline was investigated. These drugs were chosen for study on the basis of their lipophilicity, charge, and extent of ionization at pH 7.4. The fraction of drug unbound in the microsomal mixture, fu(mic), was determined by equilibrium dialysis against 0.1 M phosphate buffer, pH 7.4. The data were fitted to a standard saturable binding model defined by the binding affinity KD, and the maximum binding capacity Bmax. The derived binding parameters, KD and Bmax, were used to simulate the effects of saturable nonspecific binding on in vitro enzyme kinetics.
Results:
The acidic drugs caffeine, tolbutamide and naproxen did not bind appreciably to the microsomal membrane. Phenytoin, a lipophilic weak acid which is mainly unionized at pH 7. 4, was bound to a small extent (fu(mic) = 0.88) and the binding did not depend on drug concentration over the range used. The three weak bases amiodarone, amitriptyline and nortriptyline all bound extensively to the microsomal membrane. The binding was saturable for nortriptyline and amitriptyline. Bmax and KD values for nortriptyline at 1 mg ml-1 microsomal protein were 382 +/- 54 microM and 147 +/- 44 microM, respectively, and for amitriptyline were 375 +/- 23 microM and 178 +/- 33 microM, respectively. Bmax, but not KD, varied approximately proportionately with the microsome concentration. When KD is much less than the Km for a reaction, the apparent Km based on total drug can be corrected by multiplying by fu(mic). When the substrate concentration used in a kinetic study is similar to or greater than the KD (Km >/= KD), simulations predict complex effects on the reaction kinetics. When expressed in terms of total drug concentrations, sigmoidal reaction velocity vs substrate concentration plots and curved Eadie Hofstee plots are predicted.
Conclusions:
Nonspecific drug binding in microsomal incubation mixtures can be qualitatively predicted from the physicochemical characteristics of the drug substrate. The binding of lipophilic weak bases is saturable and can be described by a standard binding model. If the substrate concentrations used for in vitro kinetic studies are in the saturable binding range, complex effects are predicted on the reaction kinetics when expressed in terms of total (added) drug concentration. Sigmoidal reaction curves result which are similar to the Hill plots seen with cooperative substrate binding.
Related Concept Videos
 01:30
Drug Binding to Blood Components
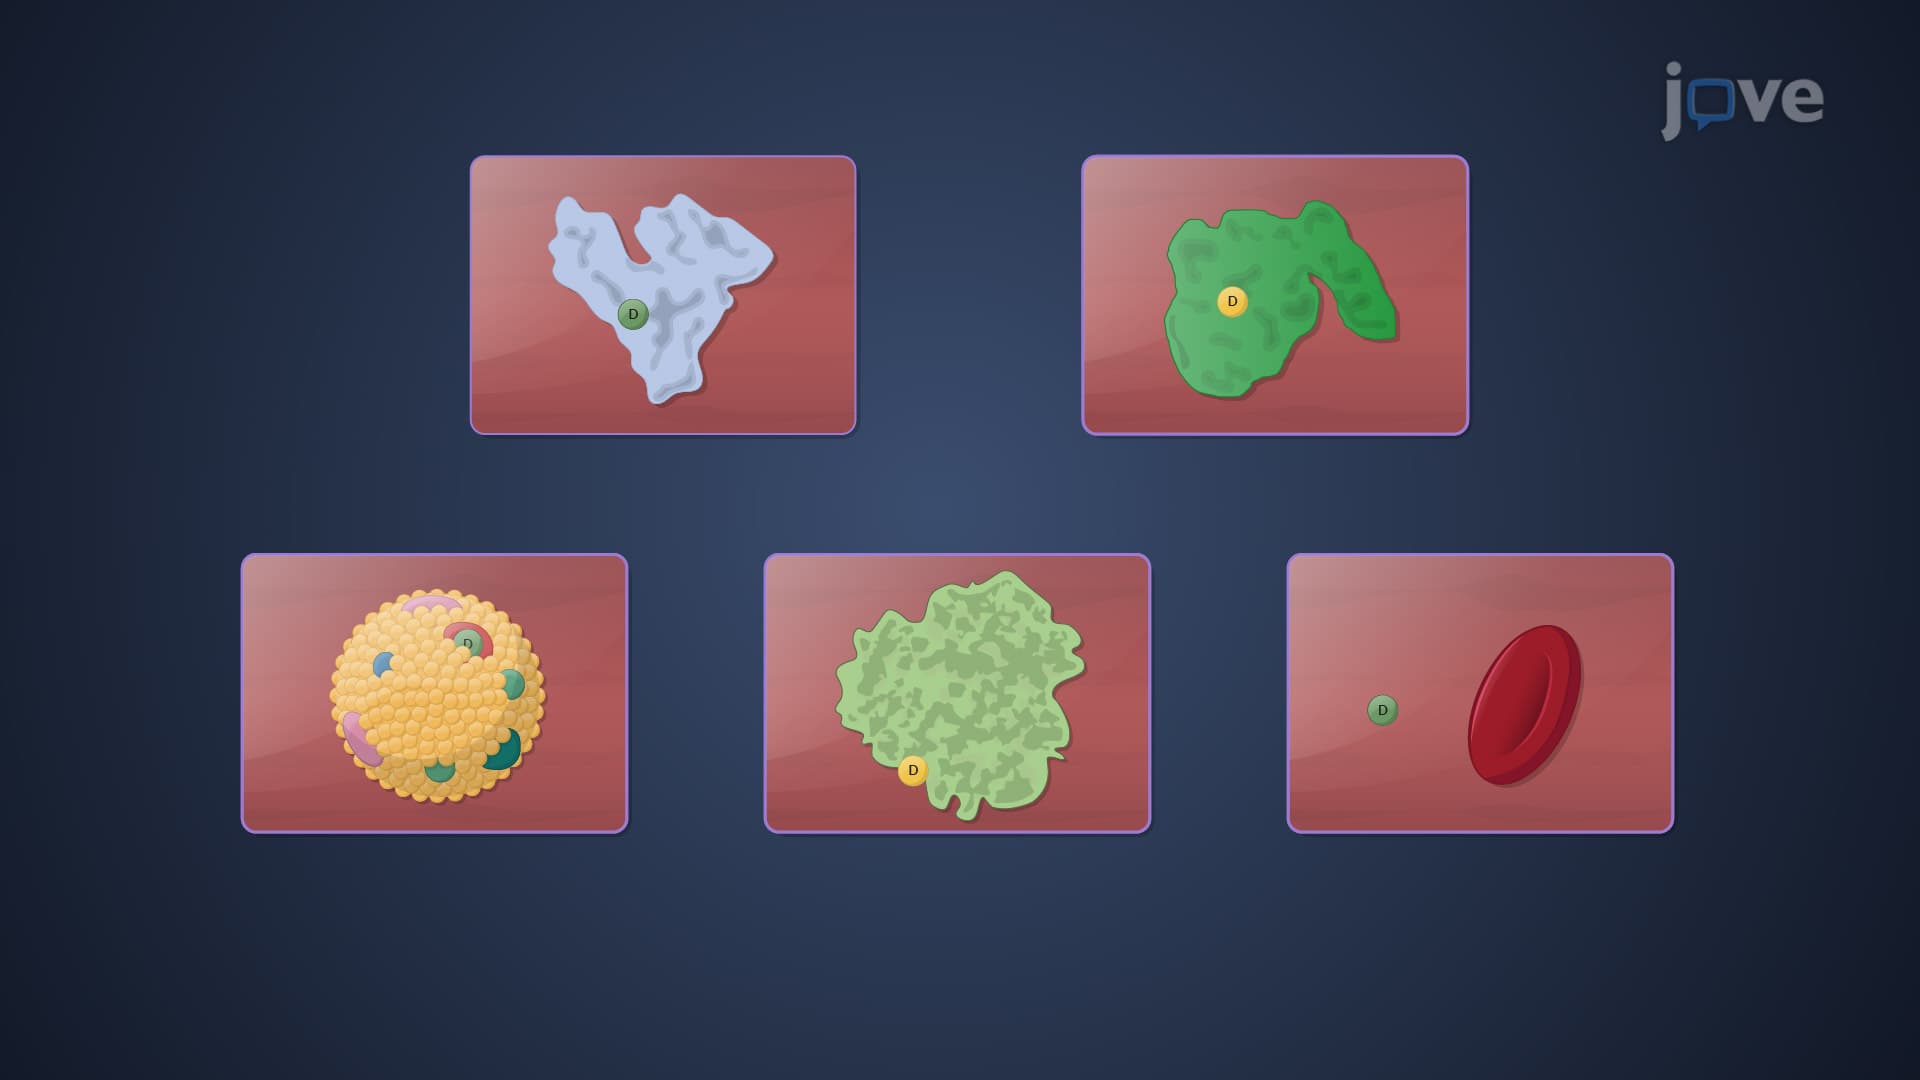
HSA is the most abundant plasma protein and is vital in drug binding. It contains distinct drug-binding sites, with different drugs exhibiting affinity for specific sites. There are three main drug-binding domains for HSA: sites I, II, and III. These domains are further...
 01:29
Drug Distribution: Plasma Protein Binding
 01:09
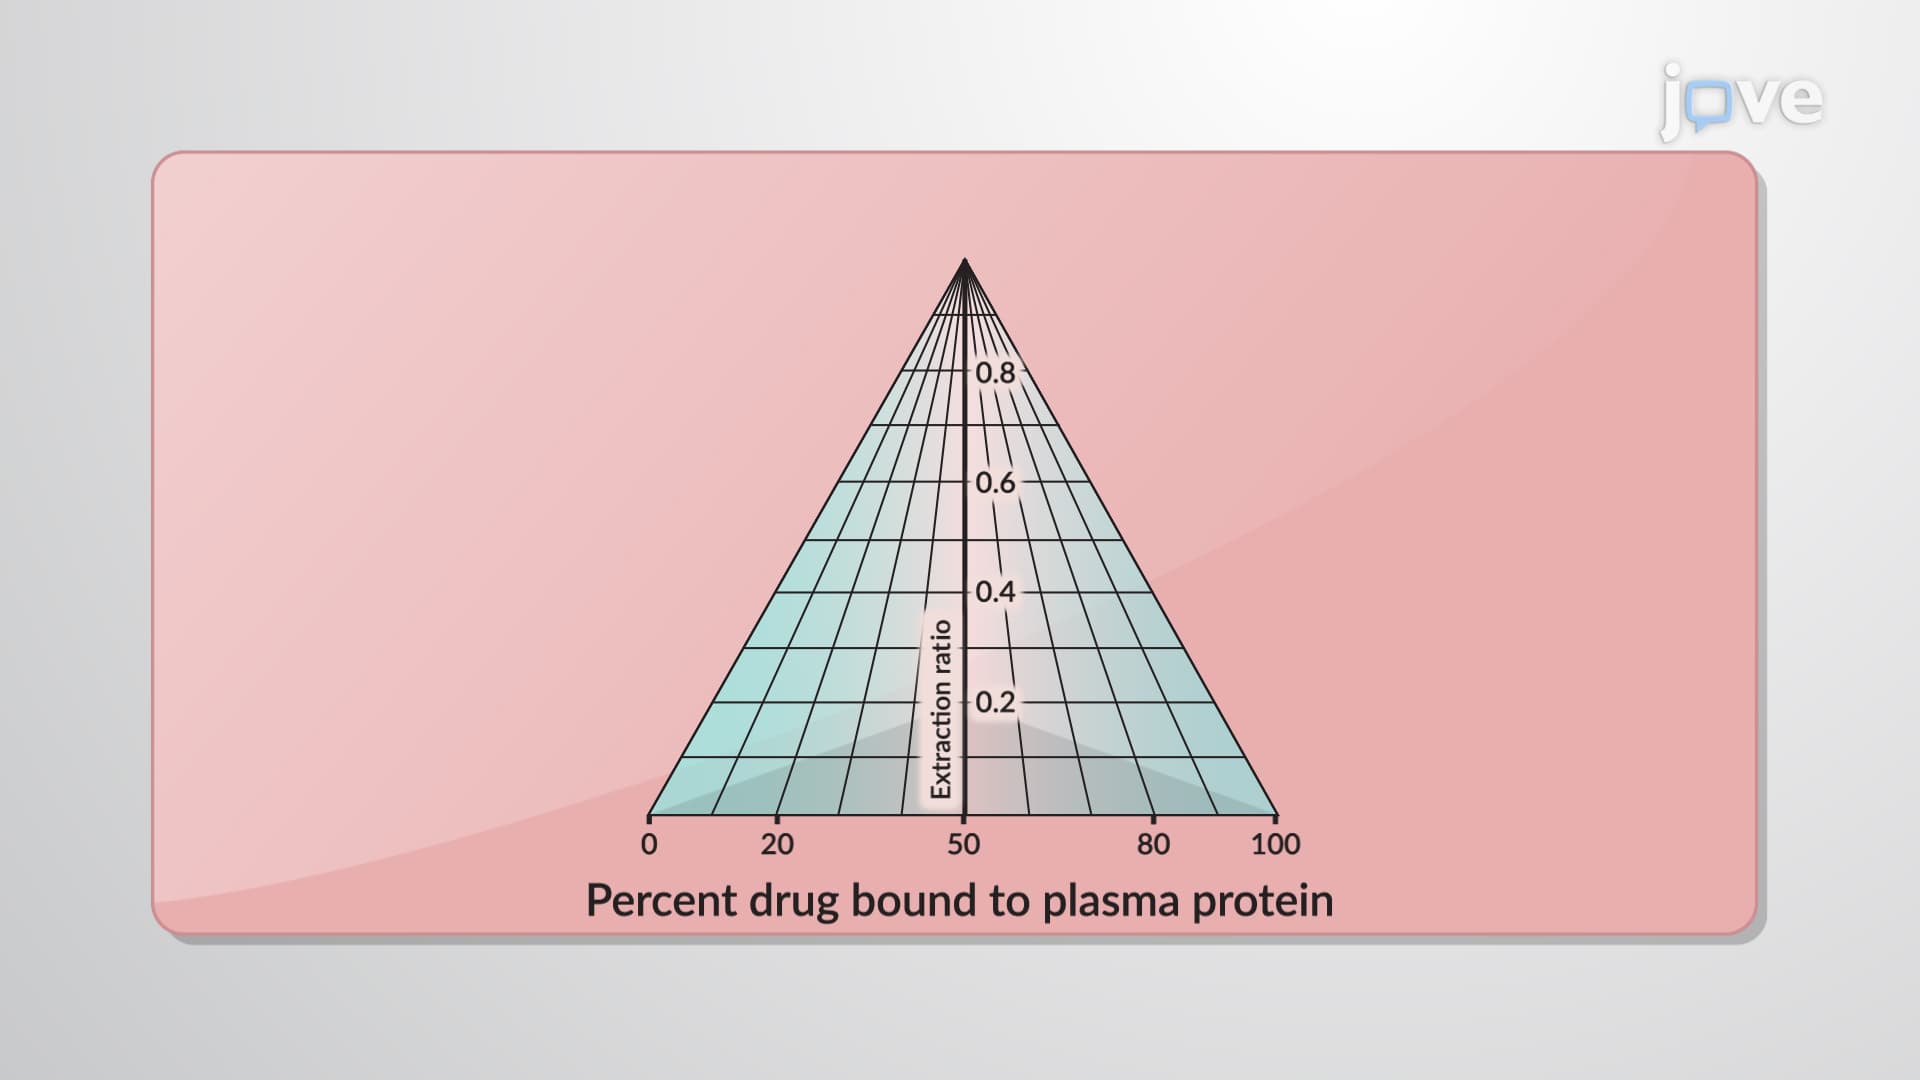
Hepatic Drug Clearance: Effect of Protein Binding
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
 01:21
Drug Distribution: Tissue Binding
For...
 01:24
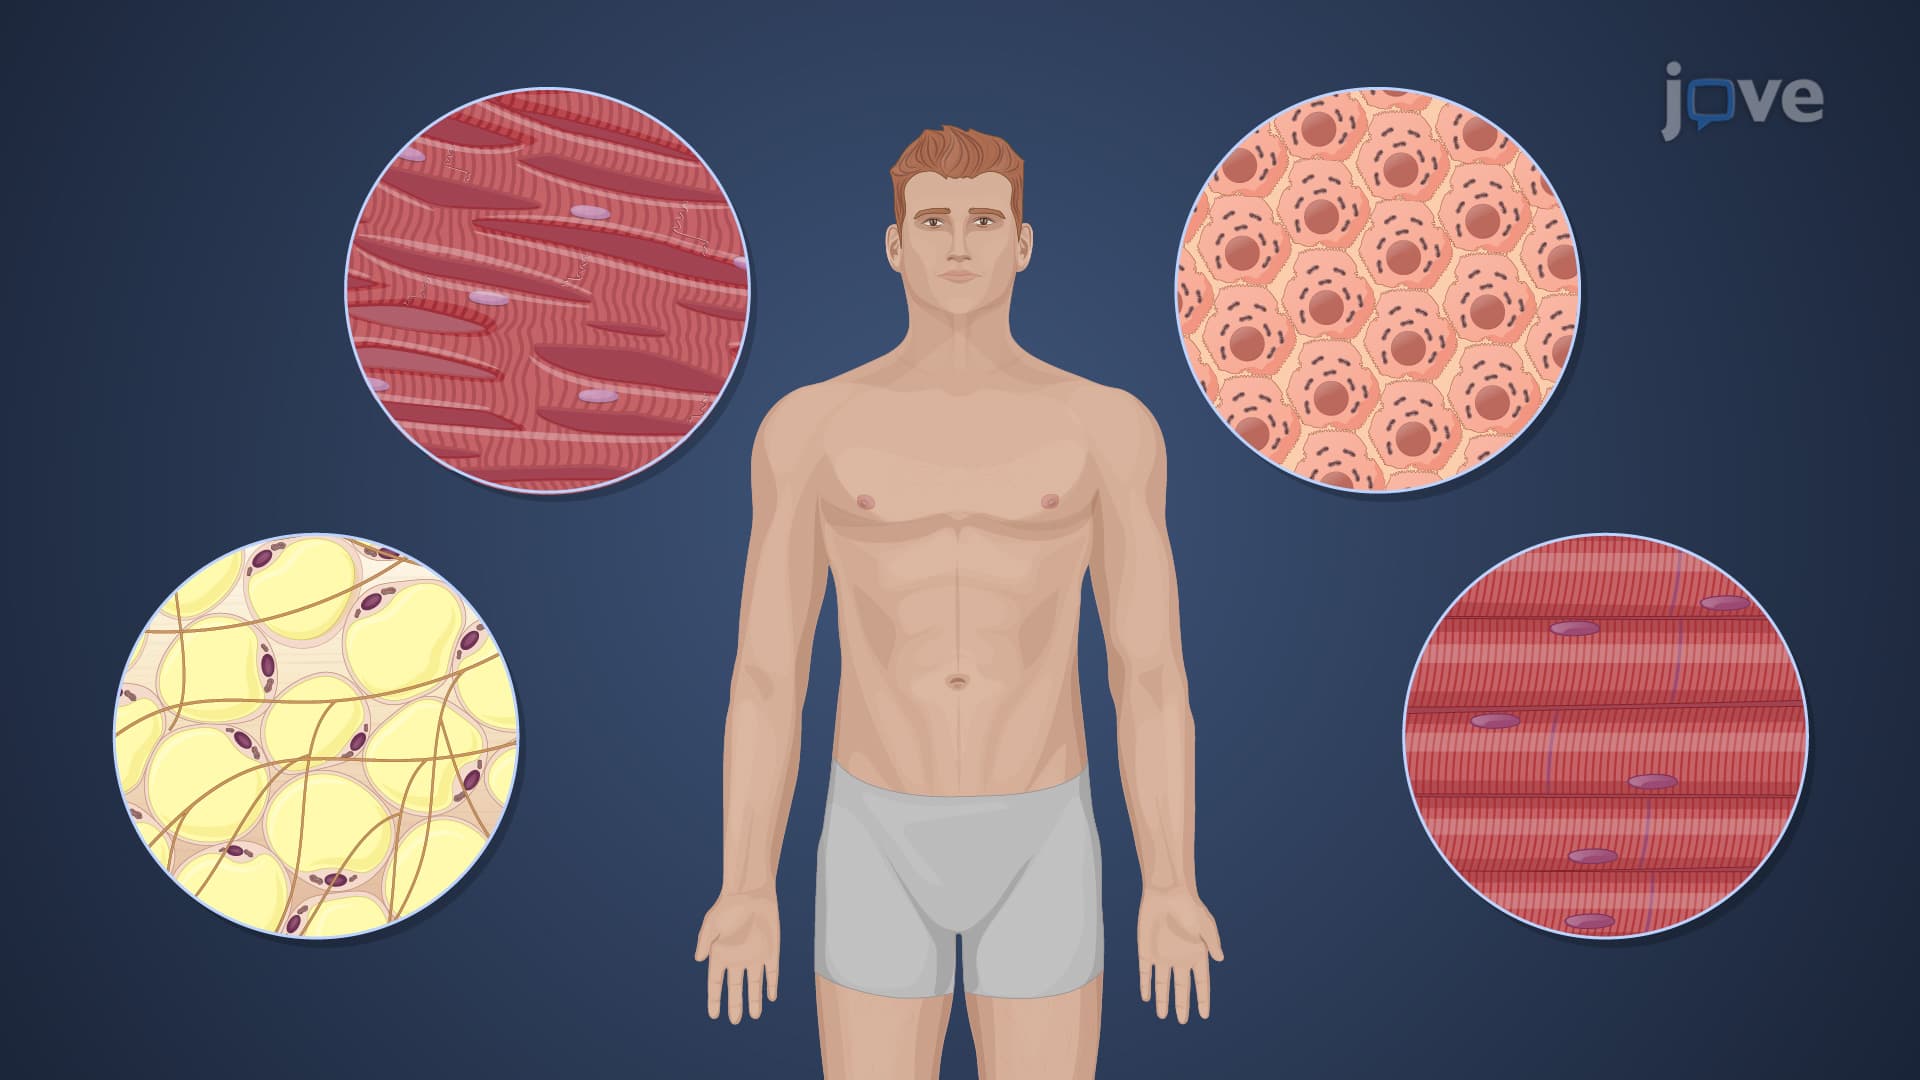
Tissue-Drug Binding: Localization of Drugs and its Significance
Drugs can bind to different tissue components, enhancing their distribution and localization. The factors influencing drug localization in tissues include the drug's lipophilicity, structural characteristics, tissue perfusion rate, and pH differences. These factors determine the...
 01:20
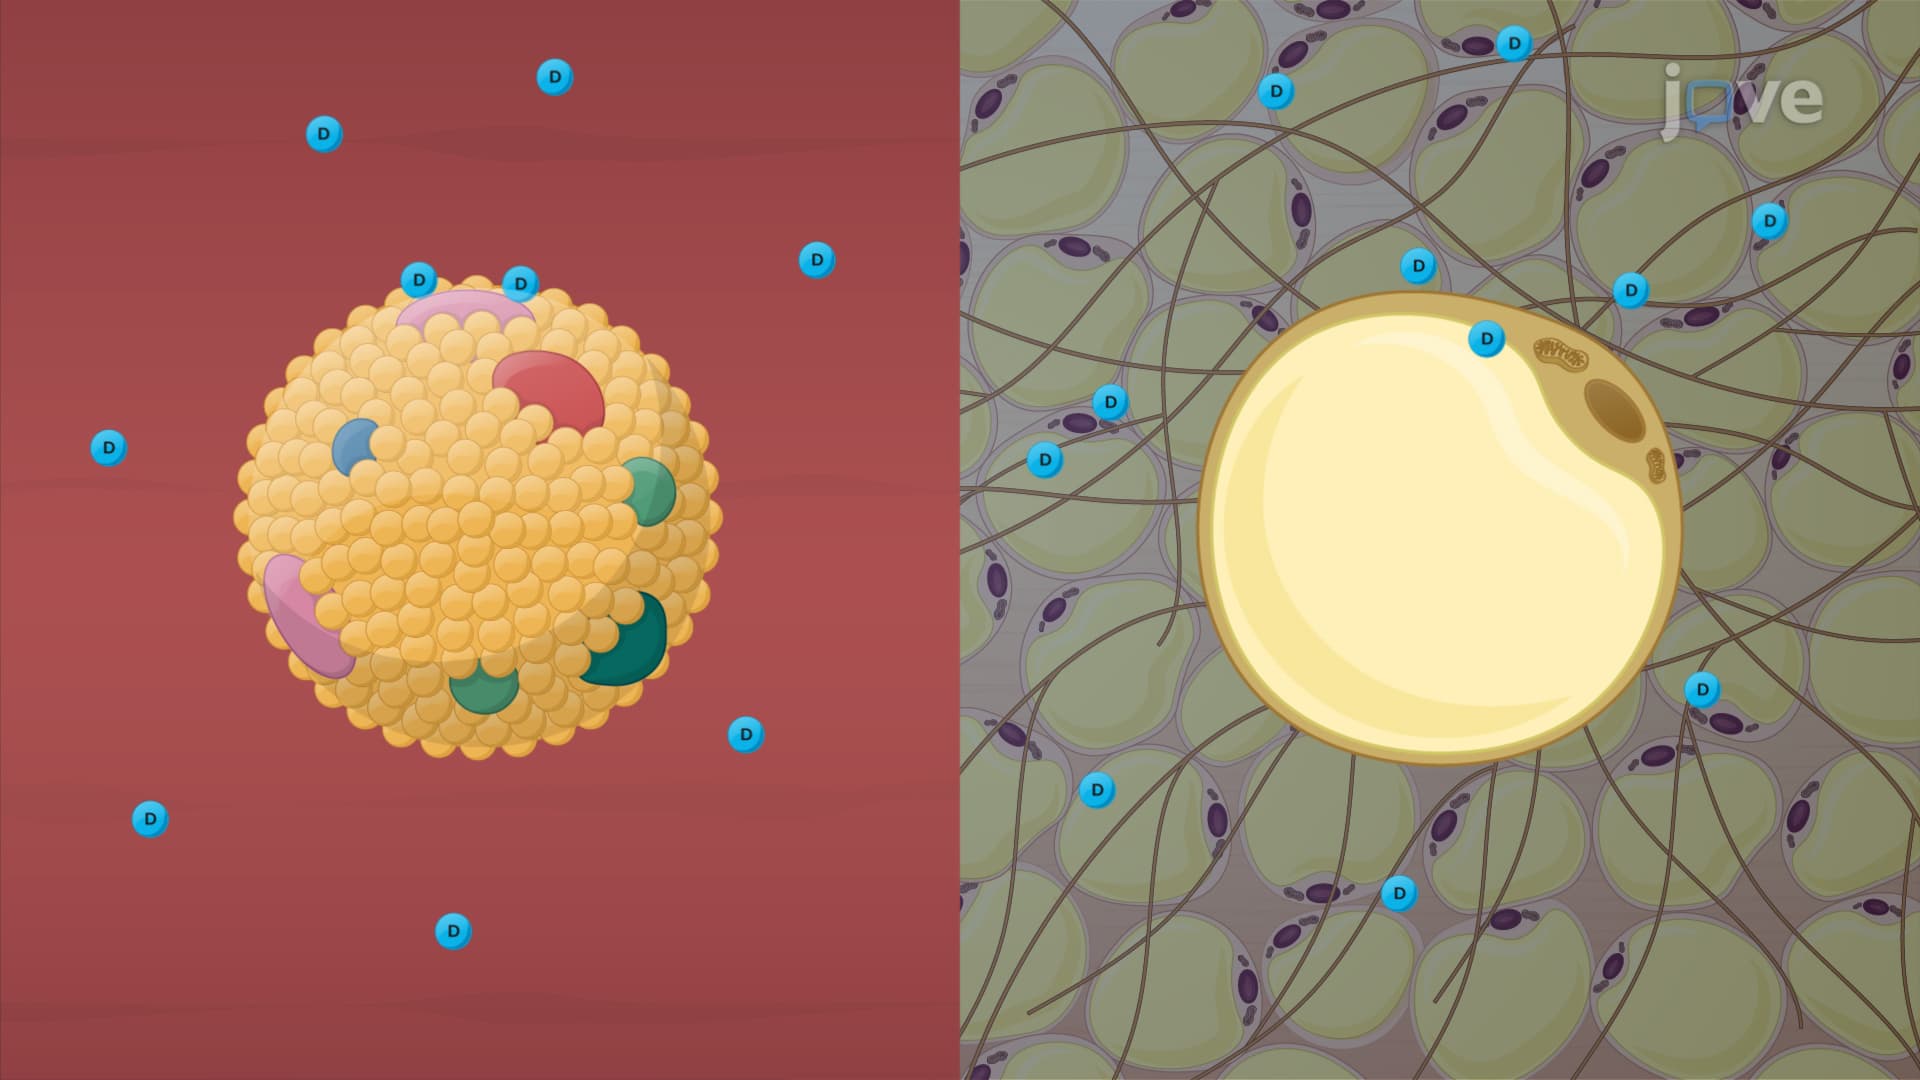
Factors Affecting Protein-Drug Binding: Protein-Related Factors
The physicochemical properties of a drug play a significant role in its ability to bind to proteins. Lipophilic drugs, which dissolve in fats, oils, and lipids, can be bound by...