Related Experiment Video
Updated: Oct 13, 2026

Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
A refined 3-dimensional QSAR of cytochrome P450 2C9: computational predictions of drug interactions

Abstract:
A ligand-based model is reported that predicts the Ki values for cytochrome P450 2C9 (CYP2C9) inhibitors. This CoMFA model was used to predict the affinity of 14 structurally diverse compounds not in the training set and appears to be robust. The mean error of the predictions is 6 microM. The experimentally measured Ki values of the 14 compounds range from 0.1 to 48 microM. Leave-one-out cross-validated partial least-squares gives a q2 value of between 0.6 and 0.8 for the various models which indicates internal consistency. Random assignment of biological data to structure leads to negative q2 values. These models are useful in that they establish a pharmacophore for binding to CYP2C9 that can be tested with site-directed mutagenesis. These models can also be used to screen for potential drug interactions and to design compounds that will not bind to this enzyme with high affinity.
Related Concept Videos
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:30
Drug toxicity: Drug–Drug Interaction
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:16
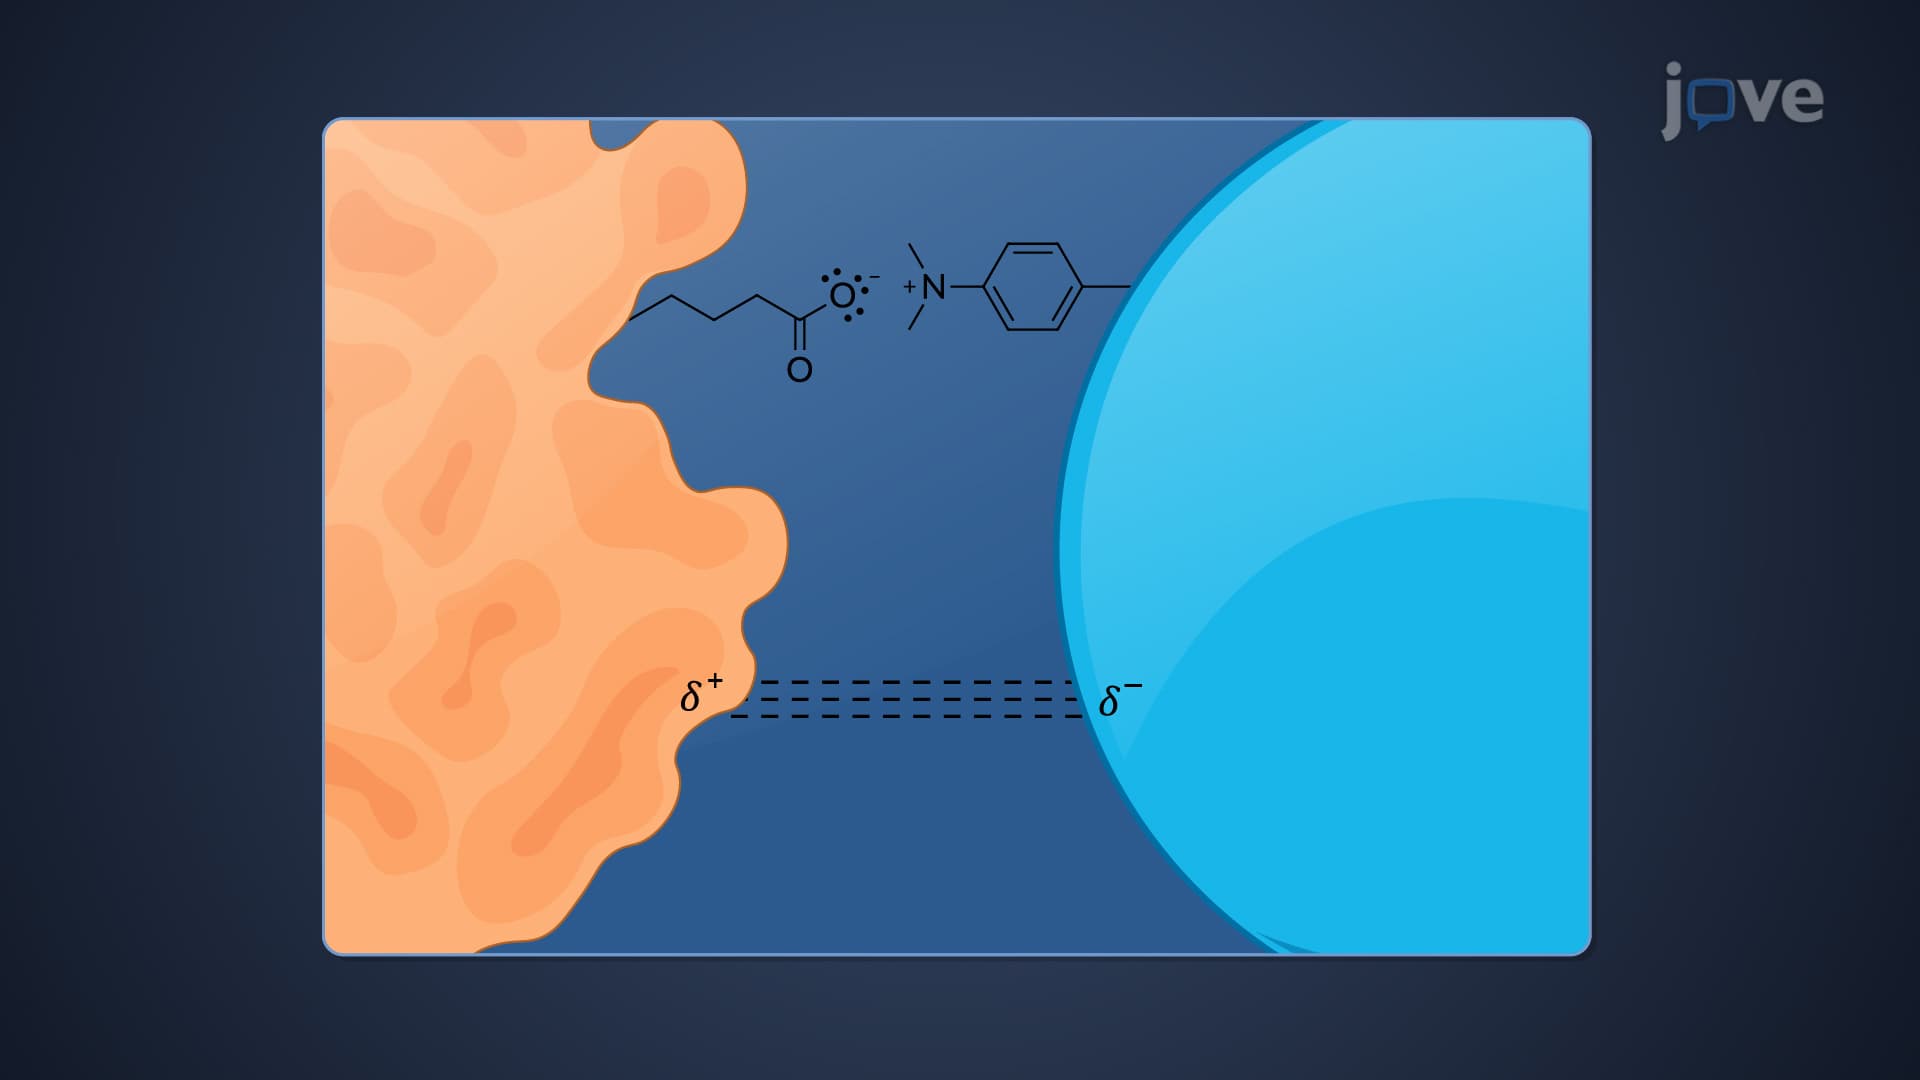
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:26
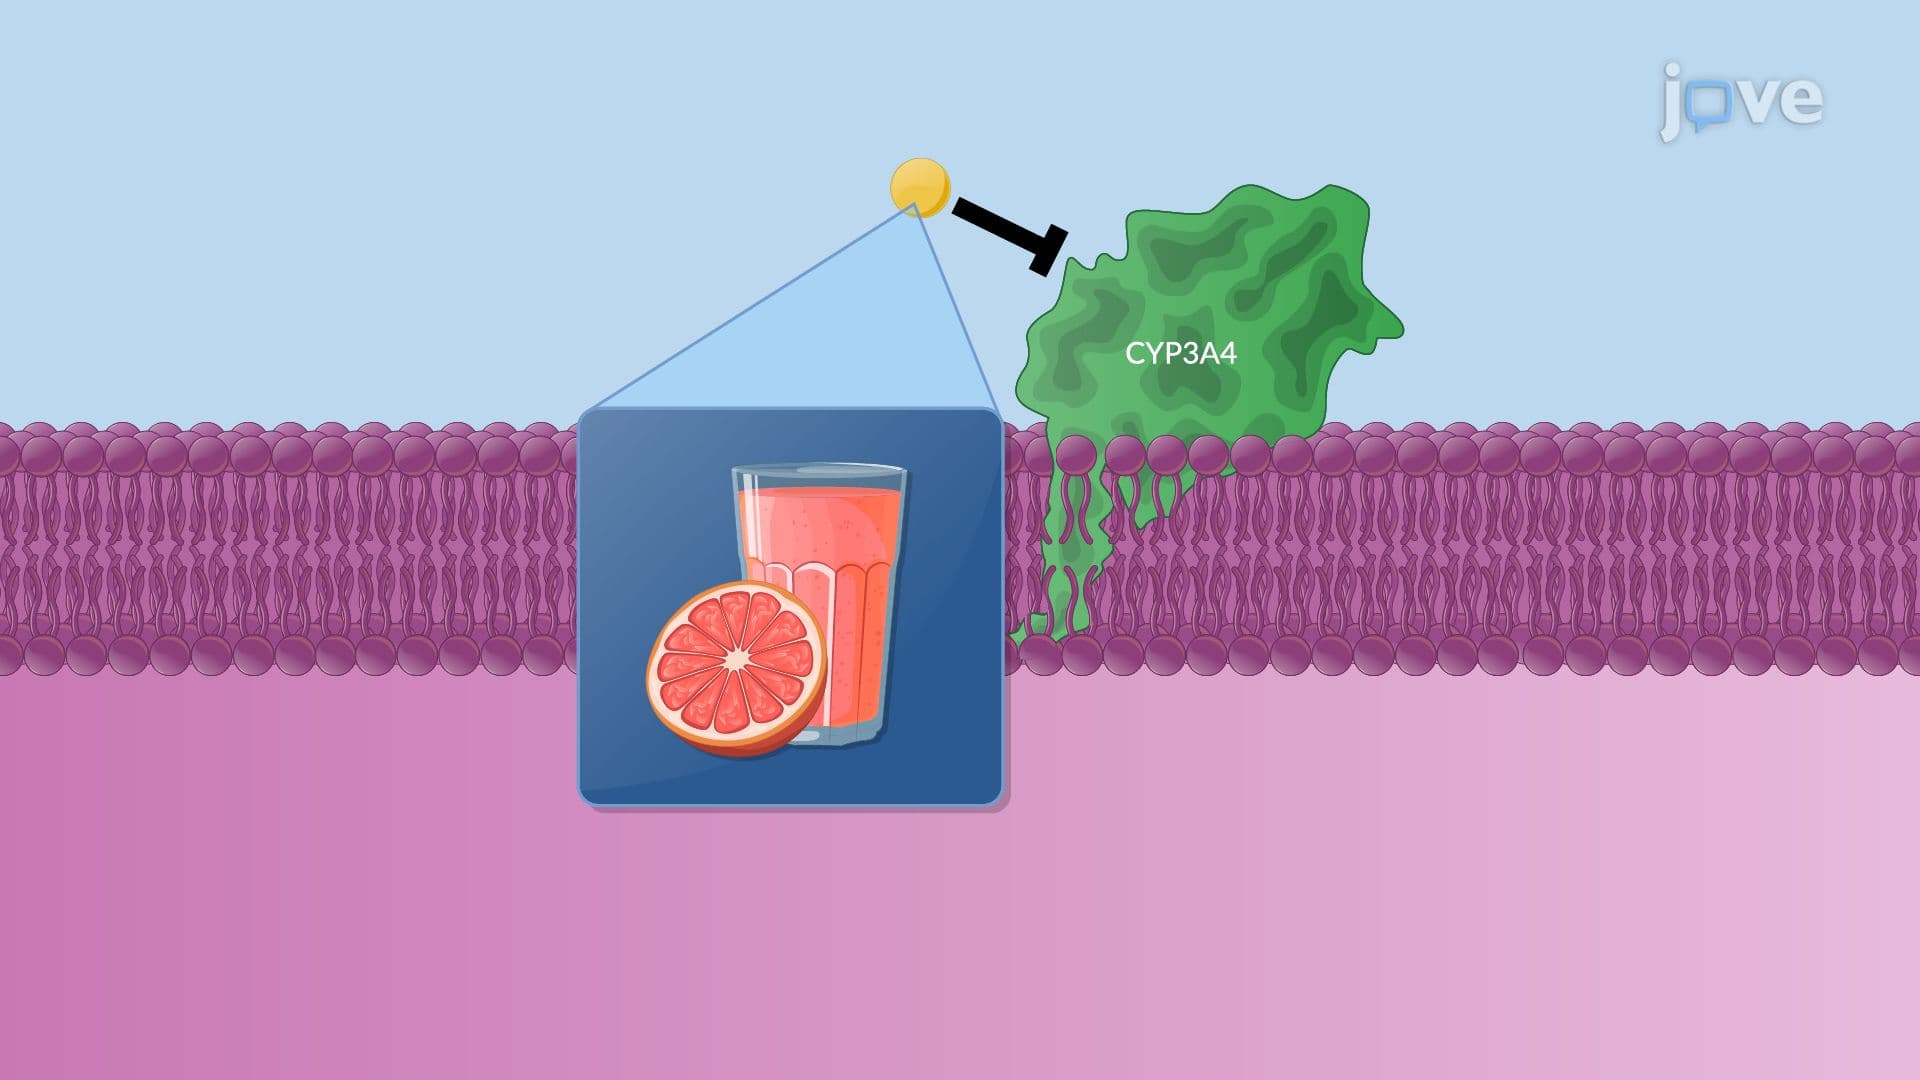
Pharmacokinetics: Drug–Food and Drug–Viral Interactions