Related Experiment Video
Updated: Aug 8, 2026
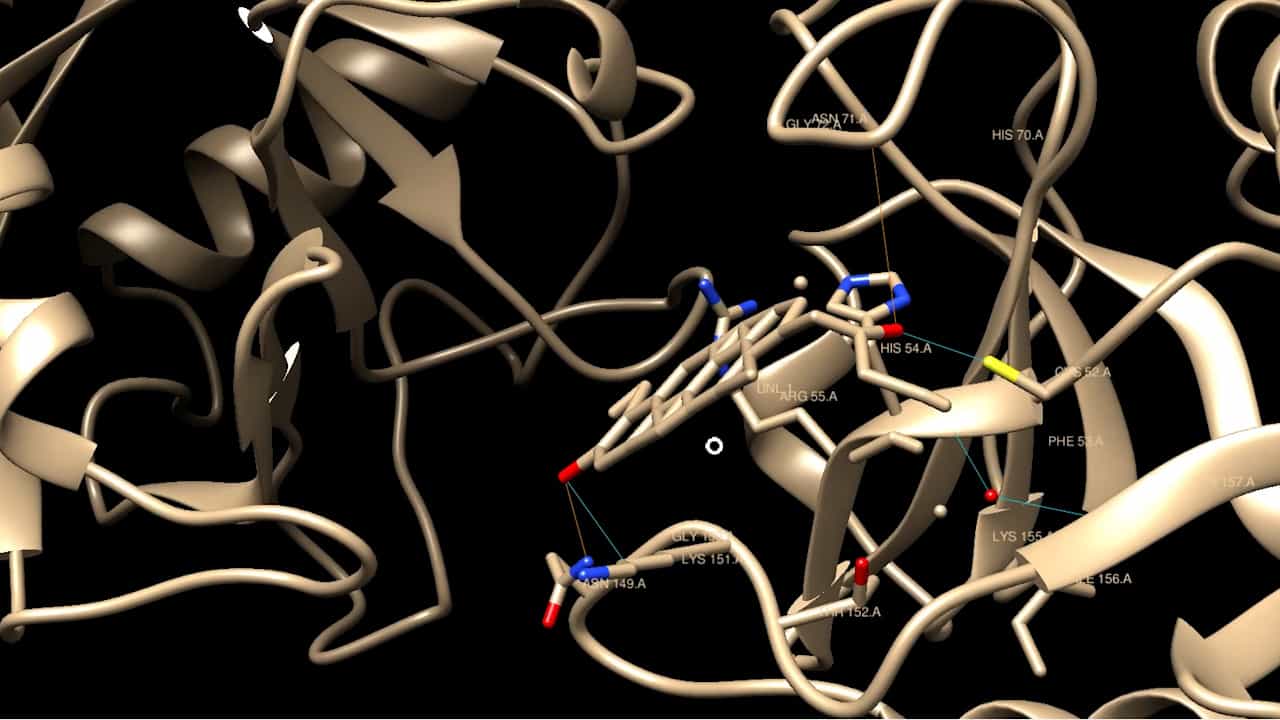

Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Can an optimization/scoring procedure in ligand-protein docking be employed to probe drug-resistant mutations in
1Department of Computational Science, National University of Singapore, Lower Kent Ridge Road, Singapore 119260. yzchen@cz3.nus.edu.sg

Abstract:
A simple ligand-protein structural optimization and binding evaluation procedure has been routinely used in high-speed ligand-protein docking studies. In this work, we examine whether such an optimization/scoring procedure is useful in indicating possible drug-resistant mutations in proteins. Crystal structures of three wild-type enzymes (HIV-1 protease, HIV-1 reverse transcriptase, and Mycobacterium tuberculosis H37Rv enoyl-ACP reductase) complexed to a variety of inhibitors are studied. Mutations are introduced into these structures by using the molecular modeling software, SYBYL. Structural optimization and scoring of a mutant complex is conducted by a procedure similar to that used in a recent docking study (Wang et al., 1999). The computed results are compared with observed drug resistance data and the profile of nonresistant mutations. Most mutations studied show an energy change in the same direction as those indicated by observed resistance data. 50% of the polar to polar or nonpolar to nonpolar mutations are found to correlate qualitatively with observed drug resistance data. Van der Waals interactions account for most of these changes, which is in agreement with conclusions from structural studies. Substantially larger deviations are found between computed results and observed data for most polar to nonpolar or nonpolar to polar mutations, which result from deficiency in modelling and scoring ligand-protein interactions in our procedure. Our results suggest that an optimization/docking scoring procedure is useful for qualitatively probing polar to polar or nonpolar to nonpolar resistant mutations in addition to its application in screening active compounds. More accurate description of ligand-protein interactions and the use of methods such as free energy perturbation and Poisson-Boltzmann may be needed to further improve the quality of prediction.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:18
The Equilibrium Binding Constant and Binding Strength
 00:49
Ligand Binding and Linkage
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...