Related Experiment Video
Updated: Oct 3, 2026

A Rapid and Quantitative Fluorimetric Method for Protein-Targeting Small Molecule Drug Screening
Published on: October 16, 2015
Cheminformatic models to predict binding affinities to human serum albumin
G Colmenarejo1, A Alvarez-Pedraglio, J L Lavandera
1Structural Chemistry Department, GlaxoSmithKline, Parque Tecnológico de Madrid, E-28760 Tres Cantos, Madrid, Spain.

Abstract:
Models to predict binding affinities to human serum albumin (HSA) should be very useful in the pharmaceutical industry to speed up the design of new compounds, especially as far as pharmacokinetics is concerned. We have experimentally determined through high-performance affinity chromatography the binding affinities to HSA of 95 diverse drugs and druglike compounds. These data have allowed us the derivation of quantitative structure-activity relationship models to predict binding affinities to HSA of new compounds on the basis of their structure. Simple linear, one-variable models have been derived for specific families of compounds (r(2) > or = 0.80; q(2) > or = 0.62): beta-adrenergic antagonists, steroids, COX inhibitors, and tricyclic antidepressants. Also, global models have been derived to be applicable to the whole medicinal chemical space by using the full database of HSA binding constants described above. For this aim, a genetic algorithm has been used to exhaustively search and select for multivariate and nonlinear equations, starting from a large pool of molecular descriptors. The resulting models display good fits to the experimental data (r(2) > or = 0.78; LOF < or = 0.12). In addition, both internal (cross validation and randomization) and external validation tests have demonstrated that these models have good predictive power (q(2) > or = 0.73; PRESS/SSY < or = 0.23; r(2) > or = 0.82 for the external set). Statistical analysis of the equation populations indicates that hydrophobicity (as measured by the ClogP) is the most important variable determining the binding extent to HSA. In addition, structural factors (especially the topological (6)chi(ring) index and some Jurs descriptors) also frequently appear as descriptors in the best equations. Therefore, binding to HSA turns out to be determined by a combination of hydrophobic forces together with some modulating shape factors. This agrees with X-ray structures of HSA alone or bound to ligands, where the binding pockets of both sites I and II are composed mainly of hydrophobic residues.
Related Concept Videos
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 02:18
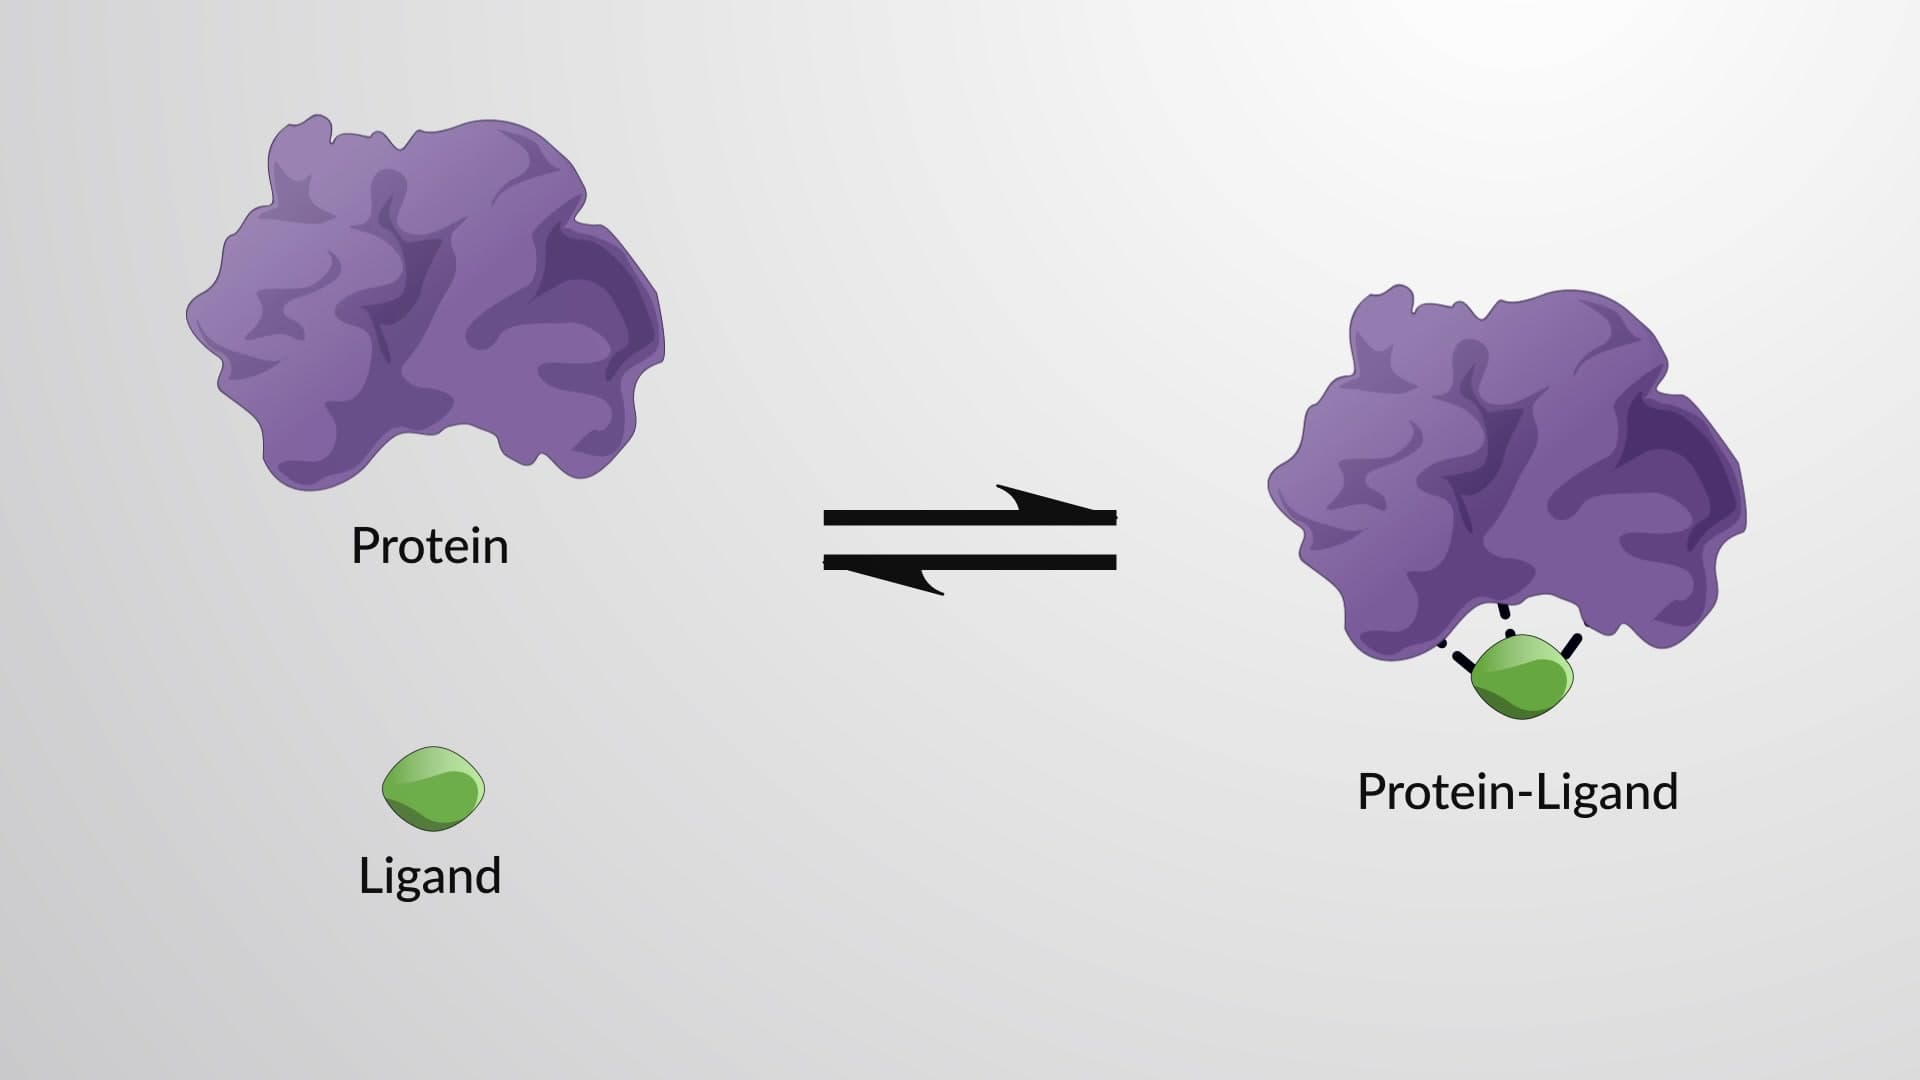
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...