Related Experiment Video
Updated: Oct 1, 2026
A Scalable, Cell-Based Method for the Functional Assessment of Ube3a Variants
Published on: October 10, 2022
No evidence for submicroscopic 22qter deletions in patients with features suggestive for Angelman syndrome
Bert B A de Vries1, Jess Tyson, Robin M Winter
1Clinical and Molecular Genetics Unit, Institute of Child Health and Great Ormond Street Hospital, London, United Kingdom, UK.

Abstract:
Patients with monosomy 22q13.3 --> qter have, in addition to (usually severe) developmental delay, hypotonia, severe expressive language delay leading to absence of speech, pervasive developmental abnormalities, and subtle facial anomalies. Thus far, it has been one of the more common submicroscopic telomere deletions seen in patients with mental retardation. Due to the phenotypic overlap between monosomy 22q13.3 and Angelman syndrome (AS), 44 patients with AS features but without one of the characteristic molecular 15q abnormalities were tested for 22qter deletions. In the study group, 31/44 (70%) were heterozygous for locus D22S163 with probe cMS607 (distance 0.125 Mb from telomere). The remaining 13/44 (30%) patients were heterozygous for one or more of four microsatellite markers centromeric from D22S163 in the 22qter region (distances 1.5-4.3 Mb from telomere). Based on the present study, there is no evidence that patients with an "Angelman-like" phenotype are more likely to have a 22qter deletion than other individuals with mental retardation.
More Related Videos

Related Concept Videos
 01:17
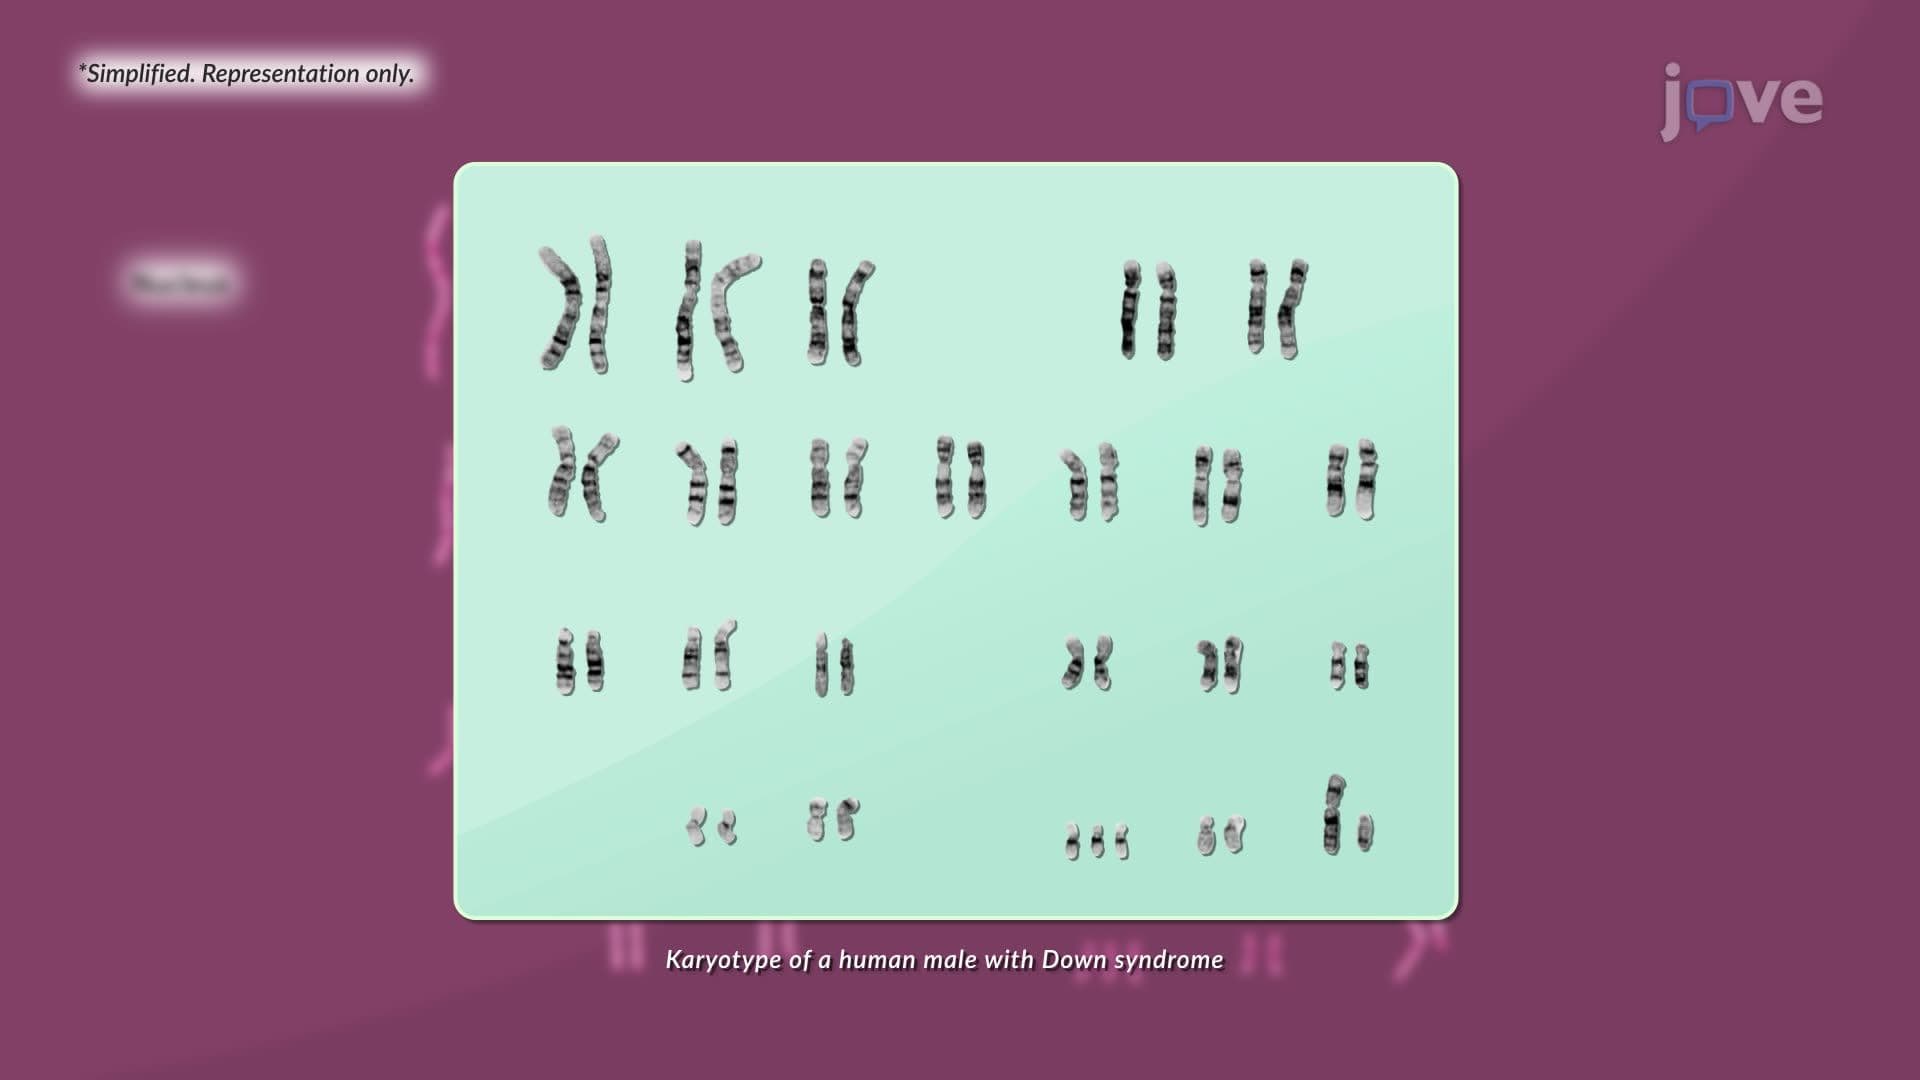
Karyotyping
 02:30
Genomic Imprinting and Inheritance
The expression of some genes depends on which parent passed the gene to the offspring, through a phenomenon known as...
 03:20
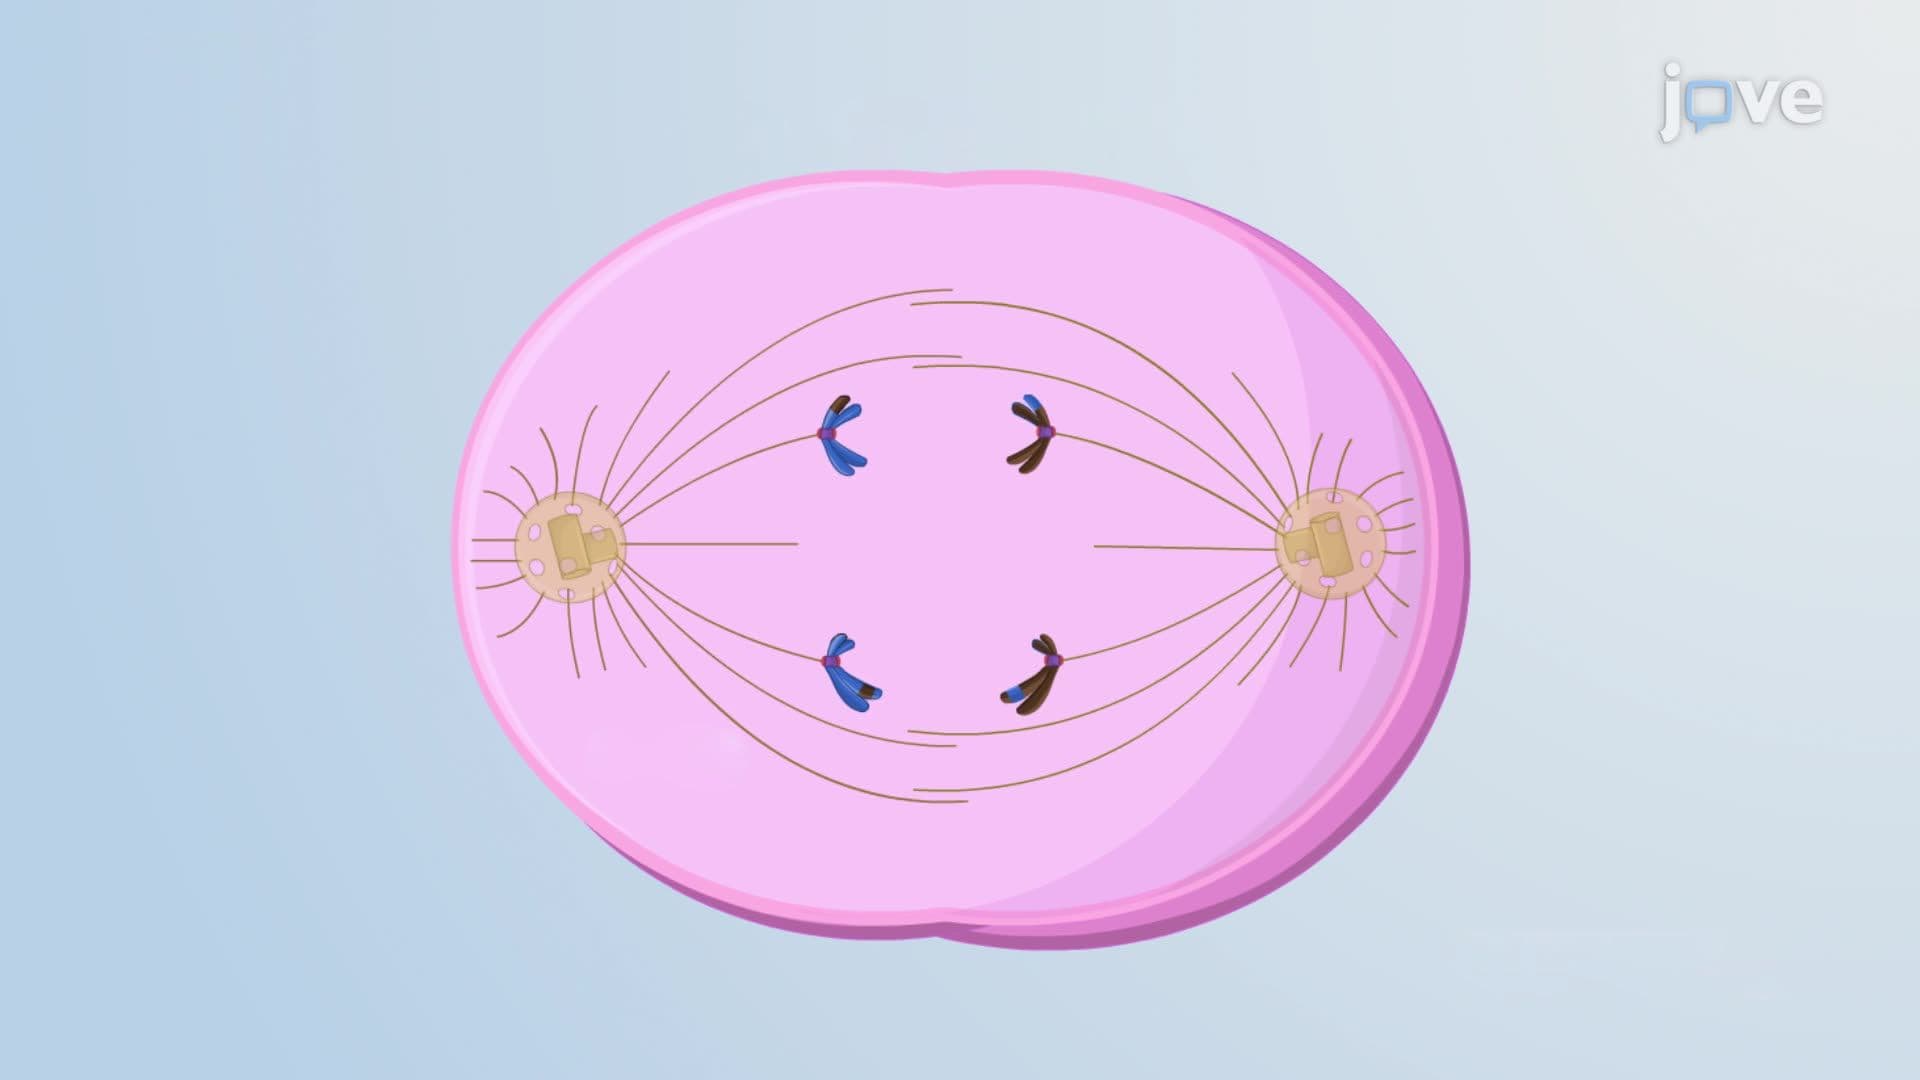
Meiosis I