
The nature of intermolecular interactions between aromatic amino acid residues
Francesco Luigi Gervasio1, Riccardo Chelli, Piero Procacci
1Dipartimento di Chimica, Università di Firenze, Firenze, Italy. procacci@chim.unifi.it

Abstract:
The nature of intermolecular interactions between aromatic amino acid residues has been investigated by a combination of molecular dynamics and ab initio methods. The potential energy surface of various interacting pairs, including tryptophan, phenilalanine, and tyrosine, was scanned for determining all the relevant local minima by a combined molecular dynamics and conjugate gradient methodology with the AMBER force field. For each of these minima, single-point correlated ab initio calculations of the binding energy were performed. The agreement between empirical force field and ab initio binding energies of the minimum energy structures is excellent. Aromatic-aromatic interactions can be rationalized on the basis of electrostatic and van der Waals interactions, whereas charge transfer or polarization phenomena are small for all intermolecular complexes and, particularly, for stacked structures. Proteins 2002;48:117-125.
Related Concept Videos
 01:18
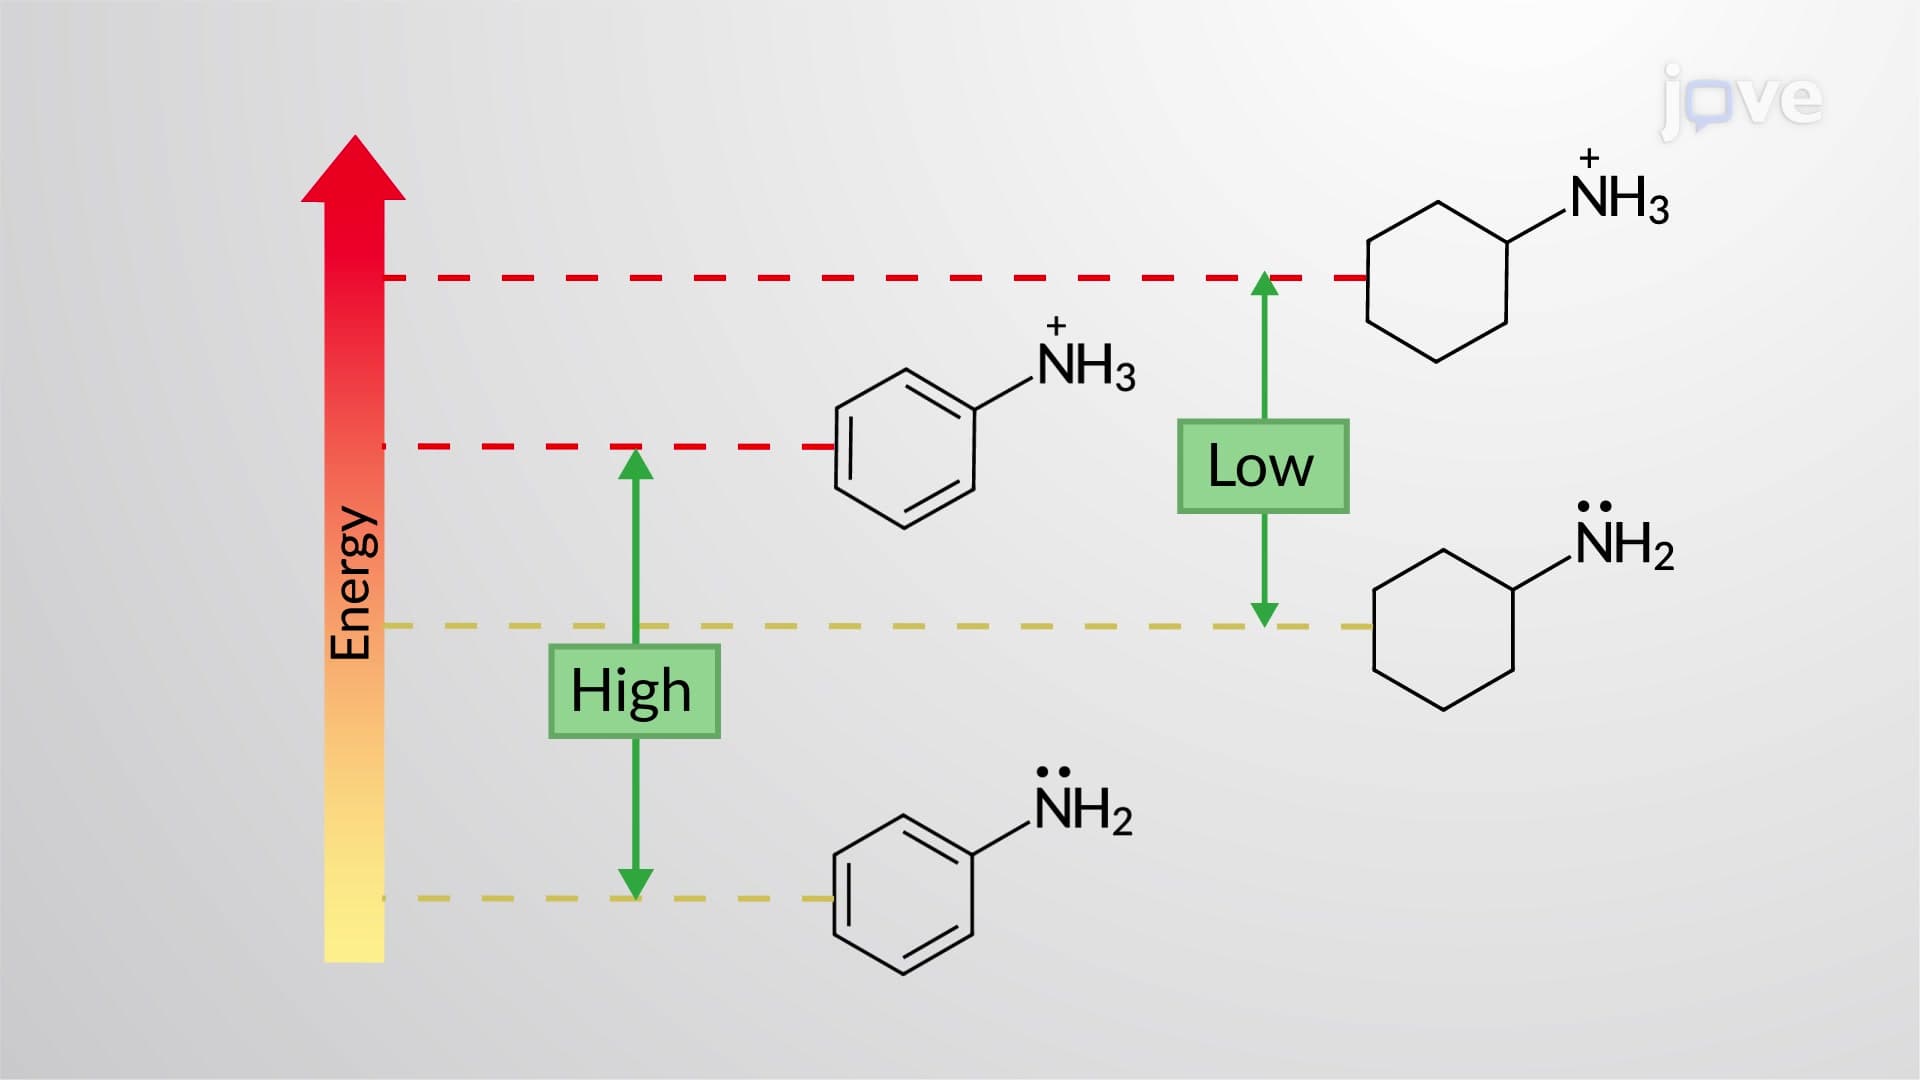
Basicity of Aromatic Amines
 03:49
Amino acids
 01:19
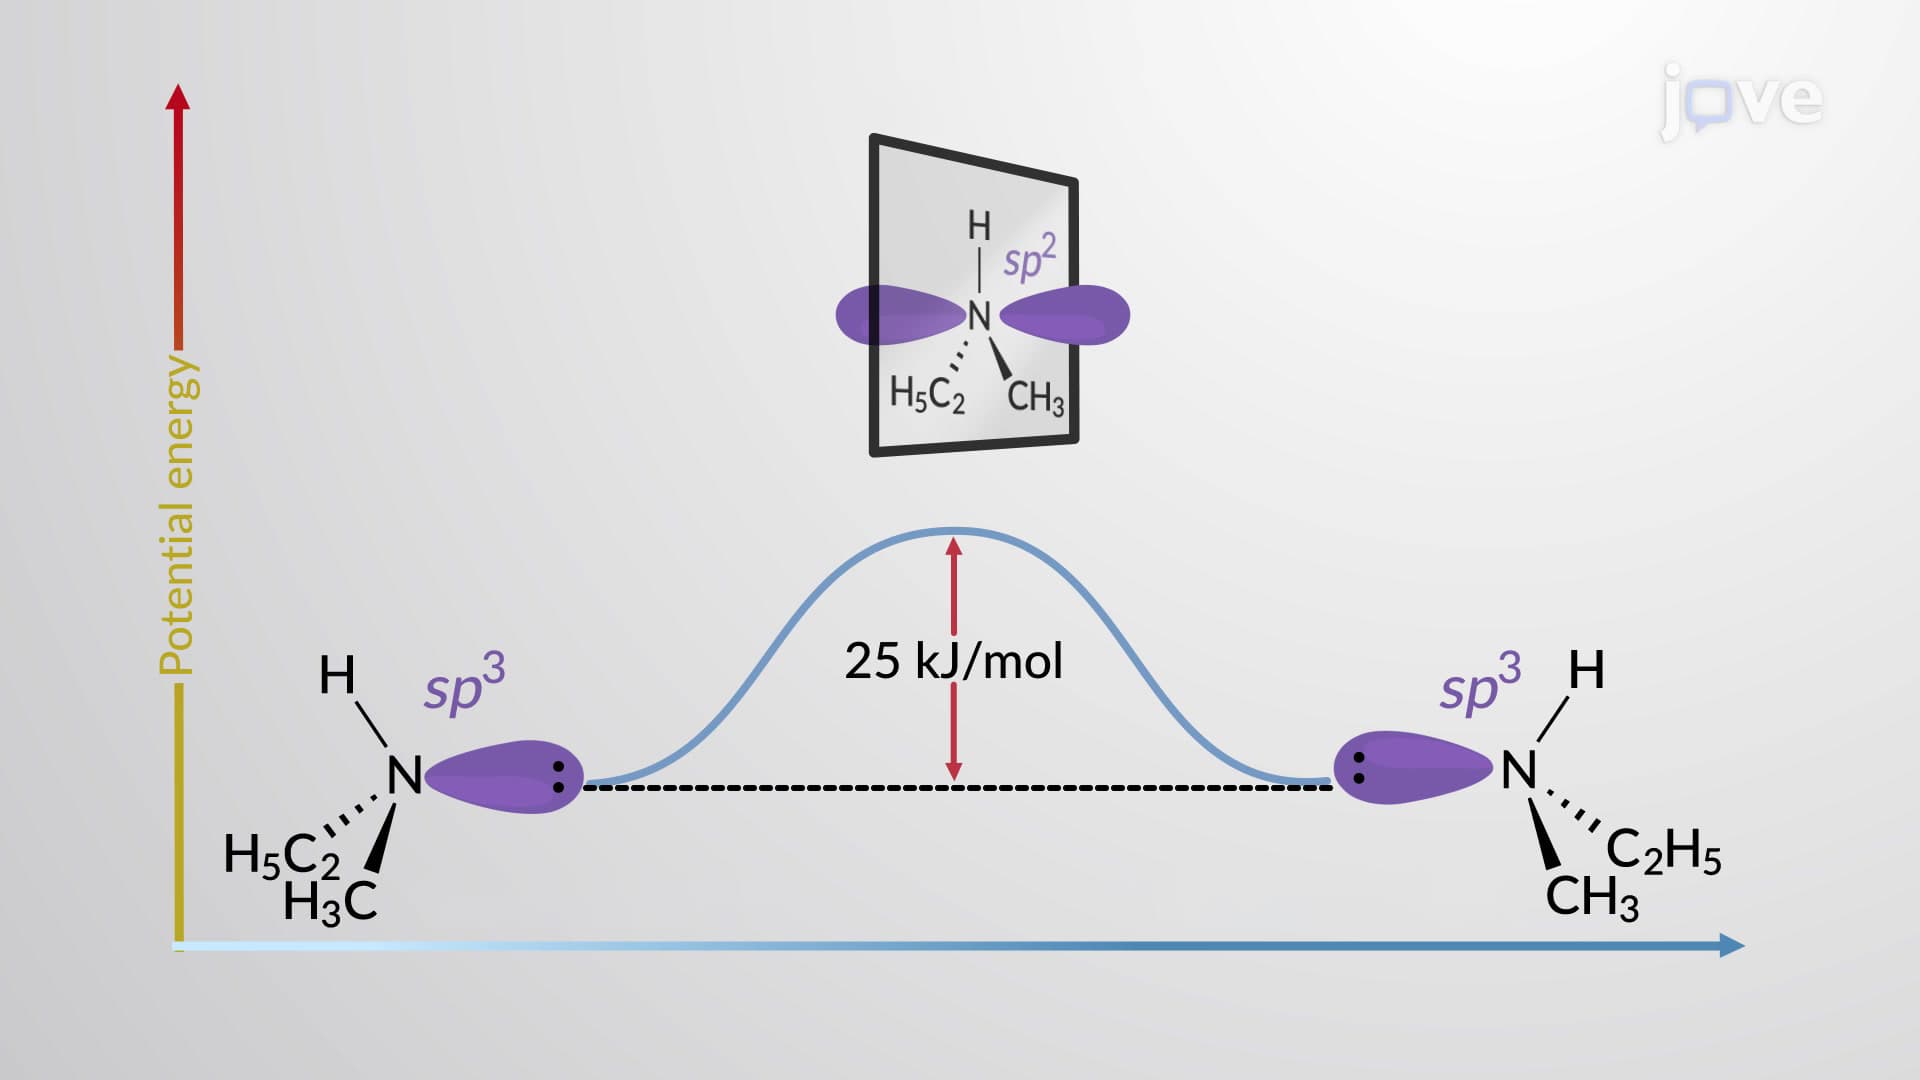
Structure of Amines
 01:26
Physical Properties of Amines
 02:35
Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
02:35Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...