Related Experiment Video
Updated: Oct 22, 2026

Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
From GROMACS Molecular Dynamics Simulations to Electronic Absorption Spectra: A Tutorial for Small Molecules in
Chiara Picarelli1, Giuseppina Raffaini1, Matteo Tommasini1
1Dipartimento di Chimica, Materiali e Ingegneria Chimica "Giulio Natta", Politecnico di Milano, Piazza Leonardo da Vinci, 32-20133 Milano, Italy.

Abstract:
Classical molecular dynamics is a theoretical method useful for investigating noncovalent intermolecular interactions and conformational flexibility, enabling the simulation of processes such as aggregation/dissolution and adsorption. Molecular dynamics and quantum chemistry are increasingly introduced in chemistry curricula, yet students may encounter difficulties when translating theoretical concepts into robust simulation protocols, particularly outside biomolecular systems. Here, we present a tutorial designed to guide students through the preparation, execution, and analysis of the simulations of small organic molecules in nonaqueous solvents, followed by the calculation of ultraviolet-visible (UV-vis) absorption spectra using time-dependent density functional theory. This tutorial focuses on several practical issues: obtaining force-field parameters, building simulation cells, selecting ensembles, and interpreting critical simulation parameters. Custom solvent models are validated against reference density and radial distribution functions, demonstrating how selected observables can be used to assess simulation reliability. We illustrate the usefulness of molecular dynamics by showing that molecular flexibility and solvent interactions broaden the UV-vis absorption band of indigo in chloroform. This contribution is intended as a hands-on educational resource for upper-level undergraduate and graduate students enrolled in physical chemistry and computational chemistry courses, as well as for Ph.D. students working in chemistry, physics, or material science, who are approaching molecular simulations. To support dissemination, we have released a ready-to-use package on GitHub that includes solvent boxes and all the necessary files and data for incorporating new molecules into GROMACS simulations: https://github.com/cippee/SolventSafari-for-GROMACS.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:12
An Introduction to Processing, Fitting, and Interpreting Transient Absorption Data
Published on: February 16, 2024
Related Concept Videos
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:14
Molecular Spectroscopy: Absorption and Emission
 01:30
UV–Vis Spectrum
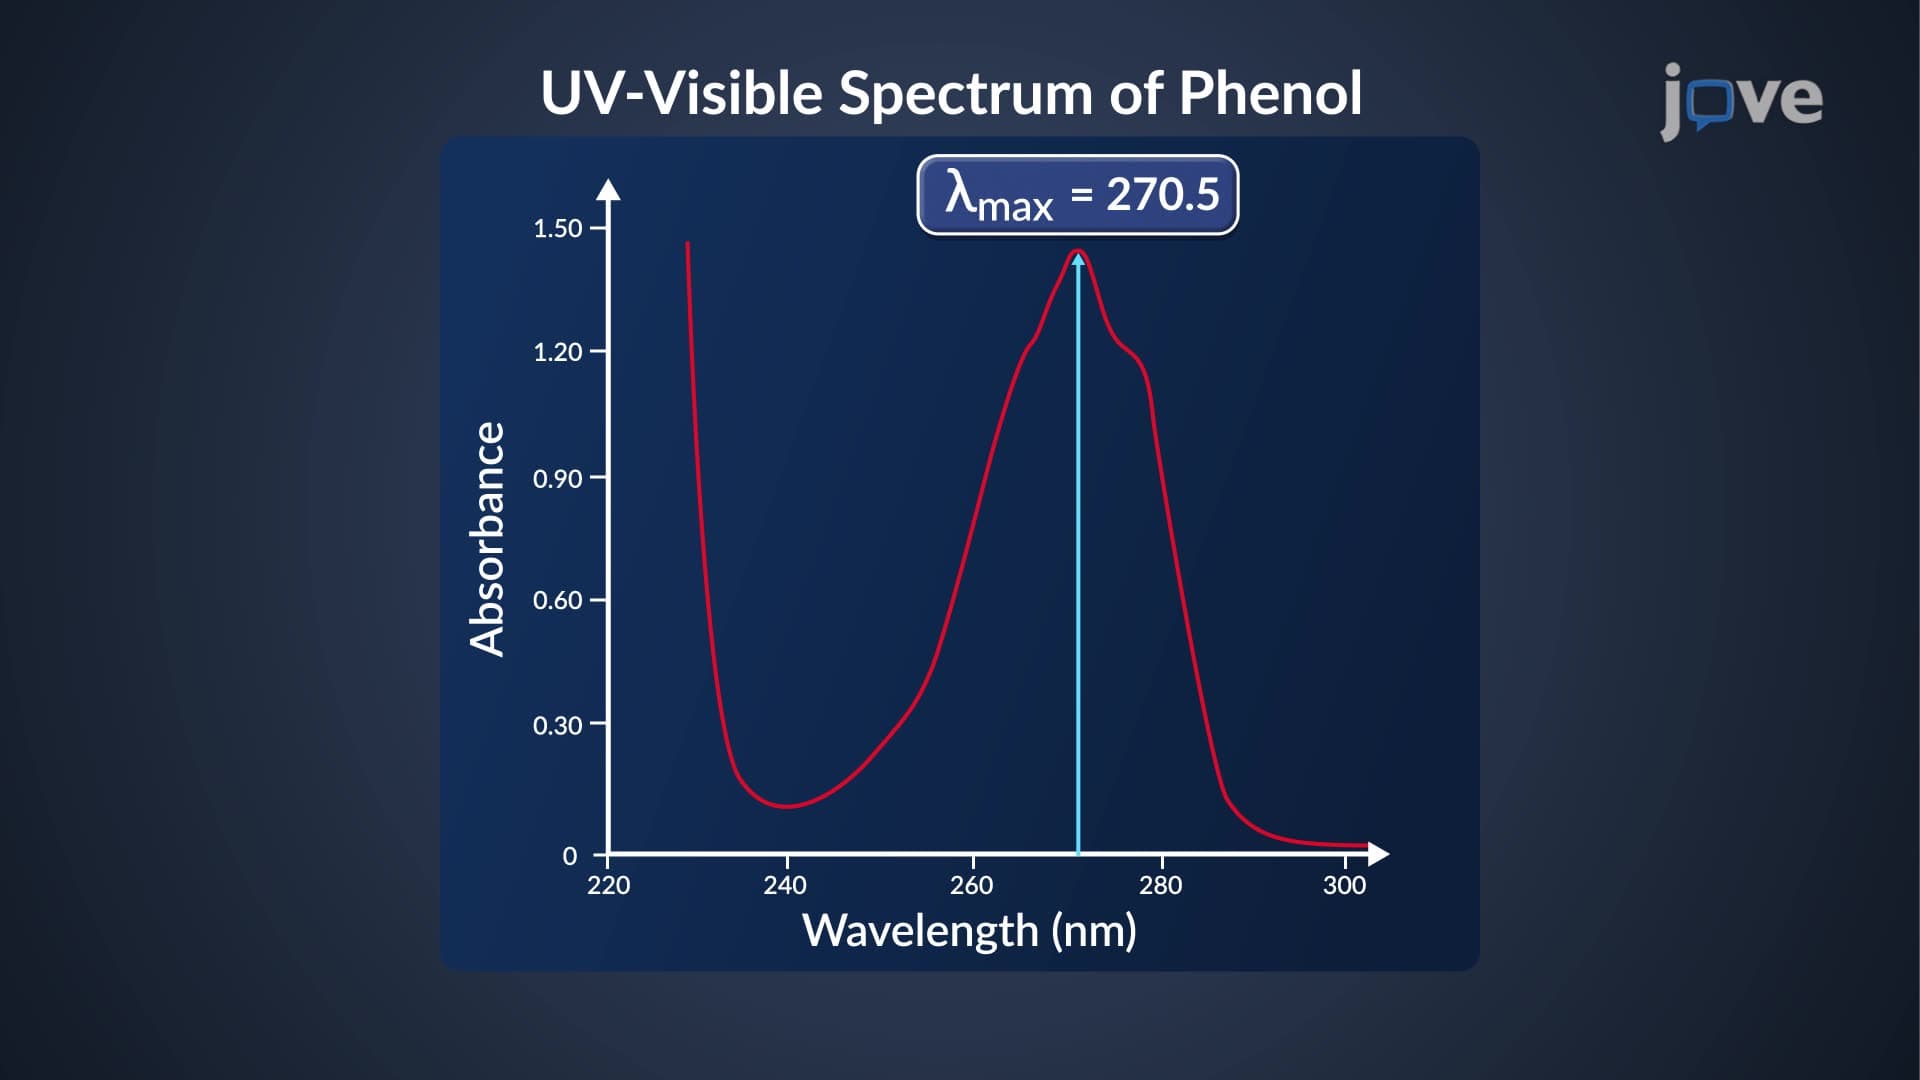
The UV–Vis spectrum of a molecule is the plot of its absorbance versus wavelength. The plot is drawn by taking molar absorptivity (ε) or log ε on the y-axis (ordinate)...
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is the extent of conjugation in the...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...