Uncontrolled C3 activation causes membranoproliferative glomerulonephritis in mice deficient in complement factor H
Related Experiment Video
Updated: Sep 30, 2026
Analyses of Proteinuria, Renal Infiltration of Leukocytes, and Renal Deposition of Proteins in Lupus-prone MRL/lpr Mice
Published on: June 8, 2022
Matthew C Pickering1, H Terence Cook, Joanna Warren
1Rheumatology Section, Faculty of Medicine, Imperial College, Hammersmith Campus, Du Cane Road, London W12 0NN, UK.

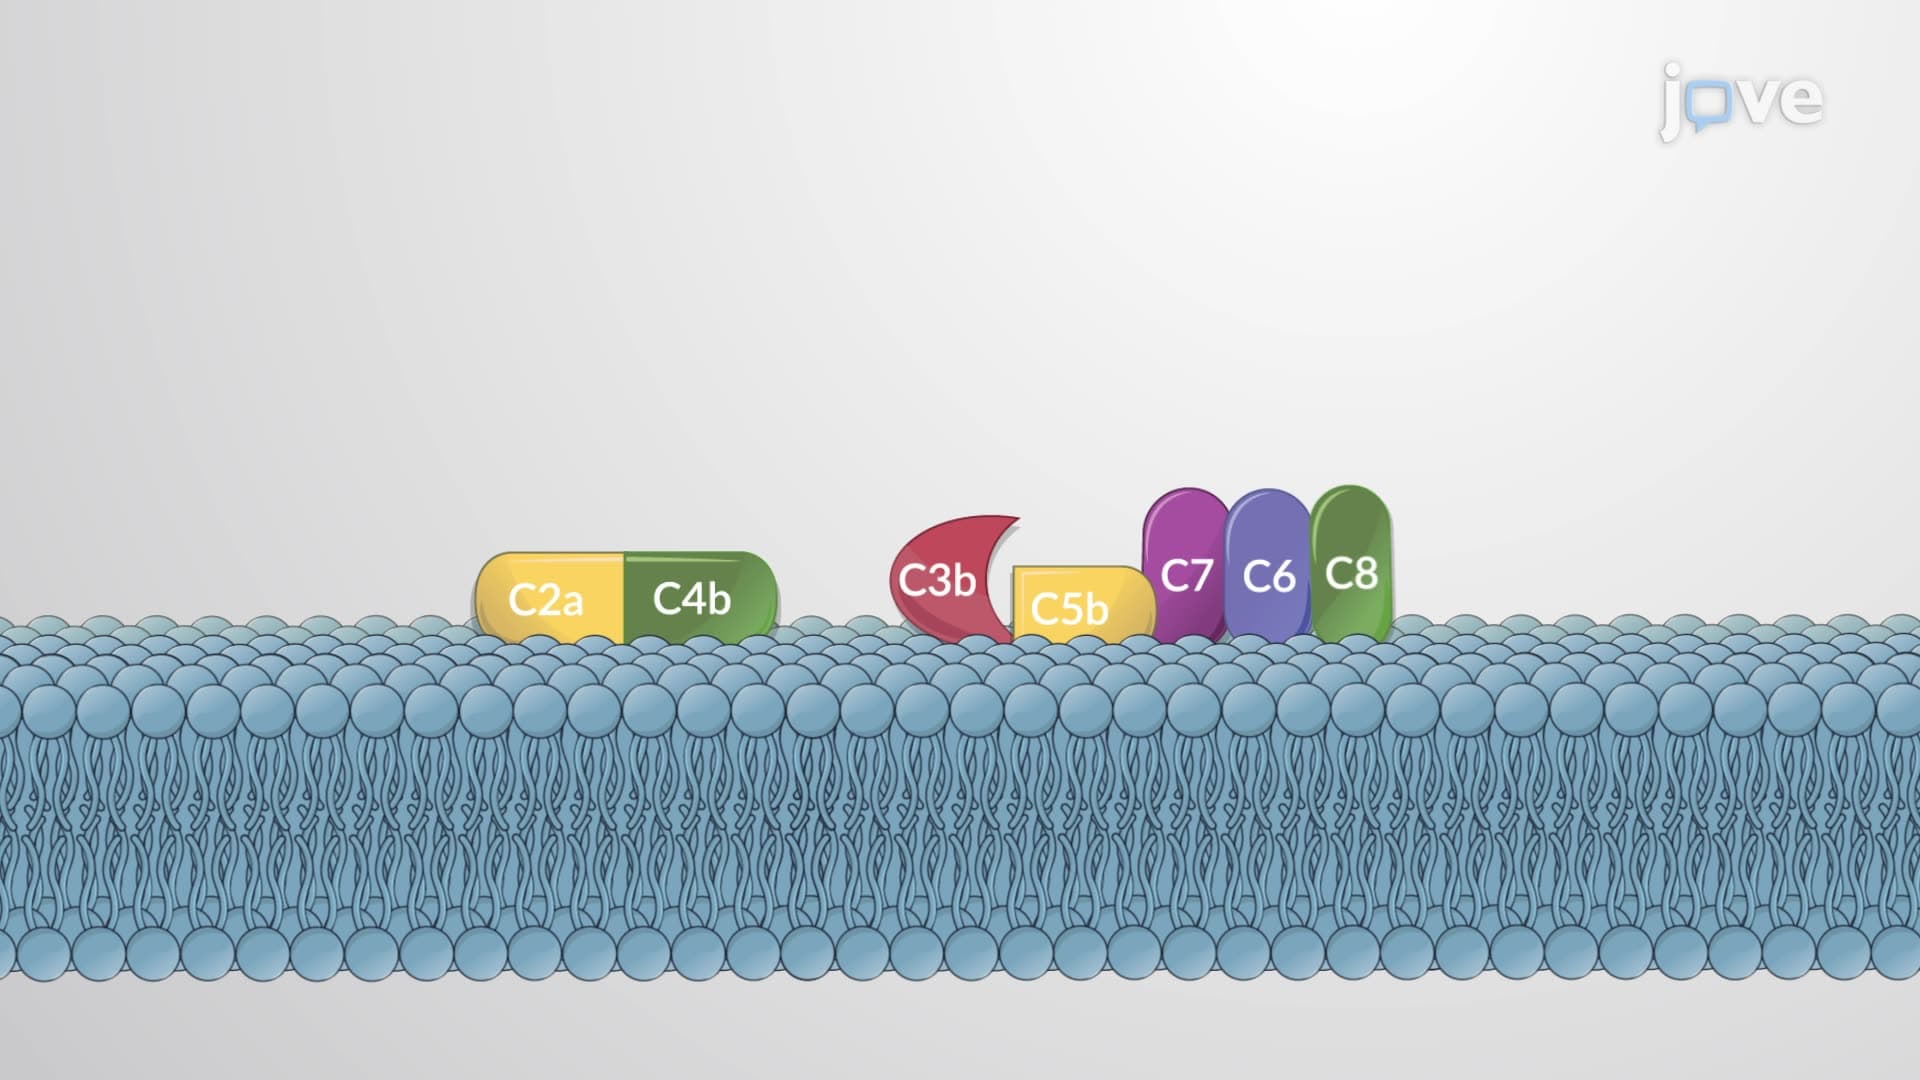
The alternative pathway of complement is activated continuously in vivo through the C3 'tick-over' pathway. This pathway is triggered by the hydrolysis of C3, resulting in the formation of C3 convertase. This, in turn, generates C3b, which mediates many of the biological functions of complement. Factor H, the main regulator of this activation, prevents formation and promotes dissociation of the C3 convertase enzyme, and, together with factor I, mediates the proteolytic inactivation of C3b. Factor H deficiency, described in 29 individuals from 12 families and in pigs, allows unhindered activation of fluid-phase C3 and severe depletion of plasma C3 (ref. 11). Membranoproliferative glomerulonephritis (MPGN) occurs in factor H-deficient humans and pigs. Although MPGN has been reported in other conditions in which uncontrolled activation of C3 occurs, the role of C3 dysregulation in the pathogenesis of MPGN is not understood. Here we show that mice deficient in factor H (Cfh(-/-) mice) develop MPGN spontaneously and are hypersensitive to developing renal injury caused by immune complexes. Introducing a second mutation in the gene encoding complement factor B, which prevents C3 turnover in vivo, obviates the phenotype of Cfh(-/-) mice. Thus, uncontrolled C3 activation in vivo is essential for the development of MPGN associated with deficiency of factor H.
The alternative pathway of complement is activated continuously in vivo through the C3 'tick-over' pathway. This pathway is triggered by the hydrolysis of C3, resulting in the formation of C3 convertase. This, in turn, generates C3b, which mediates many of the biological functions of complement. Factor H, the main regulator of this activation, prevents formation and promotes dissociation of the C3 convertase enzyme, and, together with factor I, mediates the proteolytic inactivation of C3b. Factor H deficiency, described in 29 individuals from 12 families and in pigs, allows unhindered activation of fluid-phase C3 and severe depletion of plasma C3 (ref. 11). Membranoproliferative glomerulonephritis (MPGN) occurs in factor H-deficient humans and pigs. Although MPGN has been reported in other conditions in which uncontrolled activation of C3 occurs, the role of C3 dysregulation in the pathogenesis of MPGN is not understood. Here we show that mice deficient in factor H (Cfh(-/-) mice) develop MPGN spontaneously and are hypersensitive to developing renal injury caused by immune complexes. Introducing a second mutation in the gene encoding complement factor B, which prevents C3 turnover in vivo, obviates the phenotype of Cfh(-/-) mice. Thus, uncontrolled C3 activation in vivo is essential for the development of MPGN associated with deficiency of factor H.
Related Concept Videos
 01:27
Complement System
 01:27
Cytomegalovirus Disease