[The active site of human glucocerebrosidase: structural predictions and experimental validations]
Related Experiment Video
Updated: Sep 29, 2026
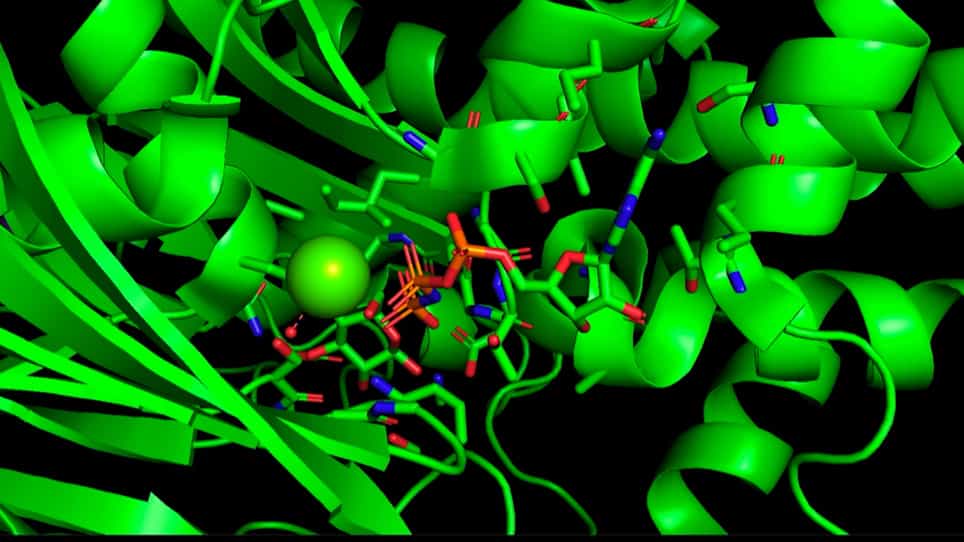
Modeling an Enzyme Active Site using Molecular Visualization Freeware
Published on: December 25, 2021
Sylvie Fabrega1, Patrick Durand, Jean-Paul Mornon
1INSERM U458, Höpital Robert Debré, 48, Bd Sérurier, 75019 Paris.

Abstract:
Gaucher disease is a lysosomal storage disorder caused by a deficiency in glucocerebrosidase which cleaves the beta-glucosidic linkage of glucosylceramide, a normal intermediate in glycolipid metabolism. Glucocerebrosidase belongs to the clan GH-A of glycoside hydrolases, a large group of enzymes which function with retention of the anomeric configuration at the hydrolysis site. Accurate three-dimensional (3D) structure data for glucocerebrosidase should help to better understand the molecular bases of Gaucher disease. As such 3D structure data were not available, we used the two-dimensional hydrophobic cluster analysis (HCA) method to make structure predictions for the catalytic domains of clan GH-A glycoside hydrolases. We found that all the enzymes of clan GH-A may share a similar catalytic domain consisting of an (alpha/beta)8 barrel with the critical acid/base and nucleophile residues located at the C-terminal ends of strands beta 4 and beta 7, respectively. In the case of glucocerebrosidase, Glu 235 was predicted to be the putative acid/base catalyst whereas the nucleophile was located at Glu 340. Next, in order to obtain experimental evidence supporting these HCA-based predictions, we used retroviral vectors to express, in murine null cells, E235A and E340A mutant proteins, in which alanine residues unable to participate in the enzymatic reaction replace the presumed critical glutamic acid residues. Both mutants were found to be catalytically inactive although they were correctly folded/processed and sorted to the lysosome. Thus, Glu 235 and Glu 340 do indeed play key roles in the active site of human glucocerebrosidase as predicted by the HCA analysis. In a broader perspective, our work points out that bioinformatics approaches may be highly useful for generating structure-function predictions based on sequence-structure interrelationships, especially in the context of a rapid increase in protein sequence information through genome sequencing.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 00:49
Ligand Binding and Linkage
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis pathway,...
 02:34
Enzymes
Enzyme deficiencies can often translate into life-threatening diseases. For example, a genetic abnormality resulting in the deficiency of the enzyme G6PD...
 01:29
Protein Folding Quality Check in the RER