Related Experiment Video
Updated: Sep 29, 2026
In Vitro Enzyme Measurement to Test Pharmacological Chaperone Responsiveness in Fabry and Pompe Disease
Published on: December 20, 2017
[Fabry's disease (alpha-galactosidase-A deficiency): physiopathology, clinical signs, and genetic aspects]
1Unité de Génétique Clinique, Hôpital Européen Georges Pompidou, 20, rue Leblanc, 75015 Paris.

Abstract:
Fabry disease (FD, OMIM 301500) is an X-linked inherited disorder of metabolism due to mutations in the gene encoding alpha-galactosidase A, a lysosomal enzyme. The enzymatic defect leads to the accumulation of neutral glycosphingolipids throughout the body, particularly within endothelial cells. Resulting narrowing and tortuosity of small blood vessels lead to tissue ischaemia and infarction. Inability to prevent the progression of glycosphingolipid deposition causes significant morbidity (acroparesthesia, angiokeratoma, autonomic dysfunction, cardiomyopathy and deafness), and mortality from early onset strokes, heart attack and renal failure in adulthood. Demonstration of alpha-galactosidase A deficiency in leukocytes or plasma is the definitive method for the diagnosis of affected hemizygous males. Most heterozygotes present with a cardiac, renal or neurological symptomatology, although to a lesser extent than what is observed in hemizygotes. Due to random X-chromosomal inactivation, enzymatic detection of carriers is often inconclusive. Molecular testing of possible carriers is therefore mandatory for accurate genetic counselling. The GLA gene has been cloned and more than 200 mutations have been identified. Medical management is symptomatic and consists of partial pain relief with analgesic drugs (gabapentin, carbamazepine), whereas renal transplantation or dialysis is available for patients experiencing end-stage renal failure. However, the ability to produce high doses of alpha-galactosidase A in vitro has opened the way to clinical studies and enzyme replacement therapy has recently been validated as a therapeutic agent for FD patients in clinical trials. Long term safety and efficacy of replacement therapy are currently being investigated.
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:27
Glucose Transporters
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes:
 01:22
Lysosomal Hydrolases
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:05
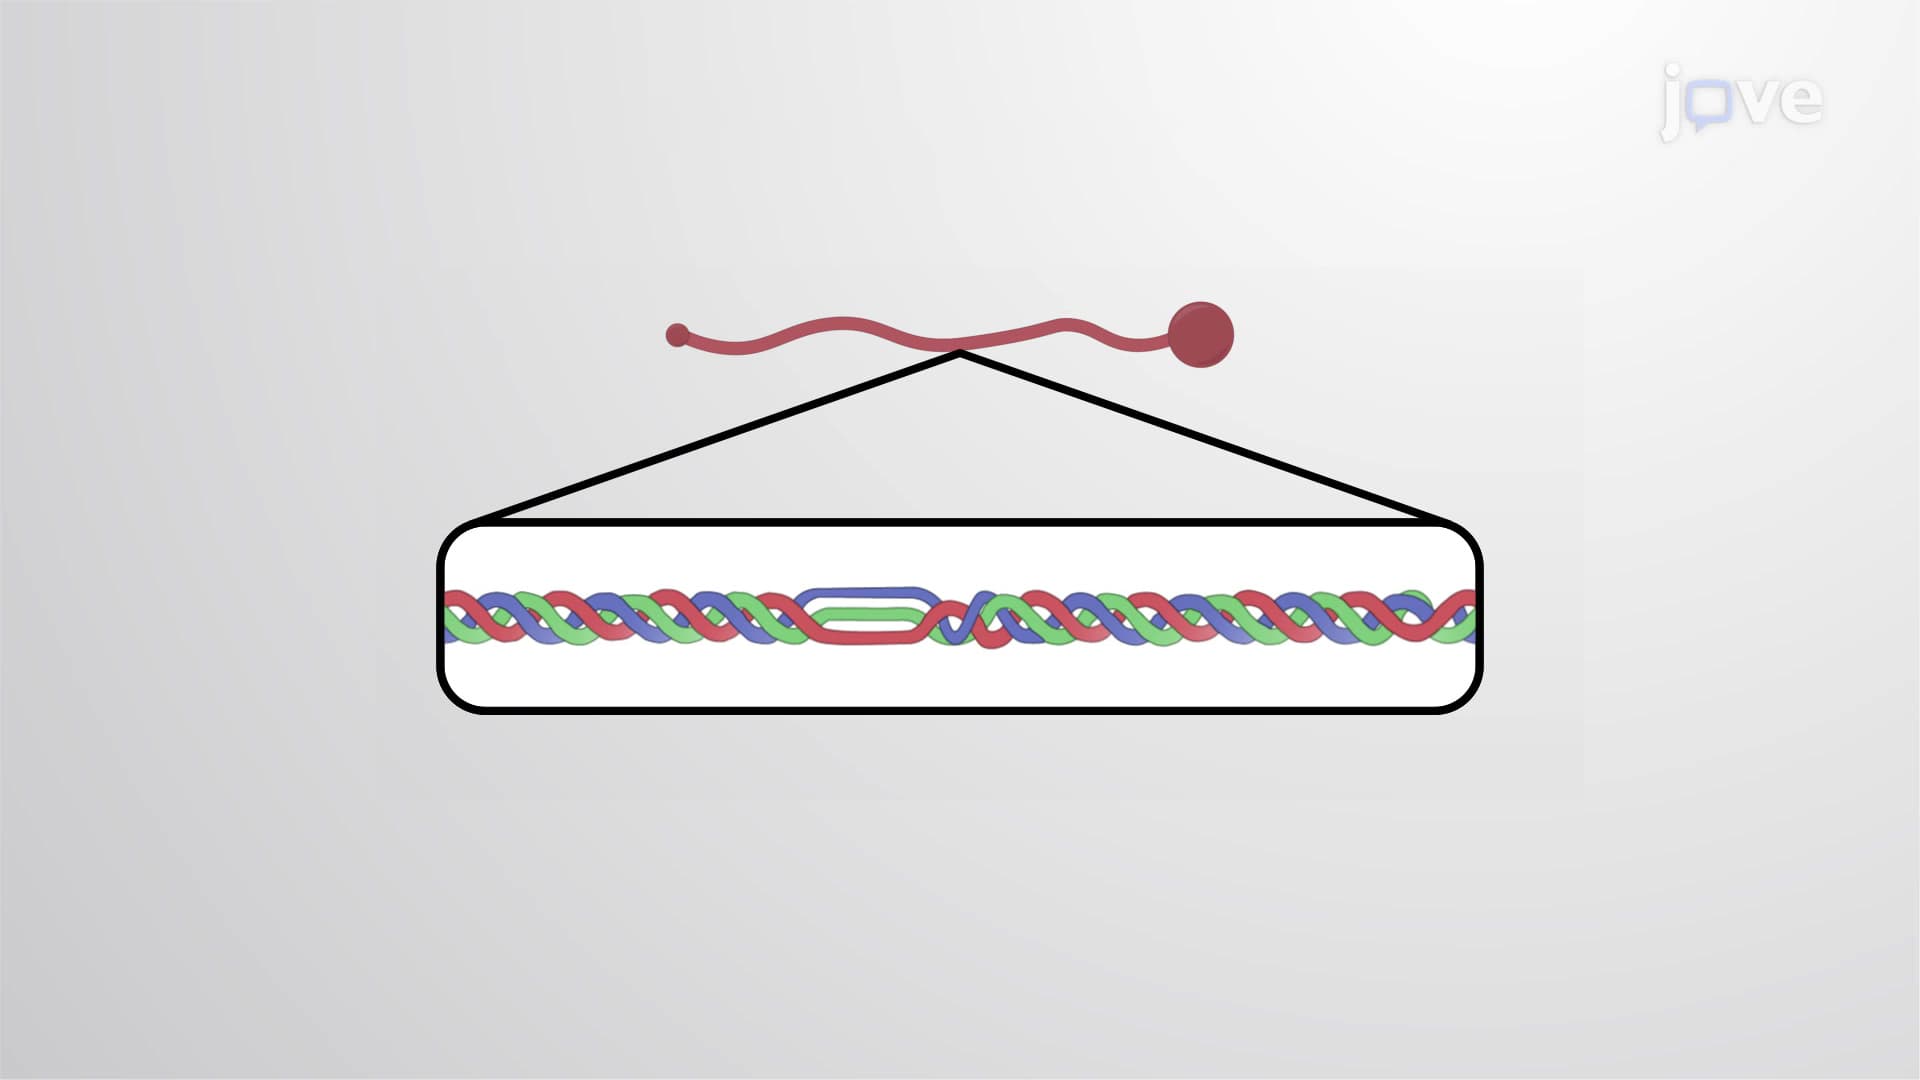
Type IV Collagen of Basal Lamina
A type IV collagen molecule has six alpha chains which can exist in...
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...