Related Experiment Video
Updated: Aug 15, 2026

Structure and Coordination Determination of Peptide-metal Complexes Using 1D and 2D 1H NMR
Published on: December 16, 2013
Conformational analysis: crystallographic, NMR, and molecular mechanics studies of flexible sulfonic esters
Orde Q Munro1, Jean M McKenzie, Sandra D Strydom
1School of Chemical and Physical Sciences, University of Natal, Pietermaritzburg, Private Bag X01, Scottsville 3209, South Africa. munroo@nu.ac.za

Abstract:
Two novel X-ray structures of the sulfonic ester derivatives 2-(6-iodo-1,3-benzodioxol-5-yl)ethyl 4-nitrobenzenesulfonate, 3, and 2-(6-iodo-1,3-benzodioxol-5-yl)ethyl 4-methylbenzenesulfonate, 4, have been obtained in a study aimed at analyzing the structures and conformations of sulfonic ester derivatives that are routinely used in alkaloid syntheses. The crystal structure of 4 is highly unusual, containing four independent molecules that belong to two distinct conformational types: (1) a hairpin conformation (stabilized mainly by intramolecular pi-stacking) and (2) a stepped conformation (stabilized mainly by intermolecular pi-stacking). Compound 3, on the other hand, crystallizes exclusively as the hairpin conformer. New MM+ force field parameters for sulfonic esters have been developed using the X-ray data, empirical rules, and DFT calculations to estimate the bond dipole parameters. Grid searches of conformational space for 3 and 4 using MM methods show that there are several gas-phase conformations within 5 kcal/mol of the global minimum and that the lowest energy conformations (by approximately 4.6 kcal/mol) are of the hairpin type. Analysis of the MM conformational energies suggests that the dominant intramolecular interaction stabilizing the hairpin conformations of 3 and 4 is van der Waals attraction. Moreover, the lattice energies for packing the hairpin conformations of 3 and 4 are approximately 4 kcal/mol more favorable than for the stepped conformations. Various intermolecular interactions contribute to the complexity of the observed crystal structures of 3 and 4, including electrostatic attraction between O and I atoms in neighboring molecules. Langevin dynamics (LD) simulations at several temperatures (6.0 ns, friction coefficient = 2.5 ps(-1)) indicate that the conformational exchange rates are approximately 10(10)-10(11) s(-1) over the temperature range 213-400 K, accounting for the temperature-independent (1)H NMR spectra of 3 and 4.
Related Concept Videos
 02:18
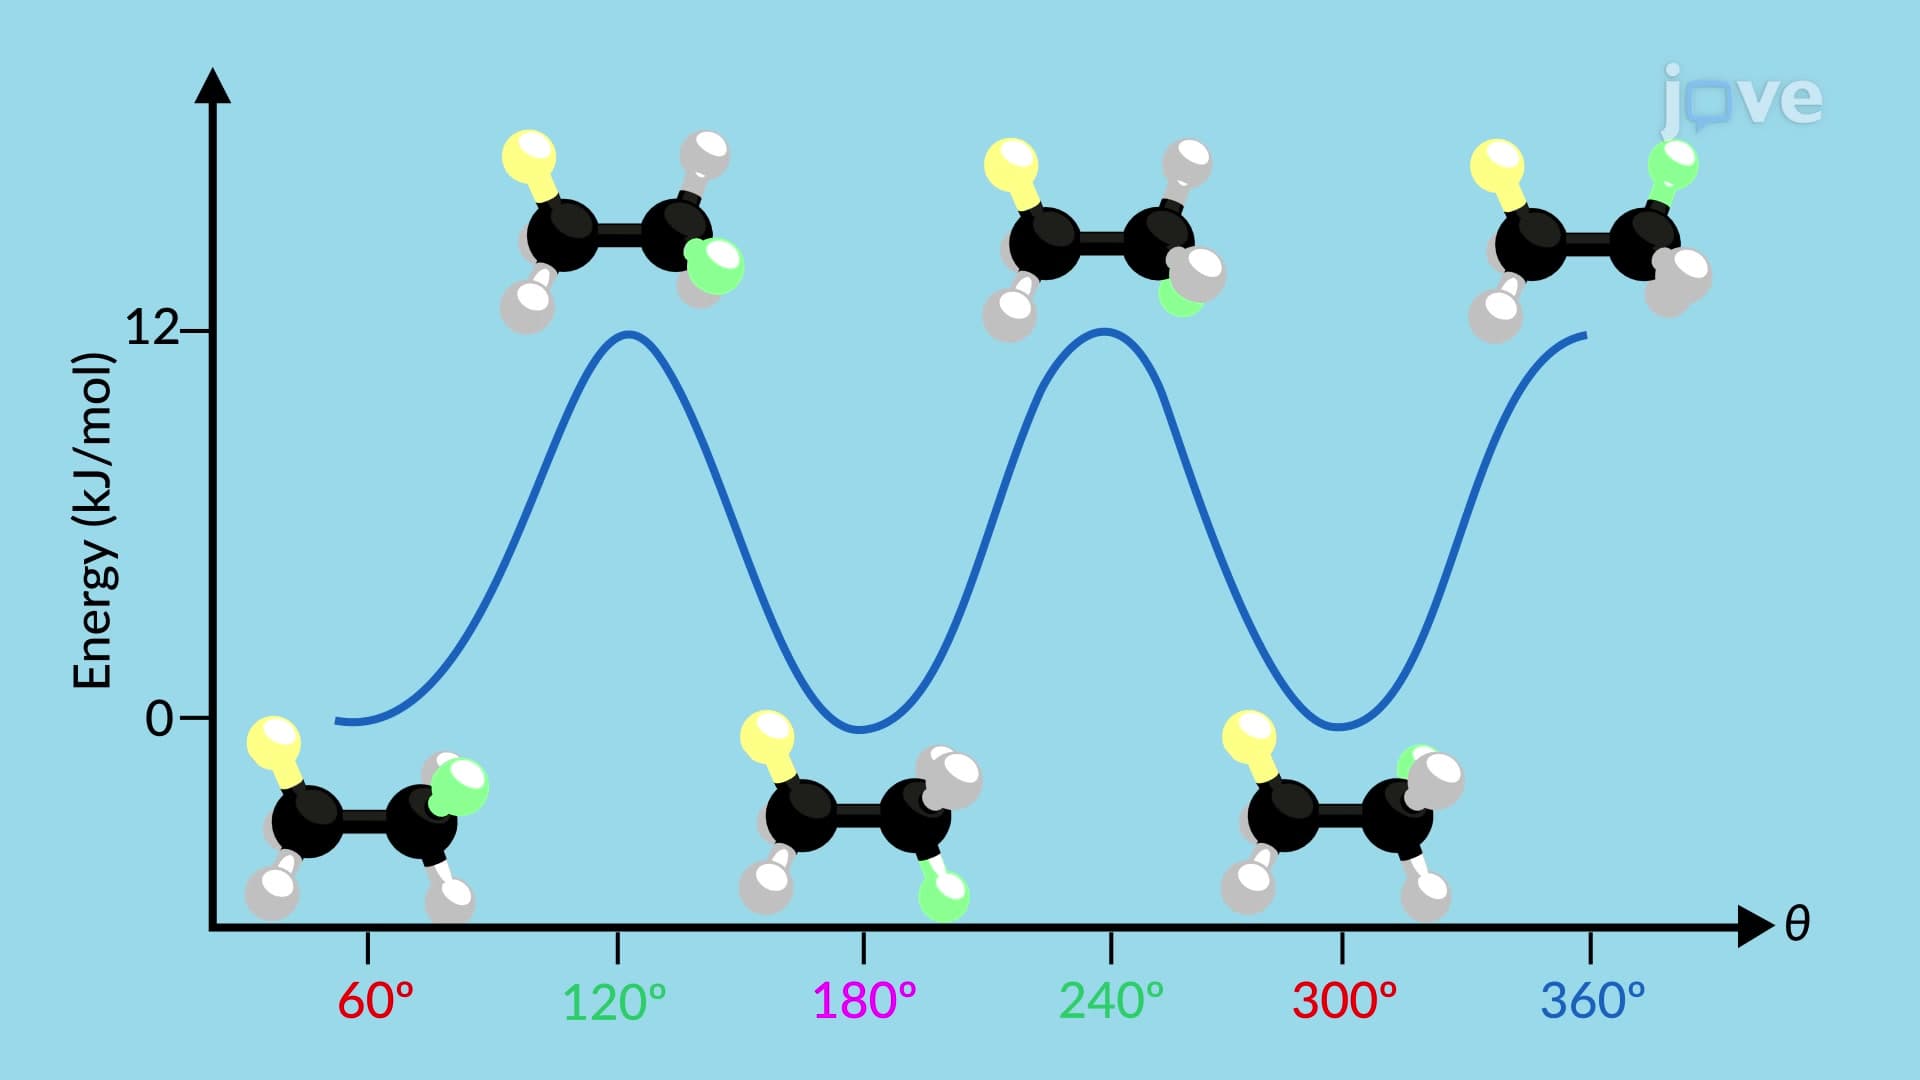
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered ethane, the...
 02:30
Chirality at Nitrogen, Phosphorus, and Sulfur
A consequence of chirality is the need for enantiomeric resolution. While this is theoretically possible for all...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:28
Structures of Carboxylic Acid Derivatives
Carboxylic acid derivatives contain an acyl group attached to a heteroatom such as chlorine, oxygen, or nitrogen. The carbonyl carbon and oxygen are both sp2-hybridized with an unhybridized p orbital.
The three sp2 orbitals of the carbonyl carbon form three σ bonds, one each with the carbonyl oxygen, the α carbon, and the heteroatom, whereas the other two sp2 orbitals of the carbonyl oxygen are occupied by the lone pairs. Further, the unhybridized p...
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR