Related Experiment Video
Updated: Sep 26, 2026
Development and Characterization of In Vitro Microvessel Network and Quantitative Measurements of Endothelial [Ca2+]i and Nitric Oxide Production
Published on: May 19, 2016
Role of Ca2+ signaling in the regulation of endothelial permeability
Chinnaswamy Tiruppathi1, Richard D Minshall, Biman C Paria
1Department of Pharmacology M/C868, College of Medicine, University of Illinois at Chicago, 835 S. Wolcott Avenue, Chicago, IL 60612, USA. tiruc@uic.edu

Abstract:
The vascular endothelial cell forms a semipermeable barrier between blood and interstitium. Inflammatory mediators such as thrombin and histamine induce vascular leakage defined as increased endothelial permeability to plasma proteins and other solutes. Increased endothelial permeability is the hallmark of inflammatory vascular edema. Inflammatory mediators that bind to heptahelical G protein-coupled receptors (GPCR) trigger increased endothelial permeability by increasing the intracellular Ca(2+) concentration ([Ca(2+)](i)). The rise in [Ca(2+)](i) activates key signaling pathways, which mediate cytoskeletal reorganization (through myosin light chain (MLC)-dependent contraction) and disassembly of VE-cadherin at the adherens junctions. The Ca(2+)-dependent protein kinase C (PKC) isoform, PKC-alpha, plays a critical role in initiating endothelial cell contraction and disassembly of VE-cadherin junctions. The increase in [Ca(2+)](i) induced by a variety of agonists is achieved by the generation of inositol 1,4,5-trisphosphate (IP3), activation of IP3 receptors (IP3R), release of stored intracellular Ca(2+), and Ca(2+) entry through plasma membrane channels. Recent findings demonstrate that IP3-sensitive Ca(2+) store depletion activates plasma membrane cation channels (i.e., store-operated cation channels (SOC) or Ca(2+) release activated channels) to cause Ca(2+) influx in endothelial cells. This mode of Ca(2+) influx is also known as capacitative Ca(2+) entry (CCE). Store-operated Ca(2+) influx signals increase in permeability and nitric oxide (NO) production and provokes changes in gene expression in endothelial cells. Recent studies have established that the Drosophila transient receptor potential (TRP) gene family of channels expressed in endothelial cells can function as SOC. Deletion of one of the TRP homologues, TRPC4, in mouse caused impairment in store-operated Ca(2+) current and Ca(2+) store release activated Ca(2+) influx in aortic and lung endothelial cells (LEC). In TRPC4 knockout (TRPC4(-/-)) mice, acetylcholine-induced endothelium-dependent smooth muscle relaxation was drastically reduced. In addition, TRPC4(-/-) mice LEC exhibited lack of actin stress fiber formation and cell retraction in response to thrombin activation of proteinase-activated receptor-1 (PAR-1) in endothelial cells. The increase in lung microvascular permeability in response to thrombin receptor activation was inhibited in TRPC4(-/-) mice. These results indicate that endothelial TRP channels such as TRPC1 and TRPC4 play an important role in signaling the increase in endothelial permeability.
Related Concept Videos
 01:16
Calmodulin-dependent Signaling
The Ca2+-CaM complex does not have enzymatic activity by itself. Instead, the complex binds downstream target proteins, including membrane proteins or enzymes,...
 01:24
Regulation of Angiogenesis and Blood Supply
 01:28
Nitric Oxide Signaling Pathway
 01:32
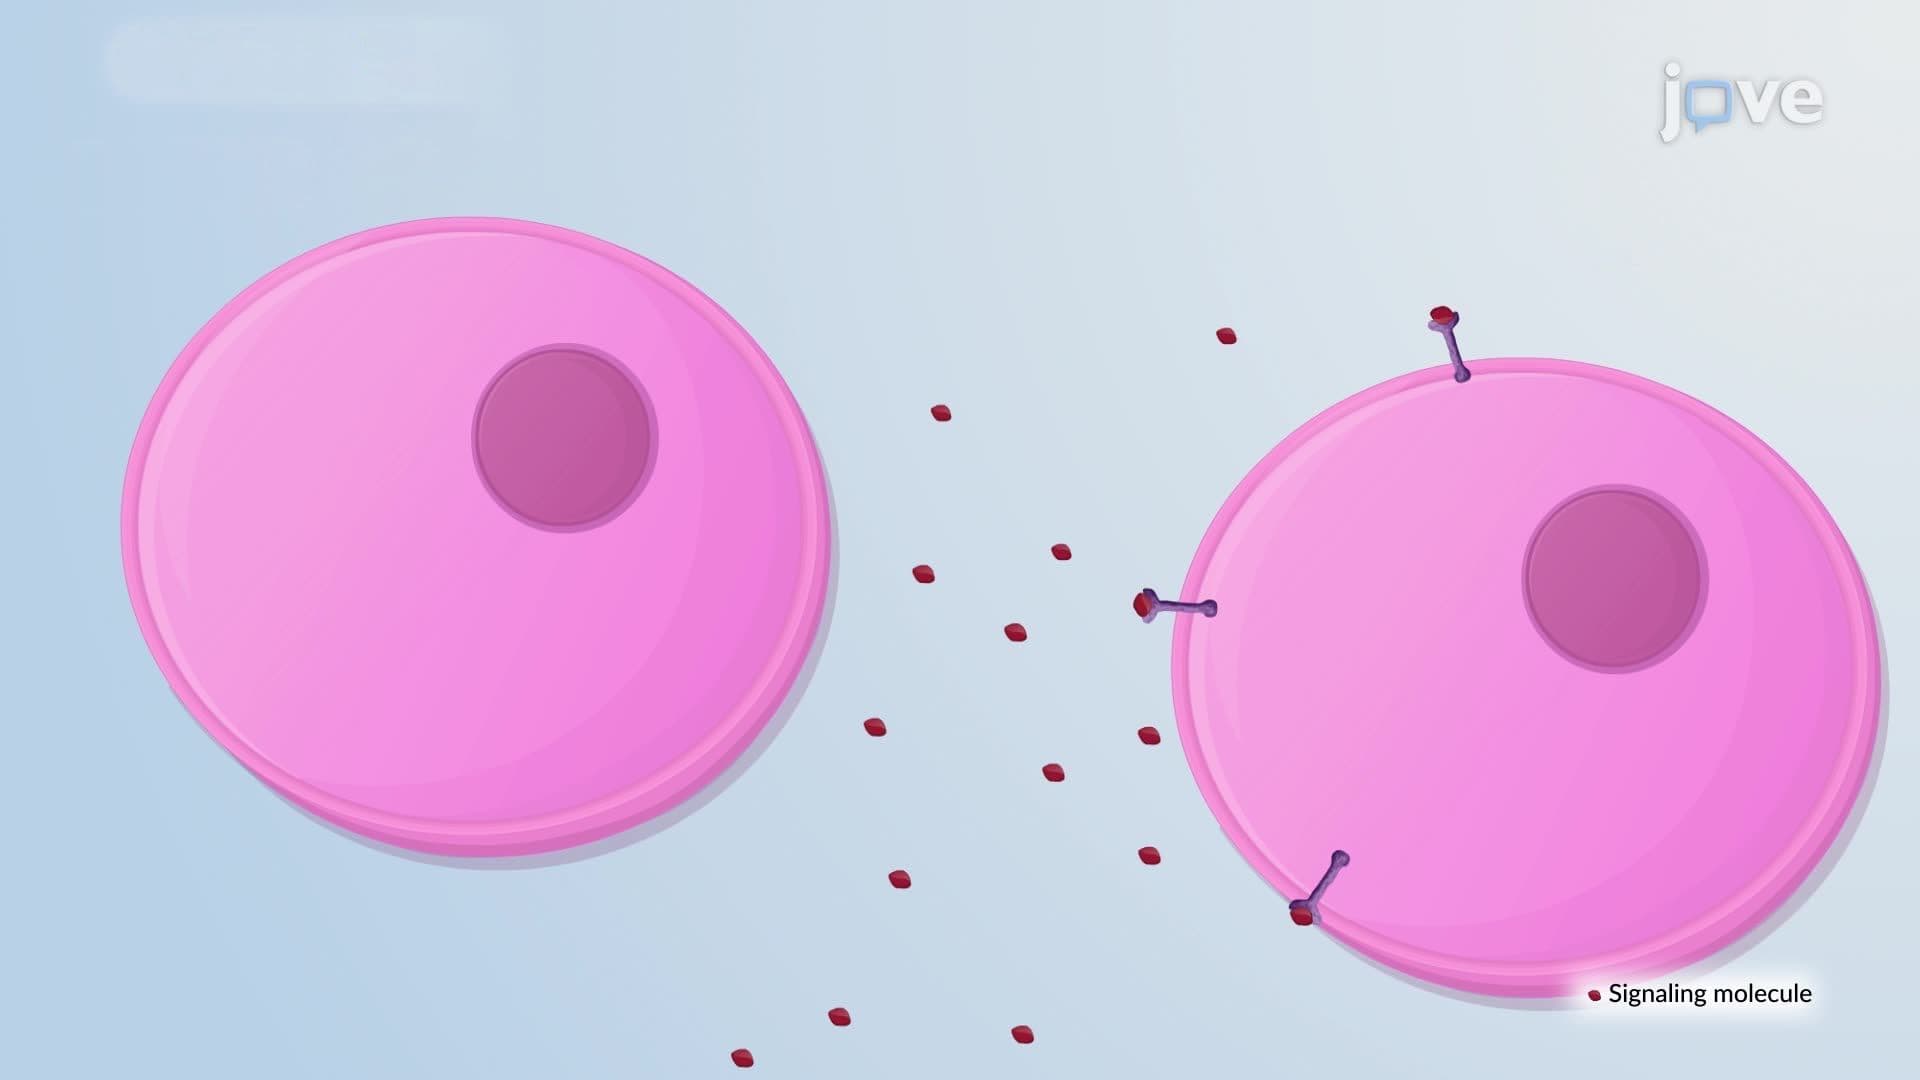
Paracrine Signaling
 01:27
Feedback Regulation of Calcium Concentration
Various transmembrane receptors, such as G protein-coupled receptors (GPCRs), elicit a response to extracellular signals by increasing cytosolic calcium. Activated GPCRs...
 01:15
Amplifying Signals via Second Messengers