Related Experiment Video
Updated: Sep 25, 2026
Diagonal Method to Measure Synergy Among Any Number of Drugs
Published on: June 21, 2018
Stereoselectivity of drug-receptor interactions
Willem Soudjin1, Ineke van Wijngaarden, A P IJzerman
1Leiden University, Leiden/Amsterdam Center for Drug Research, Division of Medicinal Chemistry, PO Box 9502, 2300 RA Leiden, the Netherlands.

Abstract:
Stereochemical aspects of drug action have intrigued researchers ever since (and even before) the introduction of the receptor concept. Easson and Stedman first formalized this by suggesting the three-point tethering of an asymmetric ligand to its macromolecular target. Today, enantioselectivity is increasingly understood in molecular and atomic detail. Through site-directed mutagenesis, the amino acids responsible for the stereoselective recognition of ligand molecules have been identified in a number of cases, in particular for the b(2)-adrenergic receptor, a prototypic G protein-coupled receptor. Novel ligands with one or more chiral centers and complicated stereochemistry continue to be developed as potential medicines, despite reservations put forward by regulatory agencies. In this review, recent achievements in this respect will be discussed for a number of receptor (sub)families, and some general thoughts will be presented on the stereoselectivity of drug action.
Related Concept Videos
 01:25
Dose-Response Relationship: Selectivity and Specificity
 01:29
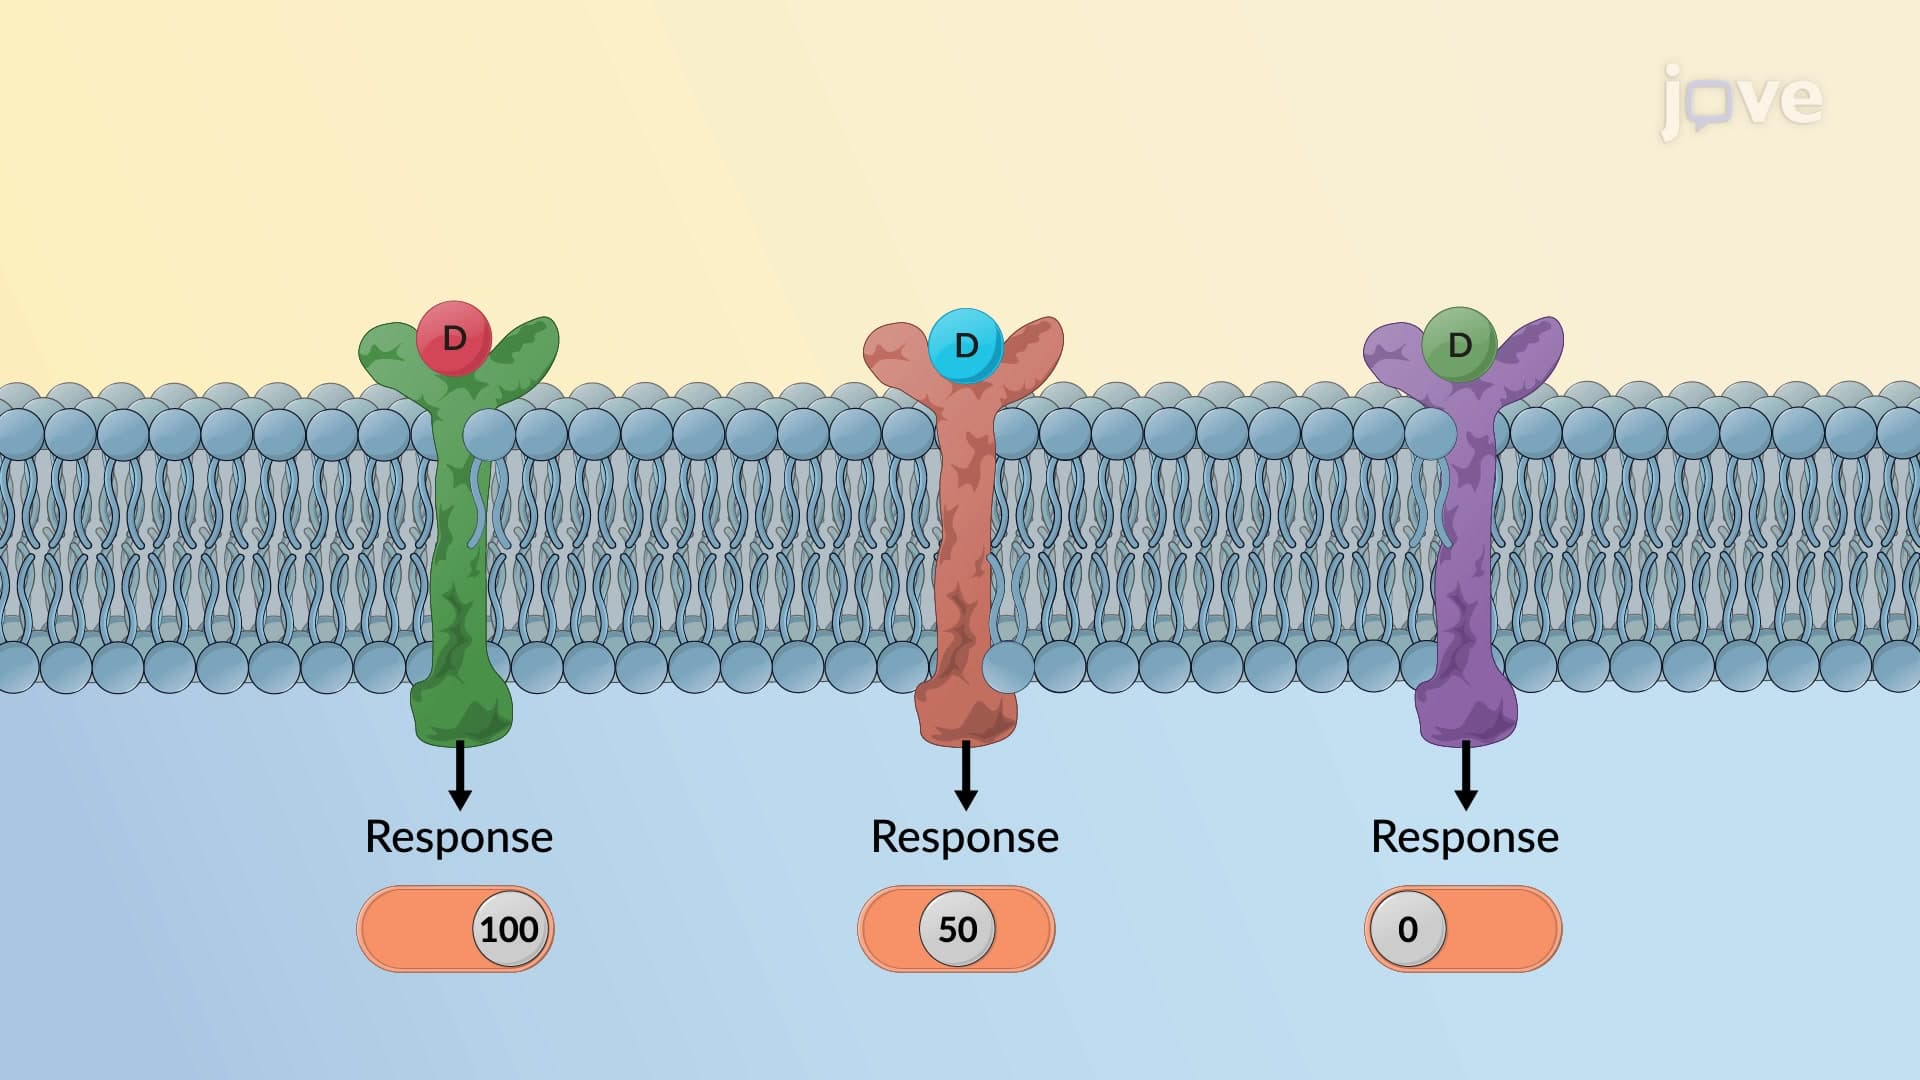
Drug-Receptor Interactions
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue.
 01:25
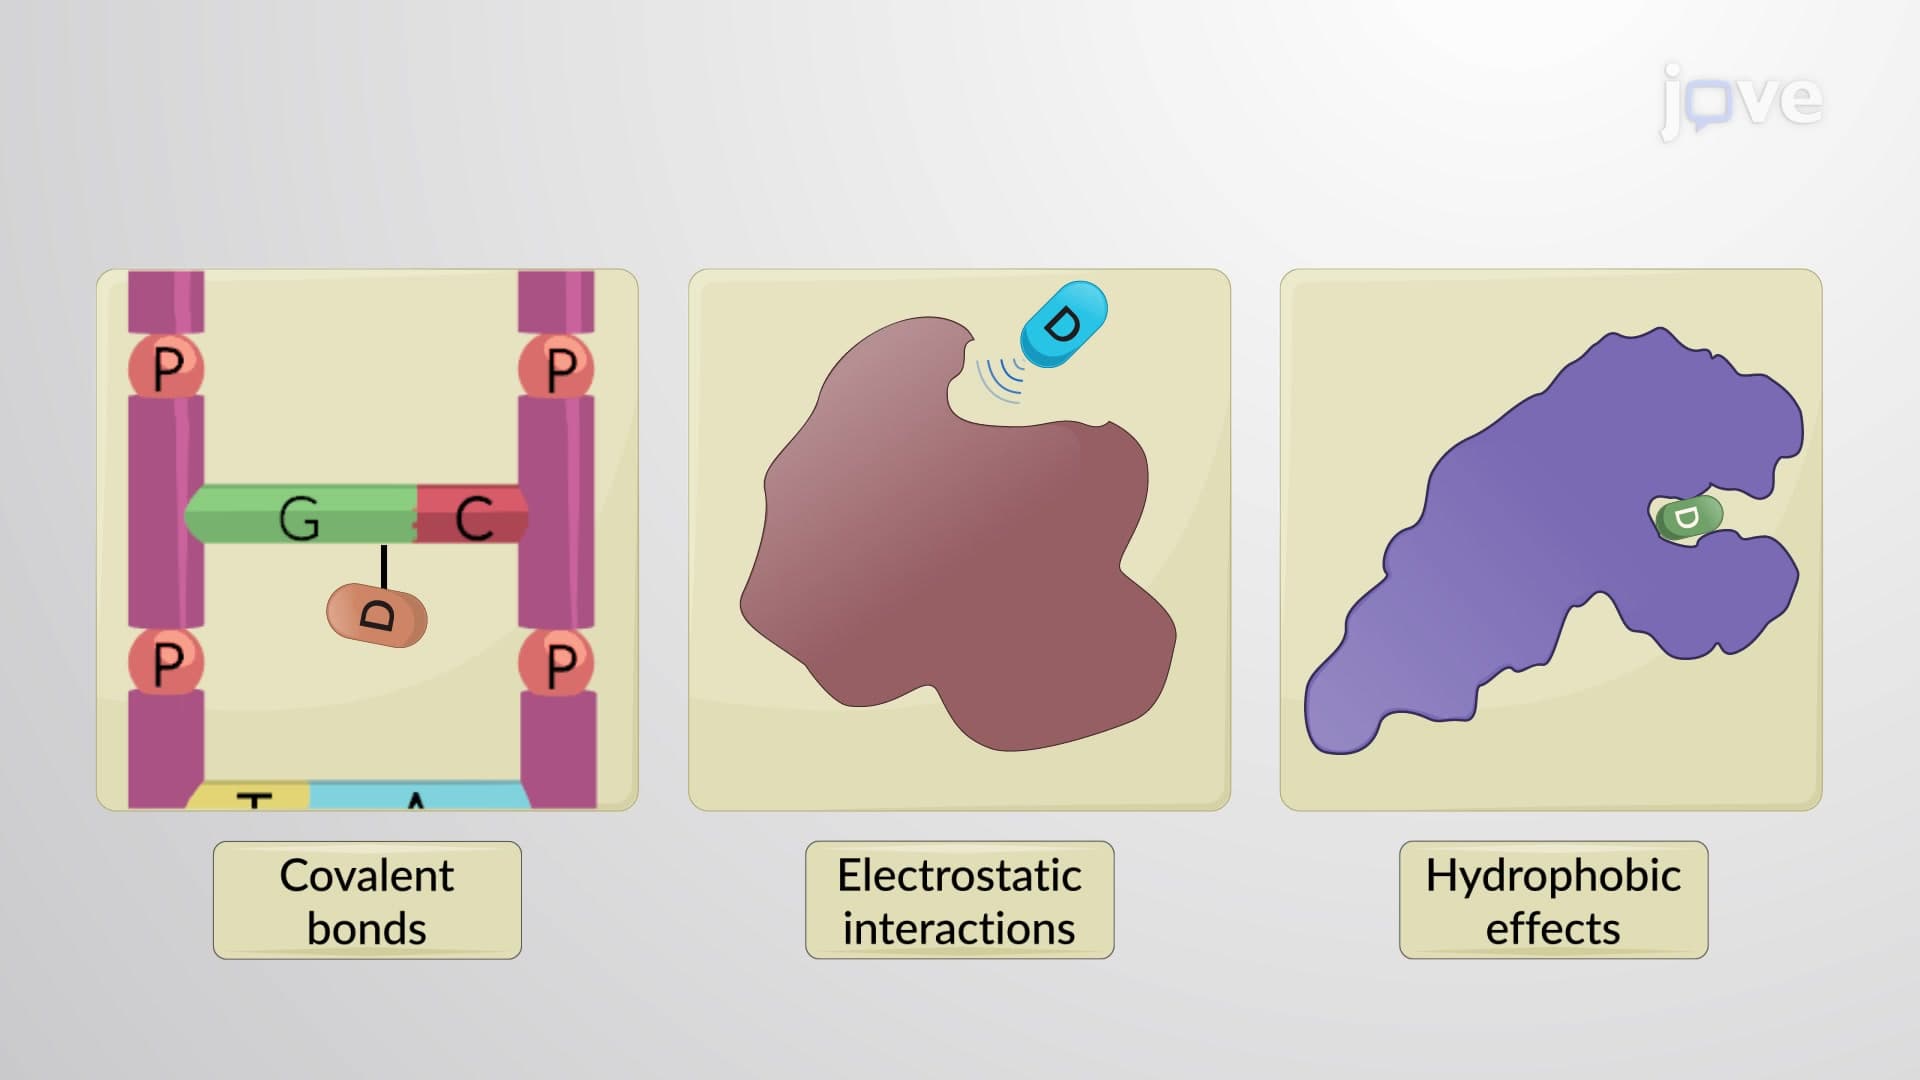
Drug-Receptor Bonds
In...
 01:20
Factors Affecting Dissolution: Drug Permeability, Stability and Stereochemistry
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with one...
 01:30
Quantitative Aspects of Drug-Receptor Interaction