Related Experiment Video
Updated: Sep 25, 2026

Modeling Encephalopathy of Prematurity Using Prenatal Hypoxia-ischemia with Intra-amniotic Lipopolysaccharide in Rats
Published on: November 20, 2015
Predicting neonatal perchlorate dose and inhibition of iodide uptake in the rat during lactation using
Rebecca A Clewell1, Elaine A Merrill, Kyung O Yu
1Geo-Centers, Inc., Wright-Patterson AFB, Ohio 45433, USA. rclewell@ciit.org

Perchlorate (ClO4-), a contaminant in drinking water, competitively inhibits active uptake of iodide (I-) into various tissues, including mammary tissue. During postnatal development, inhibition of I- uptake in the mammary gland and neonatal thyroid and the active concentration ClO4- in milk indicate a potentially increased susceptibility of neonates to endocrine disruption. A physiologically based pharmacokinetic (PBPK) model was developed to reproduce measured ClO4- distribution in the lactating and neonatal rat and predict resulting effects on I- kinetics from competitive inhibition at the sodium iodide symporter (NIS). Kinetic I- and ClO4- behavior in tissues with NIS (thyroid, stomach, mammary gland, and skin) was simulated with multiple subcompartments, Michaelis-Menten (M-M) kinetics and competitive inhibition. Physiological and kinetic parameters were obtained from literature and experiment. Systemic clearance and M-M parameters were estimated by fitting simulations to tissue and serum data. The model successfully describes maternal and neonatal thyroid, stomach, skin, and plasma, as well as maternal mammary gland and milk data after ClO4- exposure (from 0.01 to 10 mg/kg-day ClO4-) and acute radioiodide (2.1 to 33,000 ng/kg I-) dosing. The model also predicts I- uptake inhibition in the maternal thyroid, mammary gland, and milk. Model simulations predict a significant transfer of ClO4- through milk after maternal exposure; approximately 50% to 6% of the daily maternal dose at doses ranging from 0.01 to 10.0 mg ClO4-/kg-day, respectively. Comparison of predicted dosimetrics across life-stages in the rat indicates that neonatal thyroid I- uptake inhibition is similar to the adult and approximately tenfold less than the fetus.
Perchlorate (ClO4-), a contaminant in drinking water, competitively inhibits active uptake of iodide (I-) into various tissues, including mammary tissue. During postnatal development, inhibition of I- uptake in the mammary gland and neonatal thyroid and the active concentration ClO4- in milk indicate a potentially increased susceptibility of neonates to endocrine disruption. A physiologically based pharmacokinetic (PBPK) model was developed to reproduce measured ClO4- distribution in the lactating and neonatal rat and predict resulting effects on I- kinetics from competitive inhibition at the sodium iodide symporter (NIS). Kinetic I- and ClO4- behavior in tissues with NIS (thyroid, stomach, mammary gland, and skin) was simulated with multiple subcompartments, Michaelis-Menten (M-M) kinetics and competitive inhibition. Physiological and kinetic parameters were obtained from literature and experiment. Systemic clearance and M-M parameters were estimated by fitting simulations to tissue and serum data. The model successfully describes maternal and neonatal thyroid, stomach, skin, and plasma, as well as maternal mammary gland and milk data after ClO4- exposure (from 0.01 to 10 mg/kg-day ClO4-) and acute radioiodide (2.1 to 33,000 ng/kg I-) dosing. The model also predicts I- uptake inhibition in the maternal thyroid, mammary gland, and milk. Model simulations predict a significant transfer of ClO4- through milk after maternal exposure; approximately 50% to 6% of the daily maternal dose at doses ranging from 0.01 to 10.0 mg ClO4-/kg-day, respectively. Comparison of predicted dosimetrics across life-stages in the rat indicates that neonatal thyroid I- uptake inhibition is similar to the adult and approximately tenfold less than the fetus.
More Related Videos
 08:46
Dose Uptake of Platinum- and Ruthenium-based Compound Exposure in Zebrafish by Inductively Coupled Plasma Mass Spectrometry with Broader Applications
Published on: April 21, 2022
 08:08
Determination of the Transport Rate of Xenobiotics and Nanomaterials Across the Placenta using the ex vivo Human Placental Perfusion Model
Published on: June 18, 2013
Related Concept Videos
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:15
Impact of Pharmacokinetic–Pharmacodynamic Models: Regulatory Decisions
 01:29
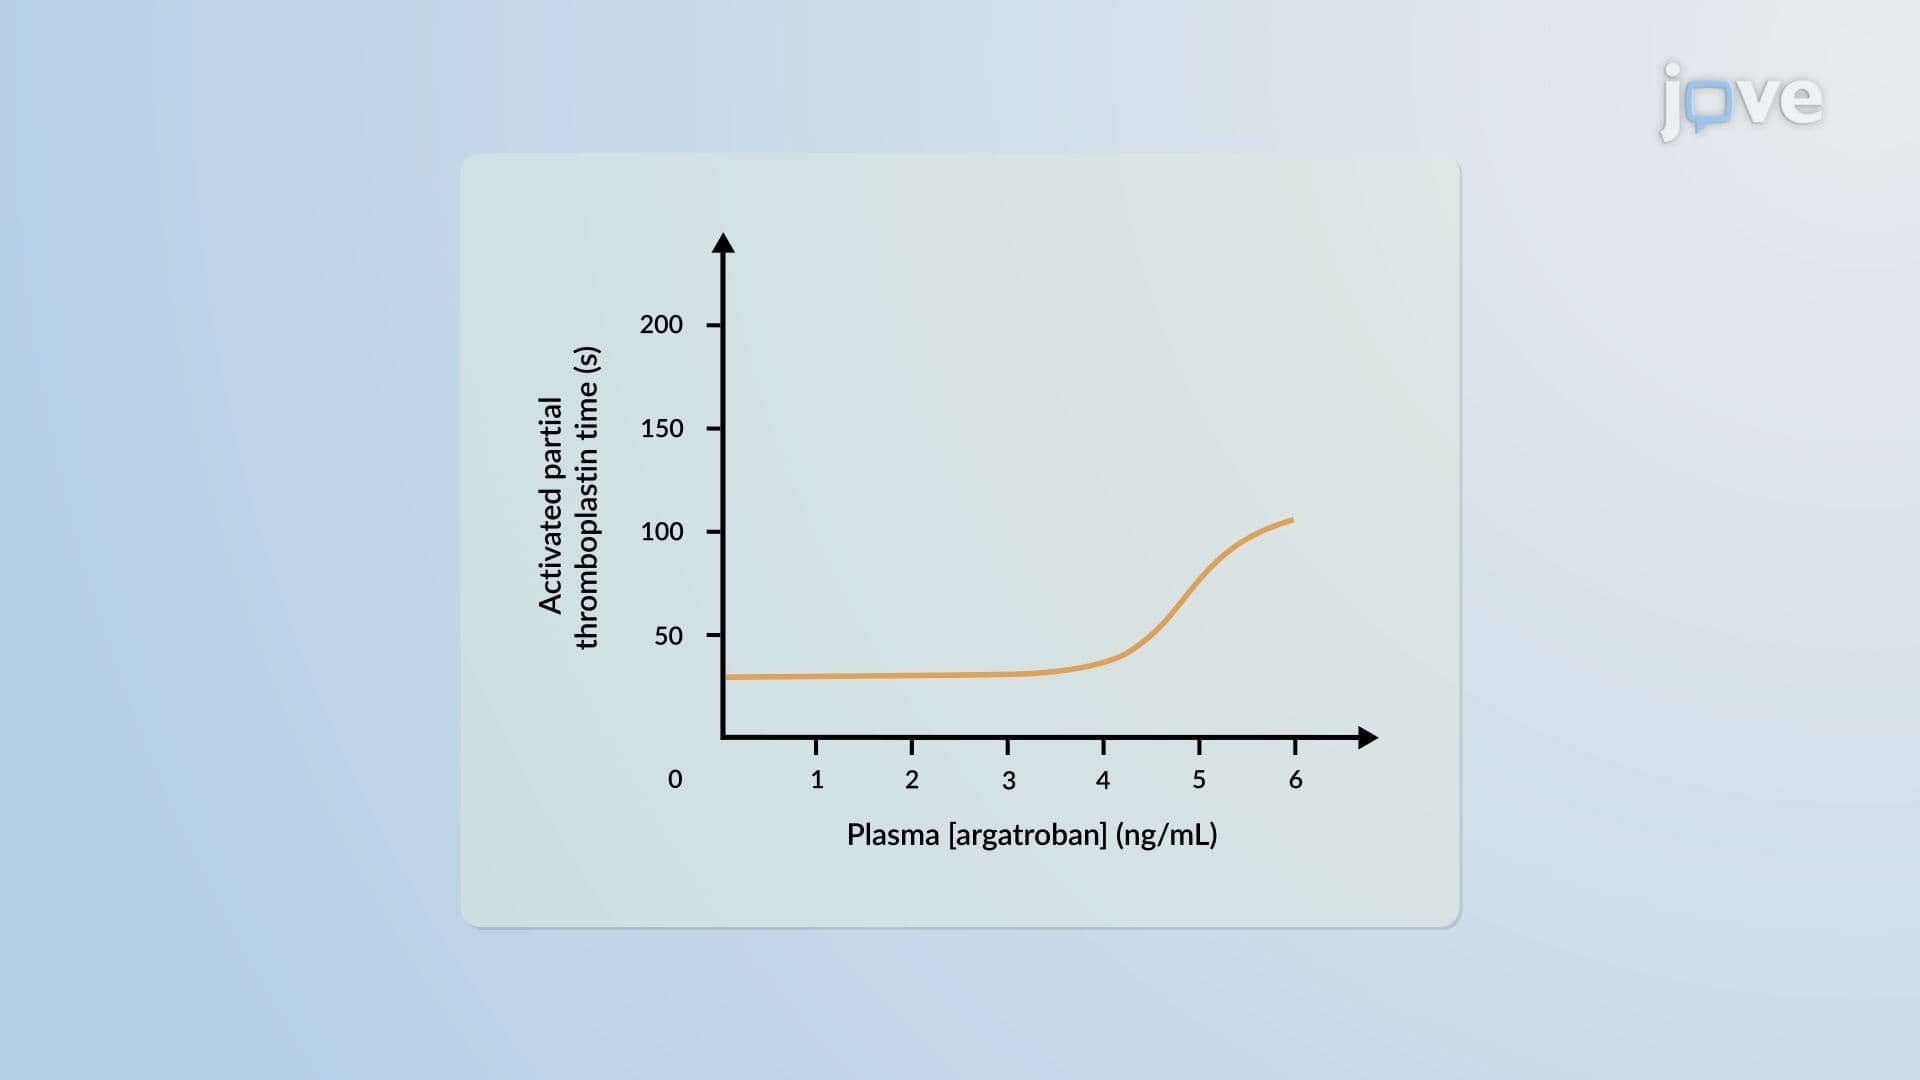
Pharmacodynamic Models: Direct Effect Model and Indirect Response Model
 01:26
Pharmacokinetic Models: Comparison and Selection Criterion
Physiological models take a detailed approach by considering specific molecular processes. They can predict drug distribution, metabolism, and elimination changes, providing a comprehensive understanding of how drugs interact with the body.
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model