Related Experiment Video
Updated: Aug 30, 2026


A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
PROSPECT II: protein structure prediction program for genome-scale applications
Dongsup Kim1, Dong Xu, Jun-tao Guo
1Protein Informatics Group, Life Sciences Division, Oak Ridge National Laboratory, Oak Ridge, TN 37831-6480, USA.

Abstract:
A new method for fold recognition is developed and added to the general protein structure prediction package PROSPECT (http://compbio.ornl.gov/PROSPECT/). The new method (PROSPECT II) has four key features. (i) We have developed an efficient way to utilize the evolutionary information for evaluating the threading potentials including singleton and pairwise energies. (ii) We have developed a two-stage threading strategy: (a) threading using dynamic programming without considering the pairwise energy and (b) fold recognition considering all the energy terms, including the pairwise energy calculated from the dynamic programming threading alignments. (iii) We have developed a combined z-score scheme for fold recognition, which takes into consideration the z-scores of each energy term. (iv) Based on the z-scores, we have developed a confidence index, which measures the reliability of a prediction and a possible structure-function relationship based on a statistical analysis of a large data set consisting of threadings of 600 query proteins against the entire FSSP templates. Tests on several benchmark sets indicate that the evolutionary information and other new features of PROSPECT II greatly improve the alignment accuracy. We also demonstrate that the performance of PROSPECT II on fold recognition is significantly better than any other method available at all levels of similarity. Improvement in the sensitivity of the fold recognition, especially at the superfamily and fold levels, makes PROSPECT II a reliable and fully automated protein structure and function prediction program for genome-scale applications.
More Related Videos
 06:50
Computational Prediction of Amino Acid Preferences of Potentially Multispecific Peptide-Binding Domains Involved in Protein-Protein Interactions
Published on: January 26, 2024
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to form...
 02:15
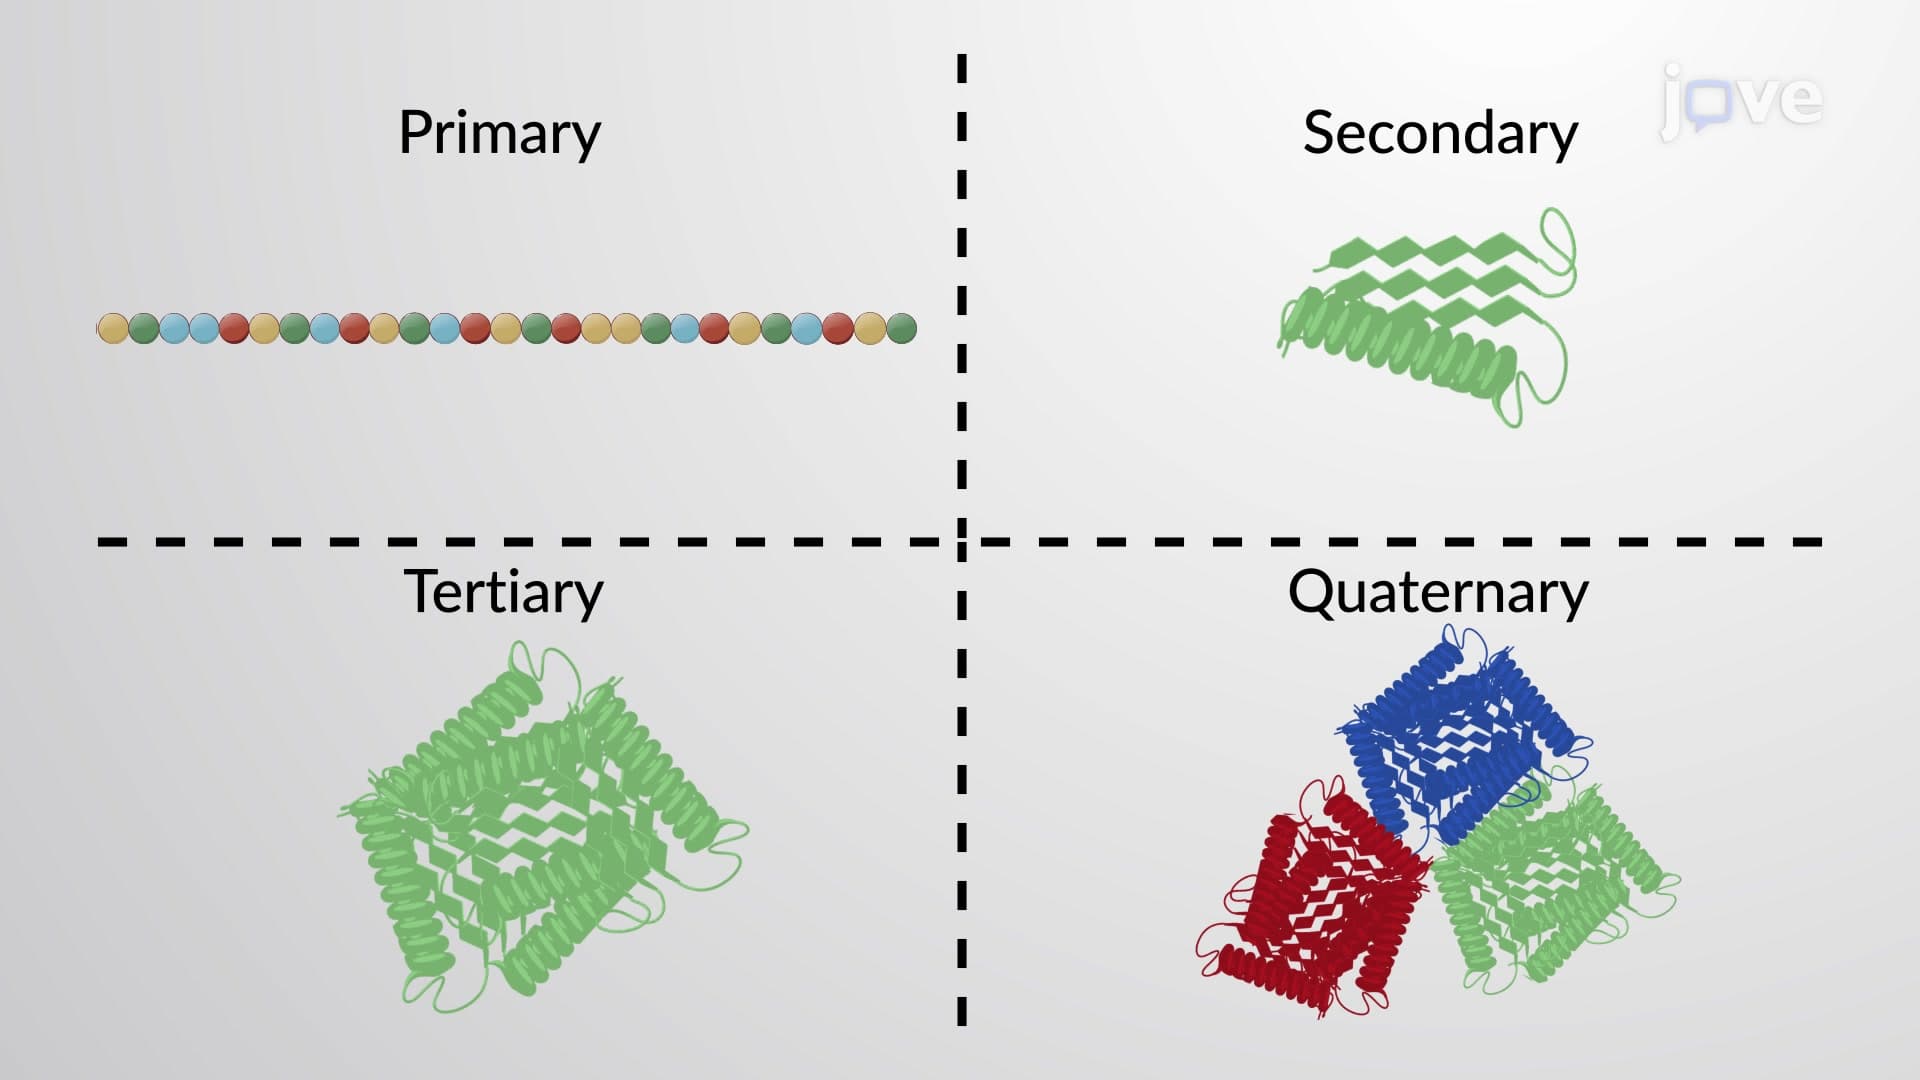
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme can...
 02:24
Ribosome Profiling
Applications of ribosome profiling
Ribosome profiling has many applications, including in vivo monitoring of translation inside a particular organ or tissue type and quantifying new protein synthesis levels.
The technique helps...
 02:04
Protein-protein Interfaces
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...