Related Experiment Video
Updated: Aug 29, 2026

Sampling Cerebrospinal Fluid and Blood from Lateral Tail Vein in Rats During EEG Recordings
Published on: September 1, 2023
Pharmacokinetic modelling of valproic acid from routine clinical data in Egyptian epileptic patients
Ehab S EL Desoky1, Eliane Fuseau, Salah EL Din Amry
1Pharmacology Department, Faculty of Medicine, Assiut University, Assiut, Egypt. ehegypt@yahoo.com

Objective:
This paper describes a developed pharmacokinetic model for the estimation of valproic acid (VPA) clearance (CL) calculated from routine clinical data taken from Egyptian epileptic patients.
Methods:
Retrospective clinical data from 81 adult and paediatric epileptic patients with one trough VPA serum concentration per patient were analysed using NONMEM to estimate drug CL and determine the influence of different covariates. A qualification group of 20 epileptic children (3-13 years old) was used to evaluate the final model.
Results:
The population CL as estimated by base model (no covariates) was 0.581 l h(-1) with inter-individual variability (C.V. %) of 17.4% and SD of residual error was 6.82 mg l(-1). Univariate selection and backward deletion of different covariates led to the development of the final regression model of CL as follows: CL(Lh-1) = 0.101 + 0.151 * CBZ + 0.000248 * VPADD + 0.0968 * age/20 + 0.0803 * INDI, in which CBZ indicates co-administration of carbamazepine, VPADD the daily dose of VPA and INDI uncontrolled epilepsy. The between-subject variability in CL was 23.6% while the standard deviation of the residual error was 5.24 mg l(-1). The model predictions in the qualification group were found to have no bias and satisfactory precision.
Conclusion:
The population pharmacokinetic model for VPA could be used for a priori recommendation and dose optimisation of that drug in the Egyptian population of epileptic patients.
Related Concept Videos
 01:20
Pharmacokinetic Models: Overview
There are three primary types of models: empirical, compartment, and physiological. Empirical models, with minimal assumptions,...
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model
 01:22
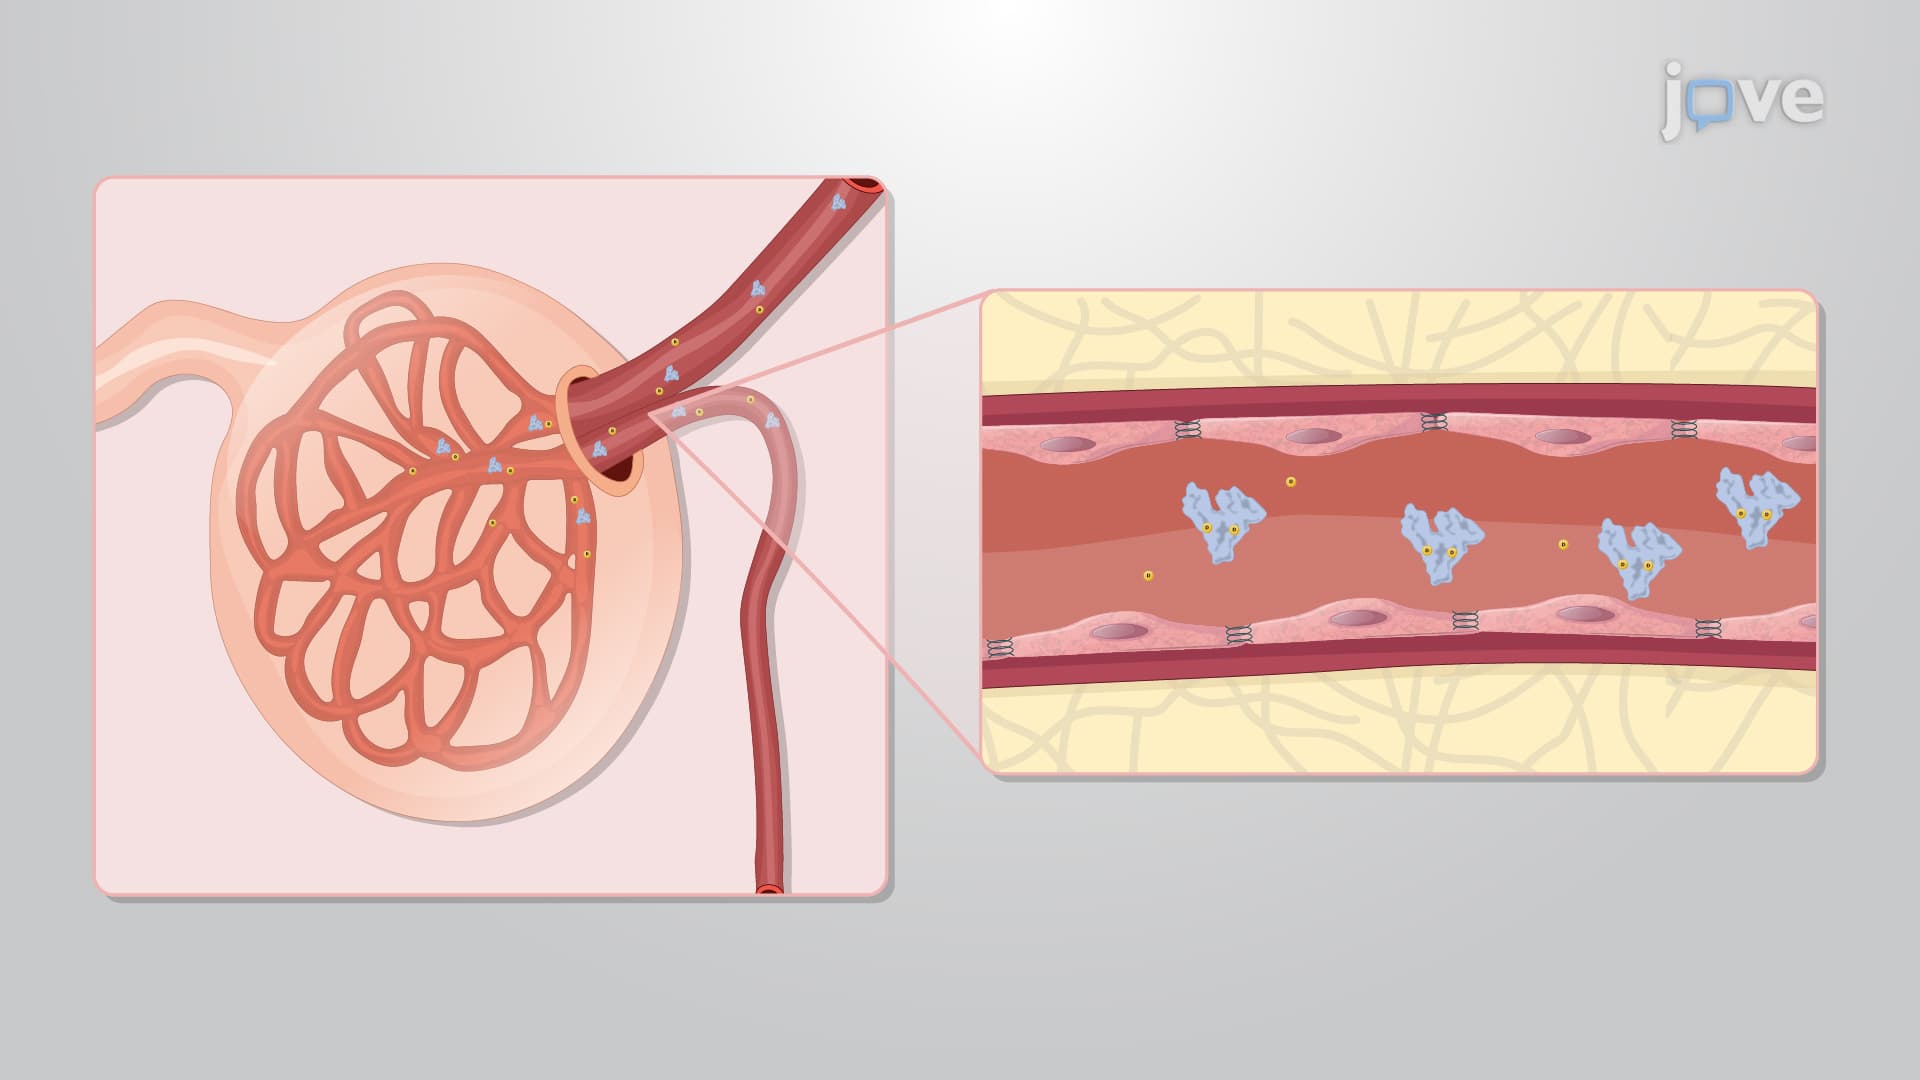
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:01
Dosage Regimens: Partial Pharmacokinetic Parameters
 01:26
Pharmacokinetic Models: Comparison and Selection Criterion
Physiological models take a detailed approach by considering specific molecular processes. They can predict drug distribution, metabolism, and elimination changes, providing a comprehensive understanding of how drugs interact with the body.