Protein structure prediction using sparse dipolar coupling data
Related Experiment Video
Updated: Aug 29, 2026

A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
Youxing Qu1, Jun-tao Guo, Victor Olman
1Computational Systems Biology Laboratory, Department of Biochemistry and Molecular Biology, University of Georgia, Athens, GA 30602, USA.

Abstract:
Residual dipolar coupling (RDC) represents one of the most exciting emerging NMR techniques for protein structure studies. However, solving a protein structure using RDC data alone is still a highly challenging problem. We report here a computer program, RDC-PROSPECT, for protein structure prediction based on a structural homolog or analog of the target protein in the Protein Data Bank (PDB), which best aligns with the (15)N-(1)H RDC data of the protein recorded in a single ordering medium. Since RDC-PROSPECT uses only RDC data and predicted secondary structure information, its performance is virtually independent of sequence similarity between a target protein and its structural homolog/analog, making it applicable to protein targets beyond the scope of current protein threading techniques. We have tested RDC-PROSPECT on all (15)N-(1)H RDC data (representing 43 proteins) deposited in the BioMagResBank (BMRB) database. The program correctly identified structural folds for 83.7% of the target proteins, and achieved an average alignment accuracy of 98.1% residues within a four-residue shift.
More Related Videos
 14:55
Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
 06:50
Computational Prediction of Amino Acid Preferences of Potentially Multispecific Peptide-Binding Domains Involved in Protein-Protein Interactions
Published on: January 26, 2024
Related Concept Videos
 02:36
Molecular Geometry and Dipole Moments
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene π orbitals.
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are slanted or...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:27
Predicting Molecular Geometry
 01:22
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
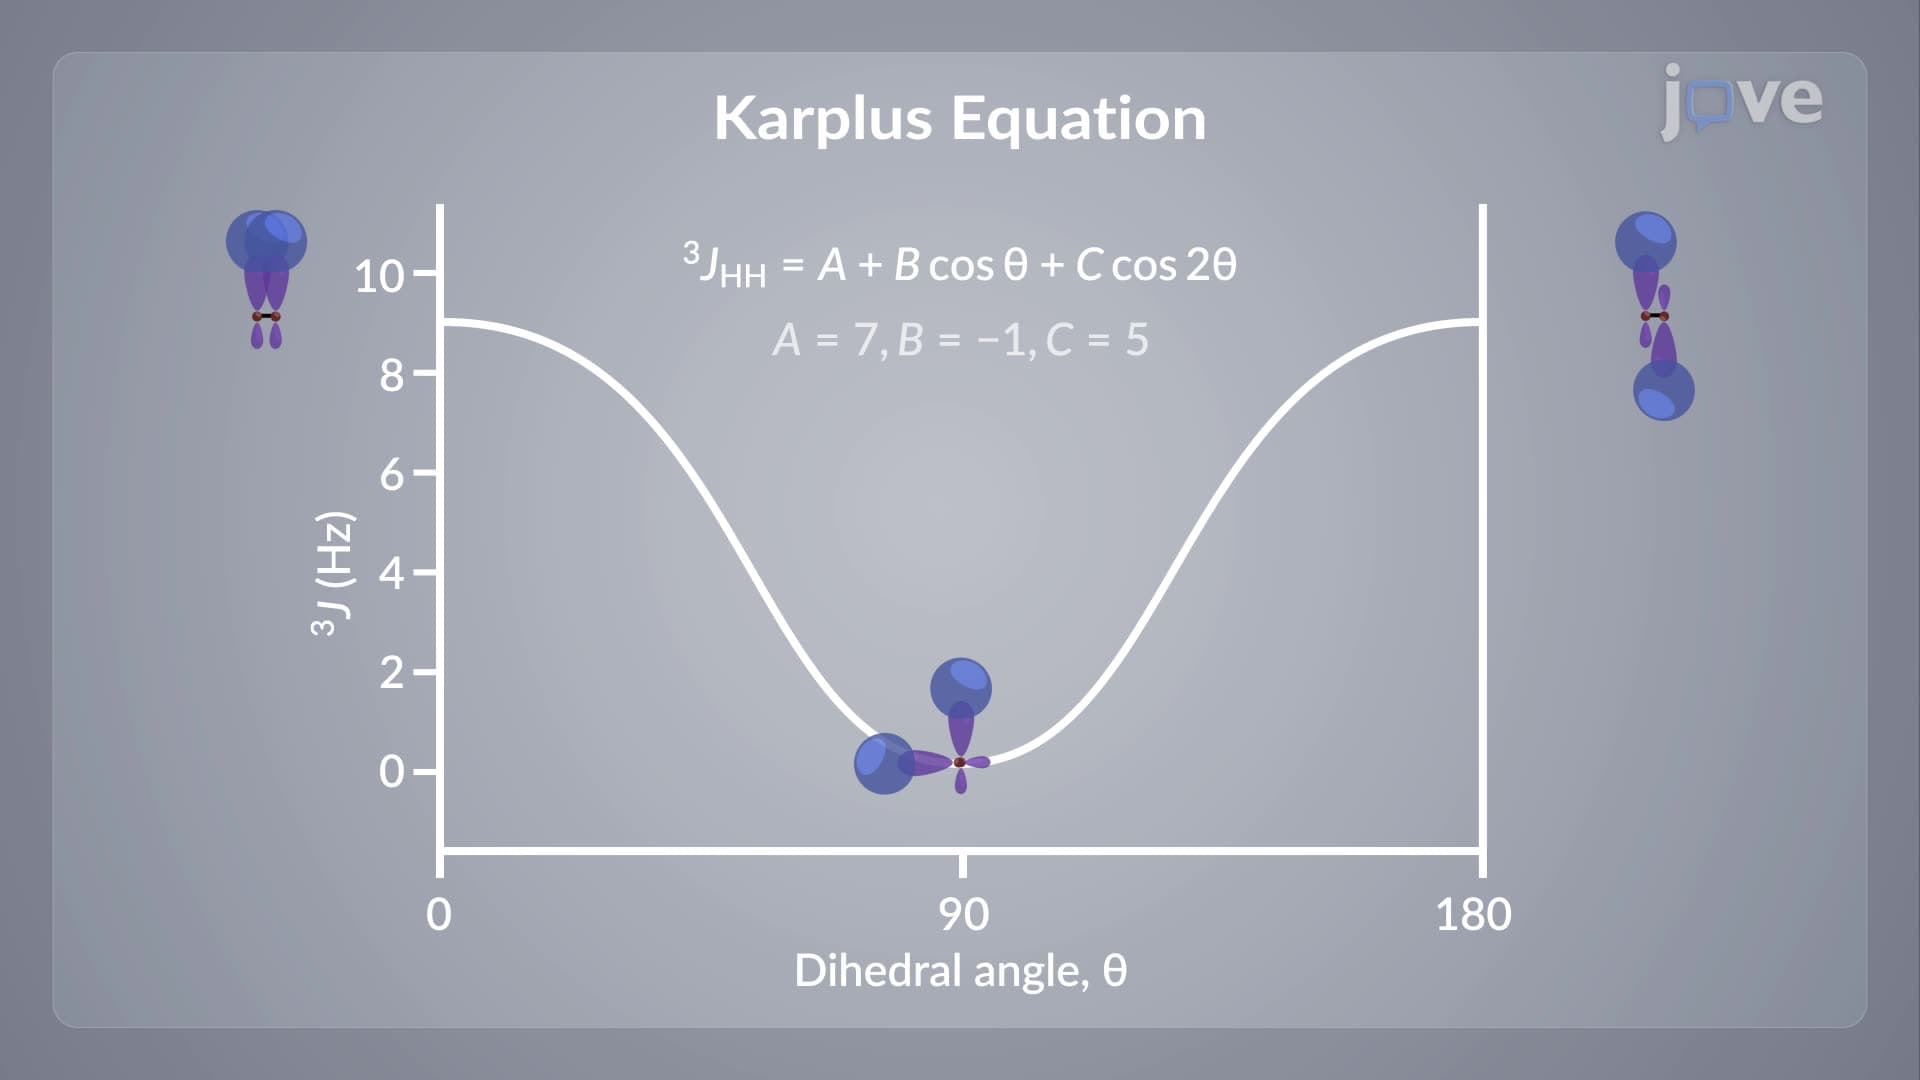
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...