Conformer- and alignment-independent model for predicting structurally diverse competitive CYP2C9 inhibitors
Related Experiment Video
Updated: Aug 29, 2026
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Lovisa Afzelius1, Ismael Zamora, Collen M Masimirembwa
1DMPK and Bioanalytical Chemistry, AstraZeneca R&D, S-431 83 Mölndal, Sweden. Afzelius@astrazeneca.com

Abstract:
A conformer- and alignment-independent three-dimensional structure-activity relationship (3D-QSAR) model has been derived that is based on flexible molecular interaction fields calculated in GRID and the subsequent description of these fields by use of alignment-independent descriptors derived in ALMOND. The training set consisted of 22 diverse and flexible competitive inhibitors of the drug-metabolizing enzyme CYP2C9 and generated a model with r(2) of 0.81 and q(2) of 0.62. The predicitive capacity of the model was externally evaluated with a test set of 12 competitive inhibitors and 11 out of 12 were predicted within 0.5 log unit. The most relevant points of interaction in the model correlated well to the amino acids involved in CYP2C9-substrate/inhibitor binding in the active site of a CYP2C9 homology model, further validating the mechanistic sense of our model. This approach offers the possibility to derive predicitve 3D-QSAR models without the need for an alignment rule for chemically diverse ligands and in the absence of target protein crystal structure information.
More Related Videos
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:26
Pharmacokinetic Models: Comparison and Selection Criterion
Physiological models take a detailed approach by considering specific molecular processes. They can predict drug distribution, metabolism, and elimination changes, providing a comprehensive understanding of how drugs interact with the body.
 01:16
Protein-Drug Binding: Mechanism and Kinetics
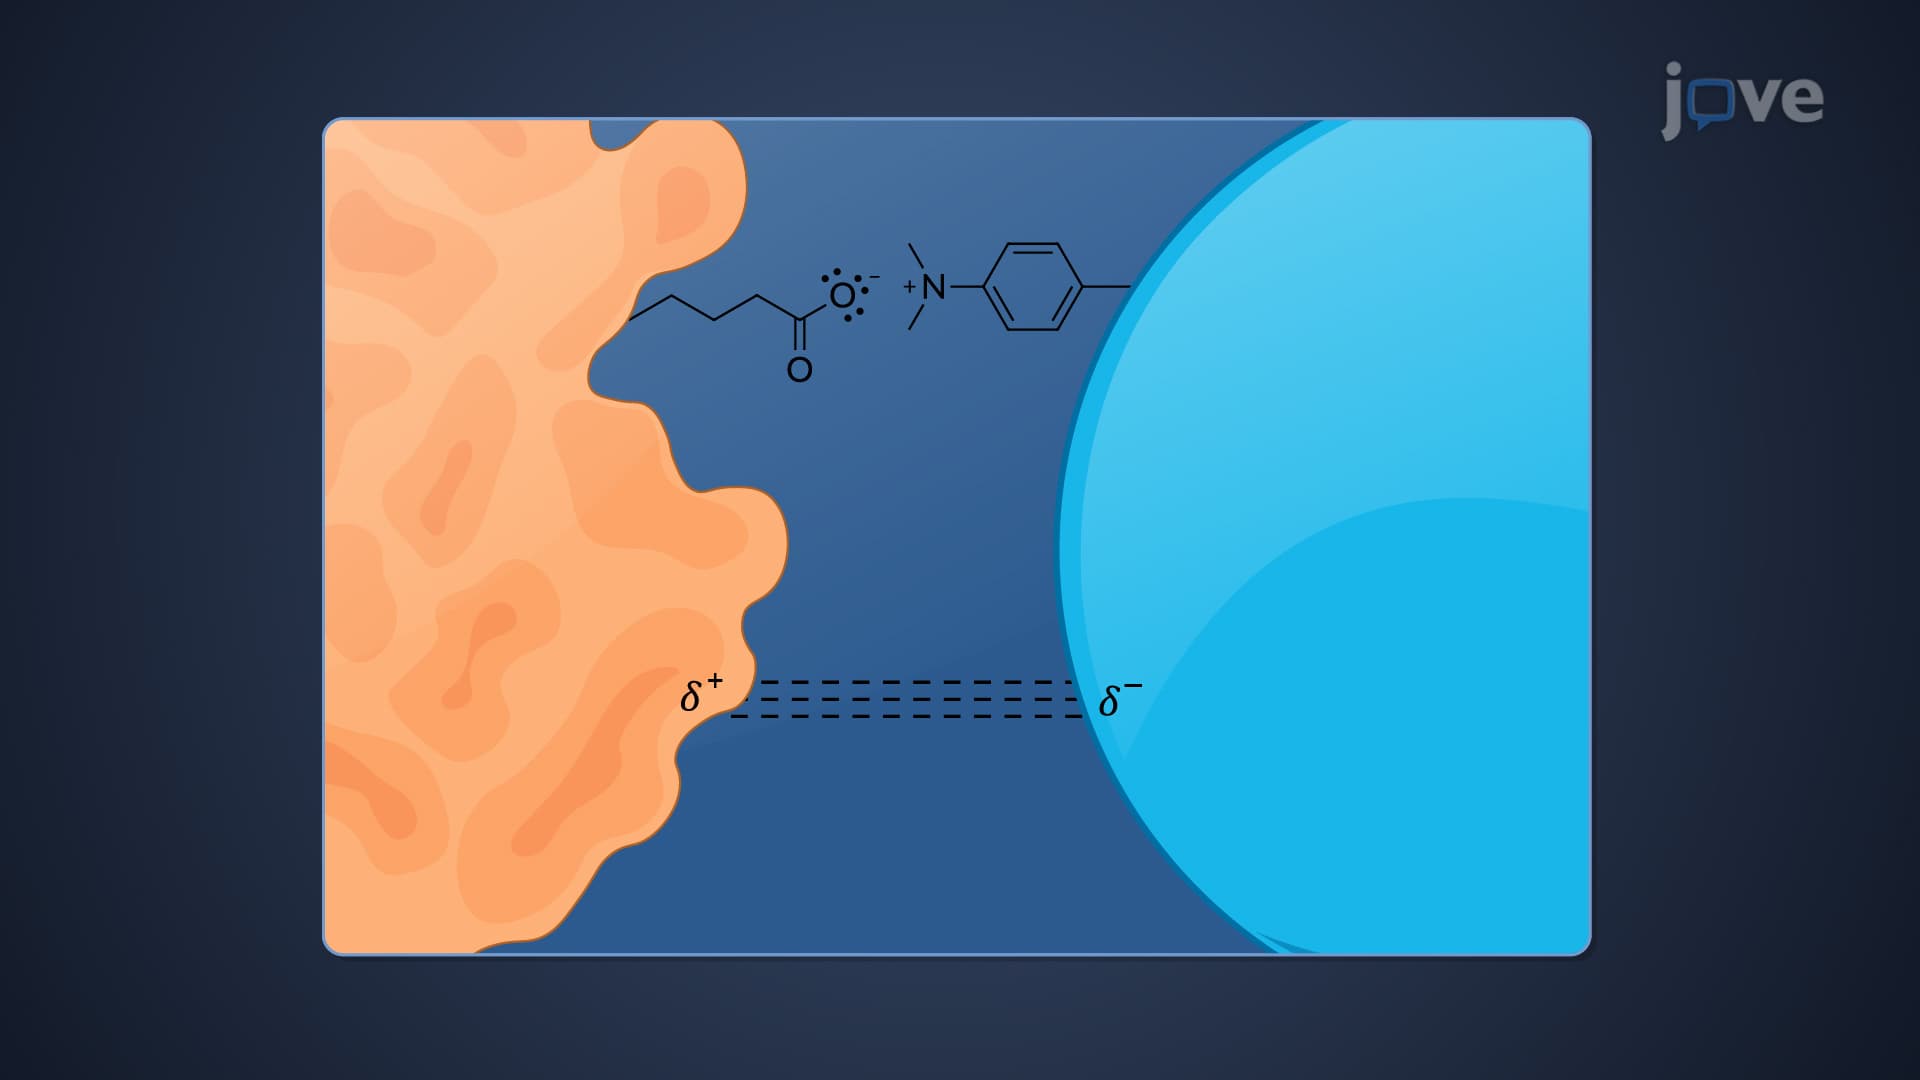
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:58
Cooperative Allosteric Transitions