
A conformation-specific interhelical salt bridge in the K+ binding site of gastric H,K-ATPase
Jan B Koenderink1, Herman G P Swarts, Peter H G M Willems
1Department of Biochemistry, Nijmegen Center for Molecular Life Sciences, University of Nijmegen, Nijmegen, The Netherlands.

Abstract:
Homology modeling of gastric H,K-ATPase based on the E2 model of sarcoplasmic reticulum Ca2+-ATPase (Toyoshima, C., and Nomura, H. (2002) Nature 392, 835-839) revealed the presence of a single high-affinity binding site for K+ and an E2 form-specific salt bridge between Glu820 (M6) and Lys791 (M5). In the E820Q mutant this salt bridge is no longer possible, and the head group of Lys791, together with a water molecule, fills the position of the K+ ion and apparently mimics the K+-filled cation binding pocket. This gives an explanation for the K+-independent ATPase activity and dephosphorylation step of the E820Q mutant (Swarts, H. G. P., Hermsen, H. P. H., Koenderink, J. B., Schuurmans Stekhoven, F. M. A. H., and De Pont, J. J. H. H. M. (1998) EMBO J. 17, 3029-3035) and, indirectly, for its E1 preference. The model is strongly supported by a series of reported mutagenesis studies on charged and polar amino acid residues in the membrane domain. To further test this model, Lys791 was mutated alone and in combination with other crucial residues. In the K791A mutant, the K+ affinity was markedly reduced without altering the E2 preference of the enzyme. The K791A mutation prevented, in contrast to the K791R mutation, the spontaneous dephosphorylation of the E820Q mutant as well as its conformational equilibrium change toward E1. This indicates that the salt bridge is essential for high-affinity K+ binding and the E2 preference of H,K-ATPase. Moreover, its breakage (E820Q) can generate a K+-insensitive activity and an E1 preference. In addition, the study gives a molecular explanation for the electroneutrality of H,K-ATPases.
More Related Videos
 10:33
Development of Inhibitors of Protein-protein Interactions through REPLACE: Application to the Design and Development Non-ATP Competitive CDK Inhibitors
Published on: October 26, 2015
 12:07
Chemical Modification of the Tryptophan Residue in a Recombinant Ca2+-ATPase N-domain for Studying Tryptophan-ANS FRET
Published on: October 9, 2021
Related Concept Videos
 01:30
ATP Driven Pumps III: V-type Pumps
The peripheral or cytosolic V1 domain with eight subunits is involved in ATP hydrolysis. The integral or transmembrane V0 domain containing at least five subunits...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis pathway,...
 01:34
ATP Driven Pumps II: P-type Pumps
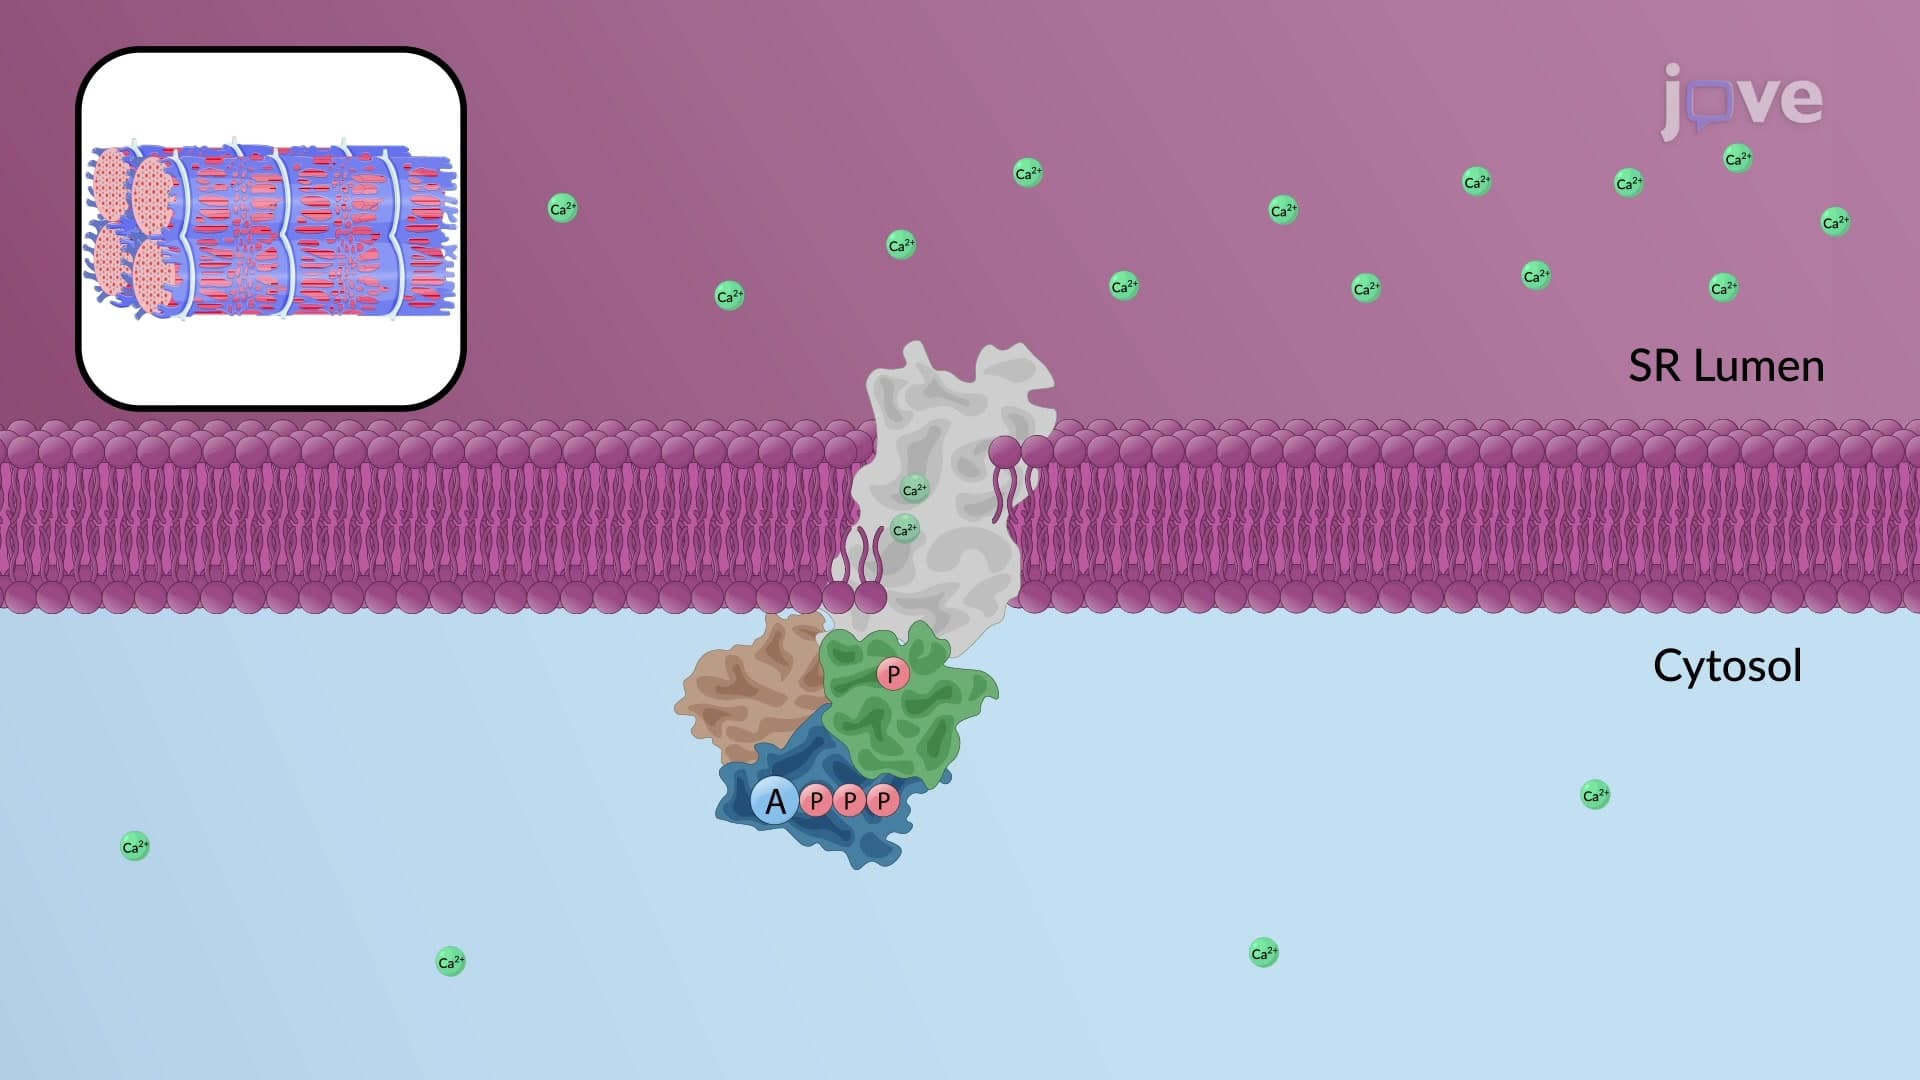
A typical P-type pump has three cytosolic domains: nucleotide-binding (N), phosphorylation (P), and activator (A) domains. These domains are connected to the membrane-spanning helices by short amino acid segments. ATP hydrolysis and covalent phosphoenzyme intermediate formation are crucial parts of the catalytic cycle. At the highly...
 02:29
Relative Strengths of Conjugate Acid-Base Pairs
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...