Related Experiment Video
Updated: Aug 29, 2026
A Practical Guide to Phylogenetics for Nonexperts
Published on: February 5, 2014
Walking tree heuristics for comparative genomic alignments
Jeffrey D Cavener1, Paul Cull, James L Holloway
1Computer Science, Oregon State University, Corvallis, OR 97339, USA. wt@cavener.com

Abstract:
Genomic sequence data is available for an ever-increasing number of organisms, but the full meaning of this data remains an enigma. String alignment is one approach for deciphering the information contained in genetic strings. Sequences which are conserved across species will help identify genes and other important structures. Similarity between species can be scored by measuring how well their sequences align. The walking tree method is an approximate string alignment method that can handle insertions, deletions, substitutions, translocations, and more than one level of inversion. We will describe this method and recent improvements which allow fast alignment of megabase strings. We will show examples in which the method located or discovered genes. We show how the method can be used to construct phylogenetic trees. We also show that the method can be used to identify essential regions for protein function.
Related Concept Videos
 02:54
Evolutionary Relationships through Genome Comparisons
 01:28
Microbial Phylogeny
 03:21
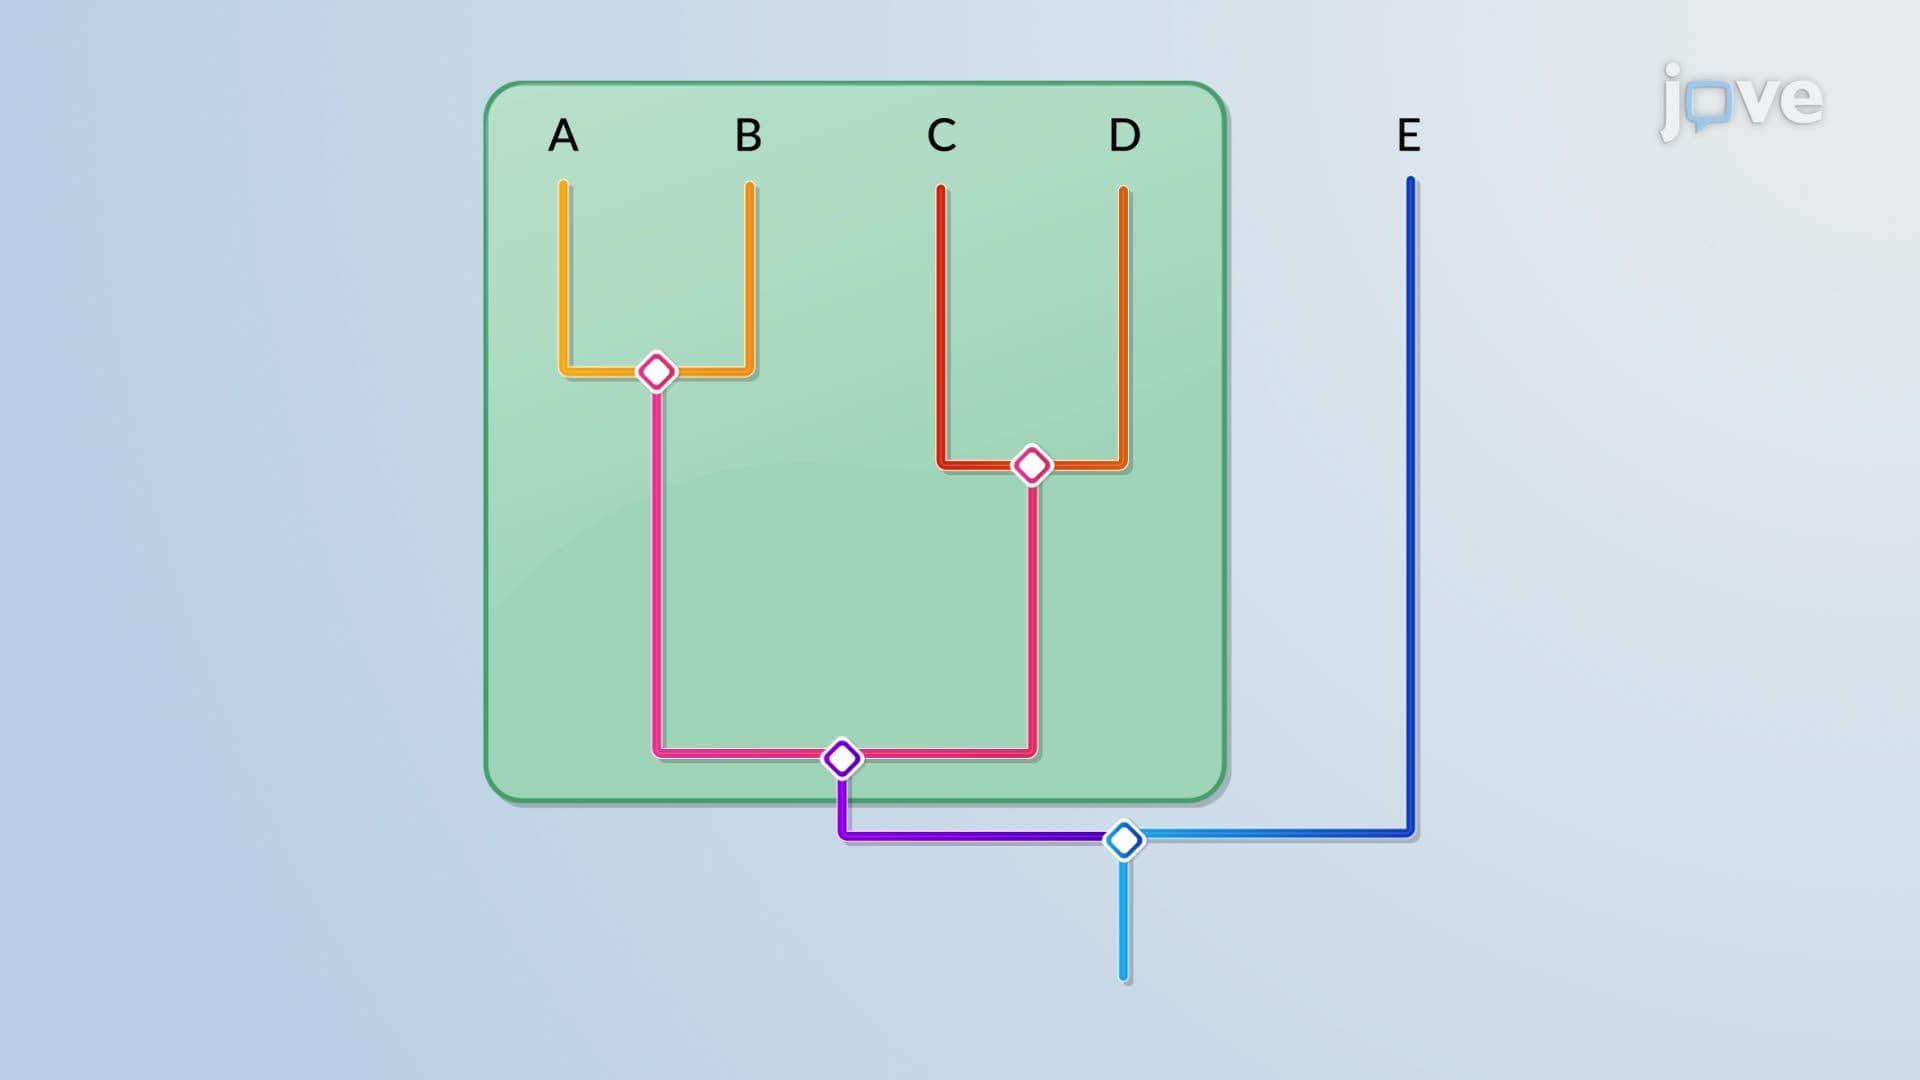
Phylogenetic Trees
03:21Phylogenetic Trees
 02:16
Comparing Mitochondrial, Chloroplast, and Prokaryotic Genomes
 02:26
Comparing Copy Number Variations and SNPs
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...