Mu-heavy chain disease associated with systemic amyloidosis

Koji Kinoshita1, Toshiaki Yamagata, Yuji Nozaki

  • 1Department of Hematology, Nephrology and Rheumatology, Kinki Univeristy School of Medicine, Osaka-Sayama, Osaka, Japan.

Related Concept Videos

Amyloid Fibrils03:03

Amyloid Fibrils

Amyloid fibrils are aggregates of misfolded proteins.  Under most circumstances, misfolded proteins are either refolded by chaperone proteins or degraded by the proteasome. However, in the case of a mutation or a disease, these proteins can accumulate to form large clusters and often further assemble to form elongated fibers, called fibrils. 
Amyloid deposits were observed as early as 1639 in the liver and the spleen.   In 1854, Rudolph Virchow performed iodine staining, normally used to...
Amyloid Fibrils03:03

Amyloid Fibrils

Amyloid fibrils are aggregates of misfolded proteins.  Under most circumstances, misfolded proteins are either refolded by chaperone proteins or degraded by the proteasome. However, in the case of a mutation or a disease, these proteins can accumulate to form large clusters and often further assemble to form elongated fibers, called fibrils. 
Amyloid deposits were observed as early as 1639 in the liver and the spleen.   In 1854, Rudolph Virchow performed iodine staining, normally used to...
Cardiomyopathy III: Hypertrophic Cardiomyopathy01:29

Cardiomyopathy III: Hypertrophic Cardiomyopathy

Hypertrophic cardiomyopathy, or HCM, is an autosomal dominant genetic disorder characterized by asymmetric left ventricular hypertrophy without ventricular dilation. It is more common in men and is typically diagnosed in young, athletic adults.EtiologyHCM is primarily genetic and is caused by mutations in genes encoding sarcomeric proteins. Researchers have identified over 1400 mutations across at least 11 different genes. Among these, the most frequently occurring mutations are found in the...
Huntington Disease l: Introduction01:21

Huntington Disease l: Introduction

Huntington disease or HD is a progressive, fatal neurodegenerative disorder inherited in an autosomal dominant pattern.PathophysiologyIt is caused by expansion of the CAG trinucleotide repeat in the HTT gene on chromosome 4 (4p16.3), producing an abnormal huntingtin protein with an expanded polyglutamine tract. This misfolded protein disrupts cellular function, leading to neuronal death. Normal alleles have ≤26 repeats, 27–35 are intermediate (risk of expansion), 36–39 show reduced penetrance,...
Cardiomyopathy IV: Restrictive Cardiomyopathy01:29

Cardiomyopathy IV: Restrictive Cardiomyopathy
Restrictive cardiomyopathy (RCM) is a rare heart muscle disease characterized by impaired ventricular filling due to stiffened ventricular walls, leading to significant diastolic dysfunction.EtiologyRestrictive cardiomyopathy can arise from both inherited and acquired diseases, many of which are systemic. It is categorized into four main types: infiltrative, storage, non-infiltrative, and endomyocardial diseases.Infiltrative diseases, such as amyloidosis, lead to RCM by depositing amyloid...
Alzheimer Disease ll: Pathophysiology01:23

Alzheimer Disease ll: Pathophysiology

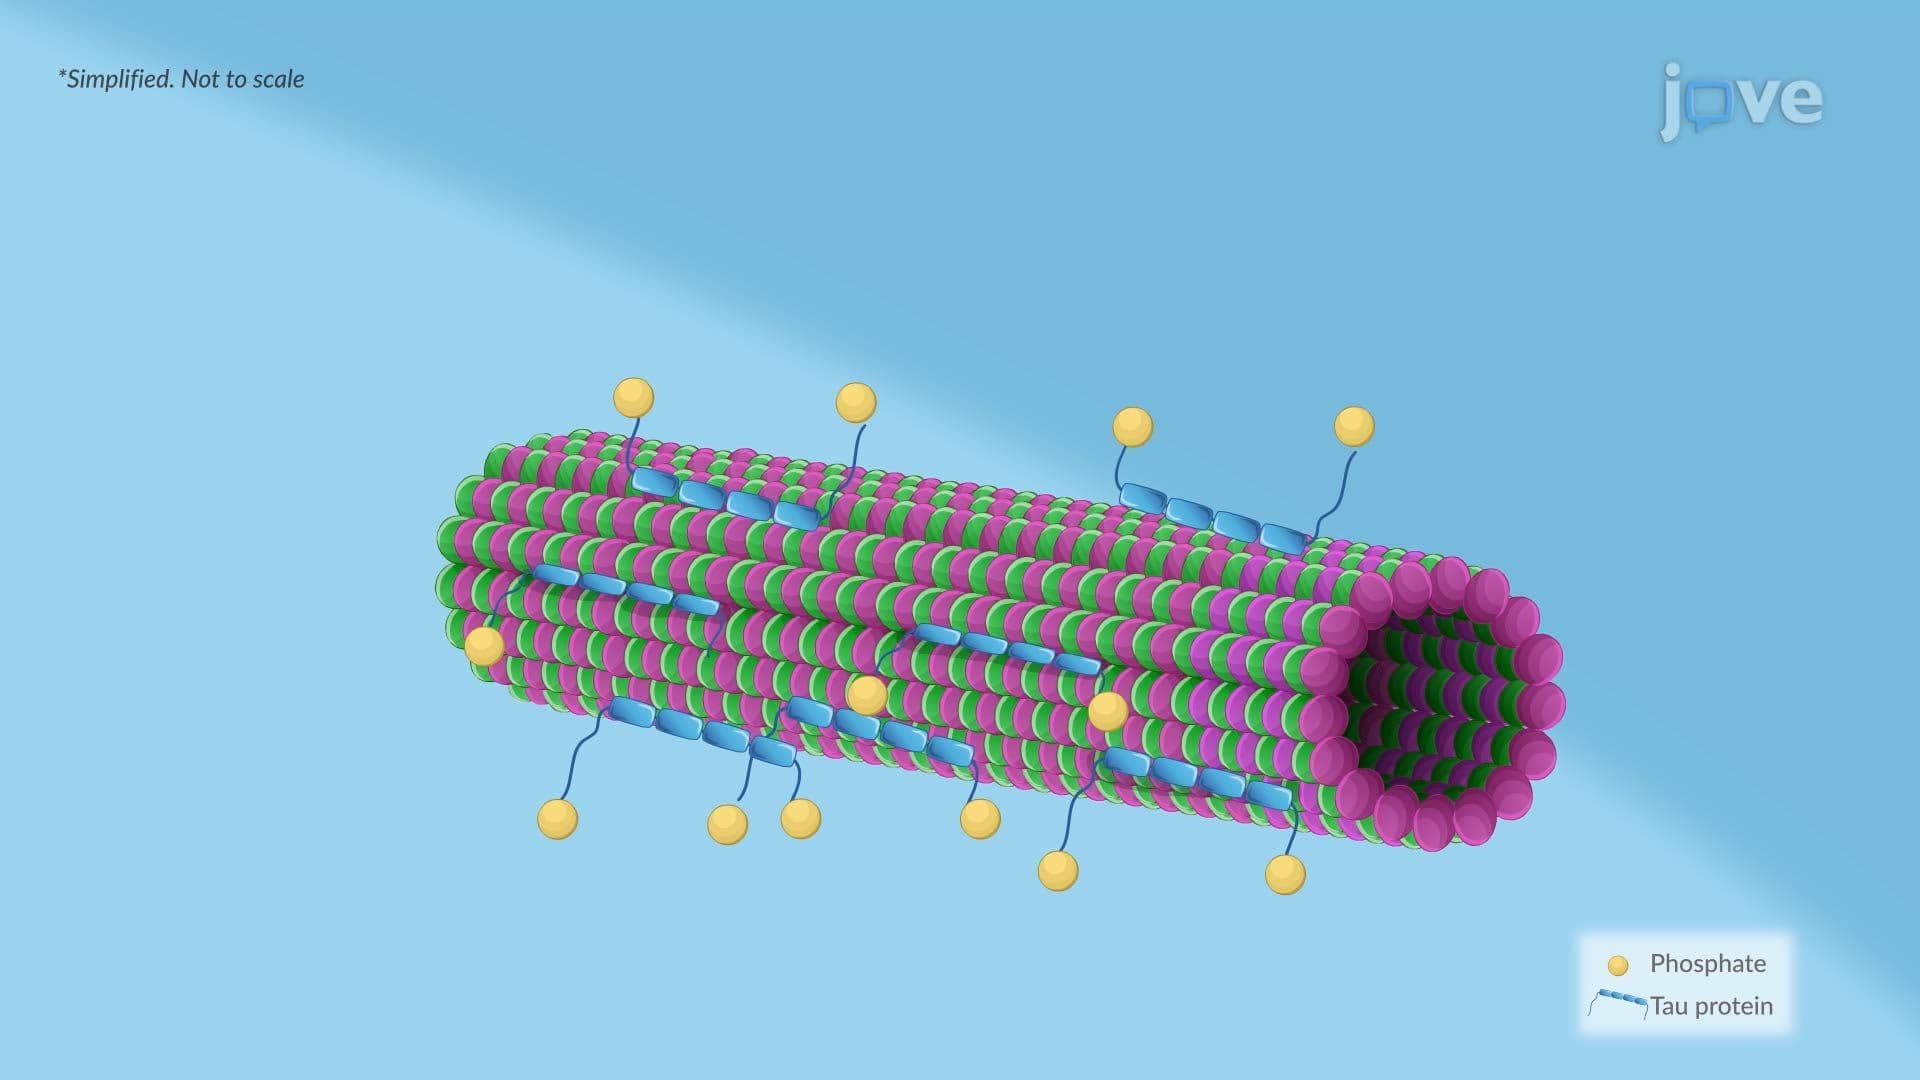
Alzheimer disease involves structural changes in the brain that begin long before symptoms appear. The most distinctive features are extracellular neuritic plaques and intracellular neurofibrillary tangles.Neuritic plaques form in the cerebral cortex and around blood vessels. These plaques contain a dense core of beta-amyloid (Aβ)—a toxic protein fragment that clumps outside neurons. The core is surrounded by damaged neuronal extensions, as well as reactive astrocytes and microglia. Abnormal...