Related Experiment Video
Updated: Aug 23, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Theoretical calculations of effective exchange integrals by spin projected and unprojected broken-symmetry methods.
Taku Onishi1, Kizashi Yamaguchi
1Department of Chemistry, Graduate School of Science, Osaka University, Toyonaka, 560-0043, Japan.

Abstract:
Previously, we have performed the spin-polarized hybrid-density functional theory (HUDFT) calculations for elucidating magnetic properties of the two-dimensional (2D) K2NiF4 and K2CuF4 solids. In Part I, it has been concluded that the half-and-half-(HH-) type HUDFT method is one of the best calculation methods for these species. On the other hand, in Part II, we have demonstrated that potential curves for cluster models of K2CuF4 and KCuF3 are reasonably calculated by the HH-type HUDFT method under the approximate spin projection, and the lattice distortion resulted by Jahn-Teller effect is expressed as the second-order polynomial. In this study, we pay attention to the three-dimensional (3D) magnetic interactions in KNiF3. Our effective exchange integral Jab schemes for 3D cluster models such as KNi8F12 (4) with and without point charges have provided the reasonable Jab values (-30.24-34.48 cm(-1)), in comparison with the experimental one (-30.58 cm(-1)). The 3D magnetic interactions have been investigated from viewpoints of the Mulliken spin density and charge density populations, the natural orbital analysis, and chemical indices. Point charges located in positions of fluorine anion adjacent to cluster model have reproduced Jab values well. Roles of potassium as counter cation in KNiF3 solid were also investigated. It was concluded that potassium has a role of stabilizing the 3D magnetic structures. Finally, the mutual relationships between broken-symmetry and symmetry-adapted approaches are discussed on the basis of chemical indices.

Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:22
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
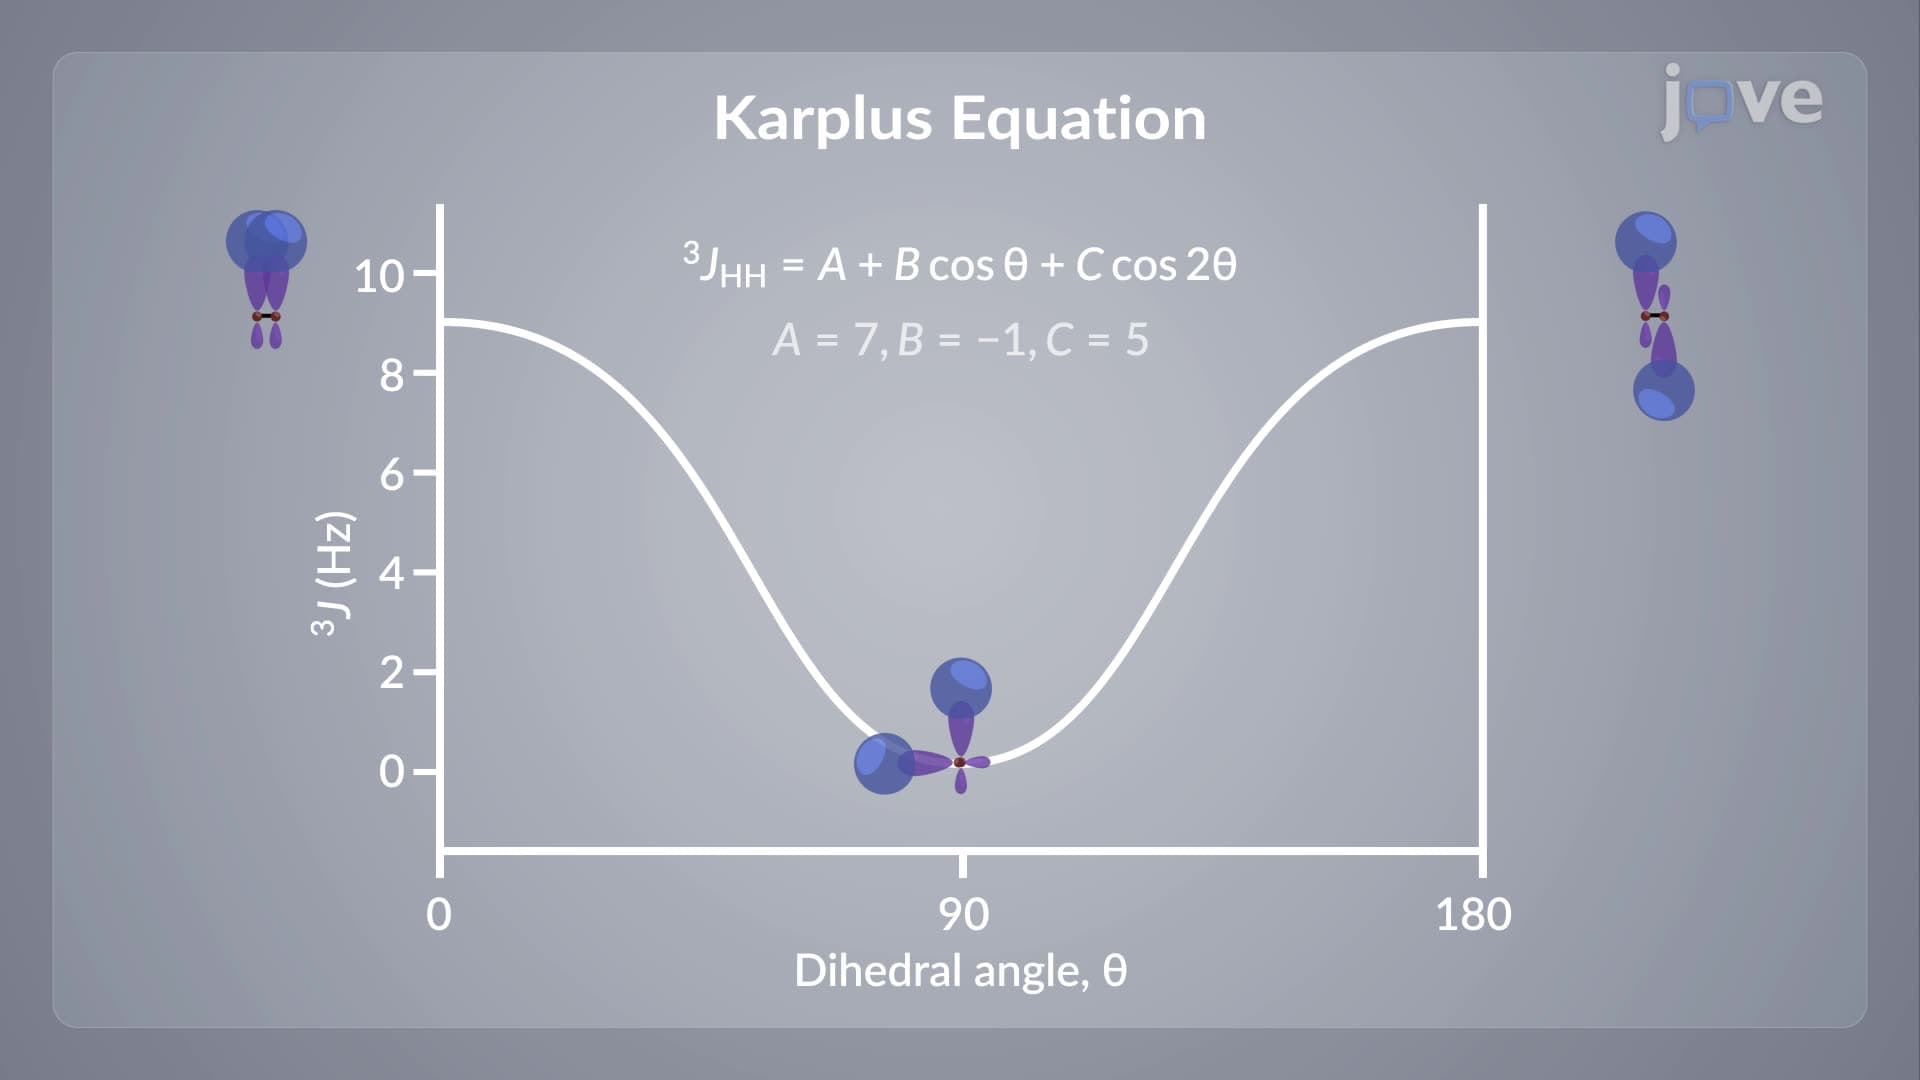
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...
 01:17
Spin–Spin Coupling: One-Bond Coupling
 02:18
Network Covalent Solids
To break or to melt a covalent network solid, covalent bonds must be broken. Because covalent bonds are relatively strong, covalent network solids are typically...
 01:20
Spin–Spin Coupling: Two-Bond Coupling (Geminal Coupling)
The central atom need not be NMR-active because its electrons are affected by the electron polarization of the spin-active atoms. However, spin information is transmitted less effectively than in one-bond coupling, and 2J values are usually weaker than 1J values. The energy of...