Distributed diffusion-clearance model for transient drug distribution within the skin
Related Experiment Video
Updated: Aug 22, 2026
A Method for Determination and Simulation of Permeability and Diffusion in a 3D Tissue Model in a Membrane Insert System for Multi-well Plates
Published on: February 23, 2018
Kosmas Kretsos1, Gerald B Kasting, Johannes M Nitsche
1Department of Chemical and Biological Engineering, State University of New York, University at Buffalo, Furnas Hall, Buffalo, New York 14260-4200, USA.

Abstract:
Quantitative predictions of molecular transport rates through the skin are key to the development of topically applied and transdermally delivered drugs, as well as risk assessment associated with dermal exposure. Most research to date has focused on correlations for the permeability of the stratum corneum, and transient diffusion models that oversimplify vascular clearance processes in terms of a perfect-removal boundary condition at an artificially introduced lower boundary. Considerations of the spatially distributed nature and action of blood vessels have usually been limited to the steady-state case. This article describes a more comprehensive transient model of percutaneous absorption formulated in terms of volumetric dispersion and clearance coefficients reflecting the spatial distribution of vascular processes. The model was implemented through an analysis of published experimental results on in vivo permeation of salicylic acid (SA) in de-epidermized rat skin. With regard to the characterization of SA in rat dermis ("de") in vivo, it was found that: (i) SA is likely to have a dermal effective partition coefficient (relative to pH 7.4 aqueous buffer "pH7.4") around unity (K(de/pH7.4) = 0.9 +/- 0.3); (ii) vascular processes seem not to increase drug dispersion significantly beyond molecular diffusion [D(de) approximately (D(de))(mol) = (8 +/- 3) . 10(-7) cm(2) s(-1)]; and (iii) vascular clearance is characterized by a rate coefficient k(de) = (7 +/- 2) . 10(-4) s(-1). Application of a whole-skin variant of the model (including the stratum corneum and viable epidermis) allowed realistic predictions to be made of transient subsurface concentration levels after application from a finite dose.
Related Concept Videos
 00:57
Physiological Pharmacokinetic Models: Blood Flow-Limited Versus Diffusion-Limited Models
 01:11
Drug Distribution: Overview
Initially, the free drug in the...
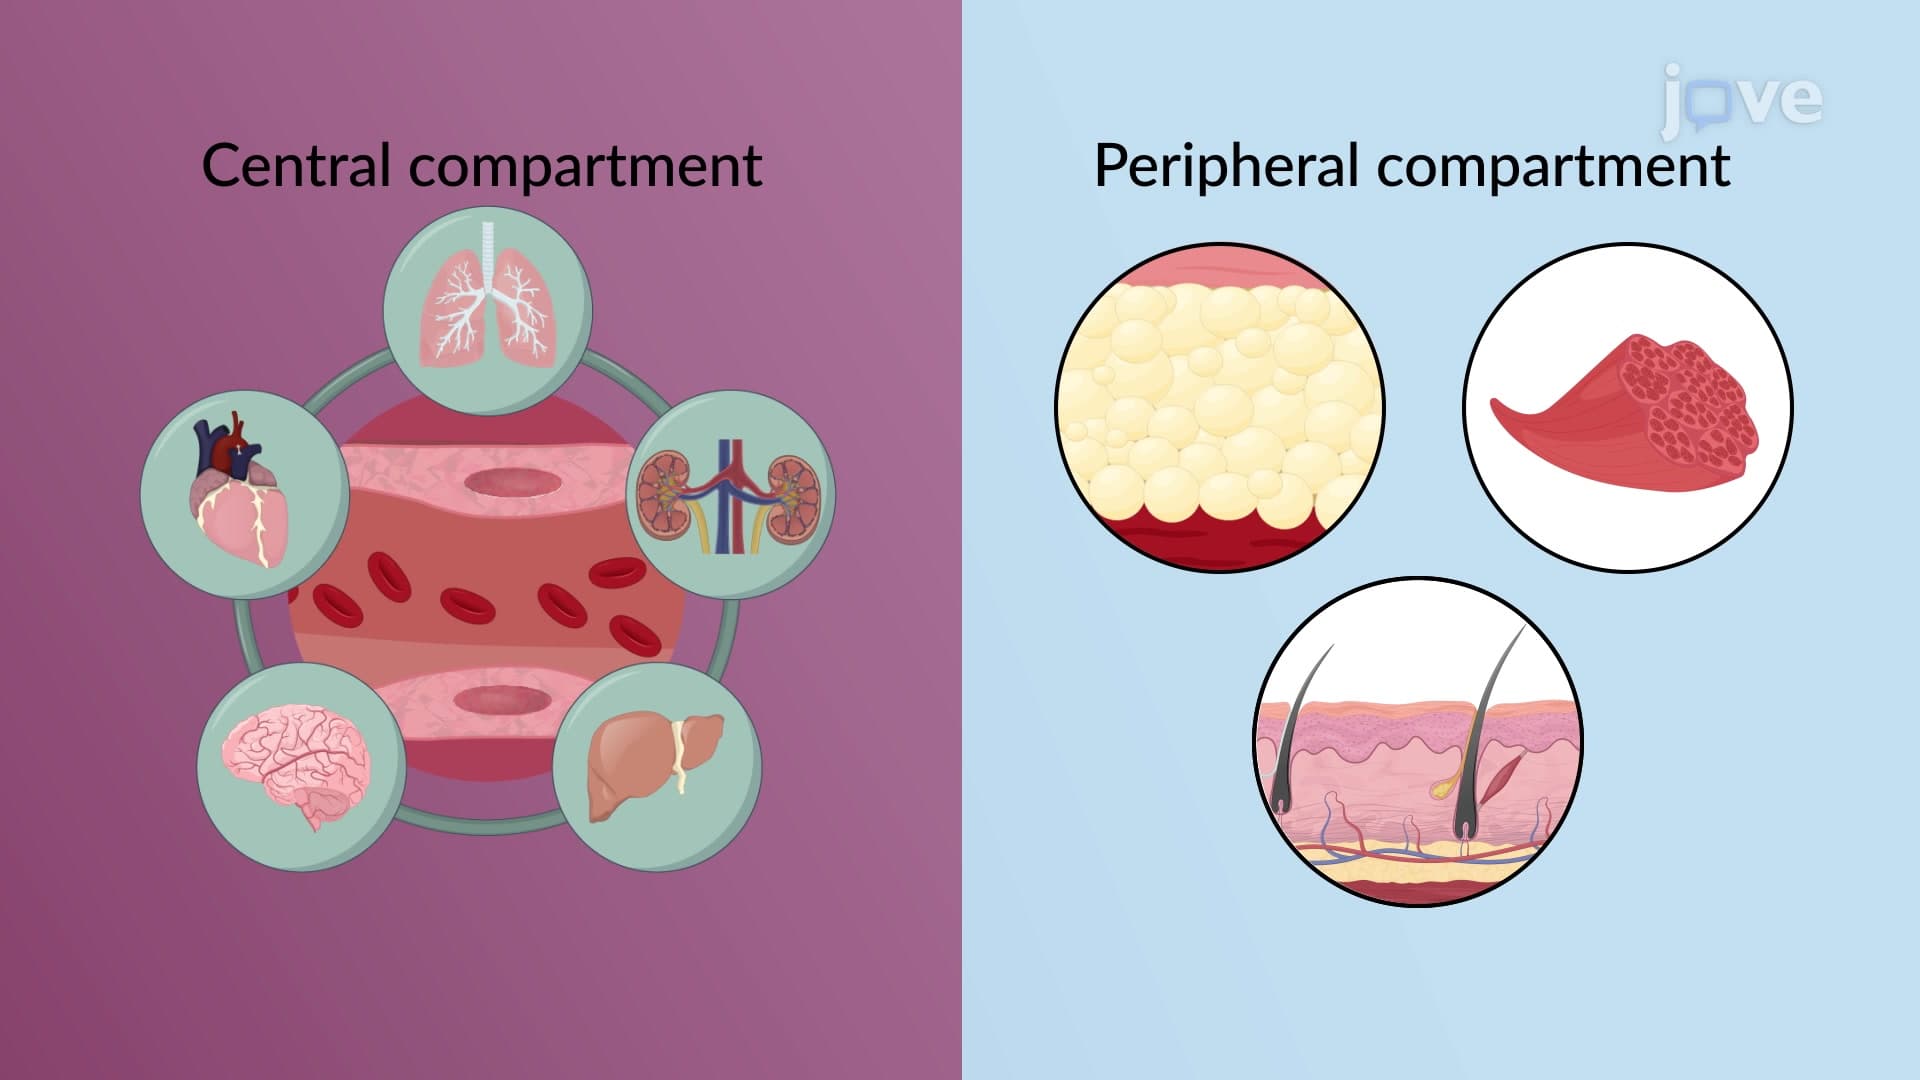 01:20
Compartment Models: Two-Compartment Model
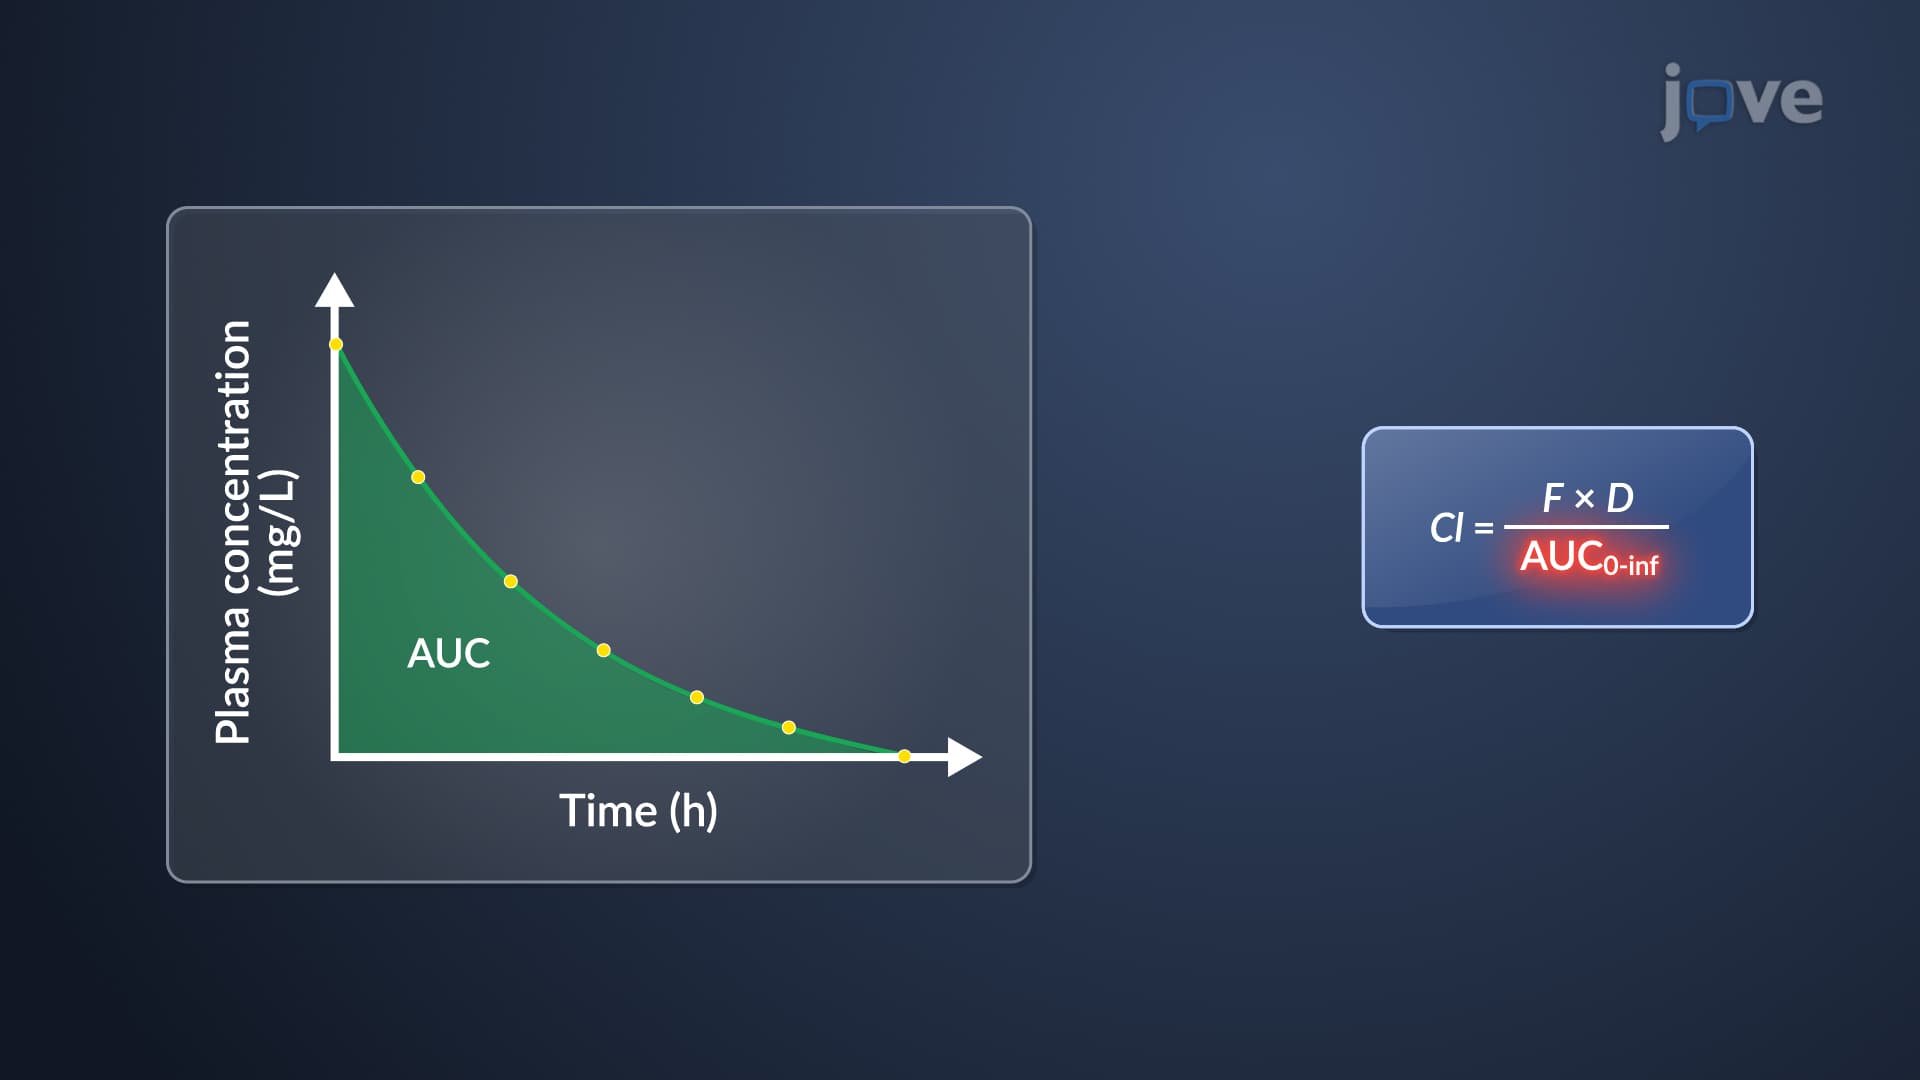 01:17
Clearance Models: Noncompartmental Models
The noncompartmental approach capitalizes on extensive sampling data, correlating the volume of distribution to systemic exposure and the administered dosage. This method enables...
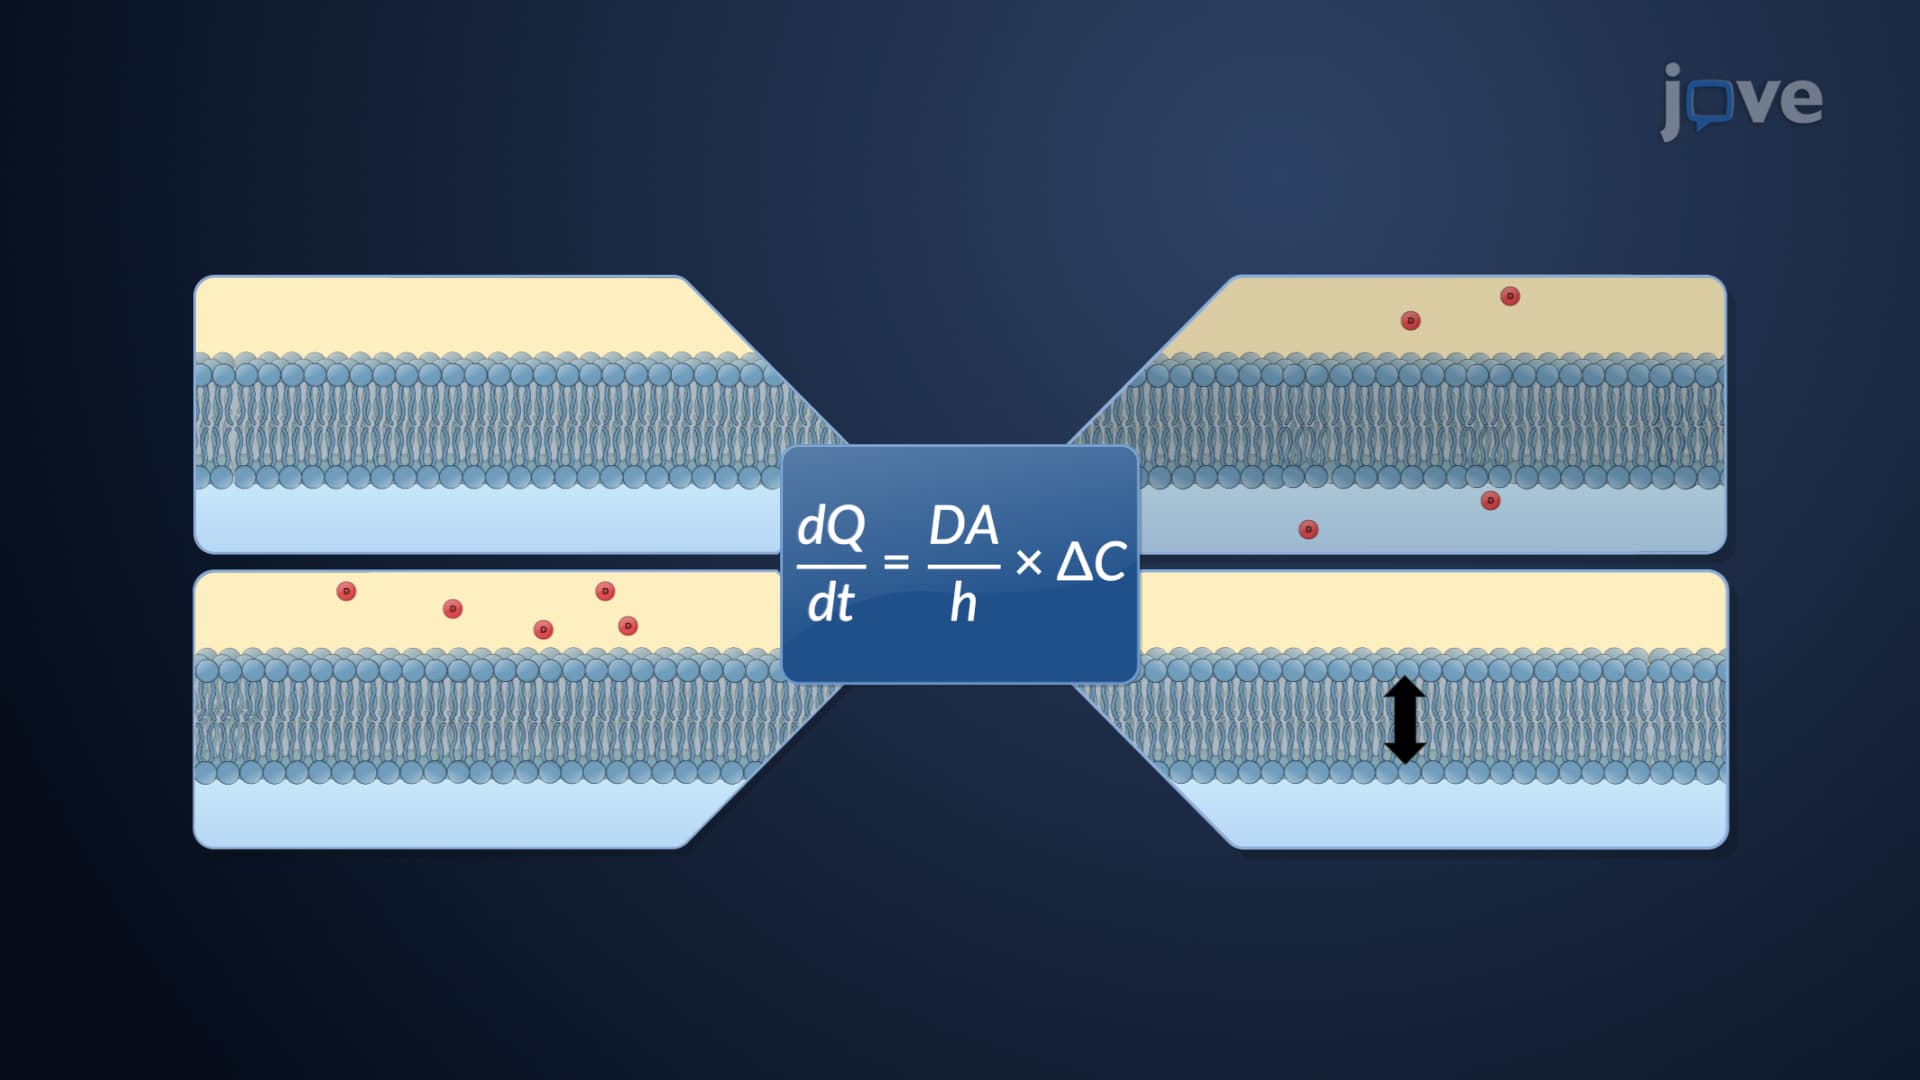 01:17
Passive Diffusion: Overview and Kinetics
When administered orally, drugs establish a substantial concentration gradient between the gastrointestinal (GI) lumen and the bloodstream, expediting their diffusion into...
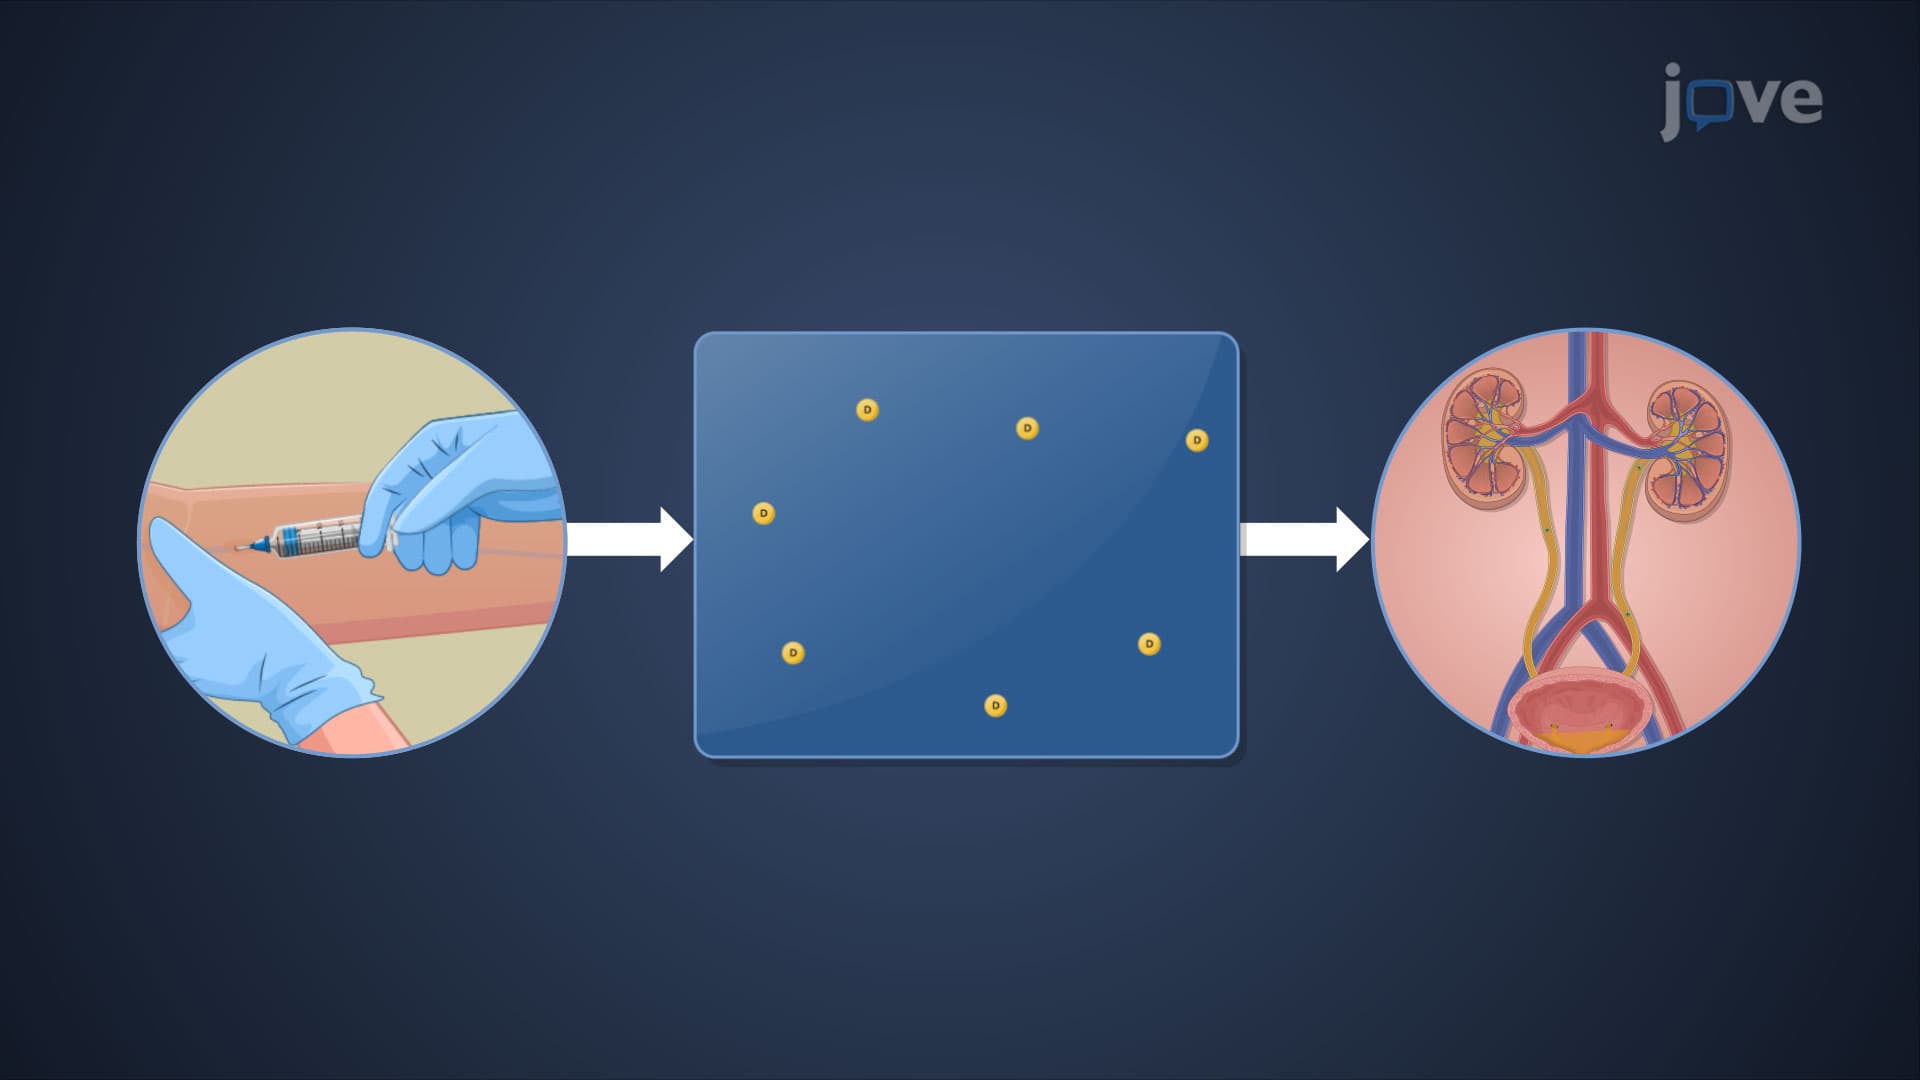 01:25
Clearance Models: Compartment Models