Related Experiment Video
Updated: Aug 14, 2026
Spatial Separation of Molecular Conformers and Clusters
Published on: January 9, 2014
Simple DFT model of clusters embedded in rare gas matrix: trapping sites and spectroscopic properties of Na embedded
B Gervais1, E Giglio, E Jacquet
1CIRIL, unité mixte CEA-CNRS-ENSICAEN et Université de Caen Basse-Normandie BP 5133, F-14070 Caen Cedex 05, France.

Abstract:
We present a theoretical model to study the dynamics of metallic clusters embedded in a rare gas matrix. We describe the active electrons of the embedded cluster using time dependent density functional theory, while the surrounding matrix is described in terms of classical molecular dynamics of polarizable atoms. The coupling between the cluster and the rare gas atoms is deduced from the work of Gross and Spiegelmann [J. Chem. Phys. 108, 4148 (1998)] and reformulated explicitly in a simple and efficient density functional form. The electron rare gas interaction takes the form of an averaged dipole fluctuation term, which retains the van der Waals long range interaction, and a short range repulsive pseudopotential, which accounts for the Pauli repulsion of the electron by the rare gas atom. We applied our model to Na clusters embedded in Ar matrix. For the latter we developed an efficient local pseudopotential, which allows studying systems containing more than 10(3) Ar atoms. We show that large systems are indeed necessary to account properly for long range polarization of the matrix, that competes with the matrix confinement effect. We focus our study on Na(2), Na(4), and Na(8). For each system, we have determined the geometry of the most favorable trapping site by means of damped molecular dynamics. We present the effect of matrix embedding on the optical absorption spectrum. For Na(2), the trapping site can be unambiguously identified by comparison of the absorption spectrum with experiment. For Na(4) the spectrum of the embedded cluster is significantly different from the free cluster spectrum, while for Na(8) differences are less pronounced.
More Related Videos

Related Concept Videos
 01:18
Electron Orbital Model
 02:44
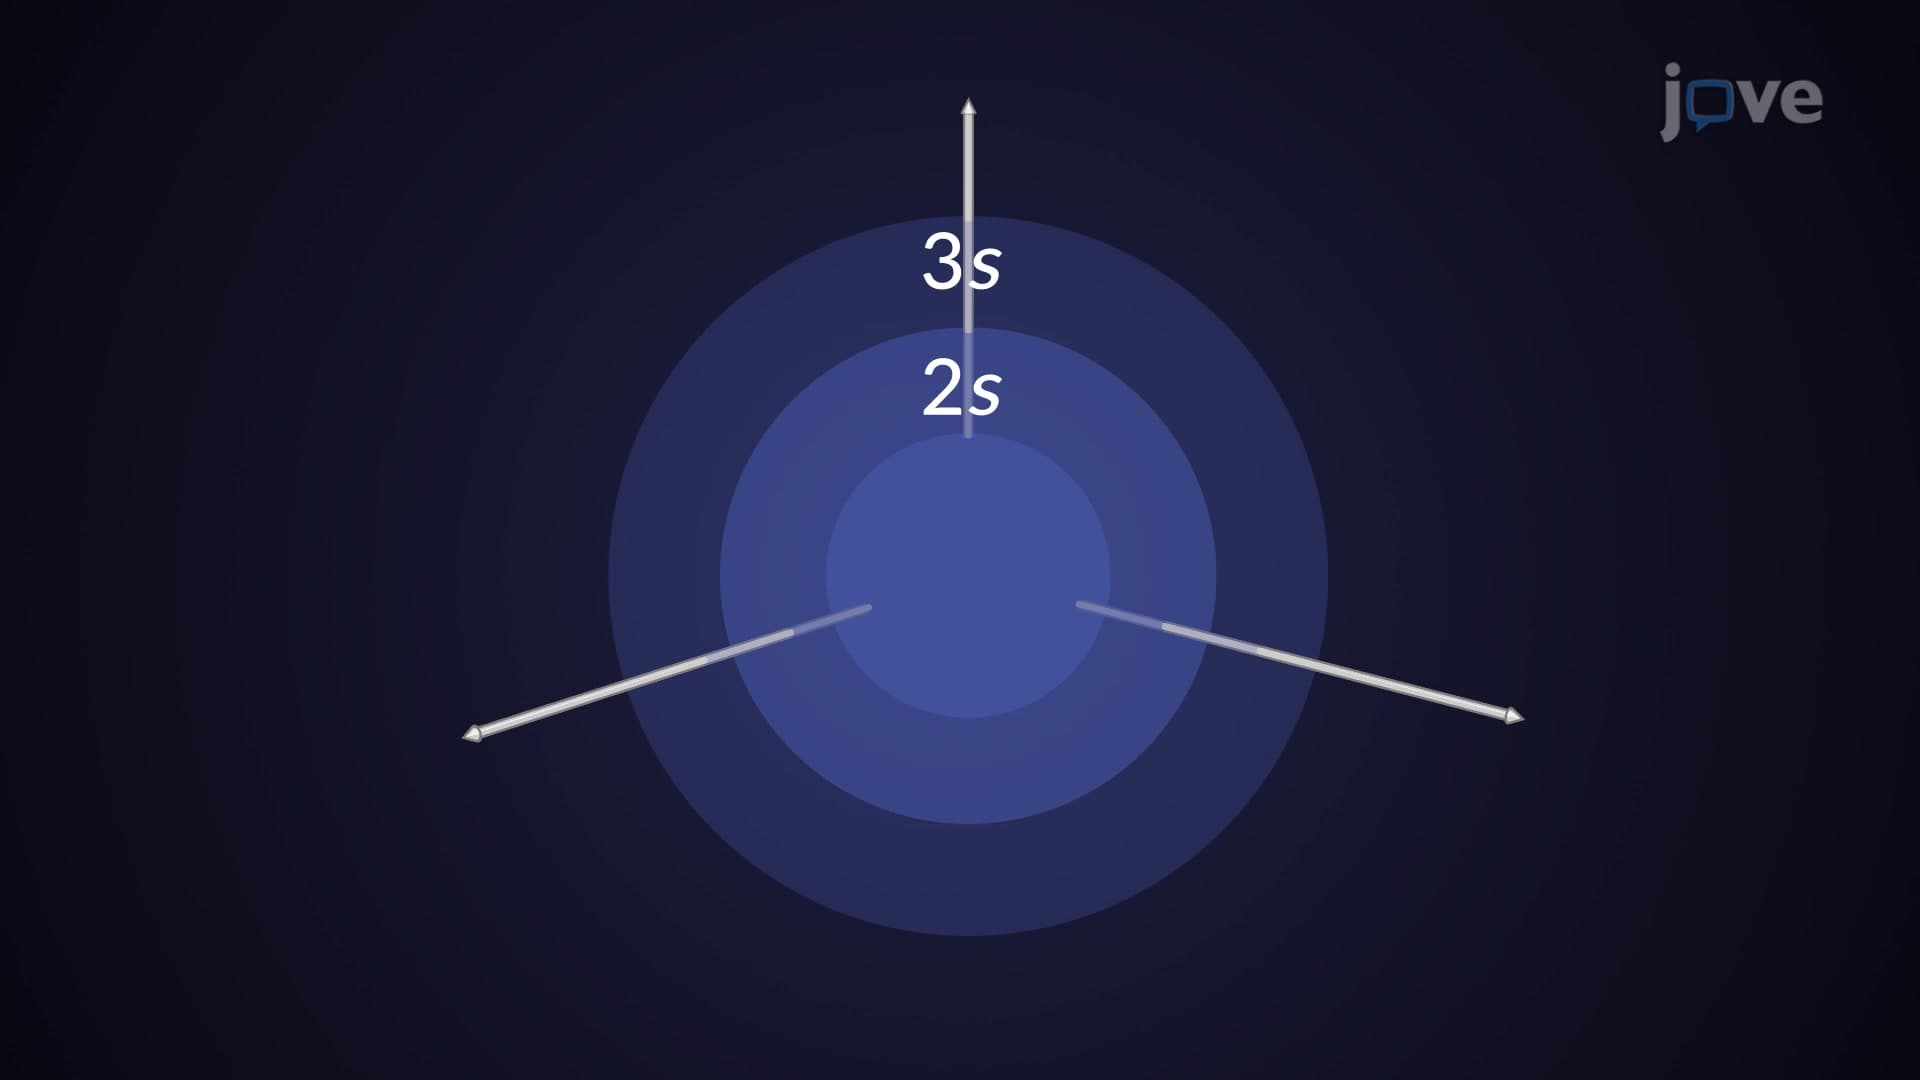
Atomic Orbitals
 03:26
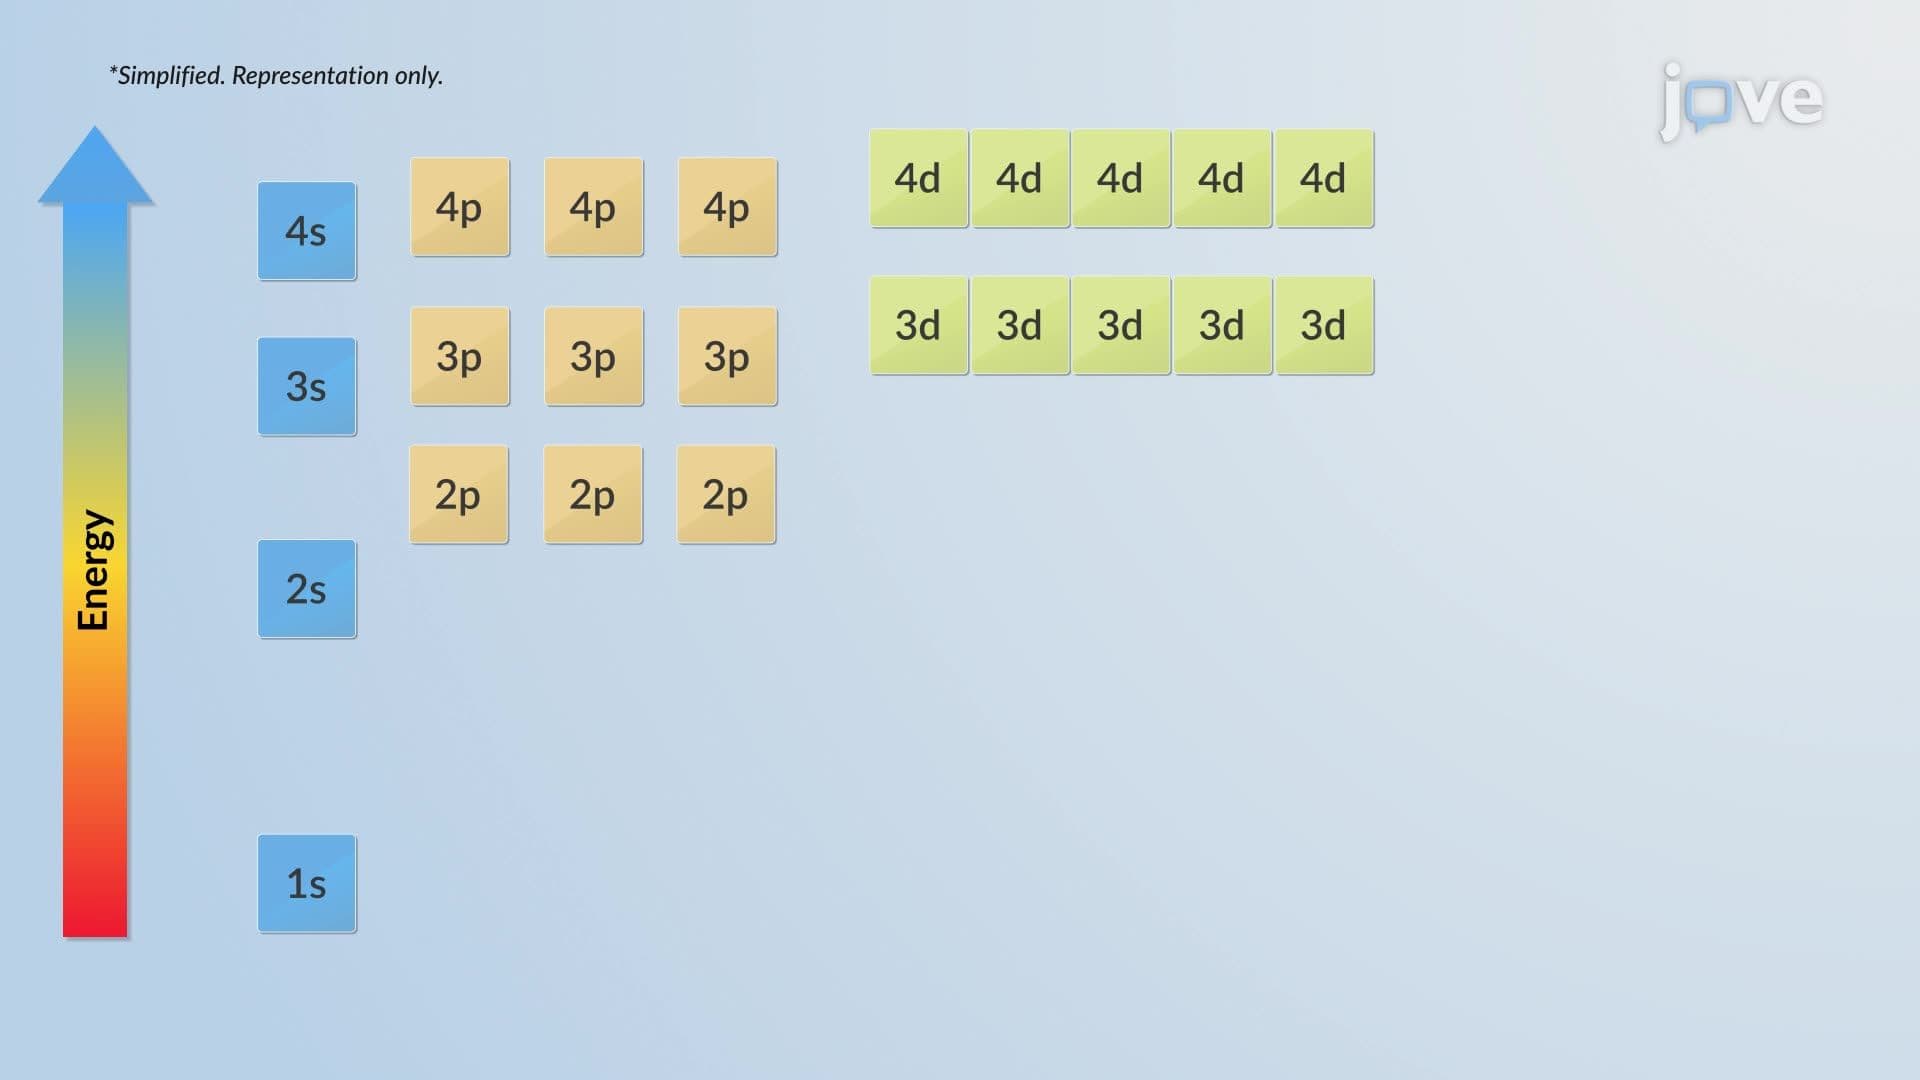
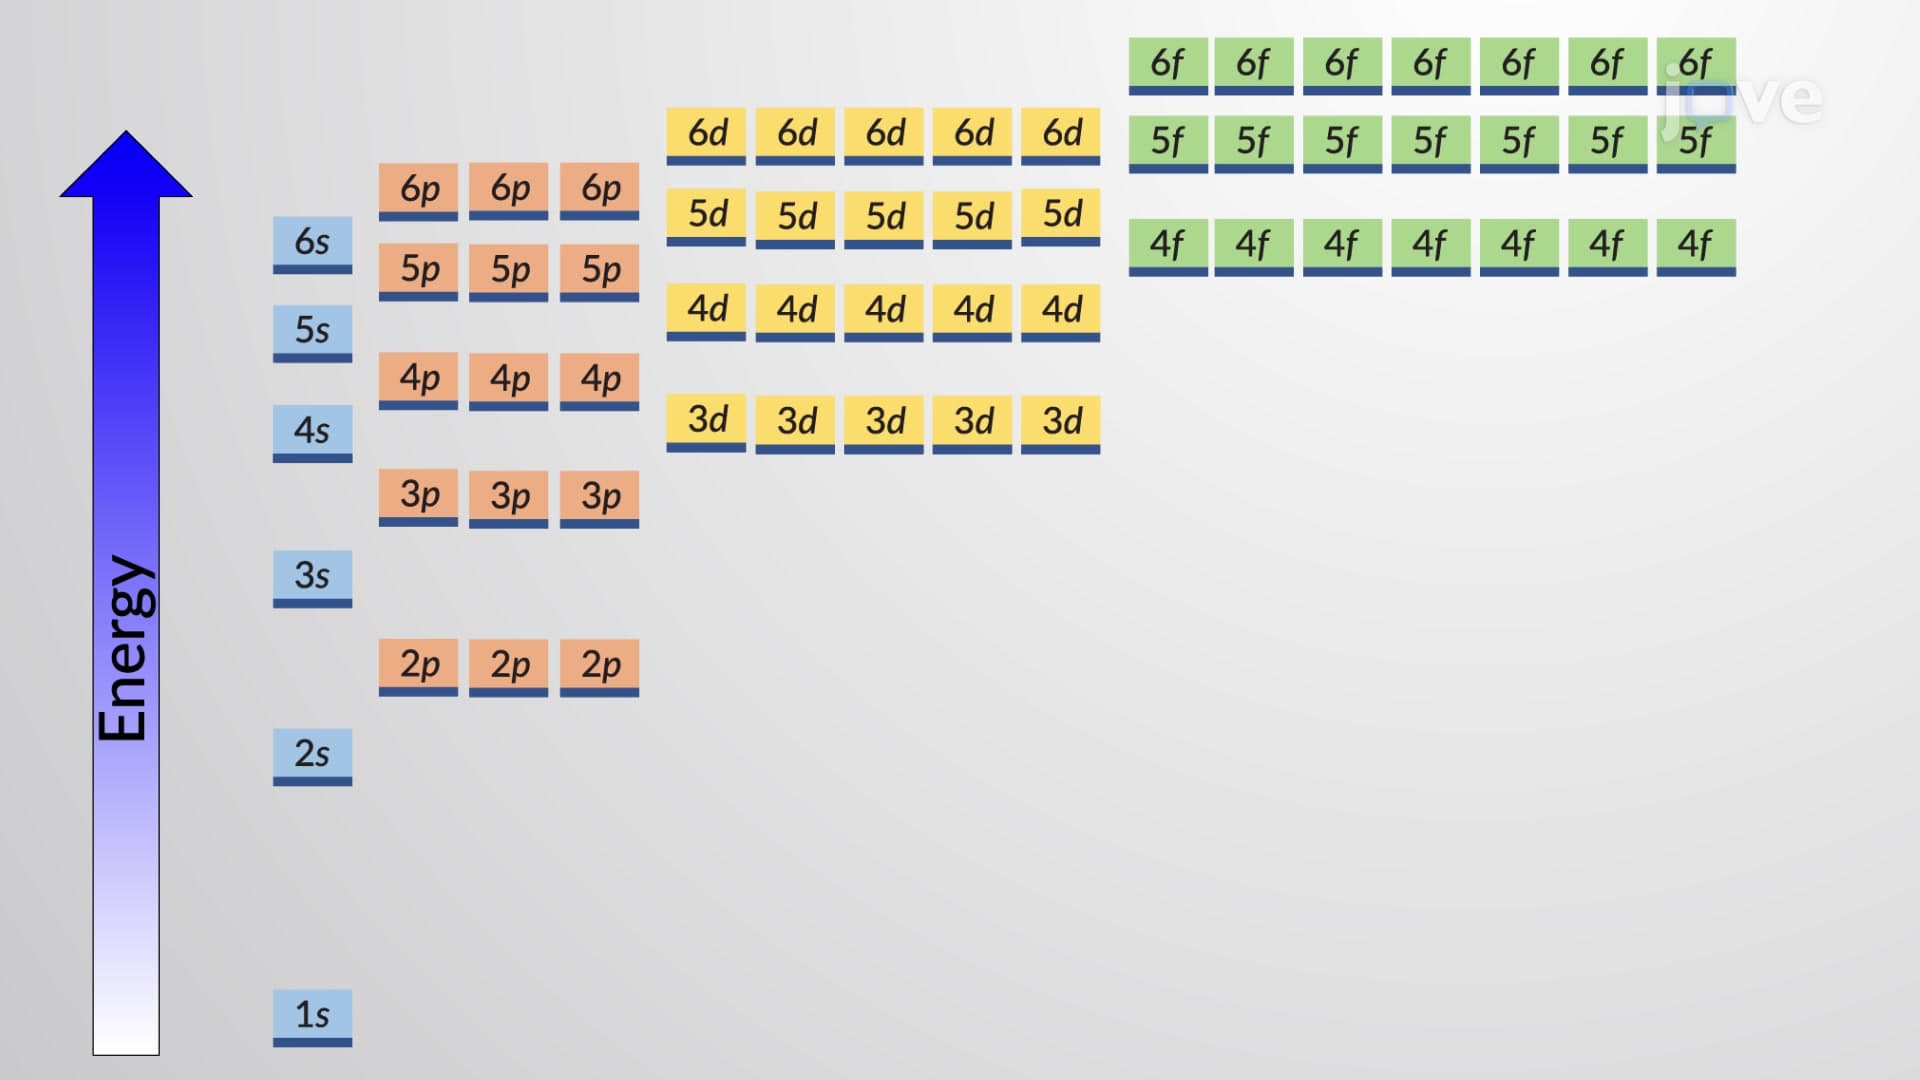
Electron Configuration of Multielectron Atoms
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:23
Atomic Spectroscopy: Absorption, Emission, and Fluorescence