Related Experiment Video
Updated: Aug 20, 2026


Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
An accurate density functional theory for the vapor-liquid interface of associating chain molecules based on the
Guy J Gloor1, George Jackson, Felipe J Blas
1Department of Chemical Engineering and Chemical Technology, Imperial College London, South Kensington Campus, London SW7 2AZ, United Kingdom.

Abstract:
A Helmholtz free energy density functional is developed to describe the vapor-liquid interface of associating chain molecules. The functional is based on the statistical associating fluid theory with attractive potentials of variable range (SAFT-VR) for the homogenous fluid [A. Gil-Villegas, A. Galindo, P. J. Whitehead, S. J. Mills, G. Jackson, and A. N. Burgess, J. Chem. Phys. 106, 4168 (1997)]. A standard perturbative density functional theory (DFT) is constructed by partitioning the free energy density into a reference term (which incorporates all of the short-range interactions, and is treated locally) and an attractive perturbation (which incorporates the long-range dispersion interactions). In our previous work [F. J. Blas, E. Martin del Rio, E. de Miguel, and G. Jackson, Mol. Phys. 99, 1851 (2001); G. J. Gloor, F. J. Blas, E. Martin del Rio, E. de Miguel, and G. Jackson, Fluid Phase Equil. 194, 521 (2002)] we used a mean-field version of the theory (SAFT-HS) in which the pair correlations were neglected in the attractive term. This provides only a qualitative description of the vapor-liquid interface, due to the inadequate mean-field treatment of the vapor-liquid equilibria. Two different approaches are used to include the correlations in the attractive term: in the first, the free energy of the homogeneous fluid is partitioned such that the effect of correlations are incorporated in the local reference term; in the second, a density averaged correlation function is incorporated into the perturbative term in a similar way to that proposed by Toxvaerd [S. Toxvaerd, J. Chem. Phys. 64, 2863 (1976)]. The latter is found to provide the most accurate description of the vapor-liquid surface tension on comparison with new simulation data for a square-well fluid of variable range. The SAFT-VR DFT is used to examine the effect of molecular chain length and association on the surface tension. Different association schemes (dimerization, straight and branched chain formation, and network structures) are examined separately. The surface tension of the associating fluid is found to be bounded between the nonassociating and fully associated limits (both of which correspond to equivalent nonassociating systems). The temperature dependence of the surface tension is found to depend strongly on the balance between the strength and range of the association, and on the particular association scheme. In the case of a system with a strong but very localized association interaction, the surface tension exhibits the characteristic "s shaped" behavior with temperature observed in fluids such as water and alkanols. The various types of curves observed in real substances can be reproduced by the theory. It is very gratifying that a DFT based on SAFT-VR free energy can provide an accurate quantitative description of the surface tension of both the model and experimental systems.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:26
The Van der Waals Equation
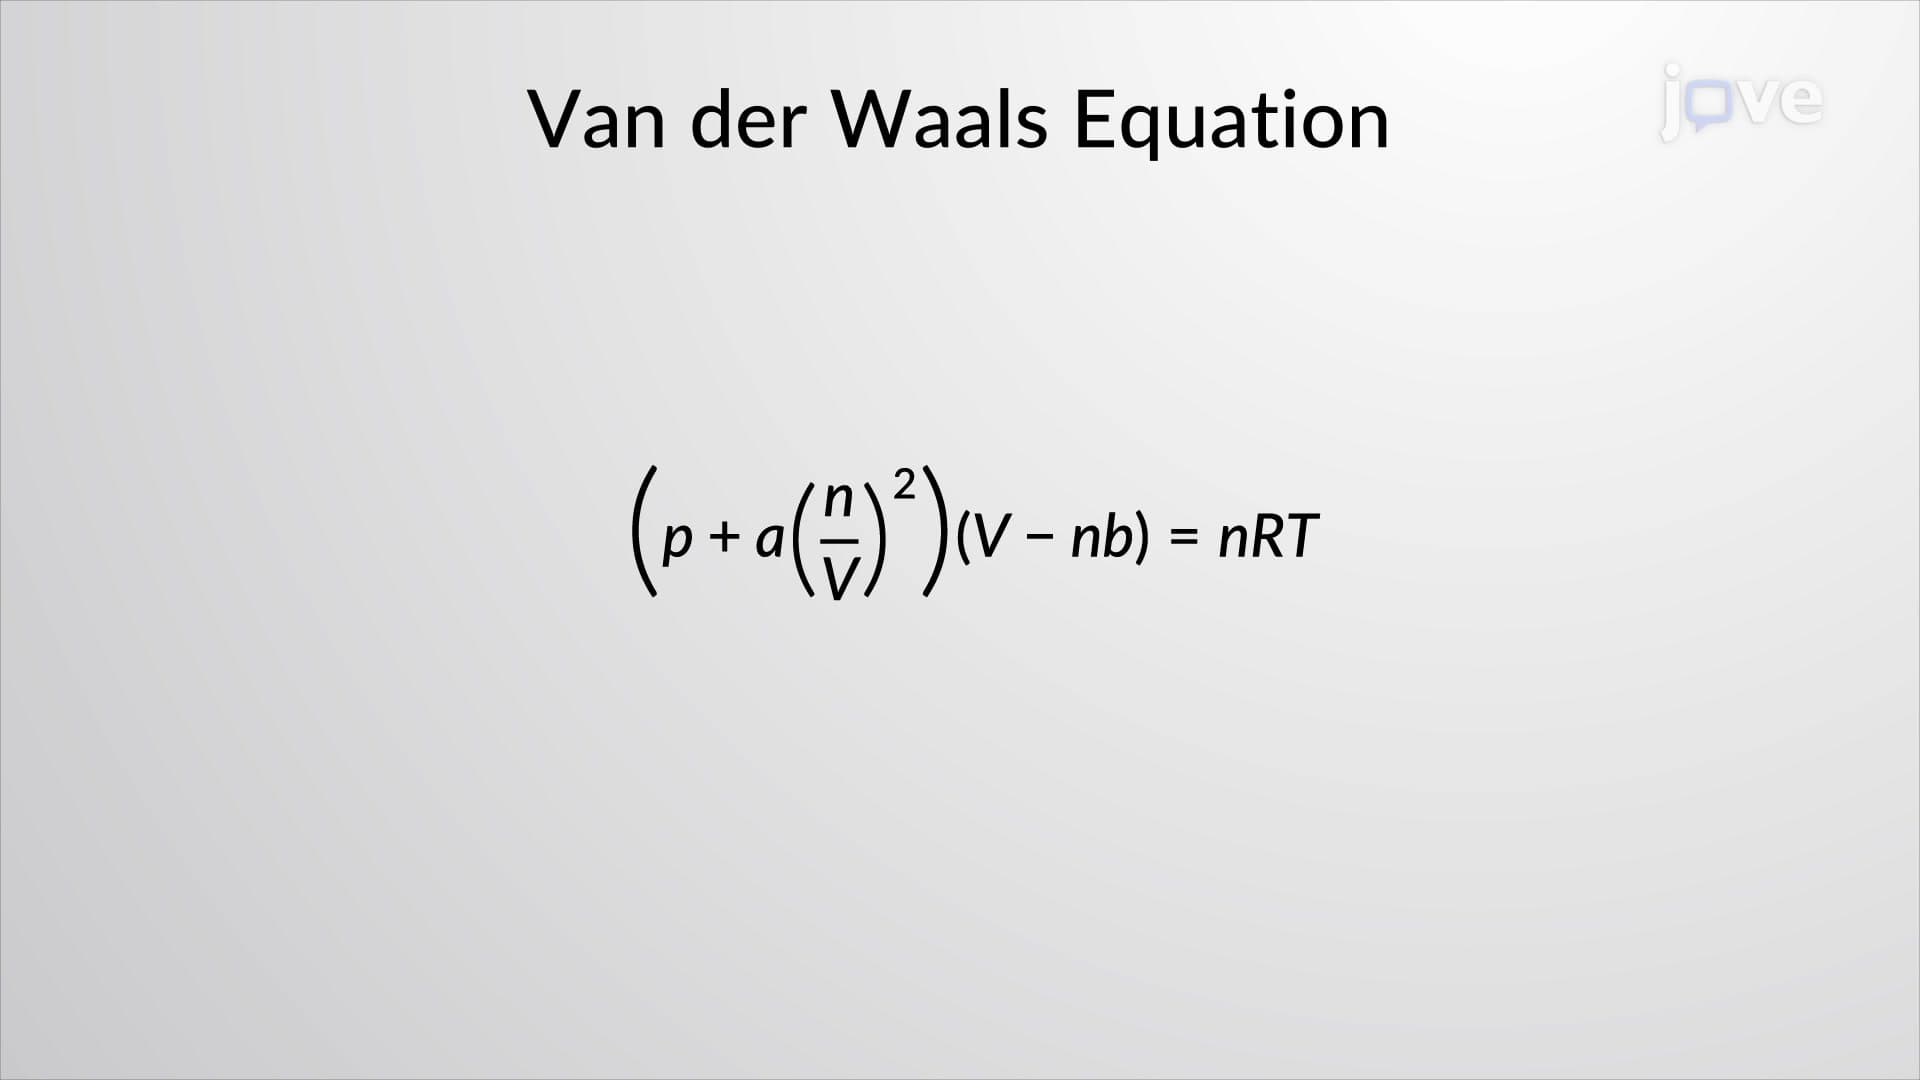 01:10
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
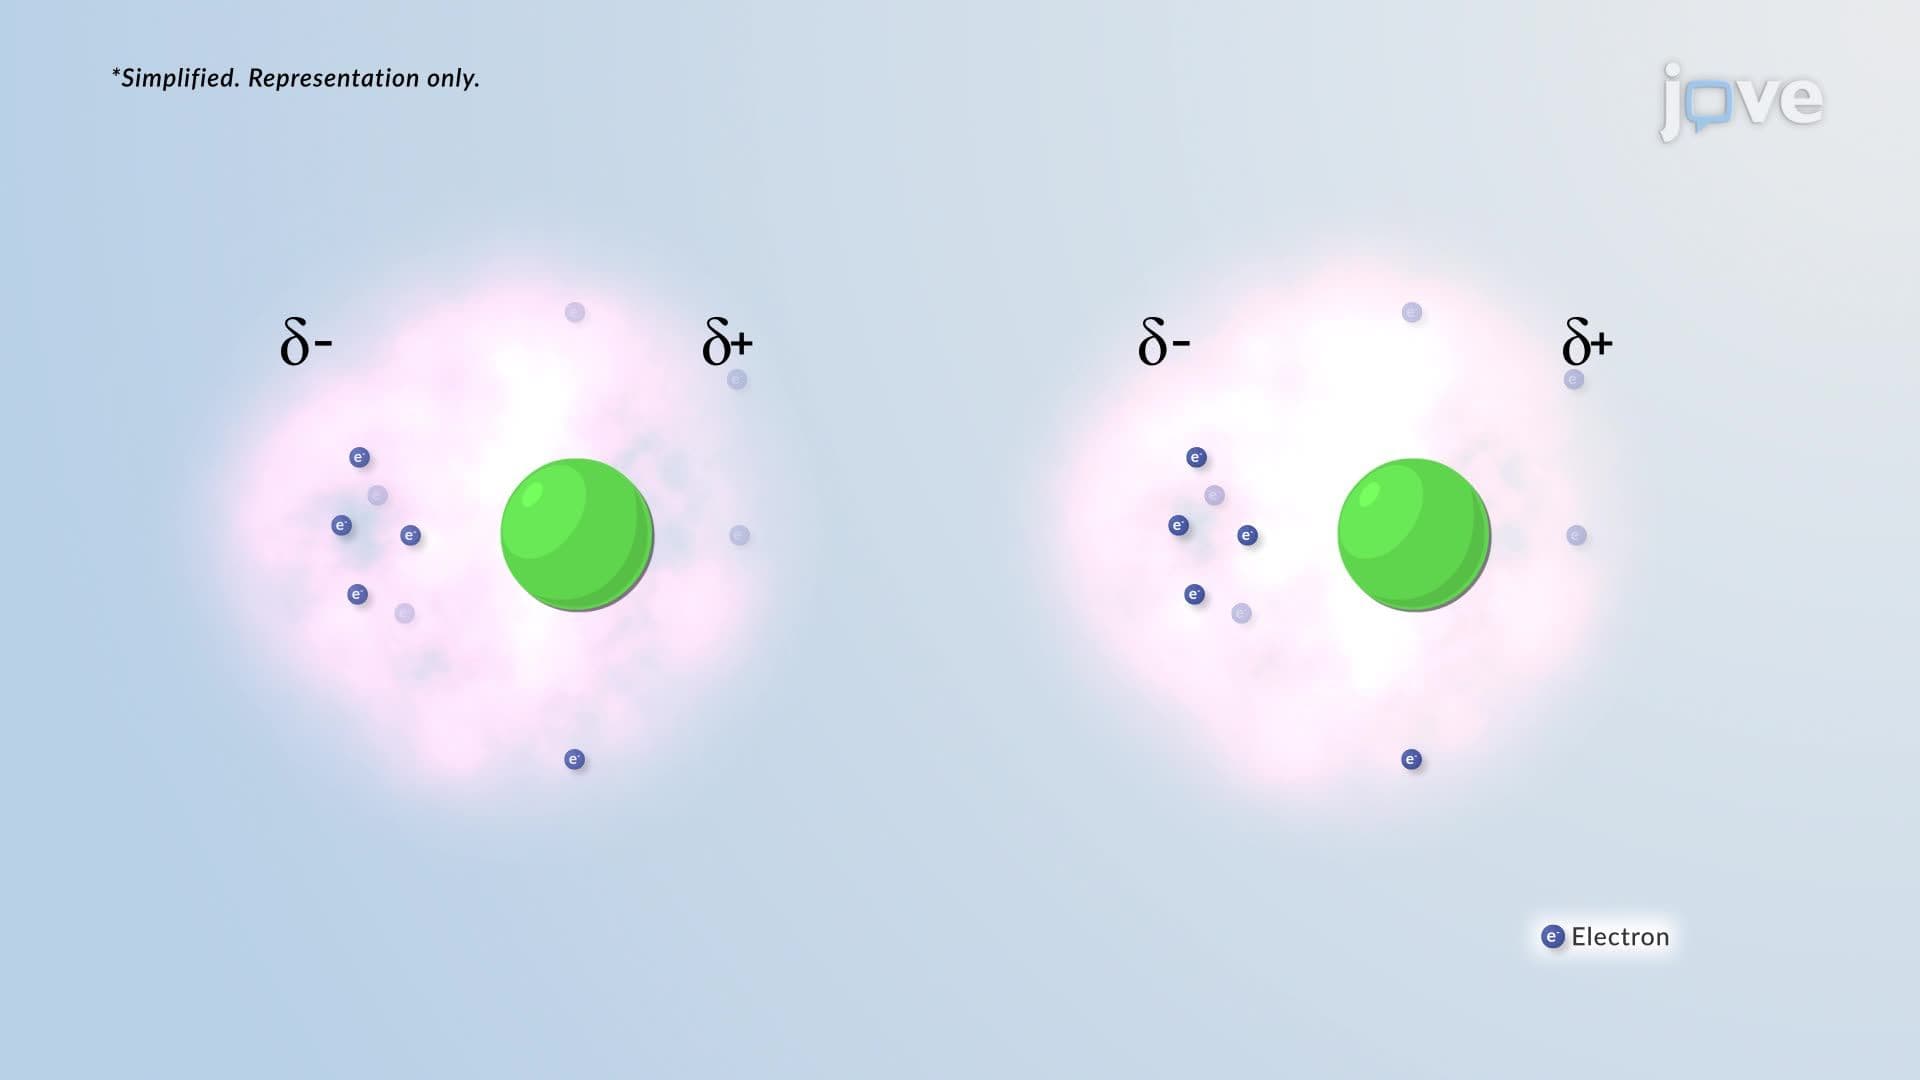 01:24
Van der Waals Interactions
 01:27
Distribution of Molecular Speeds
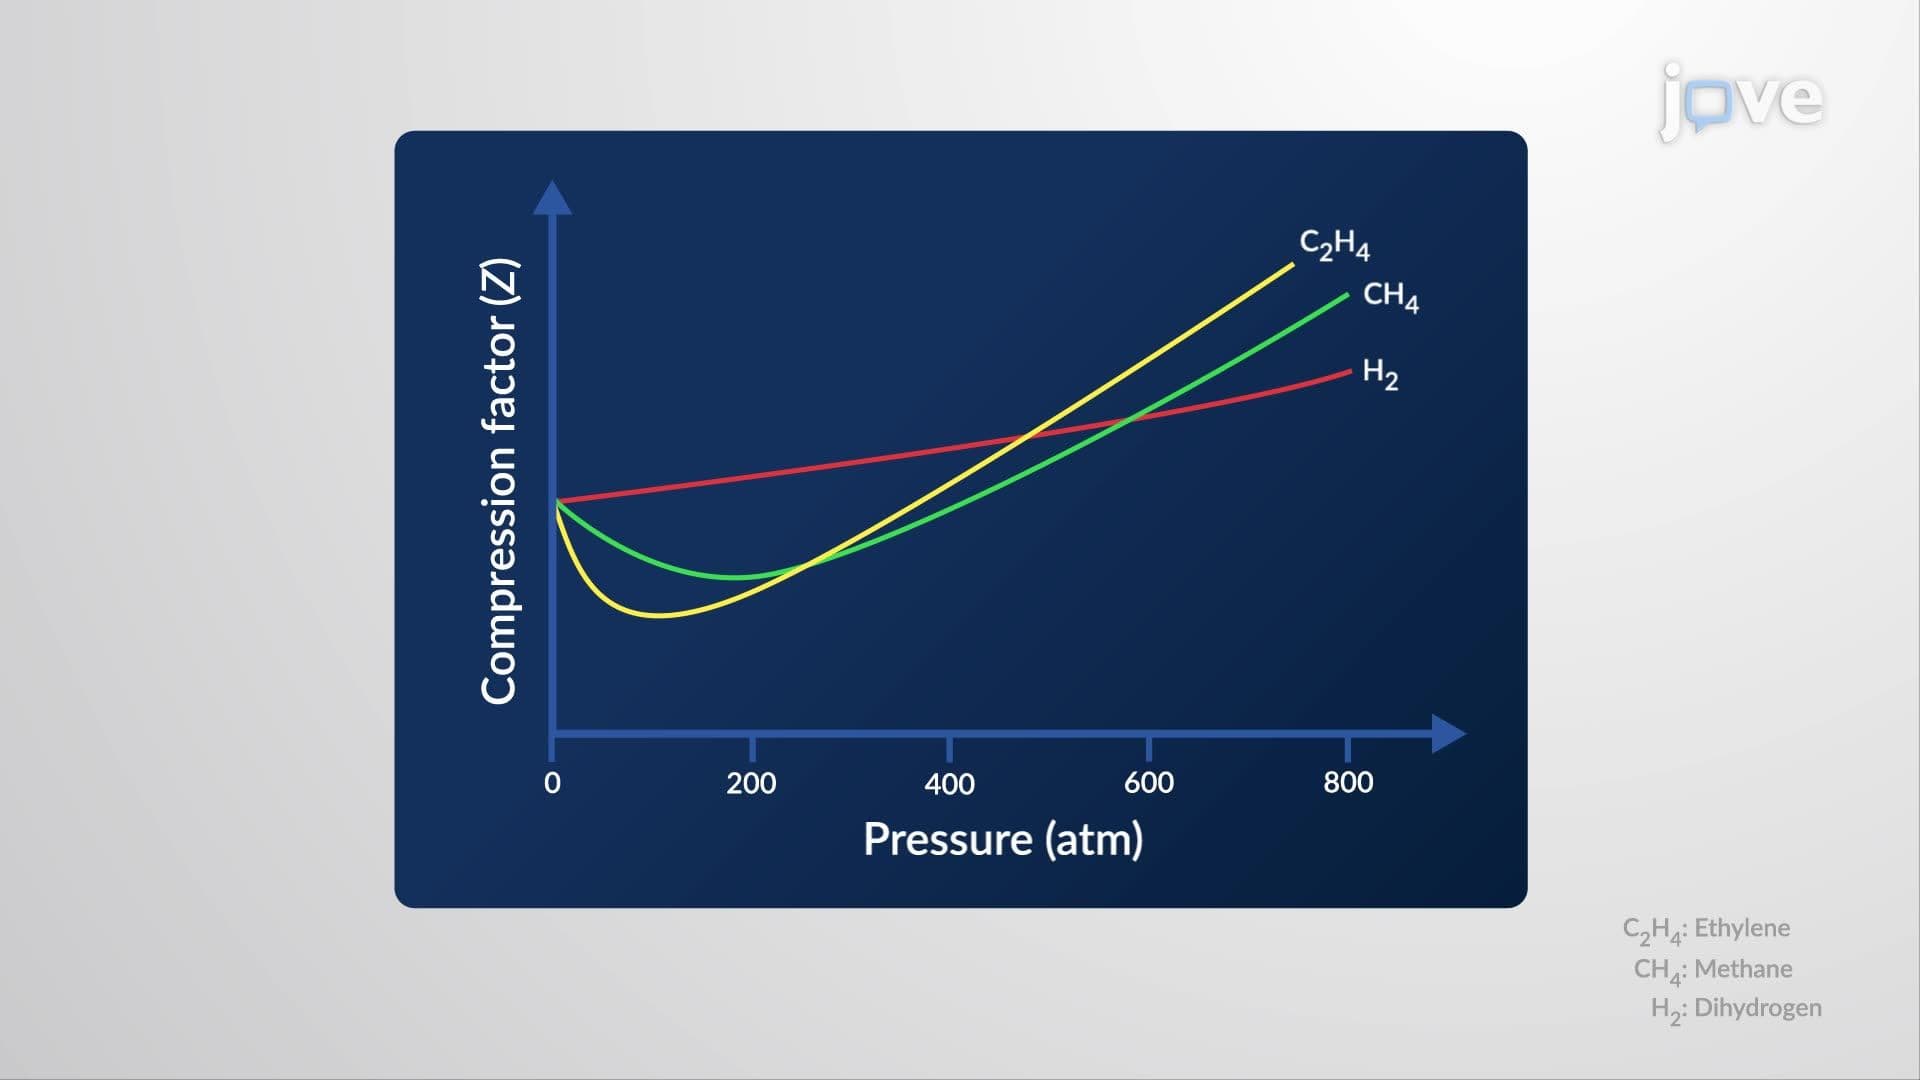 01:23
Deviation from Ideal Behaviour