Related Experiment Video
Updated: Jul 16, 2026


A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
Predictive in silico all-atom folding of a four-helix protein with a free-energy model
Alexander Schug1, Wolfgang Wenzel
1Forschungszentrum Karlsruhe, Institute for Nanotechnology, P.O. Box 3640, 76021 Karlsruhe, Germany.

Abstract:
We report the predictive all-atom folding of the 60 amino acid four-helix bacterial ribosomal protein (BRP) L20 with a stochastic evolutionary optimization method in a free-energy force field. The energetically best, as well as six of the 10 lowest conformations, converge to a near-native structure. All of the 10 best energy conformations share the secondary structure elements of the native conformation, but differ in their tertiary alignment. The best conformation has a backbone root-mean-square deviation of 4.6 A to the native conformation and reproduces most distance constraints of the NMR experiment to 1.5 A resolution. Starting from random initial conditions, the native content of the simulated population increases more than 60-fold in the course of the simulation. These data demonstrate the feasibility of predictive unbiased all-atom protein folding with present day computational resources for the BRP L20.
Related Concept Videos
 01:18
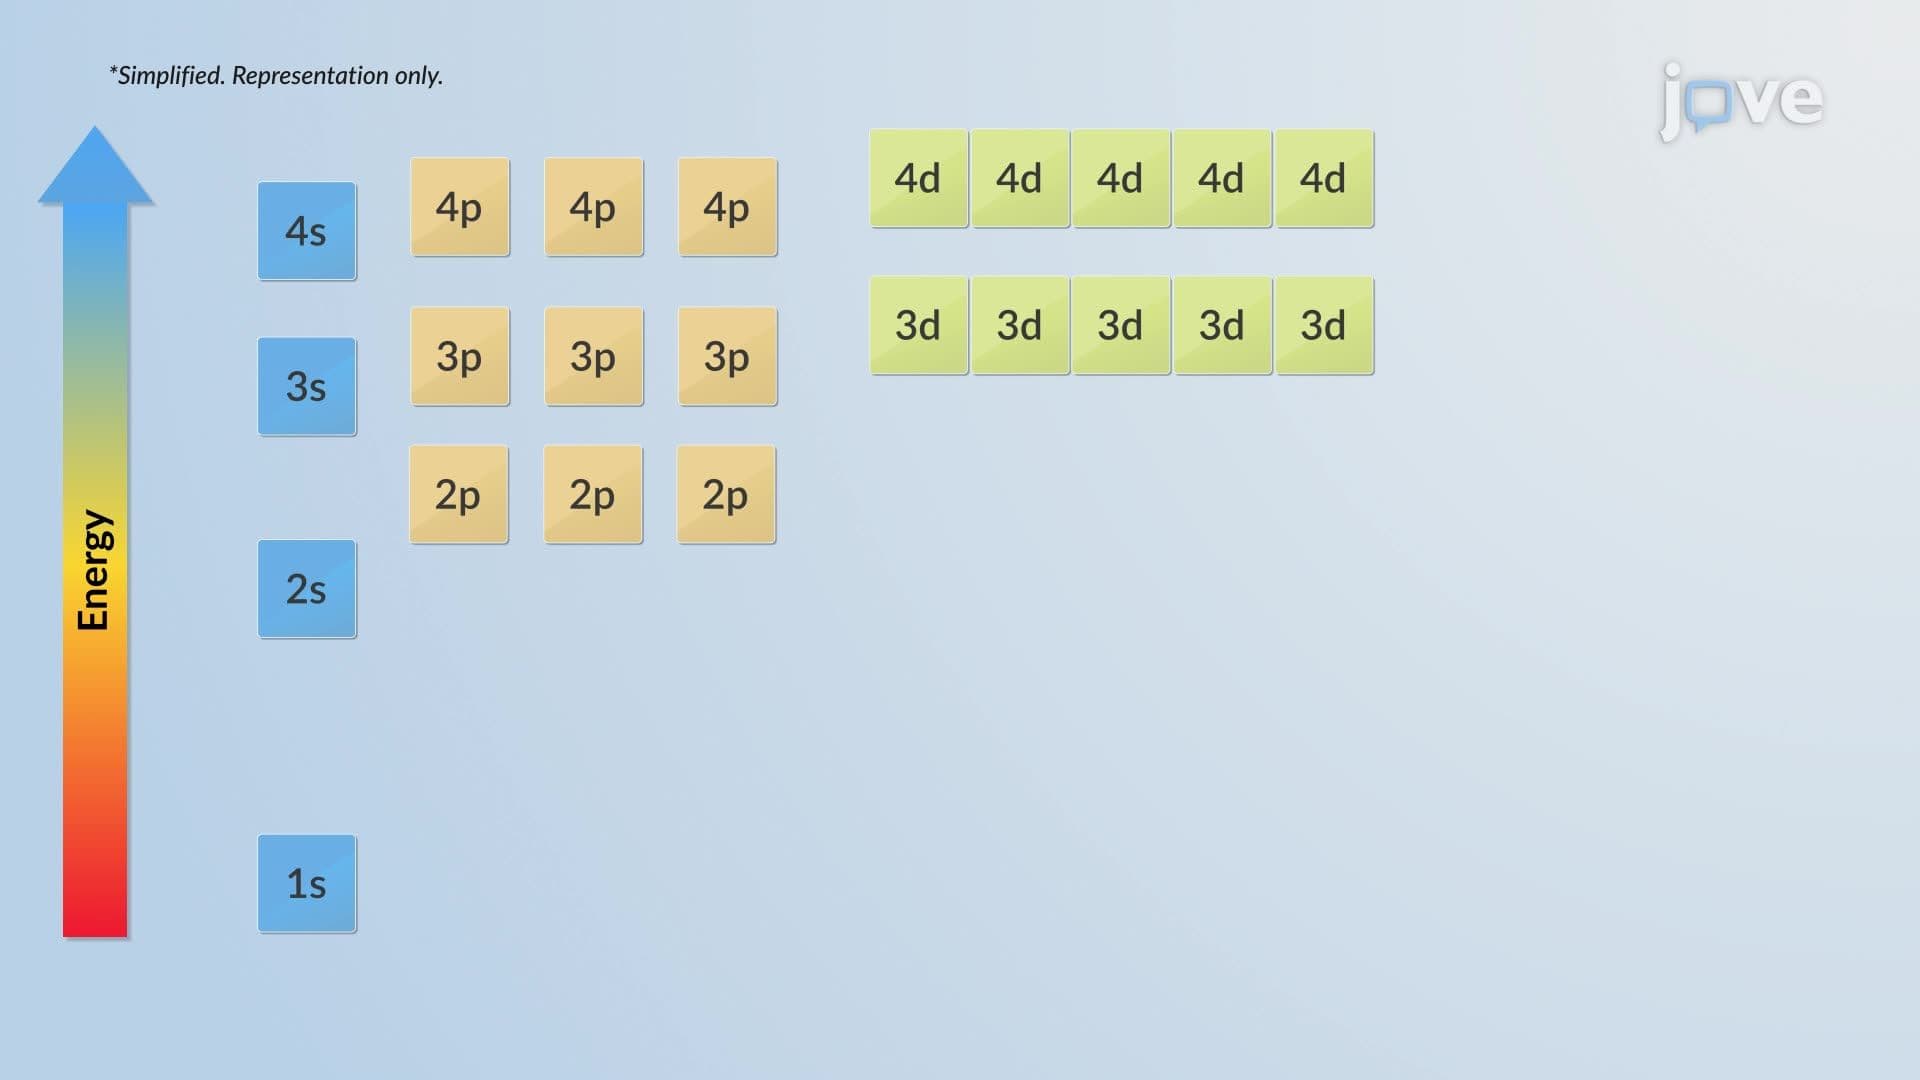
Electron Orbital Model
 02:18
The Bohr Model
 02:45
The Quantum-Mechanical Model of an Atom
 03:18
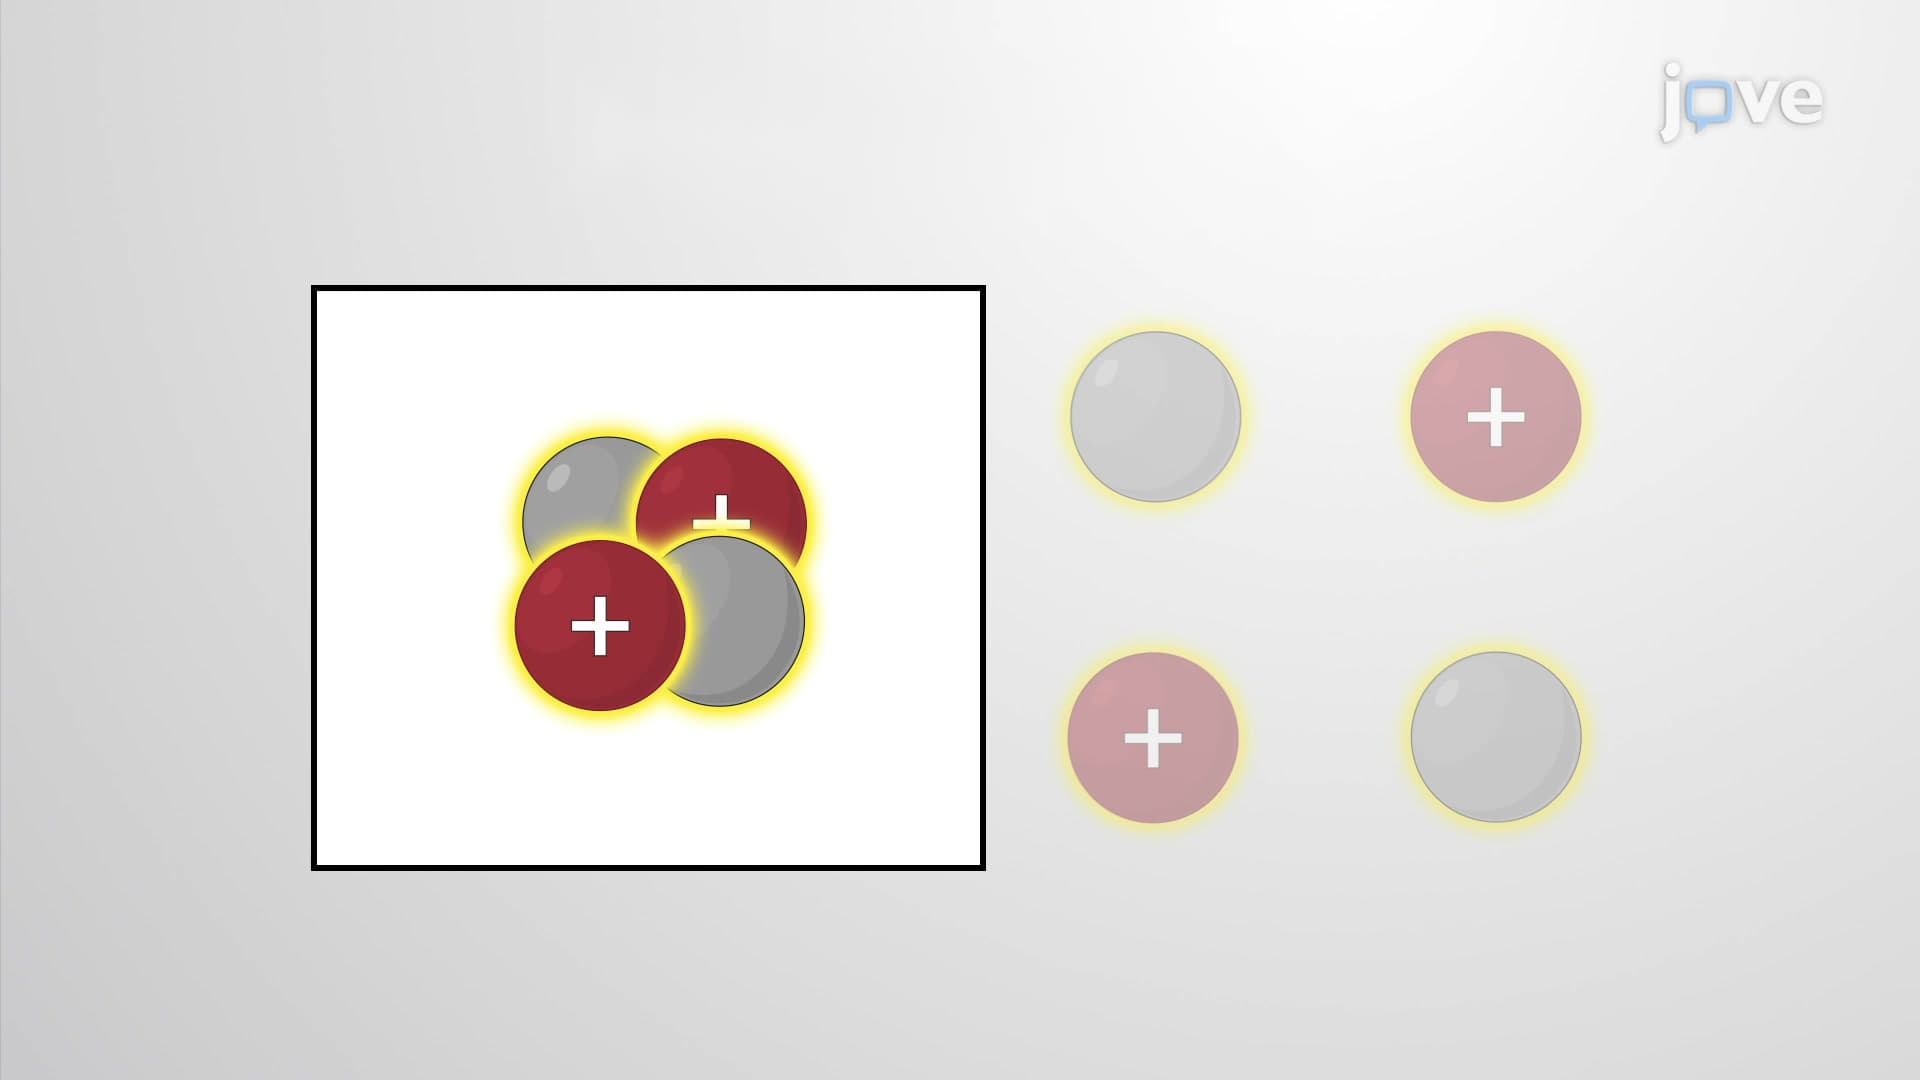
Nuclear Stability
To hold positively charged protons together in the...
 02:13
Nuclear Binding Energy
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum numbers: n, l, ml, and...