Related Experiment Video
Updated: Aug 20, 2026

Structural Biology and Analytical Chemistry Approaches for Characterizing C-Glycoside Metabolic Enzymes in Human Gut Microbiota
Published on: May 23, 2025
P-glycoprotein potentiates CYP3A4-mediated drug disappearance during Caco-2 intestinal secretory detoxification
Lauretta M S Chan1, Anne E Cooper, Adam L J Dudley
1Institute for Cell and Molecular Biosciences, University of Newcastle Medical School, Newcastle upon Tyne NE2 4HH, UK.

Abstract:
Human intestinal Caco-2 cell monolayers grown in the presence of 1alpha,25-dihydroxyvitamin D3 (1,25(OH)2D3) were used to test the hypothesis that drugs which interact with the apical efflux pump P-glycoprotein (Pgp) may enhance CYP3A4-mediated disappearance of substrates. 6beta-hydroxytestosterone production, a marker of CYP3A4 activity, was approximately 3- and 7-fold greater in 1,25(OH)2D3-treated cells compared to untreated cells when incubated with 50 and 500 microM testosterone, respectively, and was unaffected by the addition of digoxin to reduce Pgp activity. In the presence of digoxin, secretory transport of vinblastine and erythromycin, substrates for both Pgp and cytochrome P450 3A4 (CYP3A4), was significantly reduced, whereas absorptive transport was unaffected. In contrast, no directional transport of testosterone, a substrate for CYP3A4 only, was observed, either in the presence or absence of digoxin. Over 2 h, disappearance of erythromycin and vinblastine from the incubation medium was significantly greater from the basolateral than from the apical compartments. In the presence of digoxin, disappearance of both compounds from the basolateral, but not from the apical compartments, was significantly reduced. In contrast, disappearance of testosterone was unaffected by the addition of digoxin, demonstrating that the effect of digoxin on erythromycin and vinblastine disappearance was via inhibition of Pgp function, rather than on CYP3A4 activity. Thus, evidence is provided for Pgp/CYP3A4 co-substrates, Pgp potentiates CYP3A4-mediated drug disappearance during intestinal secretory detoxification.
Insights
Drugs interacting with P-glycoprotein (Pgp) can enhance CYP3A4-mediated drug metabolism in the intestines. This study shows Pgp potentiates CYP3A4 activity, aiding in intestinal drug detoxification.
Area of Science:
- Pharmacology
- Drug Metabolism
- Gastrointestinal Physiology
Background:
- The intestinal epithelium plays a crucial role in drug absorption and metabolism.
- P-glycoprotein (Pgp) and Cytochrome P450 3A4 (CYP3A4) are key players in drug efflux and metabolism, respectively.
- Understanding their interplay is vital for predicting drug disposition and efficacy.
Purpose of the Study:
- To investigate whether drugs interacting with P-glycoprotein (Pgp) can enhance the activity of Cytochrome P450 3A4 (CYP3A4).
- To determine the role of Pgp in the intestinal detoxification of co-substrates.
Main Methods:
- Utilized human intestinal Caco-2 cell monolayers.
- Assessed CYP3A4 activity by measuring 6beta-hydroxytestosterone production from testosterone.
- Investigated the transport and disappearance of CYP3A4 and/or Pgp substrates (testosterone, vinblastine, erythromycin) in the presence and absence of digoxin (a Pgp inhibitor).
Main Results:
- CYP3A4 activity (testosterone metabolism) was increased by 1,25(OH)2D3 treatment but unaffected by digoxin.
- Digoxin significantly reduced the secretory transport and basolateral disappearance of Pgp/CYP3A4 co-substrates (vinblastine, erythromycin).
- Testosterone, a CYP3A4-only substrate, showed no change in transport or disappearance with digoxin, confirming digoxin's effect was Pgp-mediated.
Conclusions:
- P-glycoprotein (Pgp) potentiates CYP3A4-mediated drug disappearance.
- This interaction facilitates intestinal secretory detoxification of drugs that are co-substrates for both Pgp and CYP3A4.
- The findings highlight a significant drug-drug interaction mechanism at the intestinal level.
Related Concept Videos
 01:16
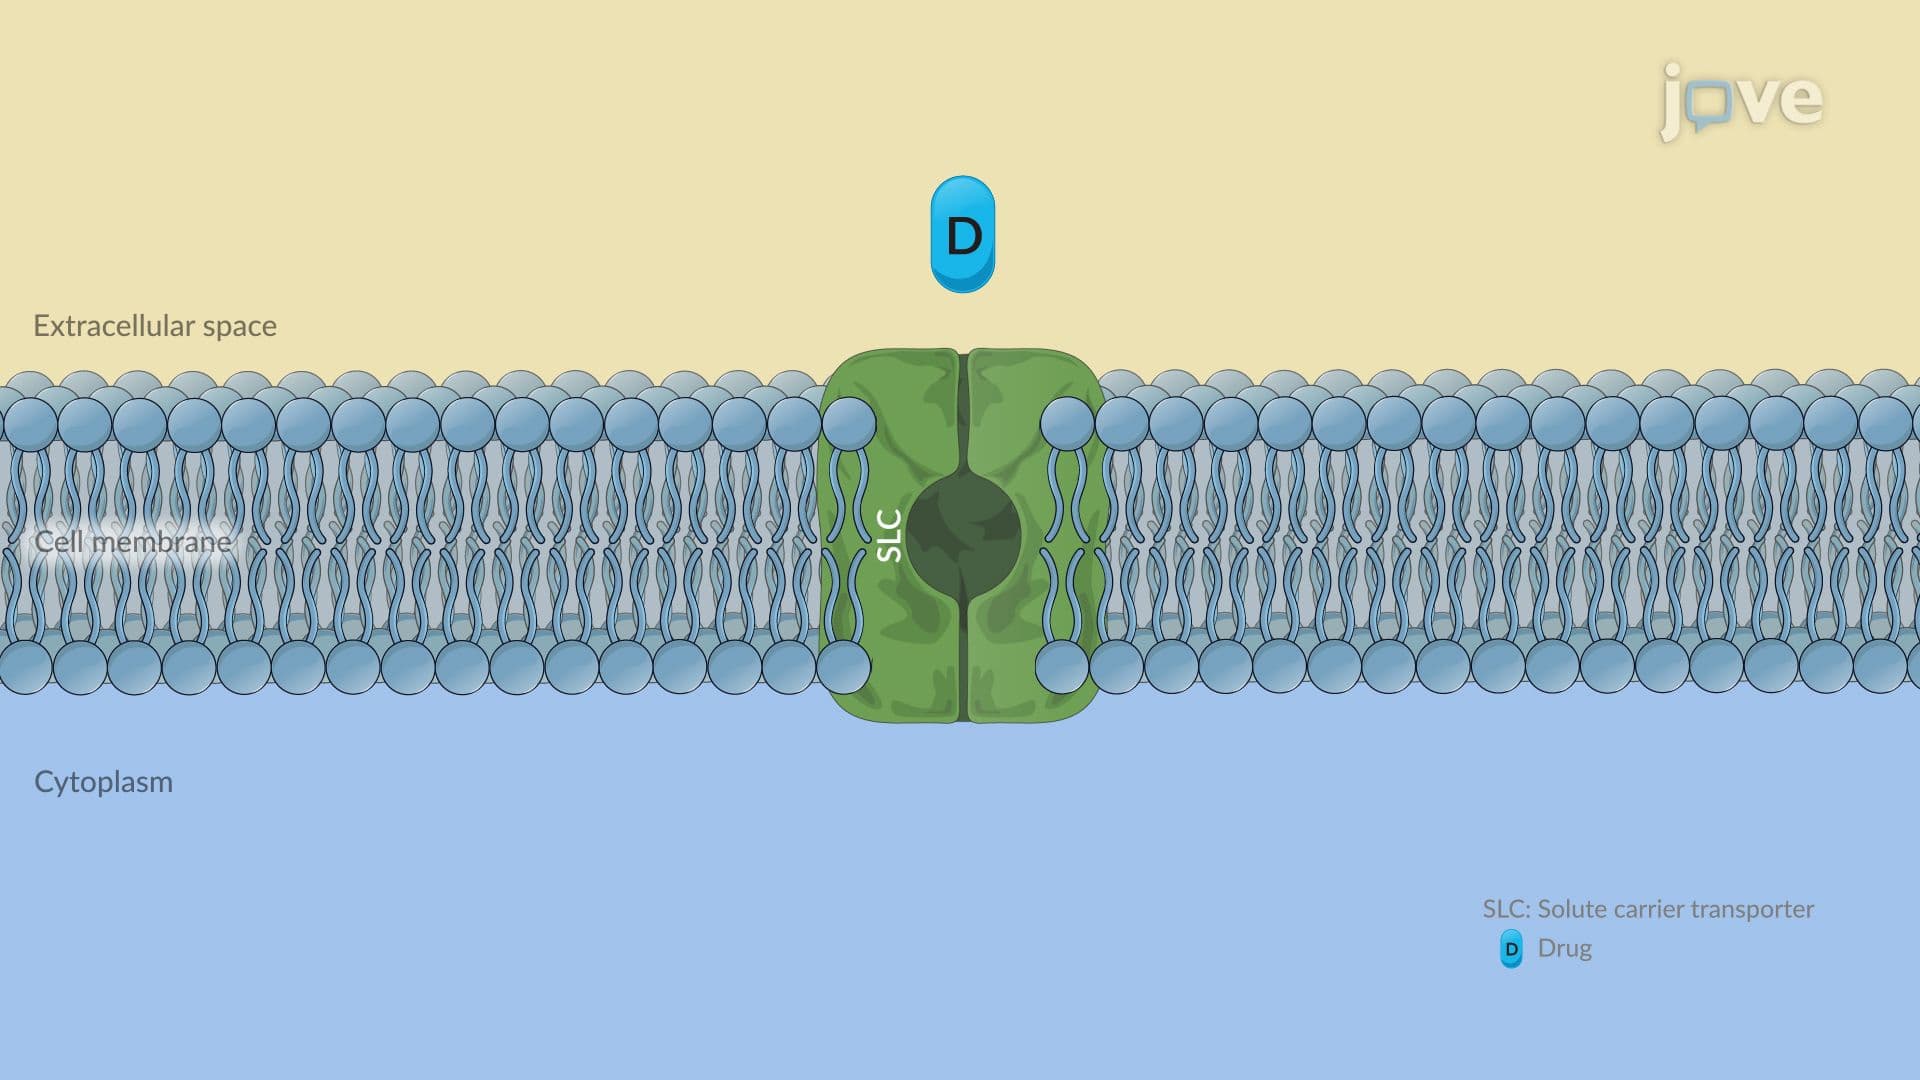
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:06
Carrier-Mediated Transport
Active transport involves two types of membrane-spanning transporters: uptake and efflux. Uptake transporters are expressed in the small...
 01:25
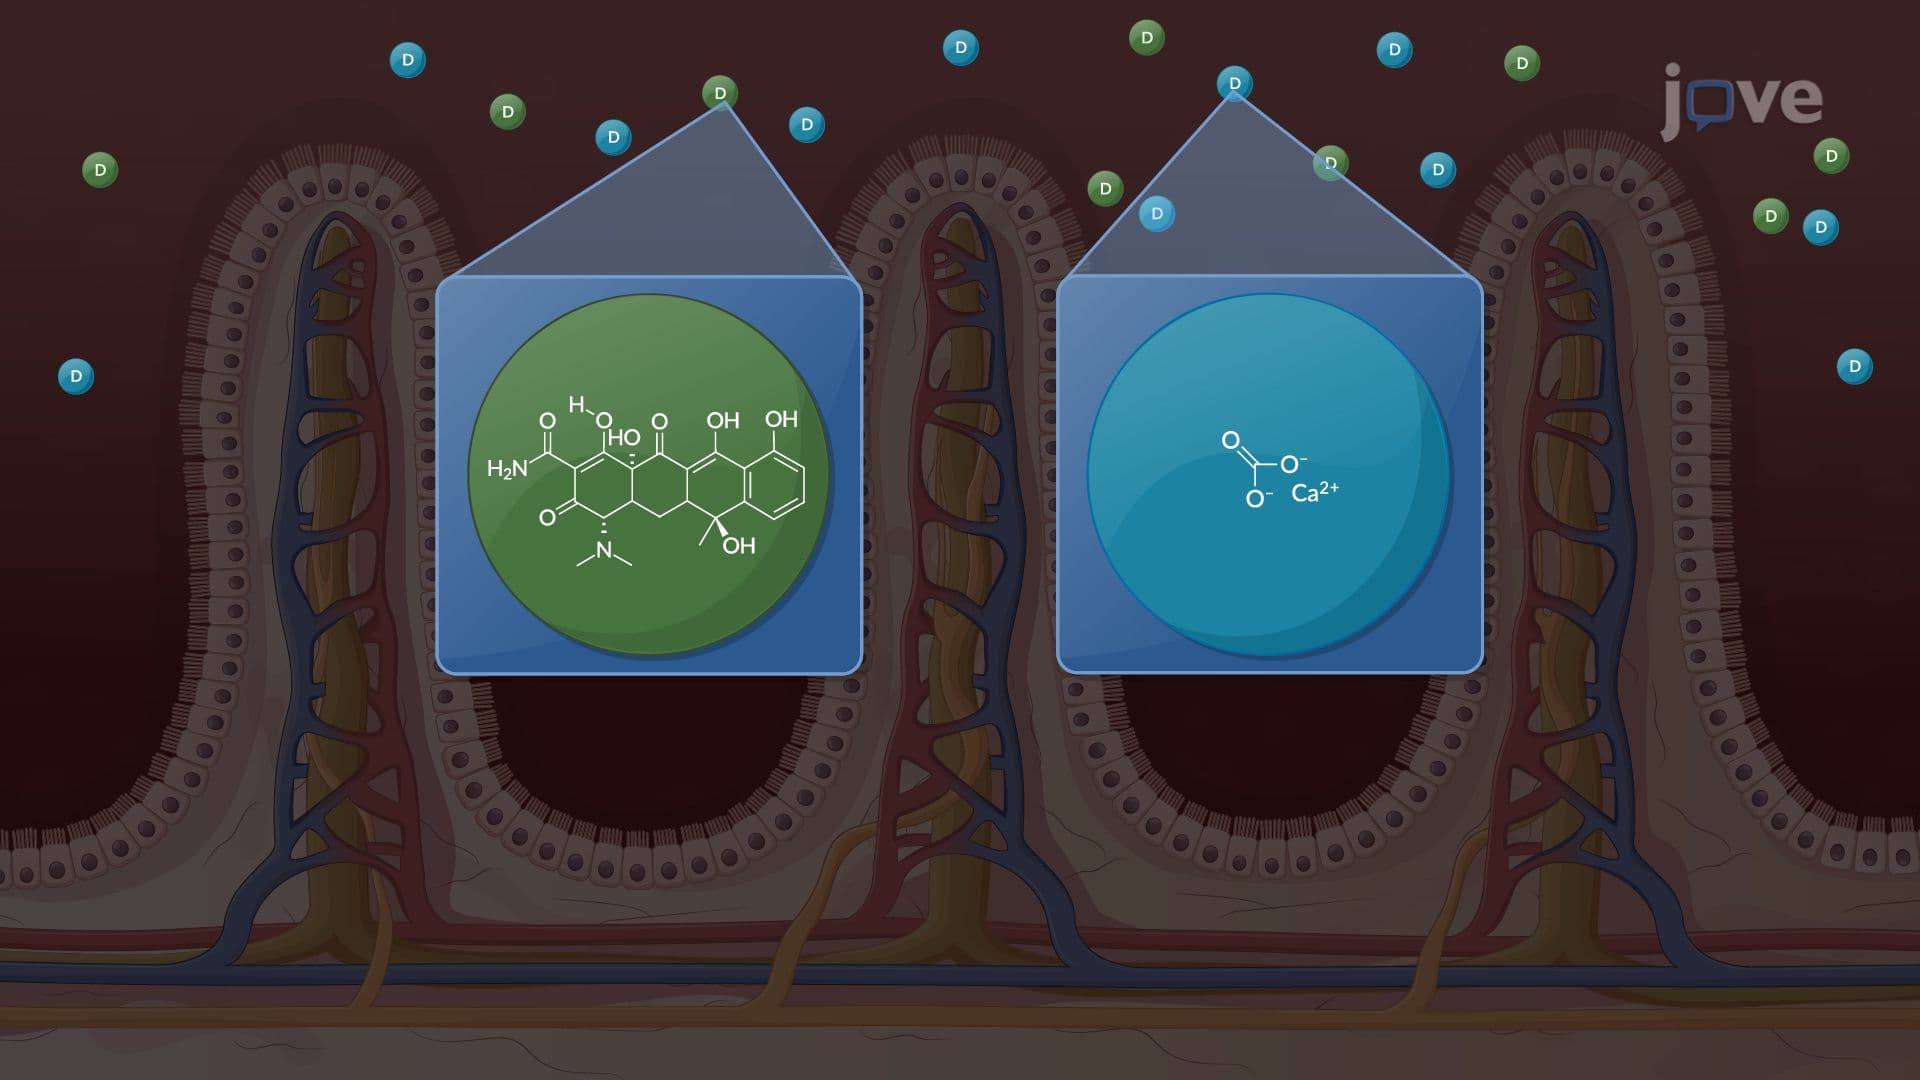
Pharmacokinetics: Drug–Drug Interactions
 01:17
Hepatic Drug Excretion: Enterohepatic Cycling
Post-release drugs and metabolites can be reabsorbed into the body from the intestine. For conjugated metabolites like glucuronides, reabsorption requires enzymatic hydrolysis by intestinal microflora. This...
 01:19
Drug Absorption: Factors Affecting GI Absorption
In...