Related Experiment Video
Updated: Aug 20, 2026
A Facile Protocol to Generate Site-Specifically Acetylated Proteins in Escherichia Coli
Published on: December 9, 2017
Lysine alpha-ketoglutarate reductase and lysine oxidation are distributed in the extrahepatic tissues of chickens
Megharaja K Manangi1, Sara F A Hoewing, Juanita G Engels
1Division of Animal and Veterinary Sciences, West Virginia University-Morgantown, WV, USA.

Abstract:
In animals, lysine oxidation is thought to occur primarily via the activity of lysine alpha-ketoglutarate reductase (LKR). This activity was reported previously in chicken liver, but no work on the tissue distribution of the enzyme in chickens has been reported. Therefore, LKR activity was assayed in liver, kidney, pancreas, heart, brain, lung, spleen, muscle, and intestinal tissues in chickens as was the in vitro ability of tissue homogenates to oxidize lysine. Additionally, the expression of LKR mRNA was assessed by RT-PCR. We found LKR to be present in all tissues studied by both enzymatic analysis and mRNA abundance. Additionally, all tissues assayed oxidized lysine. The extent of lysine oxidation differed among the tissues, consistent with the different pathways of lysine oxidation in the different tissues. These studies demonstrate that LKR is widely distributed in chicken tissues and that tissues other than liver can contribute to whole-body lysine oxidation.
Related Concept Videos
 02:41
Lethal Alleles
Lucien Cuénot discovered lethal alleles in 1905 while studying the inheritance of coat color in mice. The agouti gene is responsible for the color of the coat in mice. This gene codes for an agouti-signaling protein, which is responsible for melanin distribution in mammals. The wild-type allele gives rise to gray-brown coat color in mice, while the mutant allele gives rise to yellow coat color. In addition to coat color, the agouti gene is associated with the yellow...
 01:17
Oxidation of Phenols to Quinones
o-hydroxy phenols are oxidized to o-quinones and p-hydroxy phenols to p-quinones. Such redox reactions involve the transfer of two electrons and two protons. The reversible redox property is crucial in...
 01:29
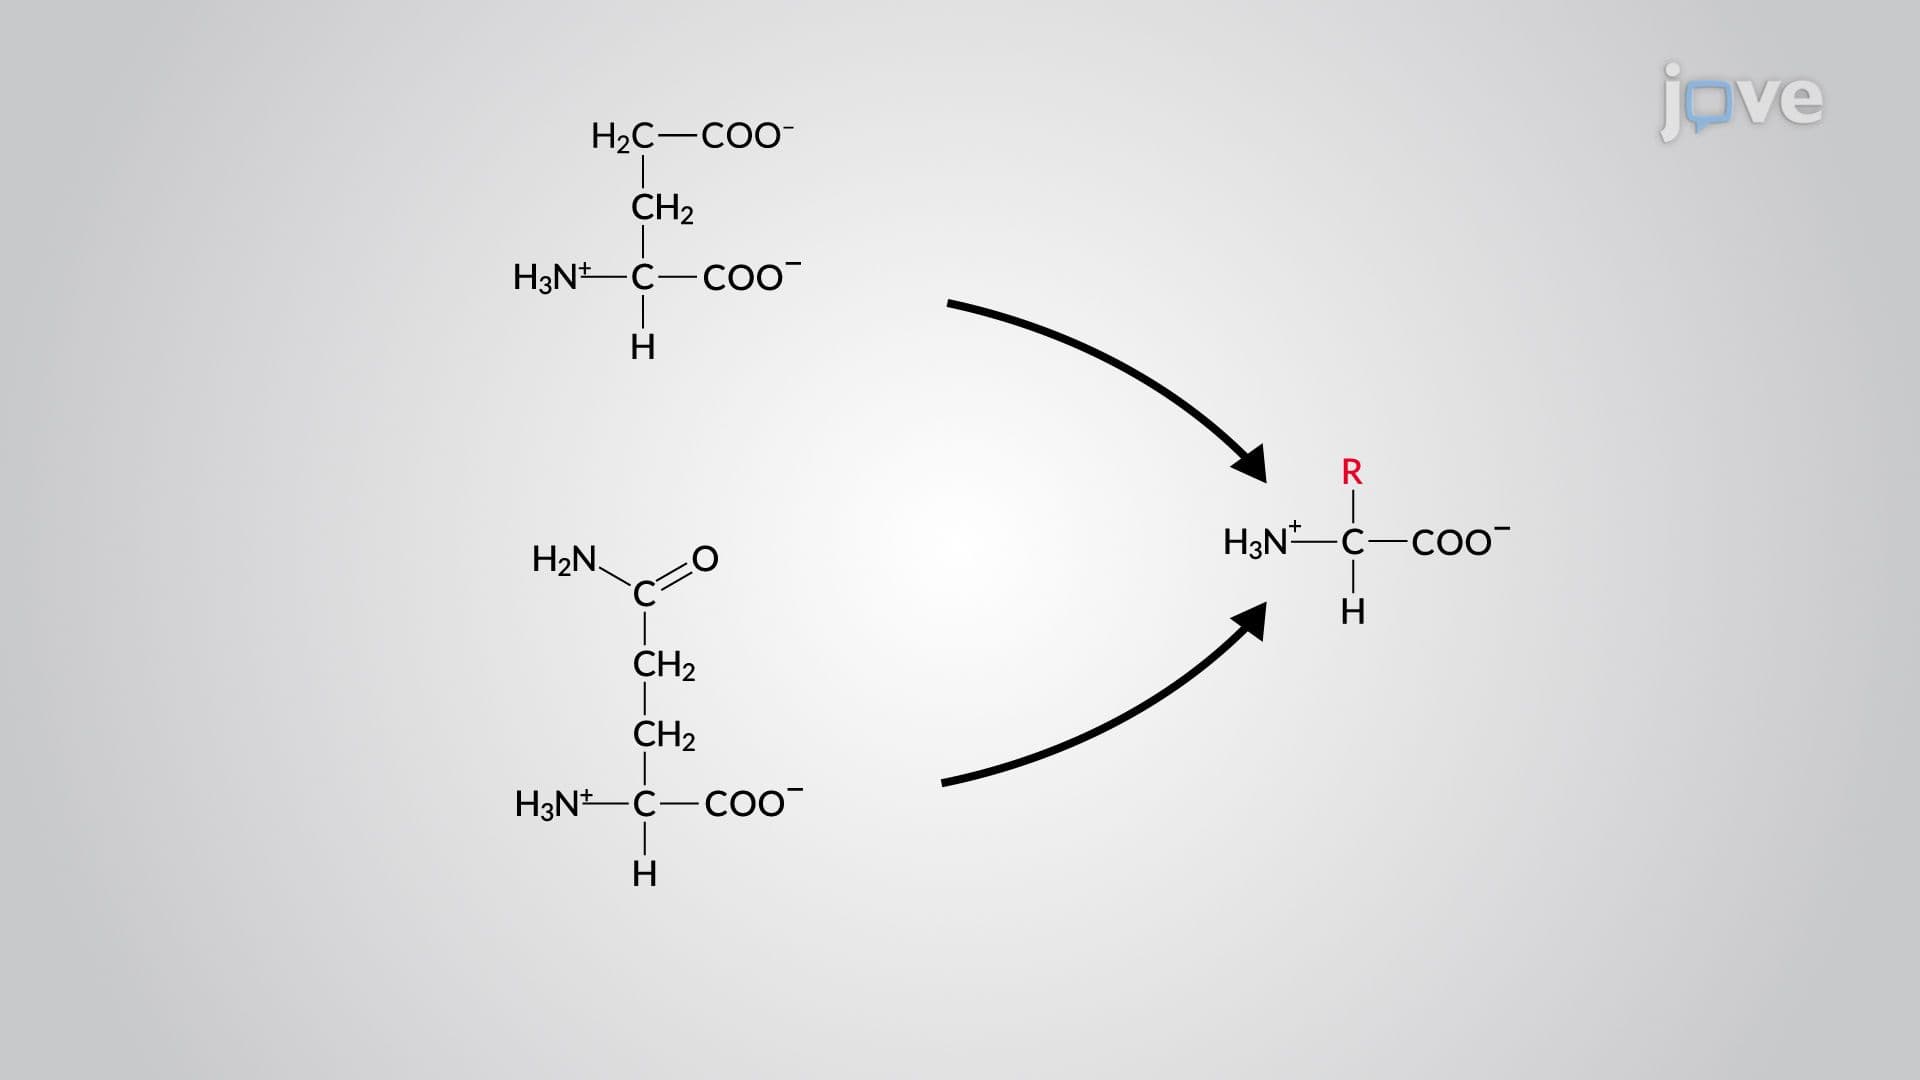
Amino Acid Biosynthetic Pathways
 01:20
Inborn Errors of Metabolism
 01:02
Protein Digestion
 01:22
Lysosomal Hydrolases