Related Experiment Video
Updated: Aug 17, 2026
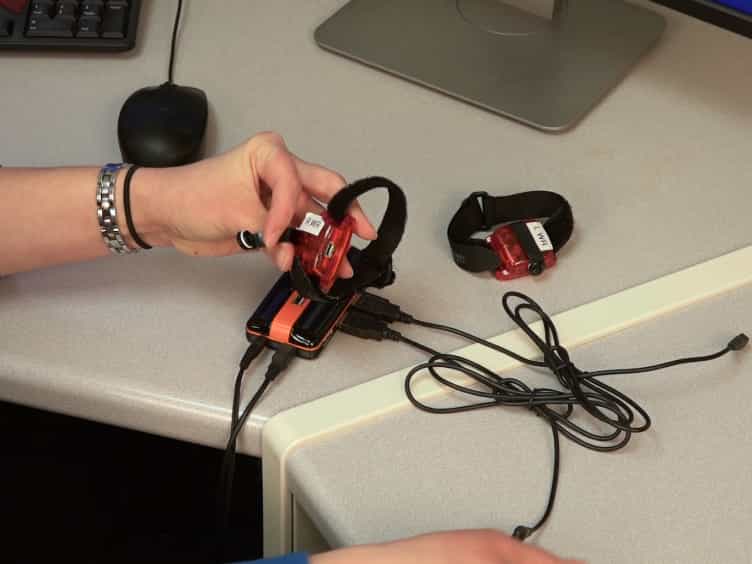

A Method for Quantifying Upper Limb Performance in Daily Life Using Accelerometers
Published on: April 21, 2017
[Measurement of daily physical activity in patients with cystic fibrosis]
C Fournier1, L Bosquet, S Leroy
1Centre de Ressources et Compétences pour la Mucoviscidose, Clinique des maladies respiratoires, Hôpital Calmette, CHRU, Lille, France. cl-fournier@chru-lille.fr

Background:
Cystic fibrosis (CF) patients often have a good level of daily activity contrasting with the impairment of pulmonary functions. The aim of this prospective study was to measure physical daily activity (PDA) in CF-patients and compare it with rest pulmonary functions and exercise tolerance.
Material And Methods:
Fifteen CF-patients and 9 control subjects have been recruted. PDA was measured during 7 days using an accelerometer: Global-PDA (daily mean counts per day), percentage of inactivity (1 to 2 METS), light activity (2 to 3 METS), and moderate-to-hard activity (greater than 3 METS). Spirometry (FEV1, Vital Capacity) and clinical exercice testing (VO2, workload) were also performed.
Results:
Global-PDA of CF-patients was lower than control subjects (p < 0.05). Light activity (15.5 +/- 3.3% vs 13.4 +/- 2.4%) and inactivity (81.2 +/- 9% vs 79.9 +/- 3.1%) were similar in both groups. Only moderate-to-hard activity was greater in control subjects than in CF-patients (6.7 +/- 1.6% vs 3.3 +/- 3.4%; p < 0.05). There were correlations between Global-PDA and moderate-to-hard activity (r = 0.94), FEV1 (r = 0.72), Vital Capacity (r = 0.77), workload peak (r = 0.88), and VO2peak (r = 0.70).
Conclusion:
CF-patients have a lower PDA than control subjects because of a lower moderate-to-hard activity (>3 METS). In clinical practice, interviewing a CF-patient on his "run-life" PDA (<3 METS) has poor relationship with his real "moderate-to-hard activity", and this is a confusing factor for clinicians.
Related Concept Videos
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:27
Chronic Obstructive Pulmonary Disease-IV: Assessement and Diagnostic Studies
Medical History
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...