Related Experiment Video
Updated: Aug 17, 2026
Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
Allosteric modulation of the adenosine family of receptors
Zhan-Guo Gao1, Soo-Kyung Kim, Adriaan P Ijzerman
1Molecular Recognition Section, Laboratory of Bioorganic Chemistry, NIDDK, National Inst. of Health, Bethesda, Maryland 20892-0810, USA. kajobs@helix.nih.gov

Abstract:
Allosteric modulators for adenosine receptors (ARs) are of an increasing interest and may have potential therapeutic advantage over orthosteric ligands. Benzoylthiophene derivatives (including PD 81,723), 2-aminothiazolium salts, and related allosteric modulators of the A(1) AR have been studied. The benzoylthiophene derivatives were demonstrated to be selective enhancers for the A(1) AR, with little or no effect on other subtypes of ARs. Allosteric modulation of the A(2A) AR has also been reported. A(3) allosteric enhancers may be predicted to be useful against ischemic conditions. We have recently characterized two classes of A(3) AR allosteric modulators: 3-(2-pyridinyl)isoquinolines (e.g. VUF5455) and 1H-imidazo-[4,5-c]quinolin-4-amines (e.g. DU124183), which selectively decreased the agonist dissociation rate at the human A(3)AR but not at A(1) and A(2A) ARs. DU124183 left-shifted the agonist conc.-response curve for inhibition of forskolin-stimulated cAMP accumulation in intact cells expressing the human A(3)AR with up to 30% potentiation of the maximal efficacy. The increased potency of A(3) agonists was evident only in the presence of an A(3) antagonist, since VUF5455 and DU124183 also antagonized, i.e. displaced binding at the orthosteric site, with K(i) values of 1.68 and 0.82 microM, respectively. A(3)AR mutagenesis studies implicated F182(5.43) and N274(7.45) in the action of the enhancers and was interpreted using a rhodopsin-based A(3)AR molecular model, suggesting multiple binding modes. Amiloride analogues, SCH-202676 (N-(2,3-diphenyl-1,2,4-thiadiazol-5(2H)-ylidene)methanamine), and sodium ions were demonstrated to be common allosteric modulators for at least three subtypes (A(1), A(2A), and A(3)) of ARs.
More Related Videos
 07:41
A Kinetic Fluorescence-based Ca2+ Mobilization Assay to Identify G Protein-coupled Receptor Agonists, Antagonists, and Allosteric Modulators
Published on: February 20, 2018
 11:29
HPLC-based Assay to Monitor Extracellular Nucleotide/Nucleoside Metabolism in Human Chronic Lymphocytic Leukemia Cells
Published on: July 20, 2016
Related Concept Videos
 01:31
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:08
Allosteric Regulation
01:08Allosteric Regulation
 01:27
Adrenergic Receptors (Adrenoceptors): Classification
α-Adrenoceptors
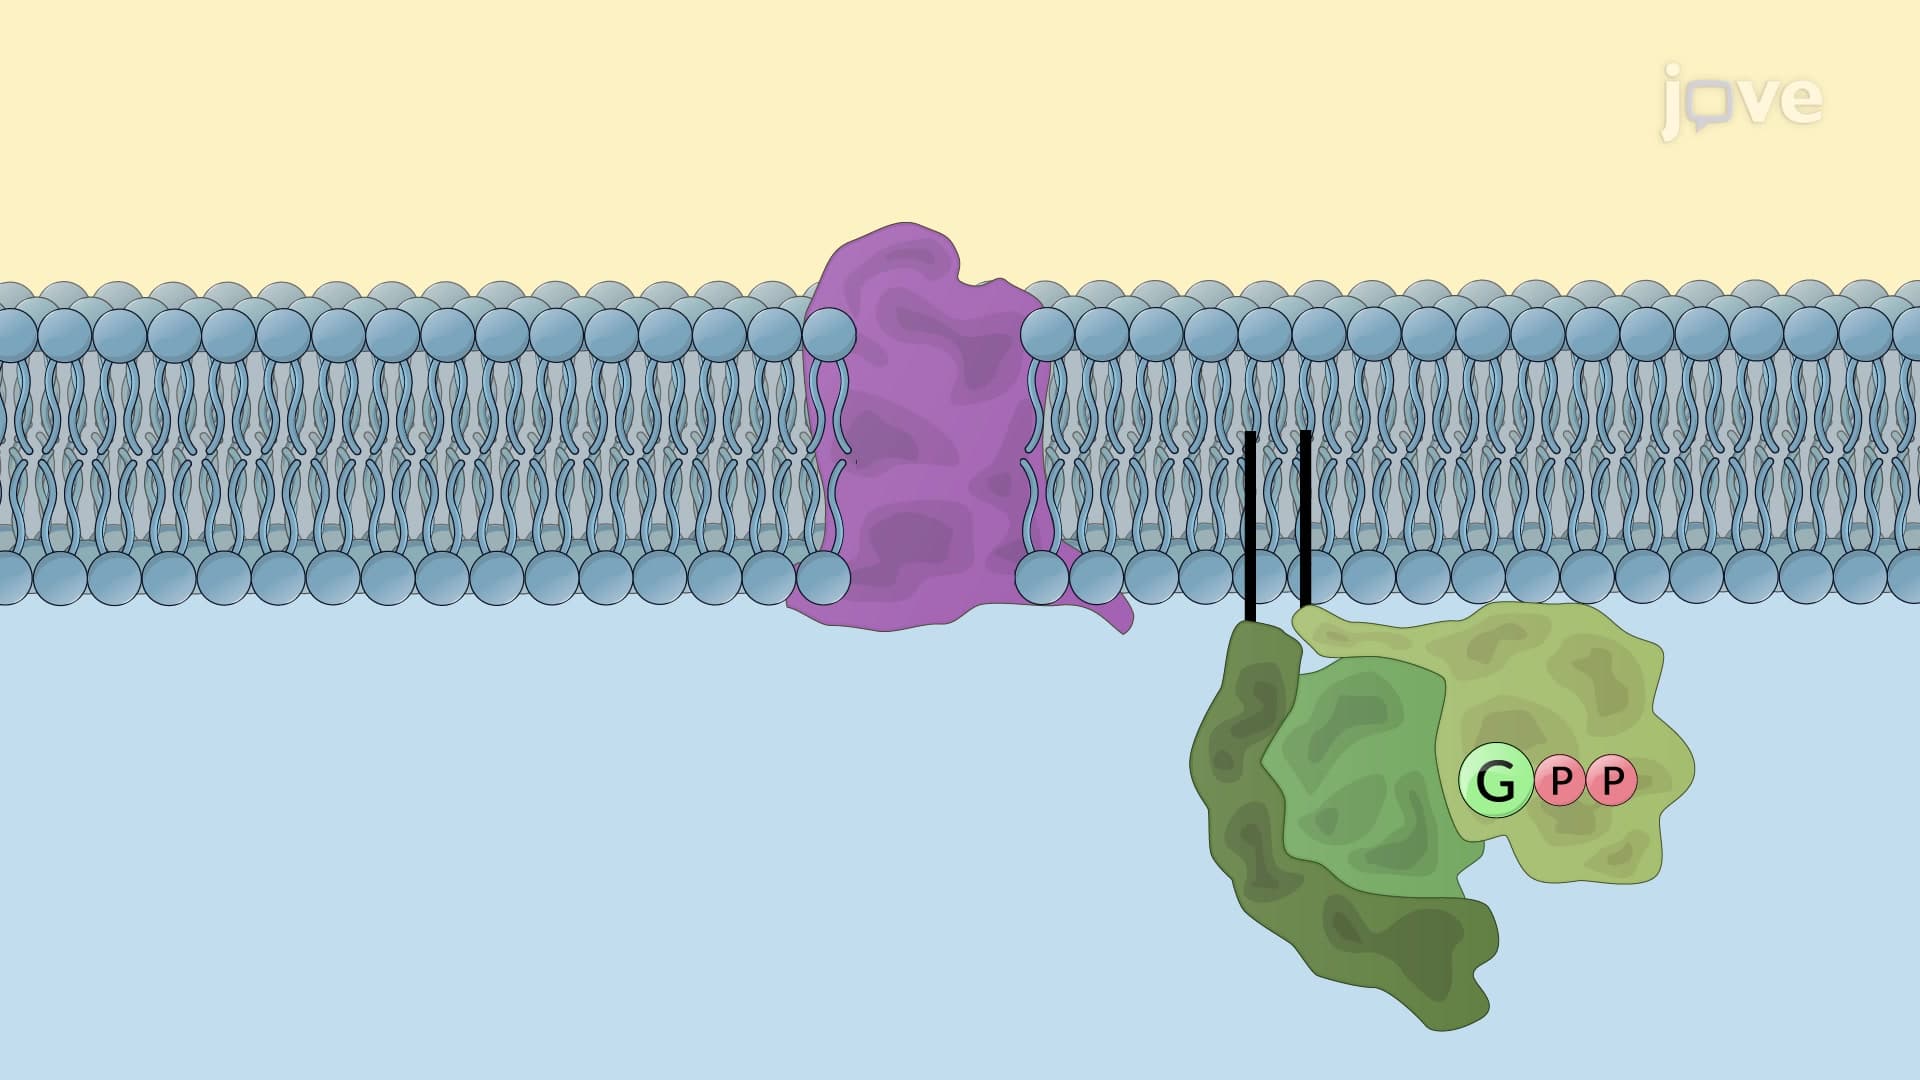
α-Adrenoceptors are classified into two main subtypes: α1 and α2. The α1 adrenoceptors, which are found on postsynaptic...
 01:17
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
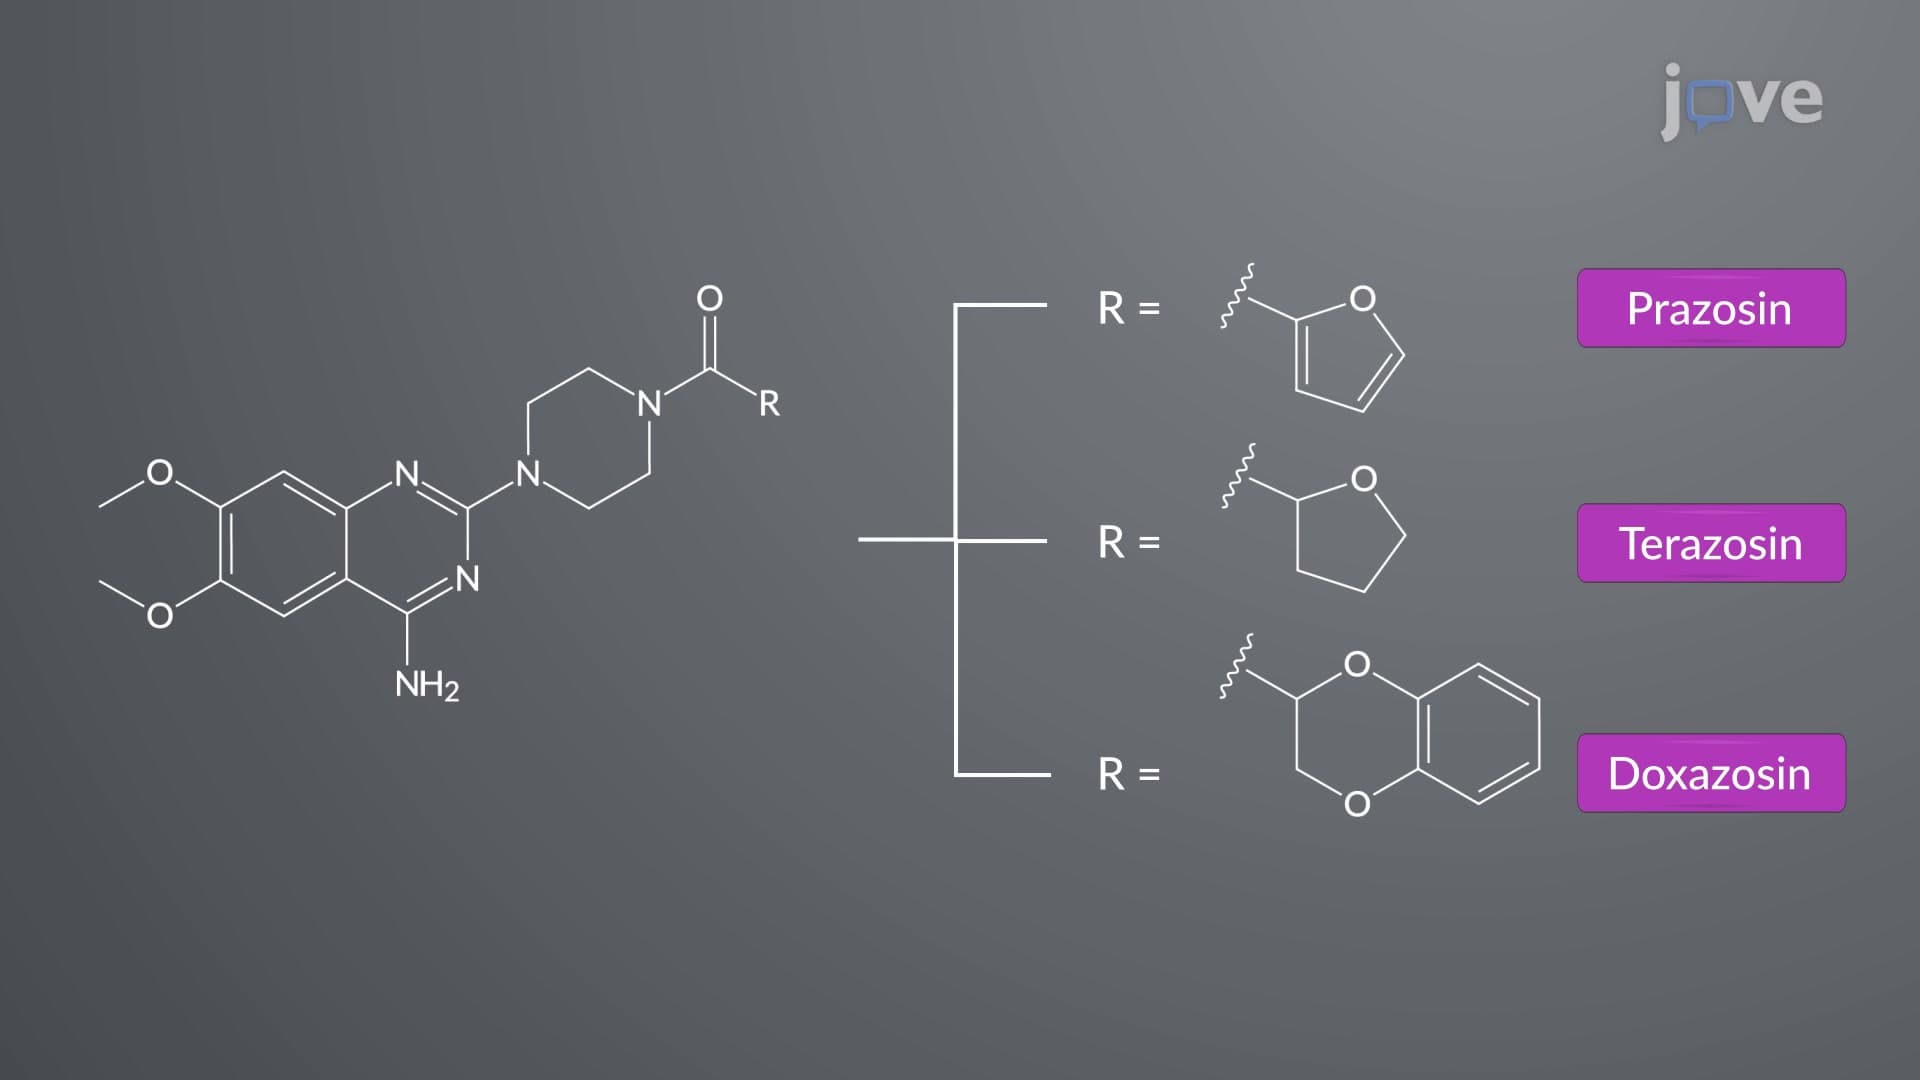
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:25
Adrenergic Agonists: Indirect-Acting Agents
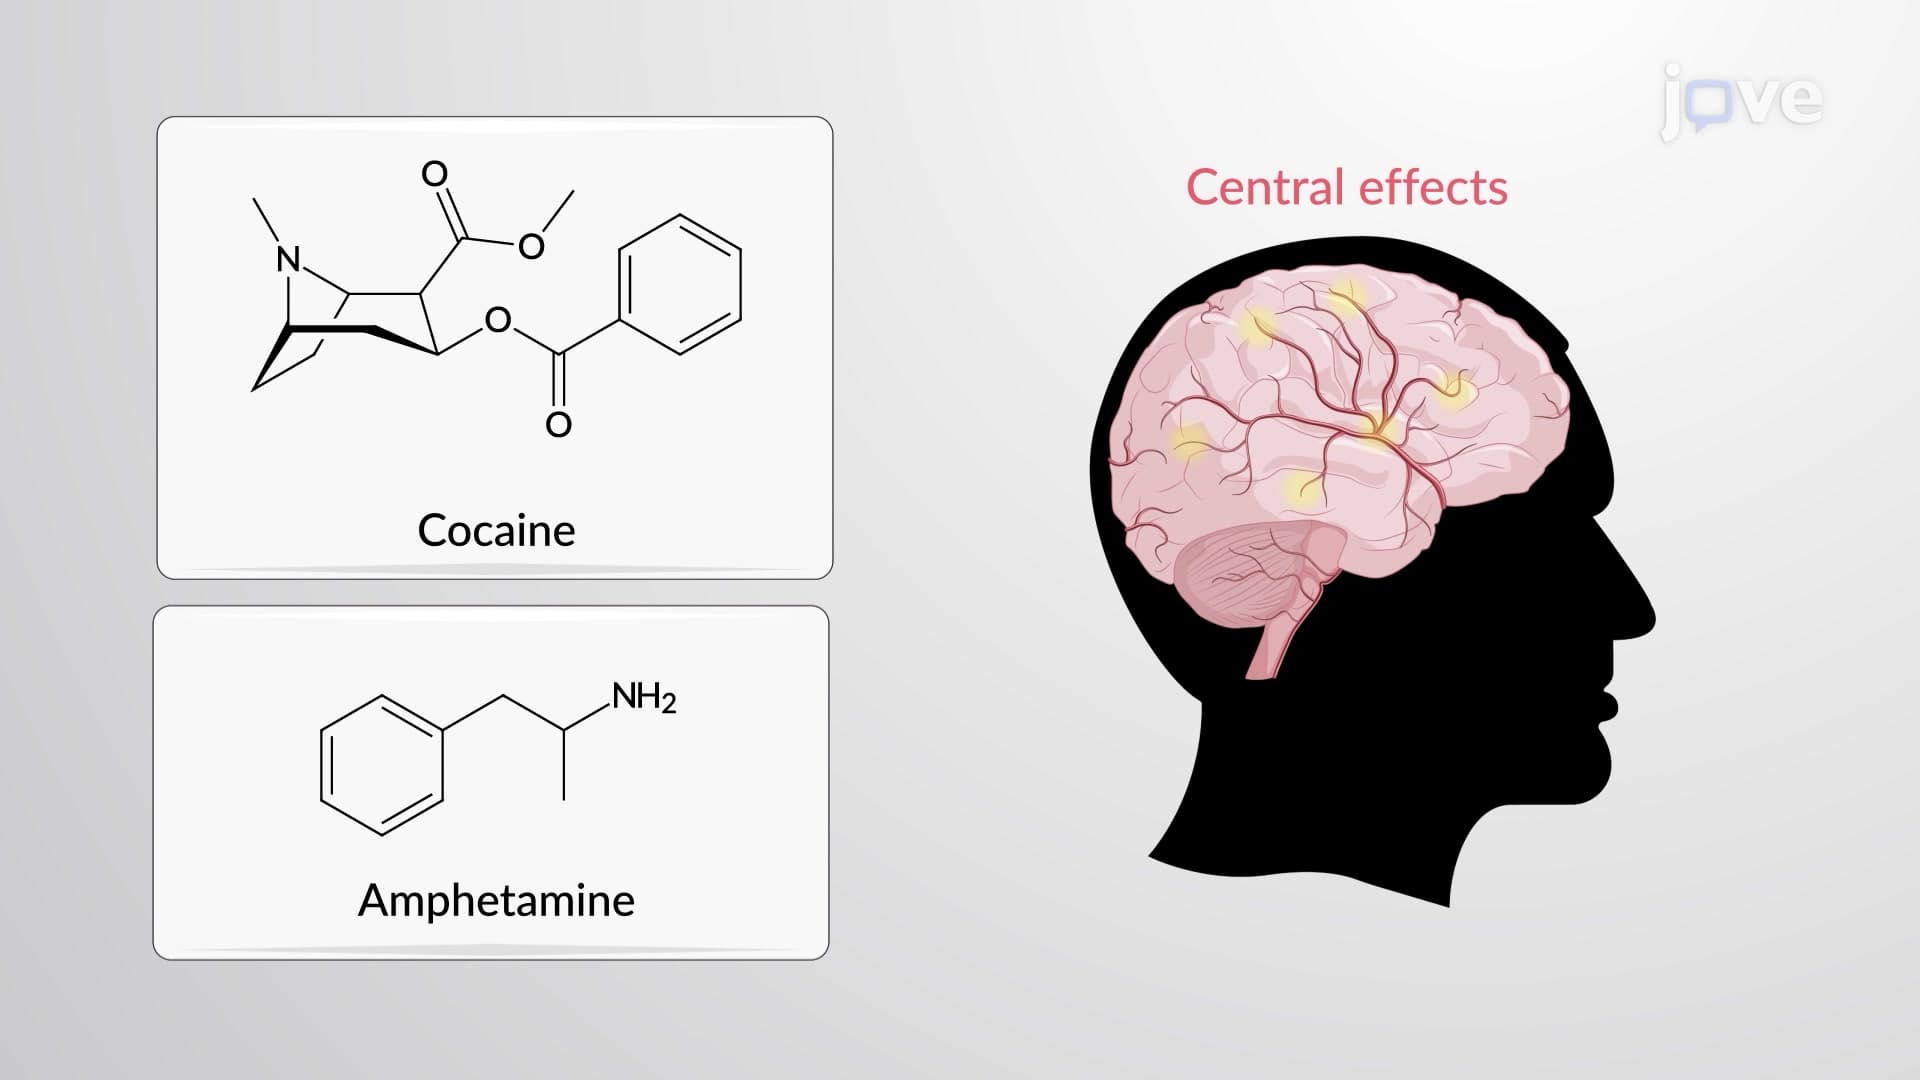
One mechanism involves depleting stored catecholamines by displacing them from synaptic vesicles. These agents, known as "displacers," are transported into vesicles at the expense of noradrenaline. Examples include amphetamine and tyramine, which lack a catechol moiety, resulting in prolonged action, improved oral bioavailability, and...