A multiple-start Monte Carlo docking method
Related Experiment Video
Updated: Aug 17, 2026
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
1Department of Medical Microbiology and Infectious Diseases, University of Alberta, Edmonton, Canada.

Abstract:
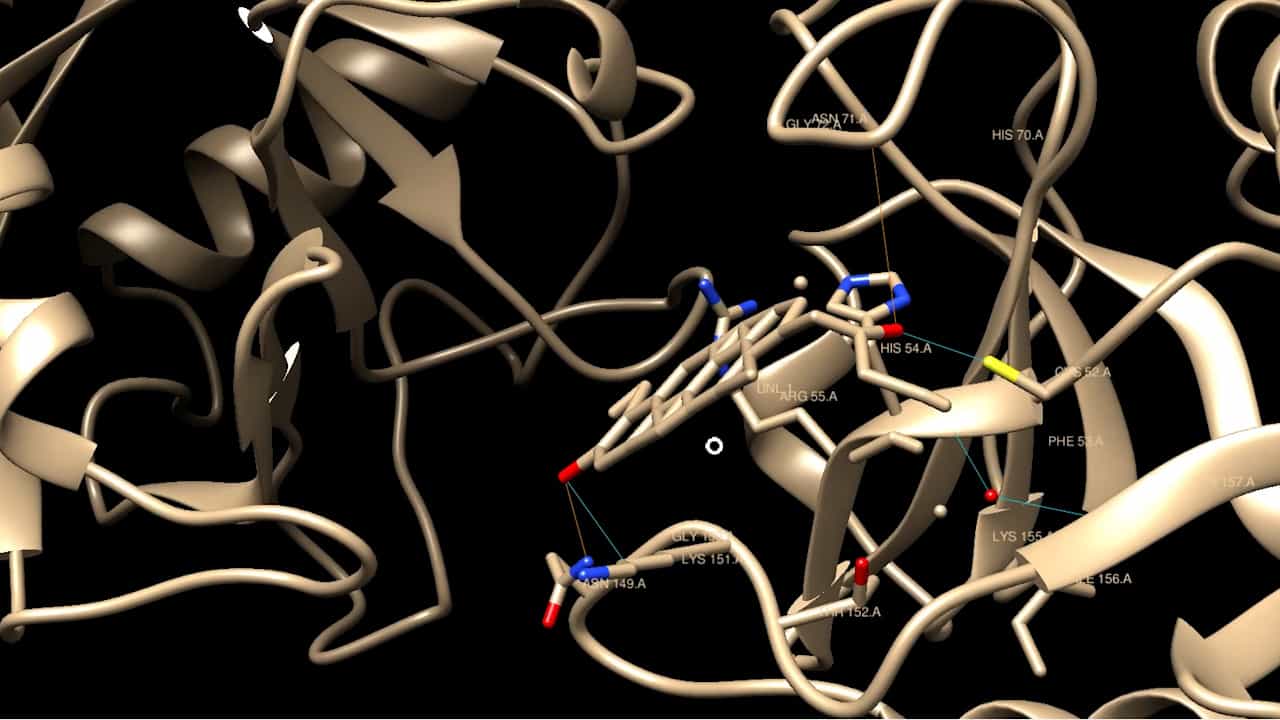
We present a method to search for possible binding modes of molecular fragments at a specific site of a potential drug target of known structure. Our method is based on a Monte Carlo (MC) algorithm applied to the translational and rotational degrees of freedom of the probe fragment. Starting from a randomly generated initial configuration, favorable binding modes are generated using a two-step process. An MC run is first performed in which the energy in the Metropolis algorithm is substituted by a score function that measures the average distance of the probe to the target surface. This has the effect of making buried probes move toward the target surface and also allows enhanced sampling of deep pockets. In a second MC run, a pairwise atom potential function is used, and the temperature parameter is slowly lowered during the run (Simulated Annealing). We repeat this procedure starting from a large number of different randomly generated initial configurations in order to find all energetically favorable docking modes in a specified region around the target. We test this method using two inhibitor-receptor systems: Streptomyces griseus proteinase B in complex with the third domain of the ovomucoid inhibitor from turkey, and dihydrofolate reductase from E. coli in complex with methotrexate. The method could consistently reproduce the complex found in the crystal structure searching from random initial positions in cubes ranging from 25 A to 50 A about the binding site. In the case of SGPB, we were also successful in docking to the native structure. In addition, we were successful in docking small probes in a search that included the entire protein surface.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025