Related Experiment Video
Updated: Aug 16, 2026

Fractionation for Resolution of Soluble and Insoluble Huntingtin Species
Published on: February 27, 2018
HD CAG repeat implicates a dominant property of huntingtin in mitochondrial energy metabolism
Ihn Sik Seong1, Elena Ivanova, Jong-Min Lee
1Molecular Neurogenetics UNIT, Center for Human Genetic Research, Massachusetts General Hospital, Richard B. Simches Research Center, Boston, MA 02114, USA.

Abstract:
The 'expanded' HD CAG repeat that causes Huntington's disease (HD) encodes a polyglutamine tract in huntingtin, which first targets the death of medium-sized spiny striatal neurons. Mitochondrial energetics, related to N-methyl-d-aspartate (NMDA) Ca2+-signaling, has long been implicated in this neuronal specificity, implying an integral role for huntingtin in mitochondrial energy metabolism. As a genetic test of this hypothesis, we have looked for a relationship between the length of the HD CAG repeat, expressed in endogenous huntingtin, and mitochondrial ATP production. In STHdhQ111 knock-in striatal cells, a juvenile onset HD CAG repeat was associated with low mitochondrial ATP and decreased mitochondrial ADP-uptake. This metabolic inhibition was associated with enhanced Ca2+-influx through NMDA receptors, which when blocked resulted in increased cellular [ATP/ADP]. We then evaluated [ATP/ADP] in 40 human lymphoblastoid cell lines, bearing non-HD CAG lengths (9-34 units) or HD-causing alleles (35-70 units). This analysis revealed an inverse association with the longer of the two allelic HD CAG repeats in both the non-HD and HD ranges. Thus, the polyglutamine tract in huntingtin appears to regulate mitochondrial ADP-phosphorylation in a Ca2+-dependent process that fulfills the genetic criteria for the HD trigger of pathogenesis, and it thereby determines a fundamental biological parameter--cellular energy status, which may contribute to the exquisite vulnerability of striatal neurons in HD. Moreover, the evidence that this polymorphism can determine energy status in the non-HD range suggests that it should be tested as a potential physiological modifier in both health and disease.
More Related Videos
Related Concept Videos
 01:21
Huntington Disease l: Introduction
 01:37
Energy to Drive Translocation
Generally, polypeptides are unfolded by two distinct...
 01:48
ATP Synthase: Mechanism
 02:59
Animal Mitochondrial Genetics
 01:19
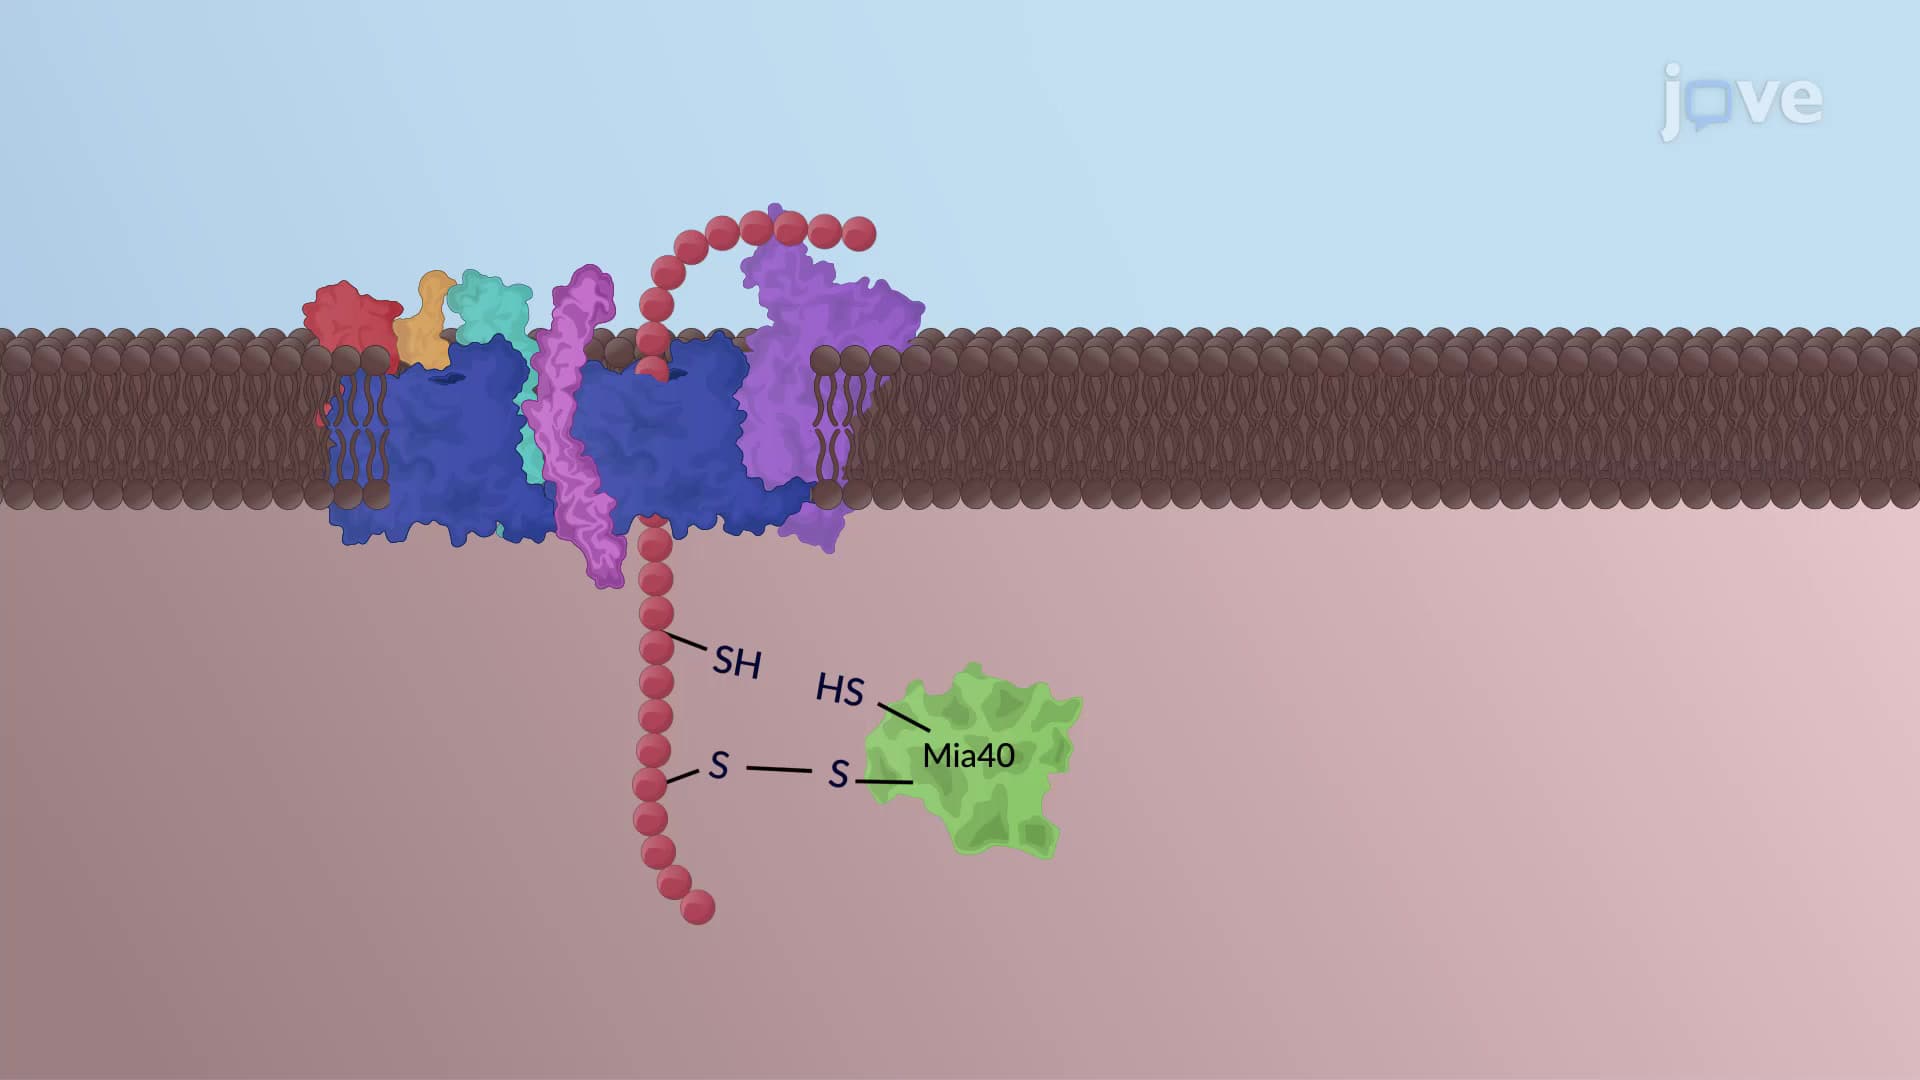
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:11
Genetic Lingo