

IR spectra of N-methylacetamide in water predicted by combined quantum mechanical/molecular mechanical molecular
1Department of Chemistry and Center for Multidimensional Spectroscopy, Division of Chemistry and Molecular Engineering, Korea University, Seoul 136-701, Korea.

Abstract:
We applied the combined quantum mechanical (QM)/molecular mechanical (MM) molecular dynamics (MD) simulation method in assessing IR spectra of N-methylacetamide and its deuterated form in aqueous solutions. The model peptide is treated at the Austin Model 1 (AM1) level and the induced dipole effects by the solvent are incorporated in fluctuating solute dipole moments, which are calculated using partial charges from Mulliken population analyses without resorting to any available high-level ab initio dipole moment data. Fourier transform of the solute dipole autocorrelation function produces in silico IR spectra, in which the relative peak intensities and bandwidths of major amide bands are quantitatively compatible with experimental results only when both geometric and electronic polarizations of the peptide by the solvent are dealt with at the same quantum-mechanical level. We cast light on the importance of addressing dynamic charge fluctuations of the solute in calculating IR spectra by comparing classical and QM/MM MD simulation results. We propose the adjustable scaling factors for each amide mode to be directly compared with experimental data.
More Related Videos
Related Concept Videos
 01:19
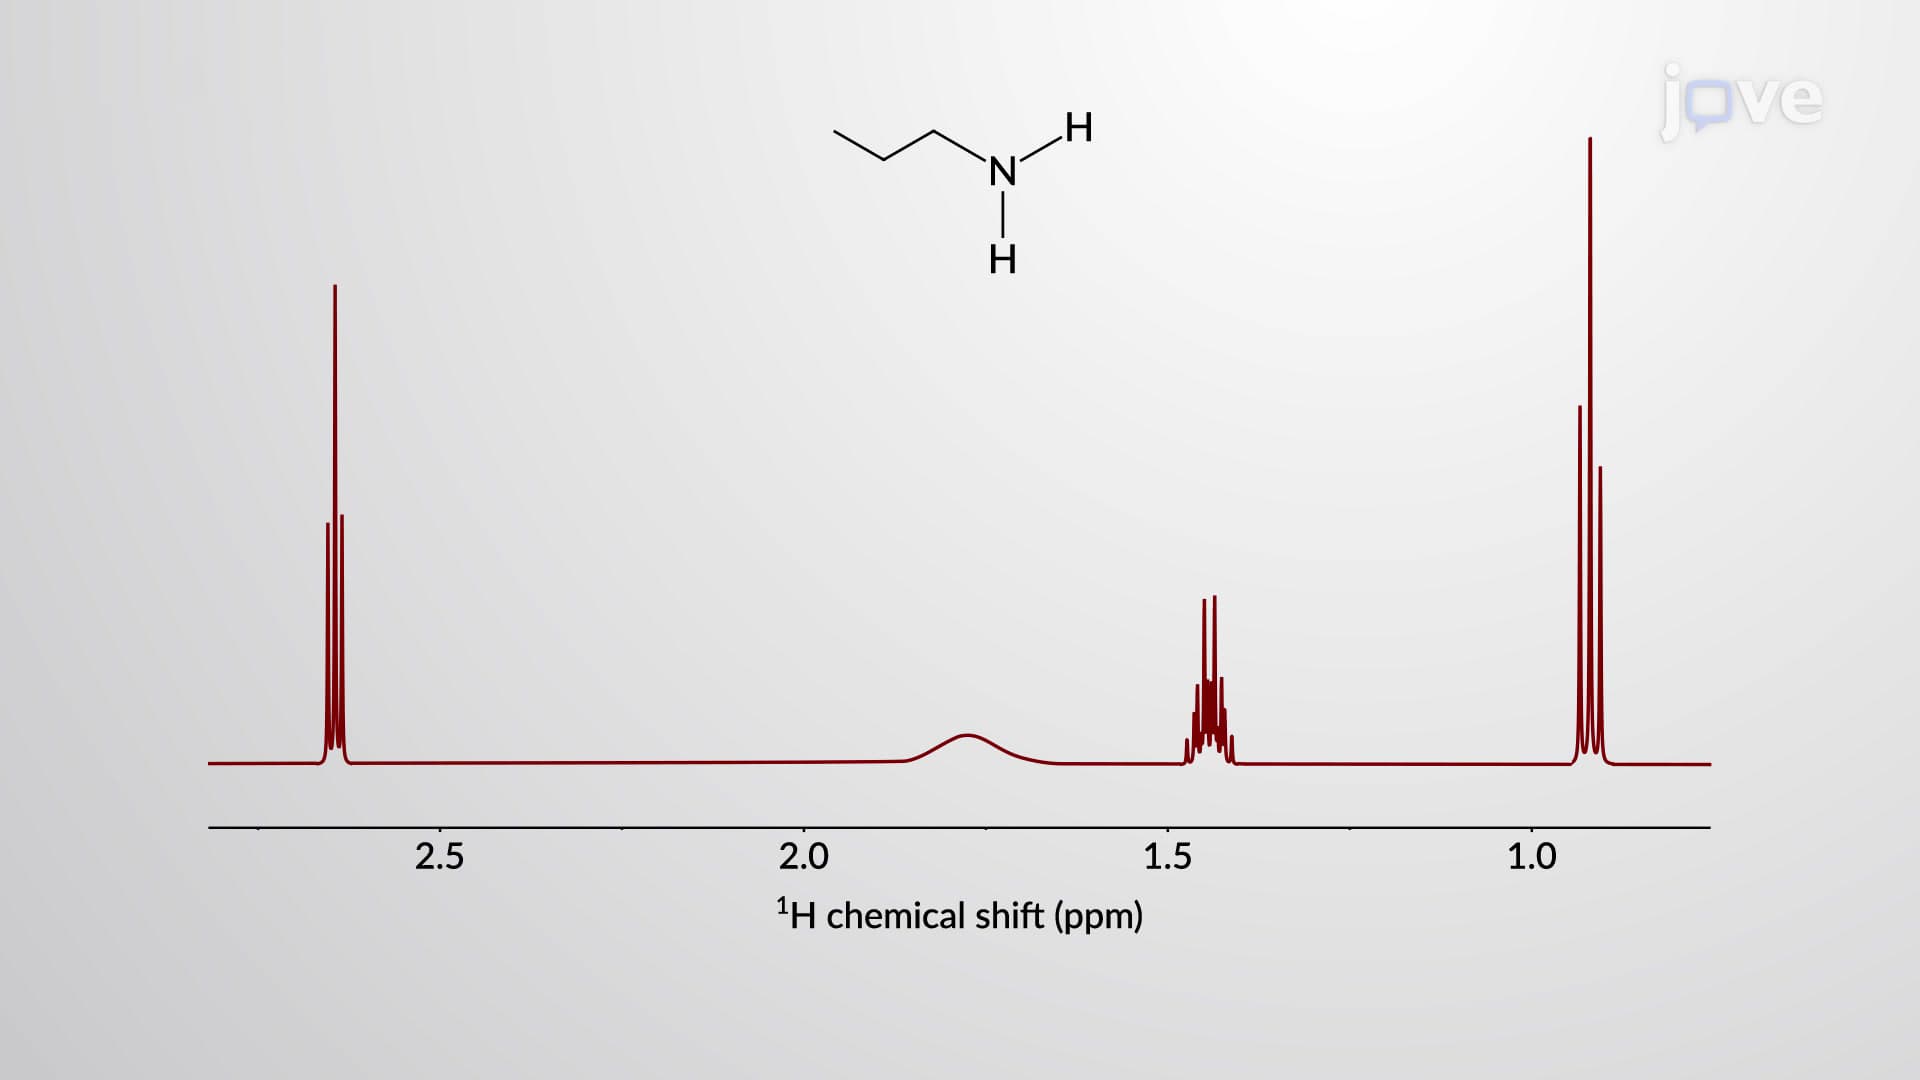
NMR Spectroscopy Of Amines
 00:58
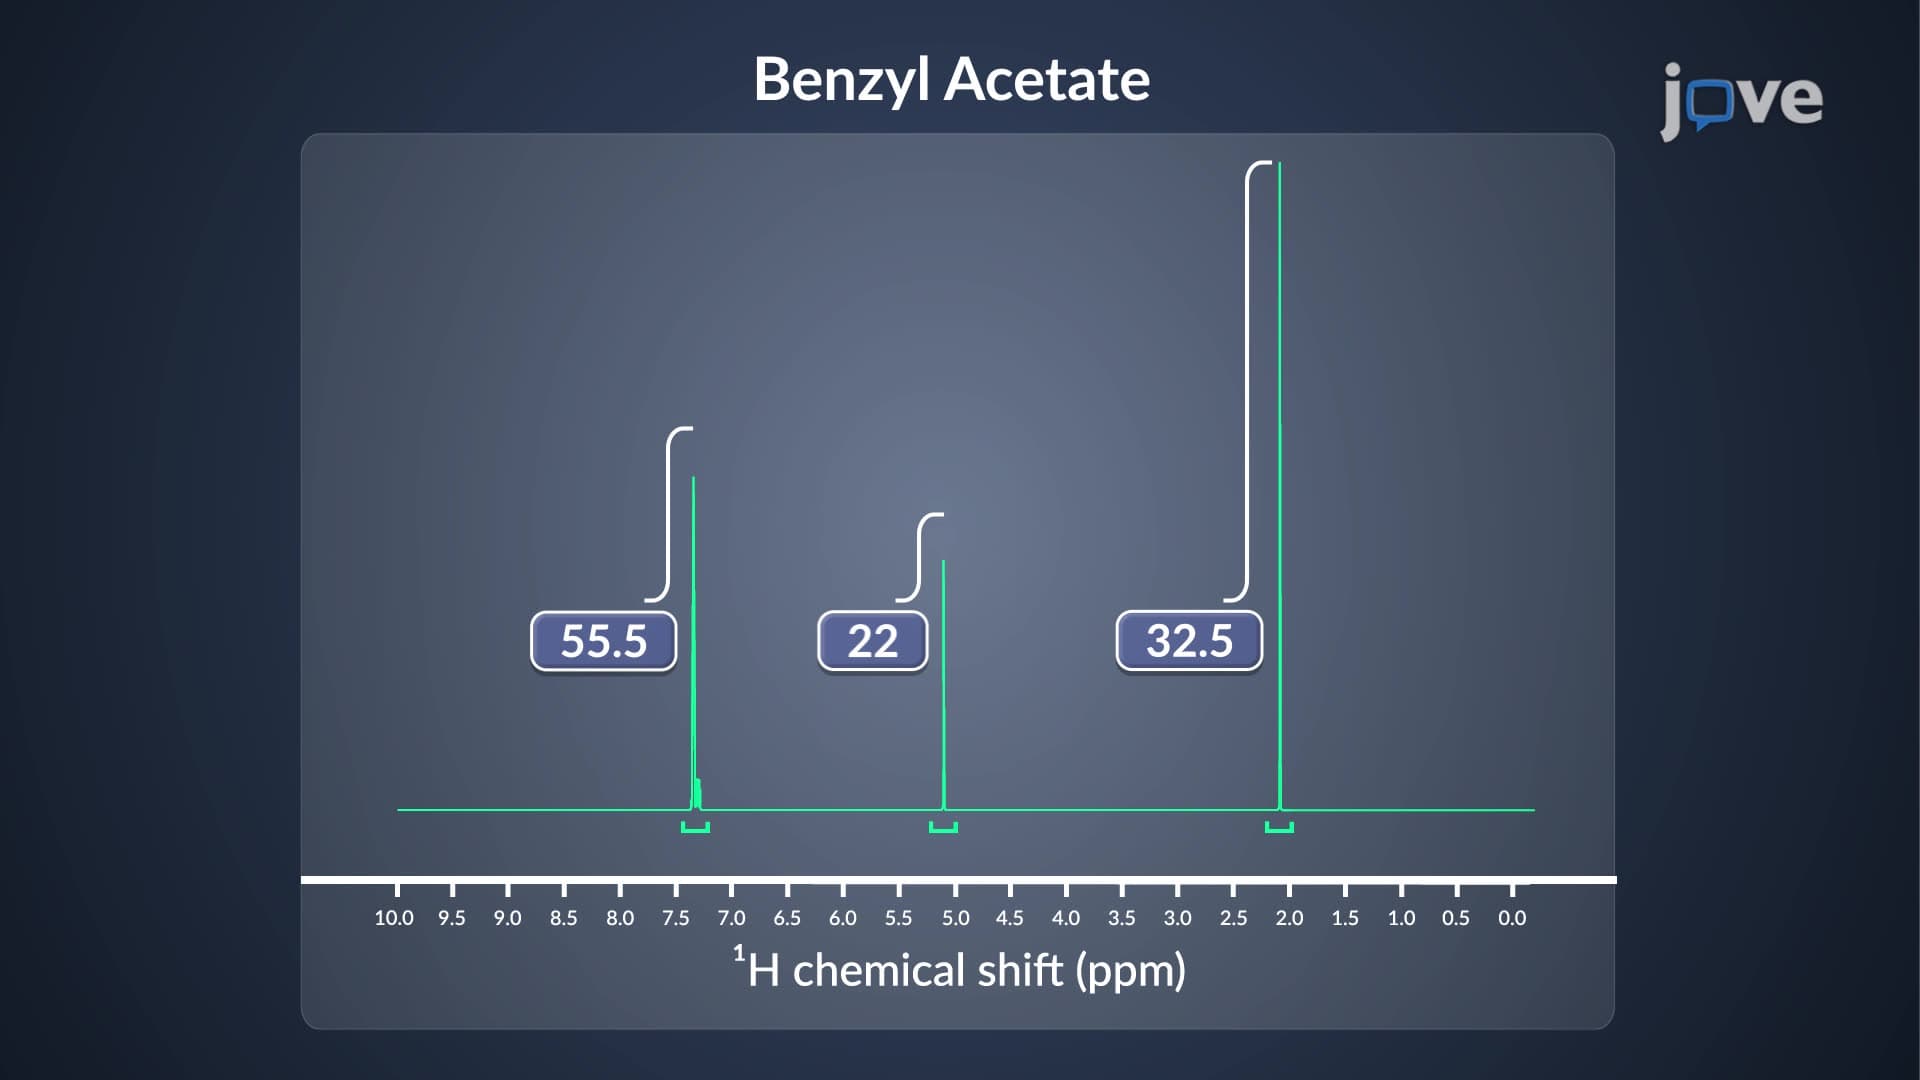
¹H NMR Signal Integration: Overview
 01:26
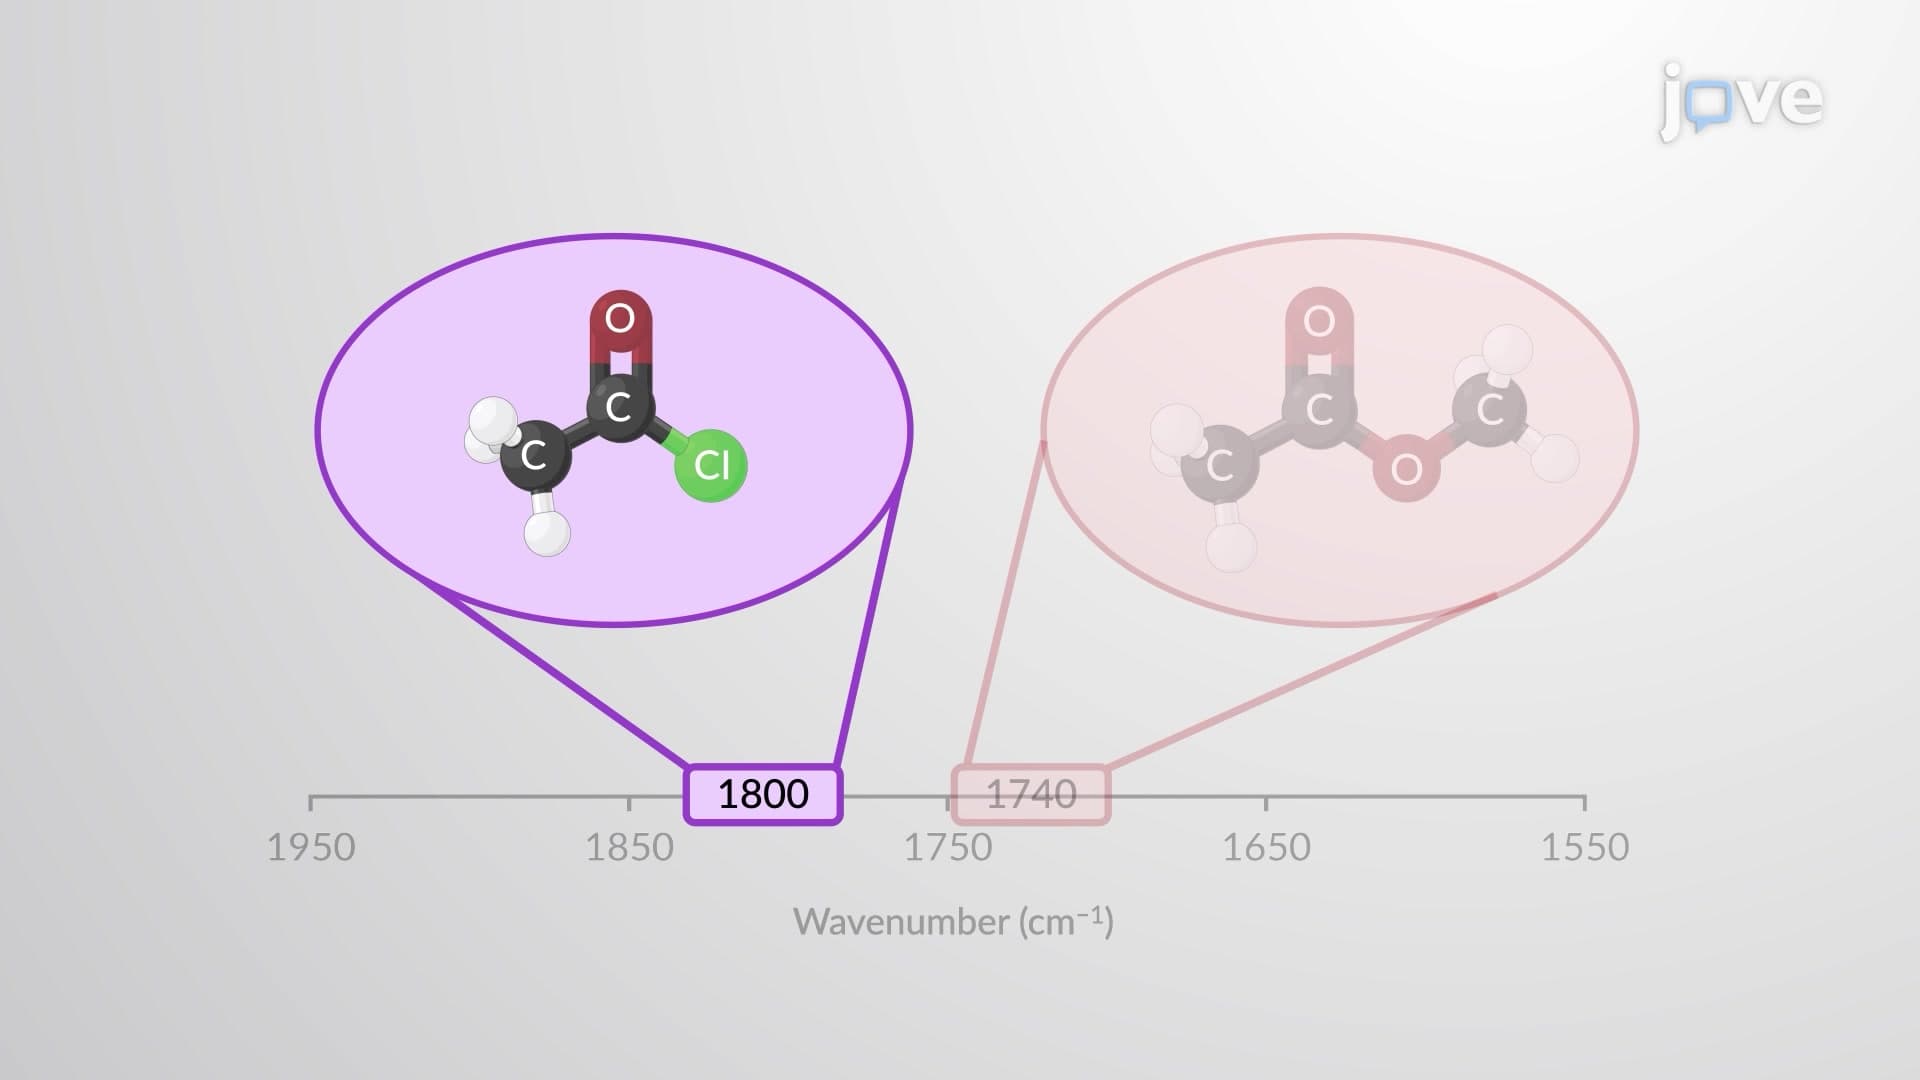
Spectroscopy of Carboxylic Acid Derivatives
In the...
 01:15
Mass Spectrometry of Amines
 01:15
NMR Spectroscopy and Mass Spectrometry of Aldehydes and Ketones
 01:13
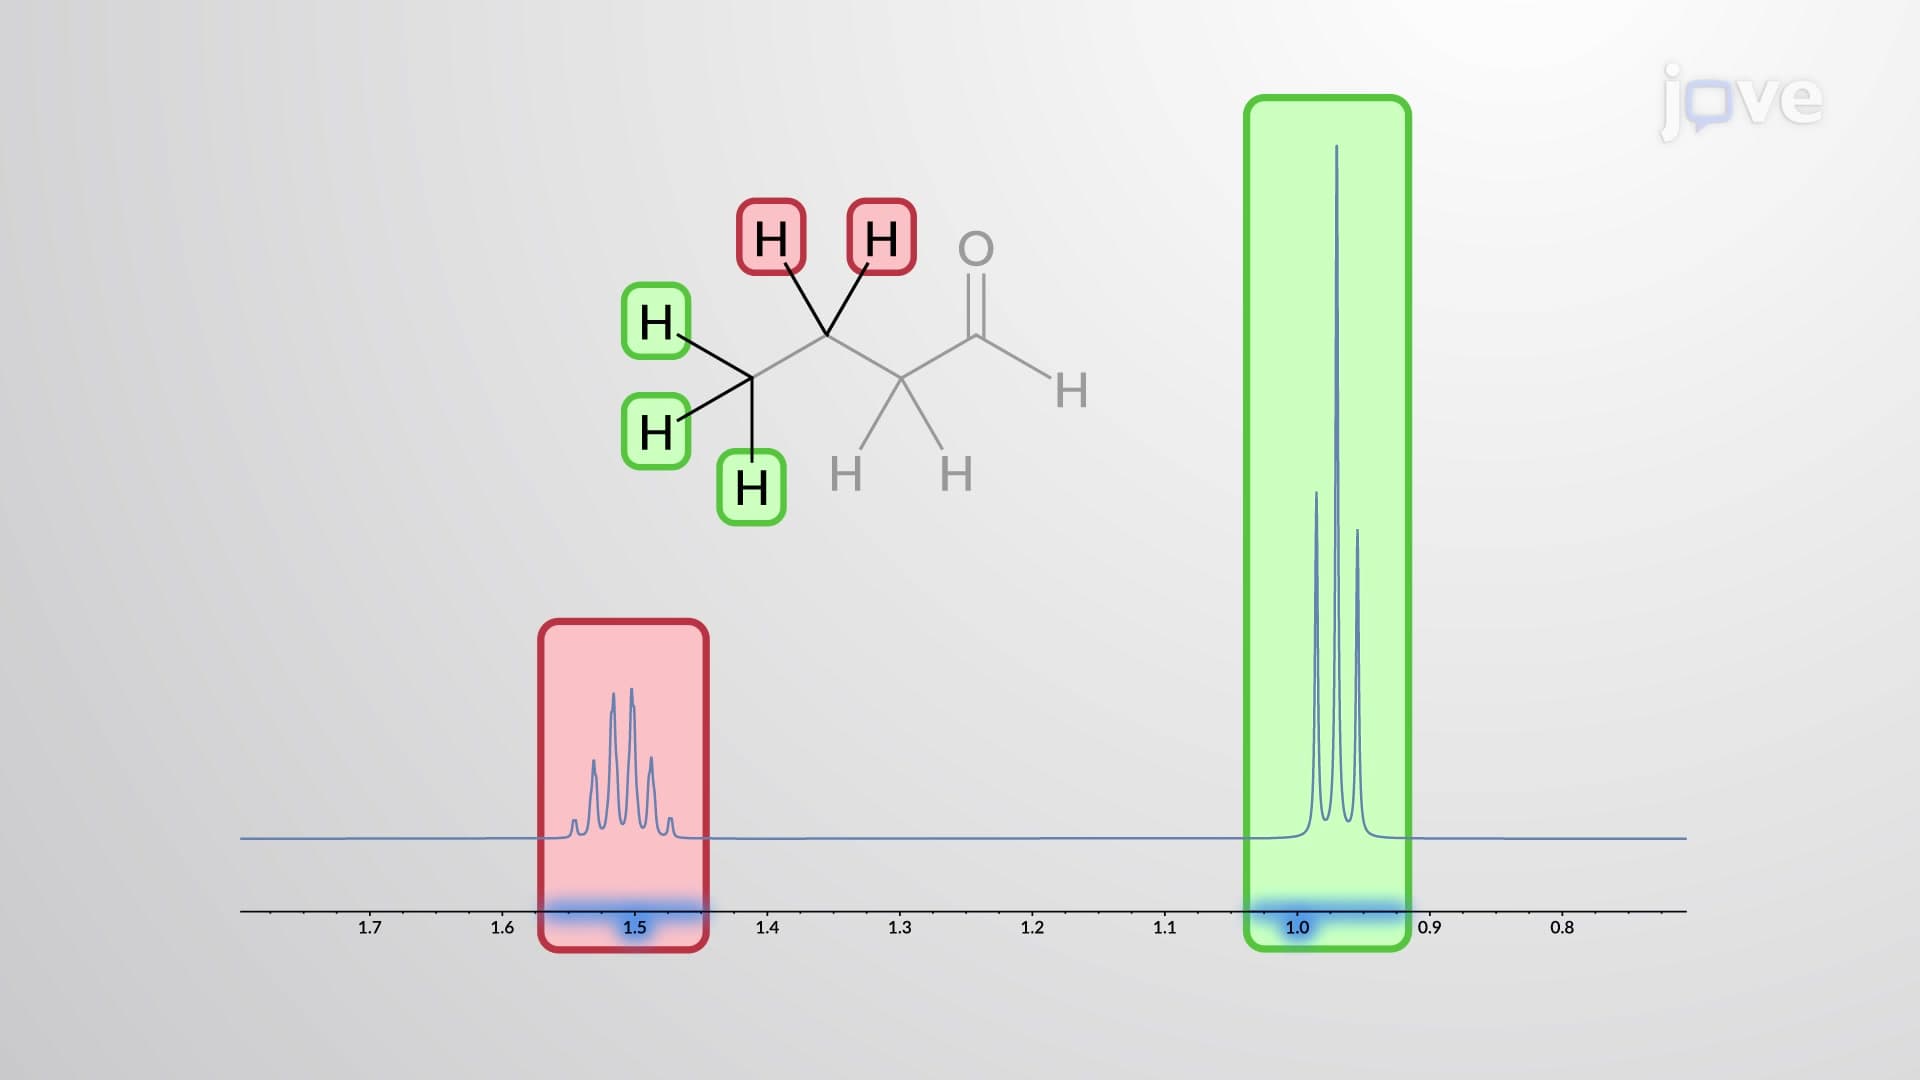
¹H NMR: Complex Splitting
Splitting diagrams or splitting tree diagrams are routinely used to depict such complex couplings. While drawing splitting diagrams, the splitting with the larger coupling constant is usually applied first.