Normal mode refinement: crystallographic refinement of protein dynamic structure applied to human lysozyme
Related Experiment Video
Updated: Aug 10, 2026
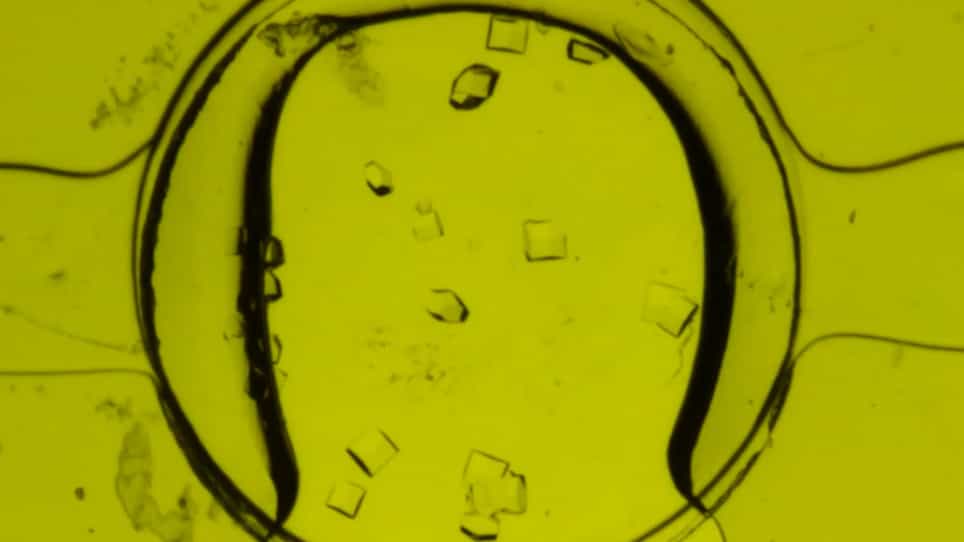
Crystallization of Proteins on Chip by Microdialysis for In Situ X-ray Diffraction Studies
Published on: April 11, 2021
A Kidera1, K Inaka, M Matsushima
1Protein Engineering Research Institute, Osaka, Japan.

Abstract:
A new method of dynamic structure refinement of protein x-ray crystallography, normal mode refinement, is developed. In this method the Debye-Waller factor is expanded in terms of the low-frequency normal modes and external normal modes, whose amplitudes and couplings are optimized in the process of crystallographic refinement. By this method, internal and external contributions to the atomic fluctuations can be separated. Also, anisotropic atomic fluctuations and their interatomic correlations can be determined experimentally even with a relatively small number of adjustable parameters. The method is applied to the analysis of experimental data of human lysozyme to reveal its dynamic structure.
More Related Videos
 10:10
Neutron Crystallography Data Collection and Processing for Modelling Hydrogen Atoms in Protein Structures
Published on: December 1, 2020
 10:45
Crystallization and Structural Determination of an Enzyme:Substrate Complex by Serial Crystallography in a Versatile Microfluidic Chip
Published on: March 20, 2021