Related Experiment Video
Updated: Aug 14, 2026

Radiochemical Assessment of Glycogen Synthase Enzyme Activity in Animal Tissue
Published on: October 24, 2025
Hepatic glycogen synthase deficiency: an infrequently recognized cause of ketotic hypoglycemia
David A Weinstein1, Catherine E Correia, Andrew C Saunders
1Division of Pediatric Endocrinology, University of Florida College of Medicine, Gainesville, USA. WeinsDA@peds.ufl.edu

Abstract:
The glycogen storage diseases comprise several inherited diseases caused by abnormalities of enzymes that regulate the synthesis or degradation of glycogen. In contrast to the classic hepatic glycogen storage diseases that are characterized by fasting hypoglycemia and hepatomegaly, the liver is not enlarged in GSD0. Patients with GSD0 typically have fasting ketotic hypoglycemia without prominent muscle symptoms. Most children are cognitively and developmentally normal. Short stature and osteopenia are common features, but other long-term complications, common in other types of GSD, have not been reported in GSD0. Until recently, the definitive diagnosis of GSD0 depended on the demonstration of decreased hepatic glycogen on a liver biopsy. The need for an invasive procedure may be one reason that this condition has been infrequently diagnosed. Mutation analysis of the GYS2 gene (12p12.2) is a non-invasive method for making this diagnosis in patients suspected to have this disorder. This mini-review discusses the pathophysiology of this disorder, use of mutation analysis to diagnose GSD0, and the clinical characteristics of all reported cases of GSD0.
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:26
Hypoglycemia
 01:15
Hypoglycemia and Glucagon
 01:22
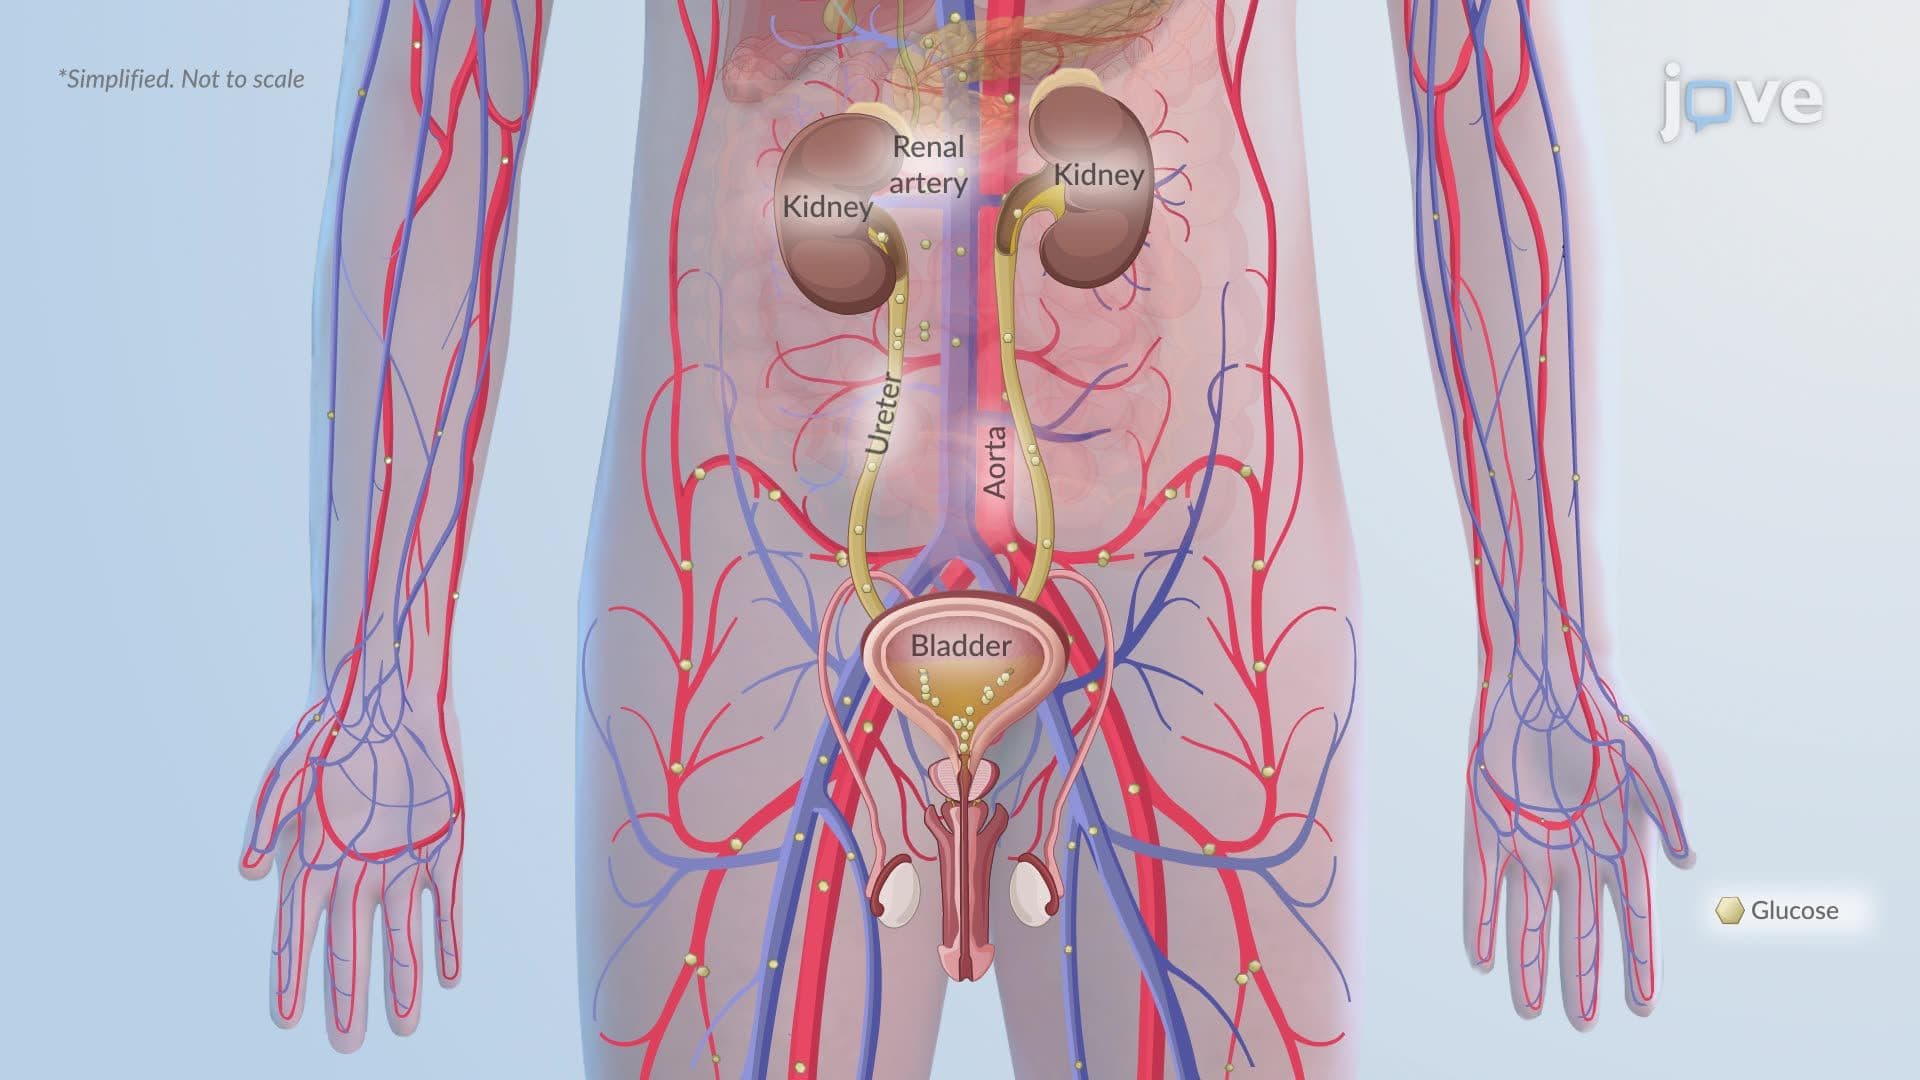
Diabetic Ketoacidosis ll: Pathophysiology
 01:27
Glucose Transporters
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes:
 01:29
Hyperglycemia