Related Experiment Video
Updated: Aug 7, 2026

Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
Docking studies of agonists and antagonists suggest an activation pathway of the A3 adenosine receptor
Soo-Kyung Kim1, Zhan-Guo Gao, Lak Shin Jeong
1Molecular Recognition Section, Laboratory of Bioorganic Chemistry, National Institute of Diabetes and Digestive and Kidney Diseases (NIDDK), National Institutes of Health (NIH), Bethesda, MD 20892, USA.

Abstract:
Structural determinants of ligand efficacy in the human A(3) adenosine receptor (AR) were studied using pharmacophore and docking analyses of various categories of A(3) selective ligands: inverse agonist, neutral antagonist (nonnucleoside and nucleoside), and agonist (partial and full). The homology modeling of GPCRs was adapted to provide two templates: the rhodopsin-based resting state for antagonist binding and a putative Meta I state, conformationally altered at a key residue (W6.48), for agonist binding. The preferential binding domains and/or local conformational changes associated with docking of three high affinity A(3)AR ligands were compared: inverse agonist PSB-11 1 ((R)-8-ethyl-4-methyl-2-phenyl-imidazo[2,1-i]purin-5-one); neutral antagonist MRE-3008F20 7 (5-[[(4-methoxyphenyl)amino]carbonyl]amino-8-methyl-2-(2-furyl)pyra-zolo[4,3-e]1,2,4-triazolo[1,5-c]pyrimidine), and full agonist Cl-IB-MECA 21 (2-chloro-N(6)-(3-iodobenzyl)-5'-N-methylcarboxamidoadenosine) to define a distinct recognition mode for each. Ribose-containing agonists were more hydrophilic than nonnucleoside antagonists, and H-bonding ability at the ribose 3'- and 5'-positions was required for agonism. From the receptor perspective, common requirements for activation included the destabilization of H-bond networks at W6.48 and H7.43, the specific interactions of the ribose moiety in its putative hydrophilic pocket at T3.36, S7.42, and H7.43, the stabilization of the complex by inward movement of F5.43, and the characteristic rotation of W6.48. By analogy, outward rotation of the W6.48 side-chain upon activation of an internally-crosslinking mutant M(3) muscarinic receptor was indicated by constrained molecular dynamics (MD). Our results are consistent with an anti-clockwise rotation (from the extracellular view) of transmembrane domains 3, 5, 6, and 7, as proposed for other Family A GPCRs. Thus, the putative conformational changes associated with A(3)AR activation indicate a shared mechanism of GPCR activation similar to rhodopsin.
Related Concept Videos
 01:25
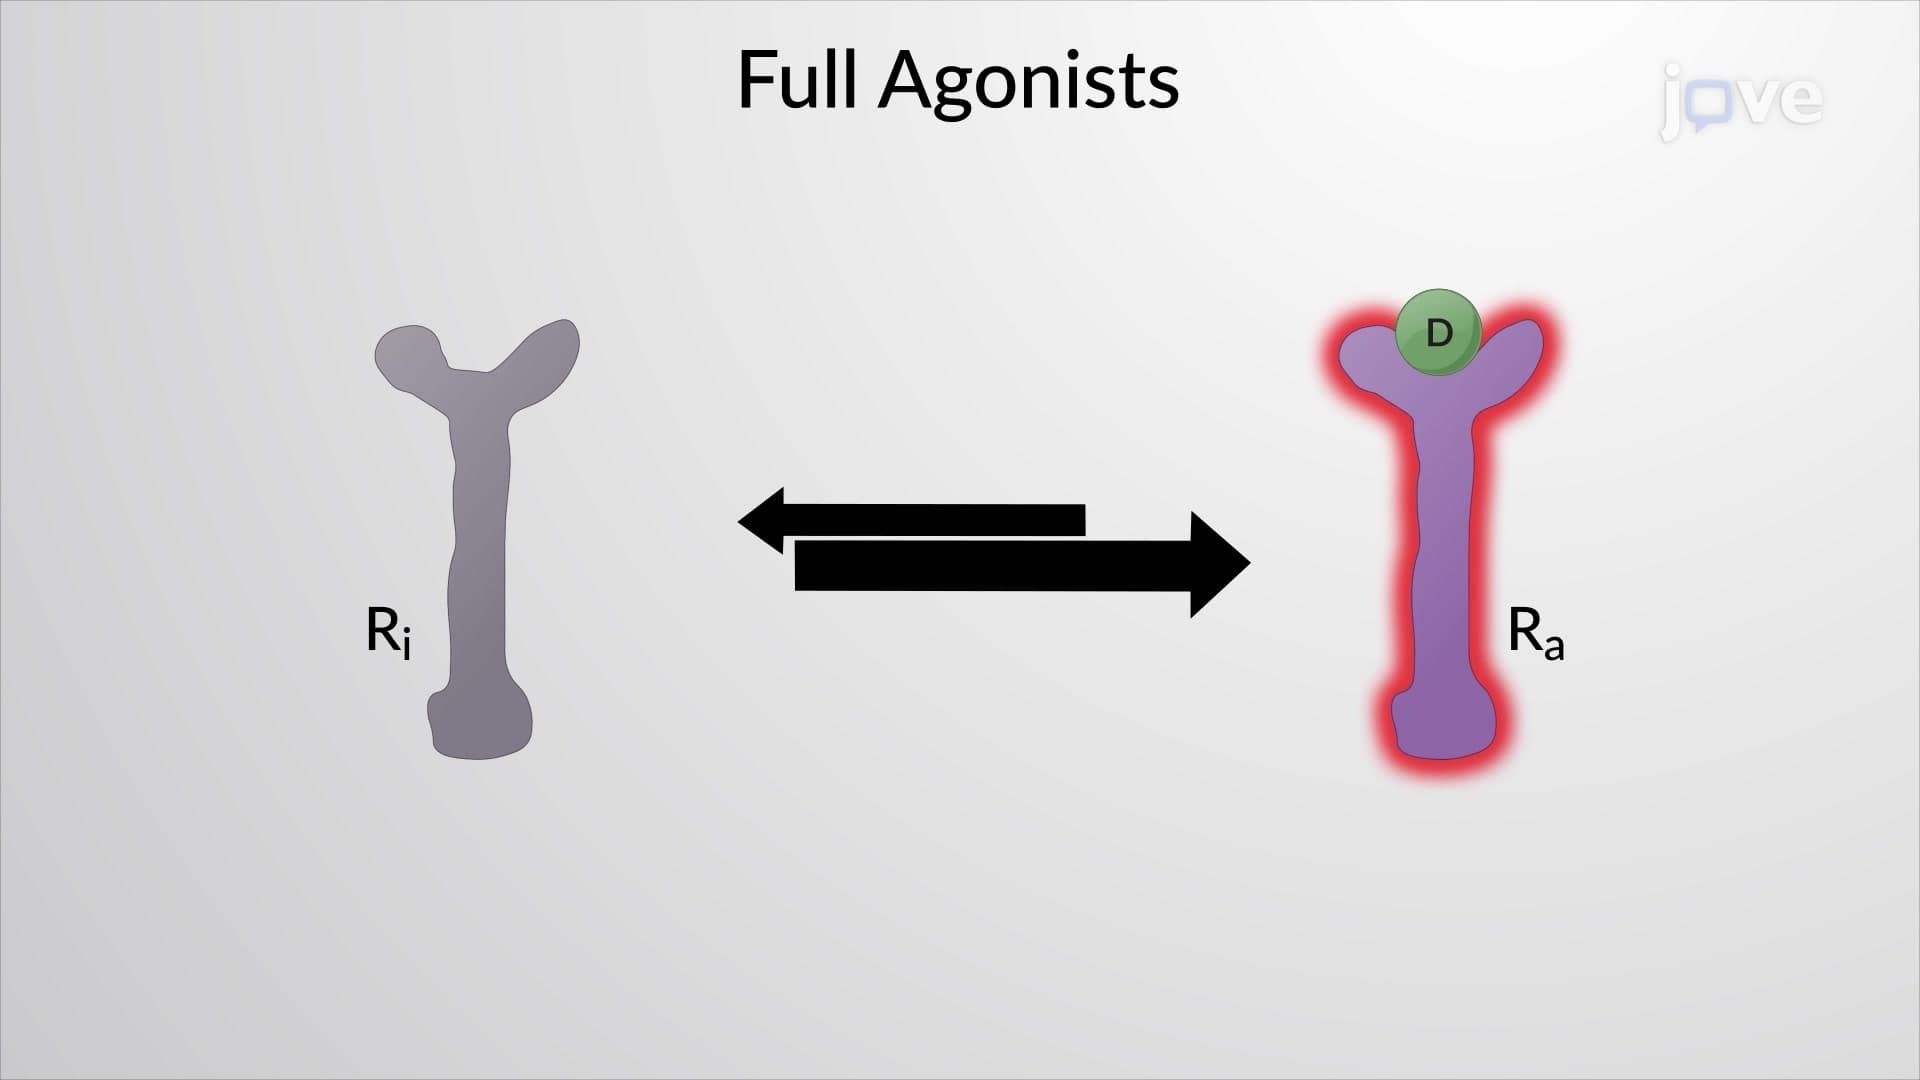
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous ligand's action.
 01:31
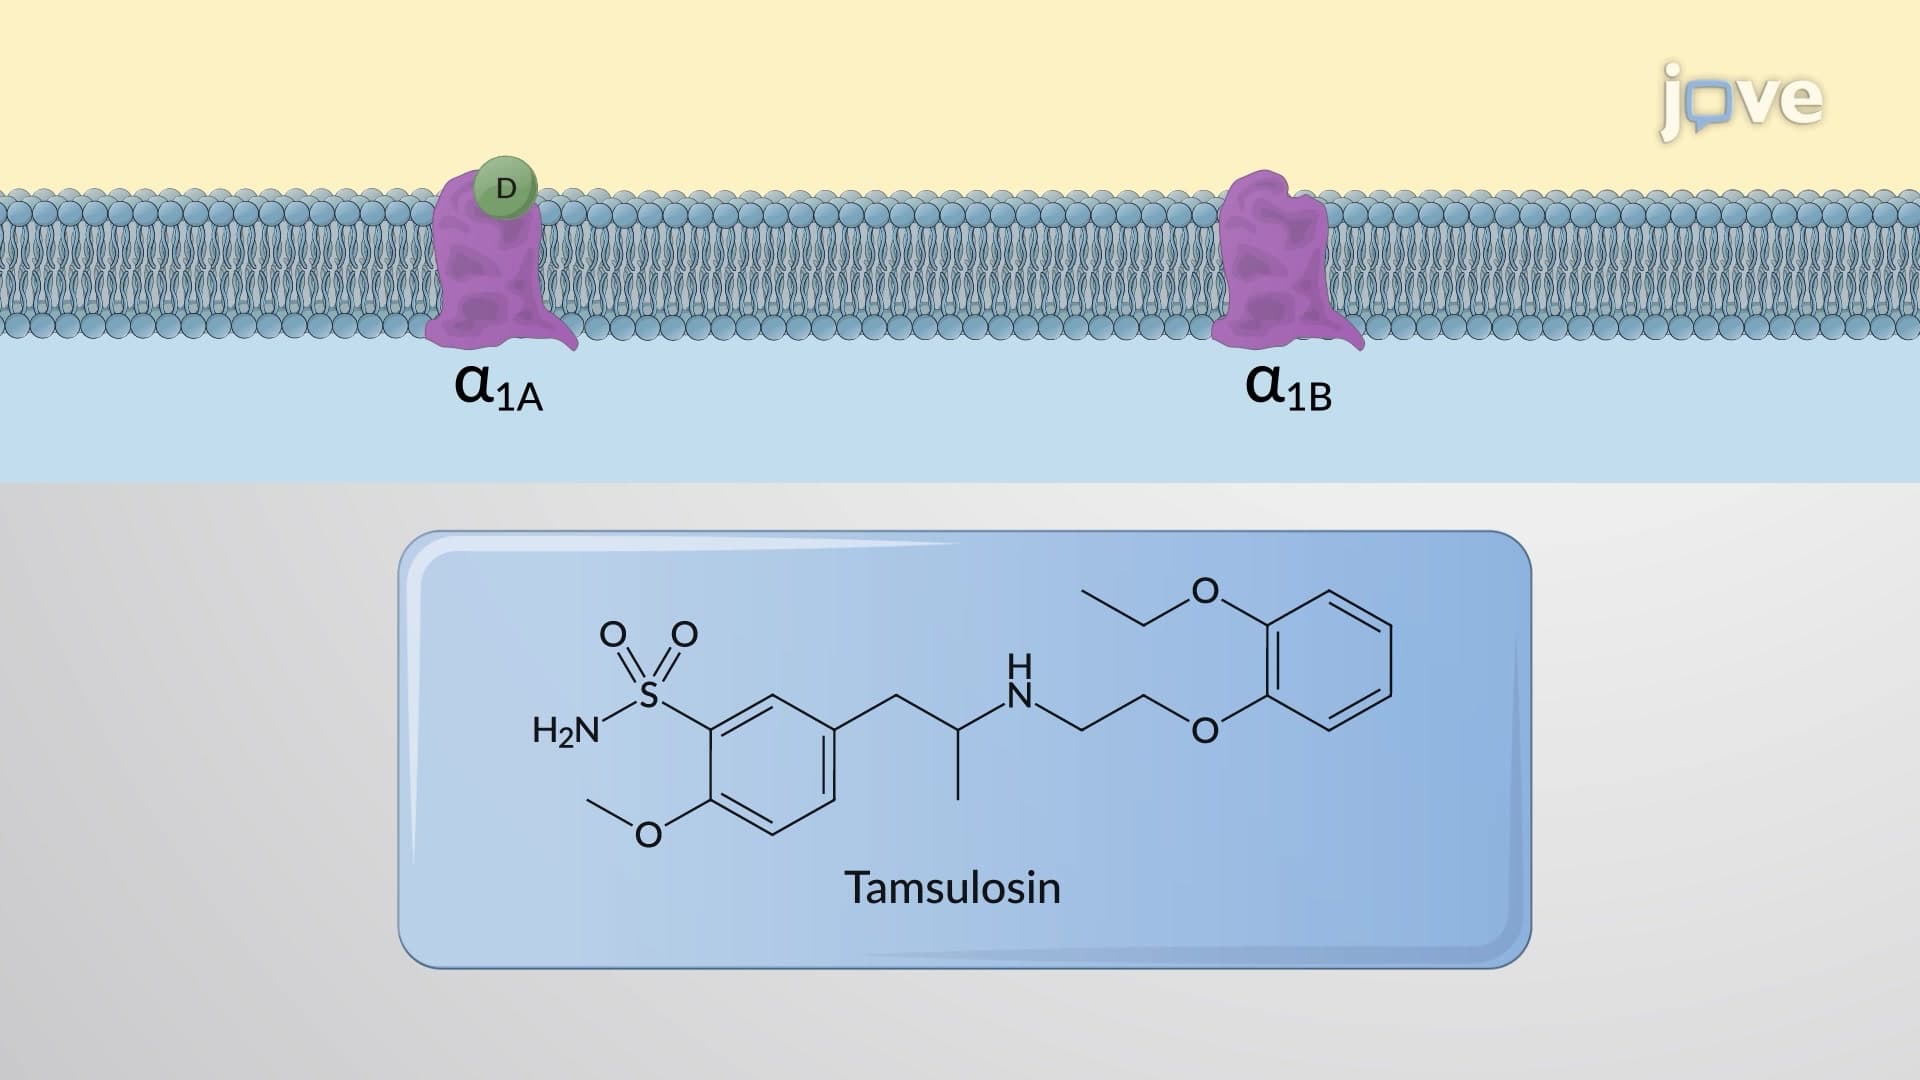
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:29
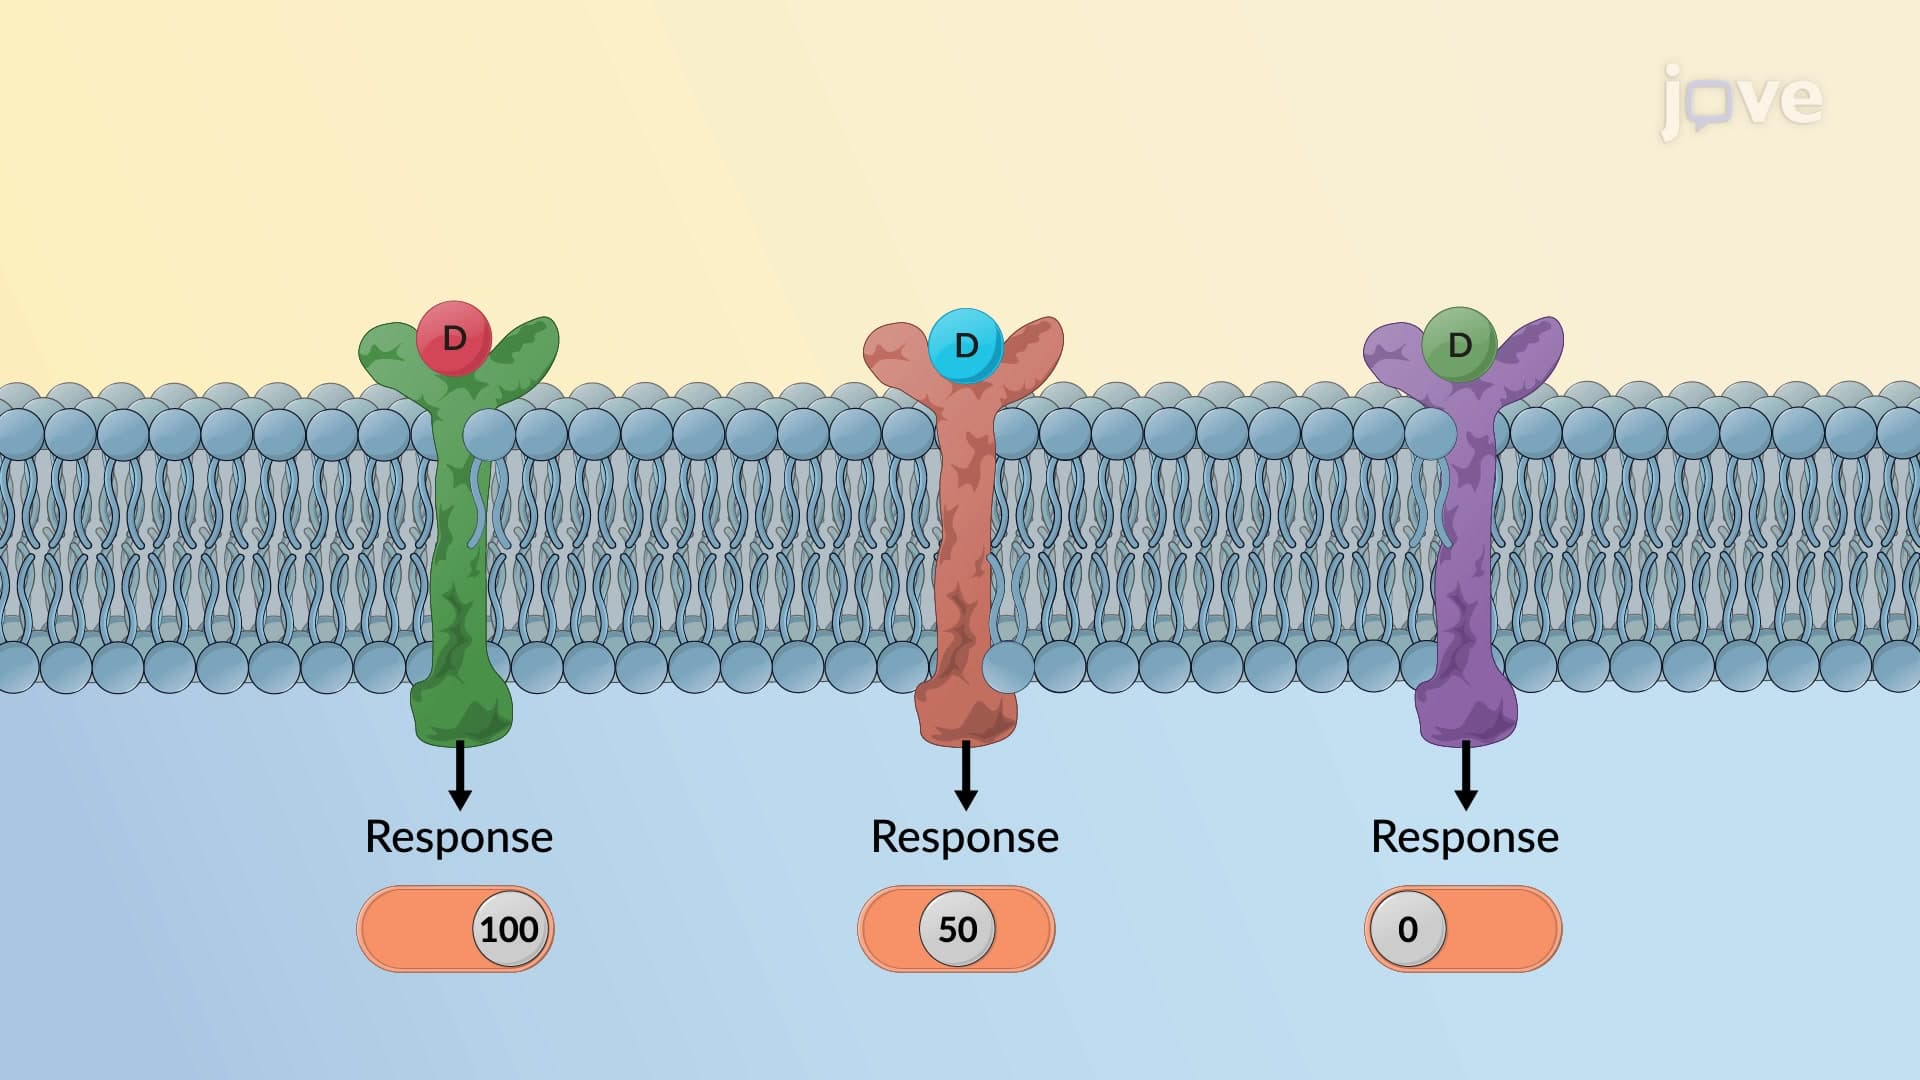
Drug-Receptor Interactions
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue.
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:27
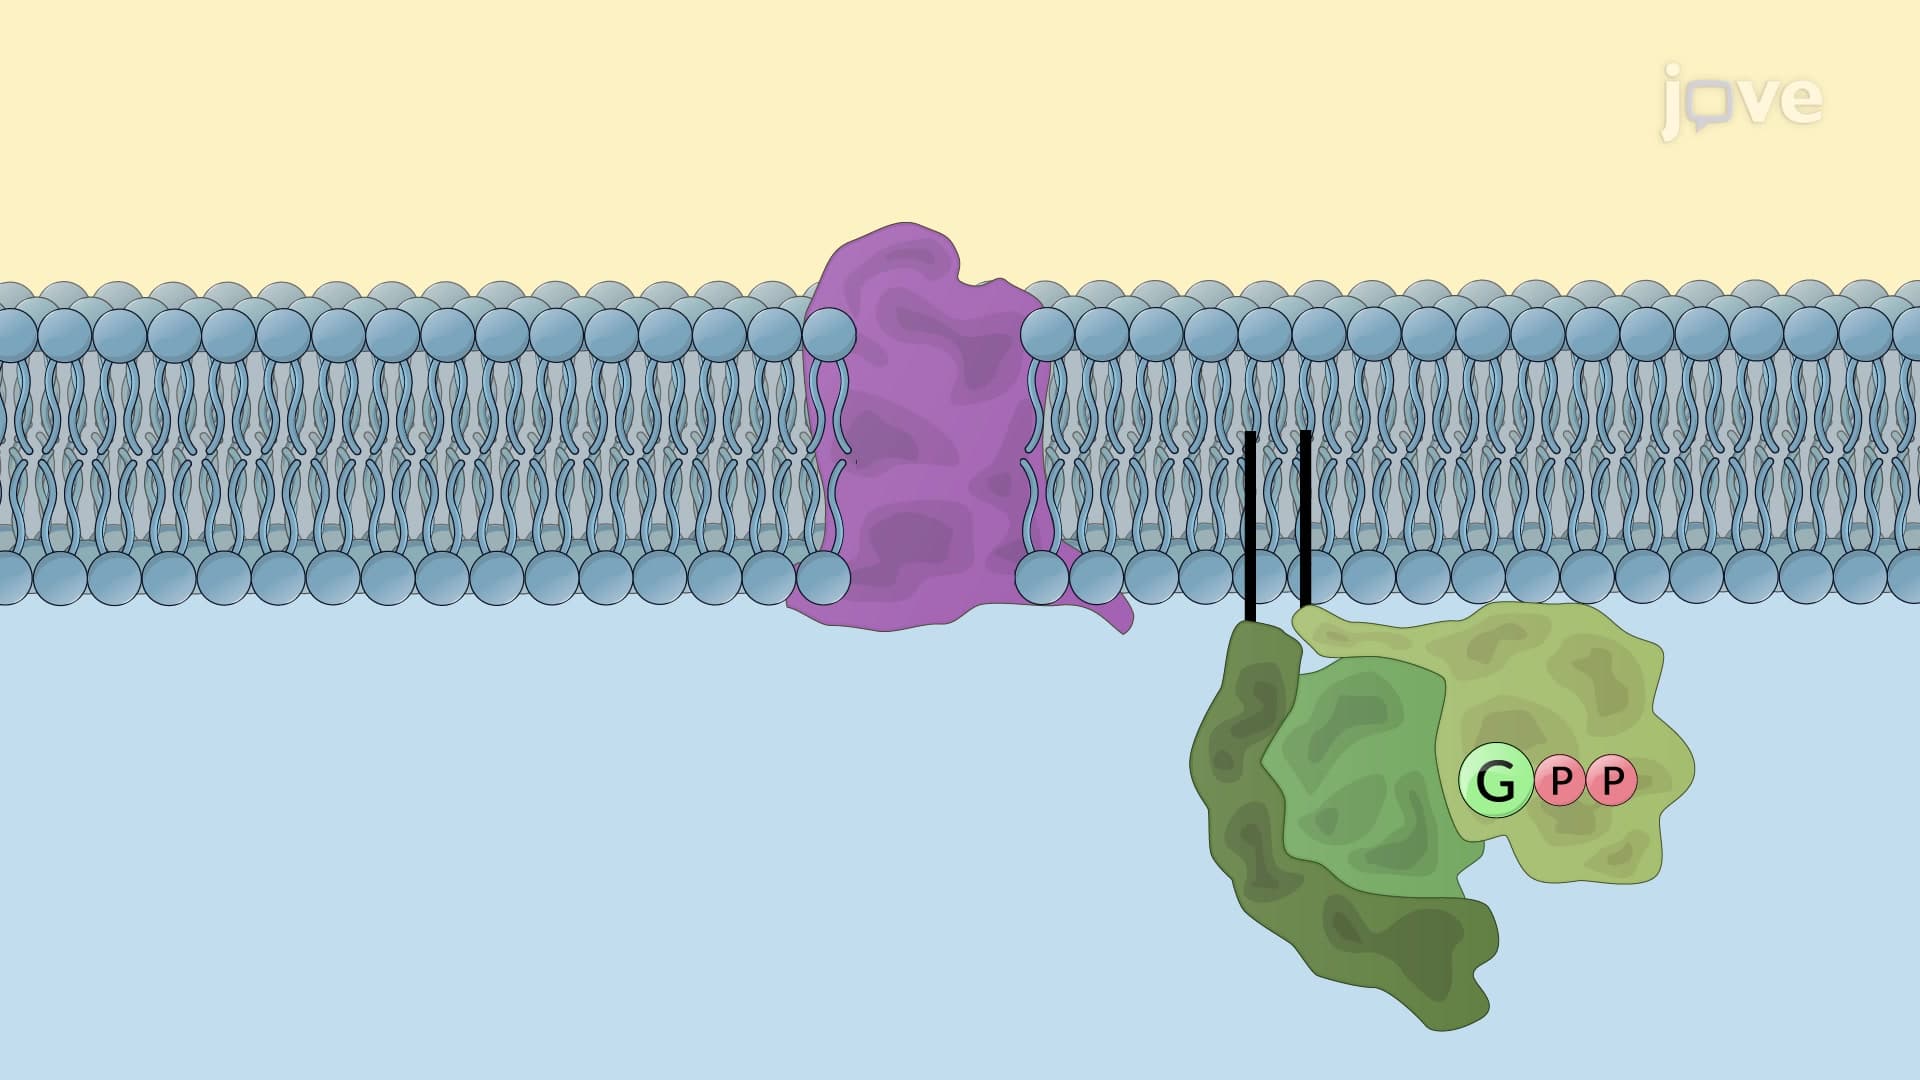
Adrenergic Receptors (Adrenoceptors): Classification
α-Adrenoceptors
α-Adrenoceptors are classified into two main subtypes: α1 and α2. The α1 adrenoceptors, which are found on postsynaptic...
 01:17
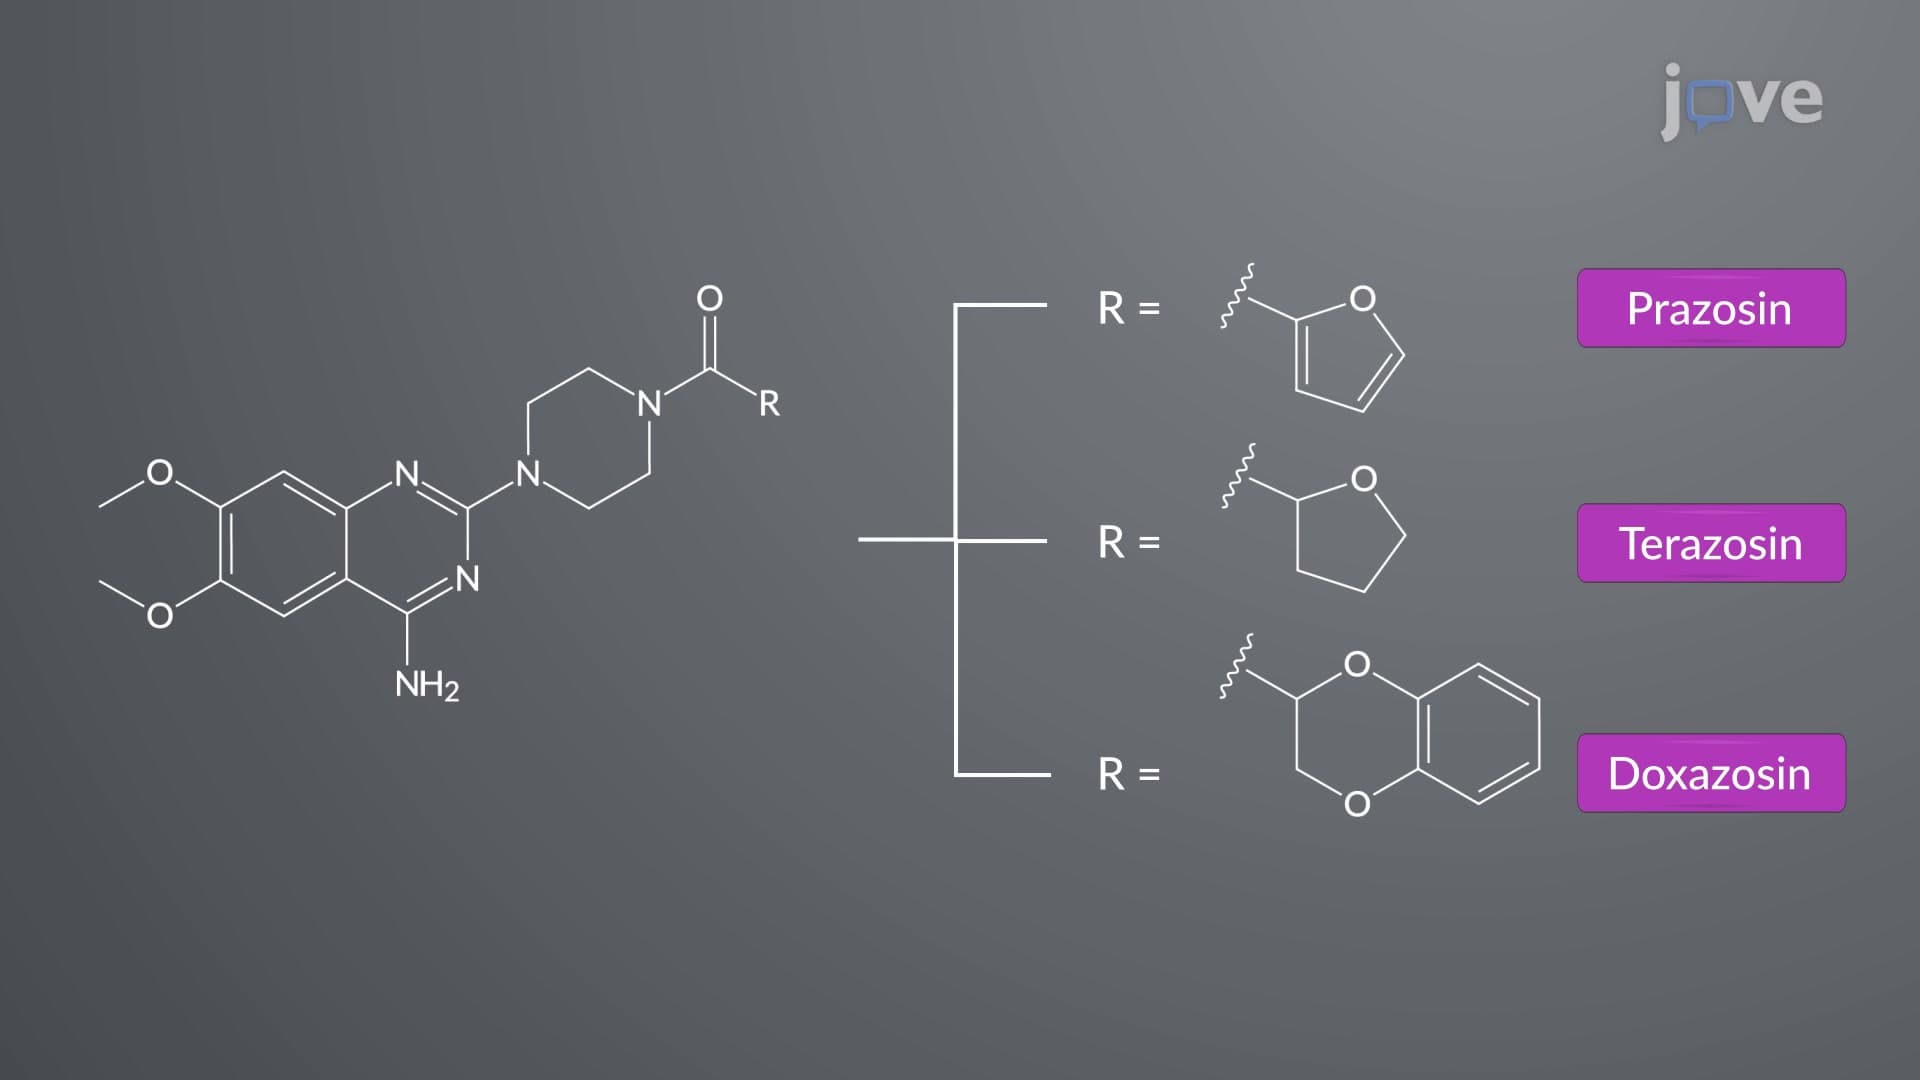
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...