Related Experiment Video
Updated: Oct 10, 2026

The Application of Open Searching-based Approaches for the Identification of Acinetobacter baumannii O-linked Glycopeptides
Published on: November 2, 2021
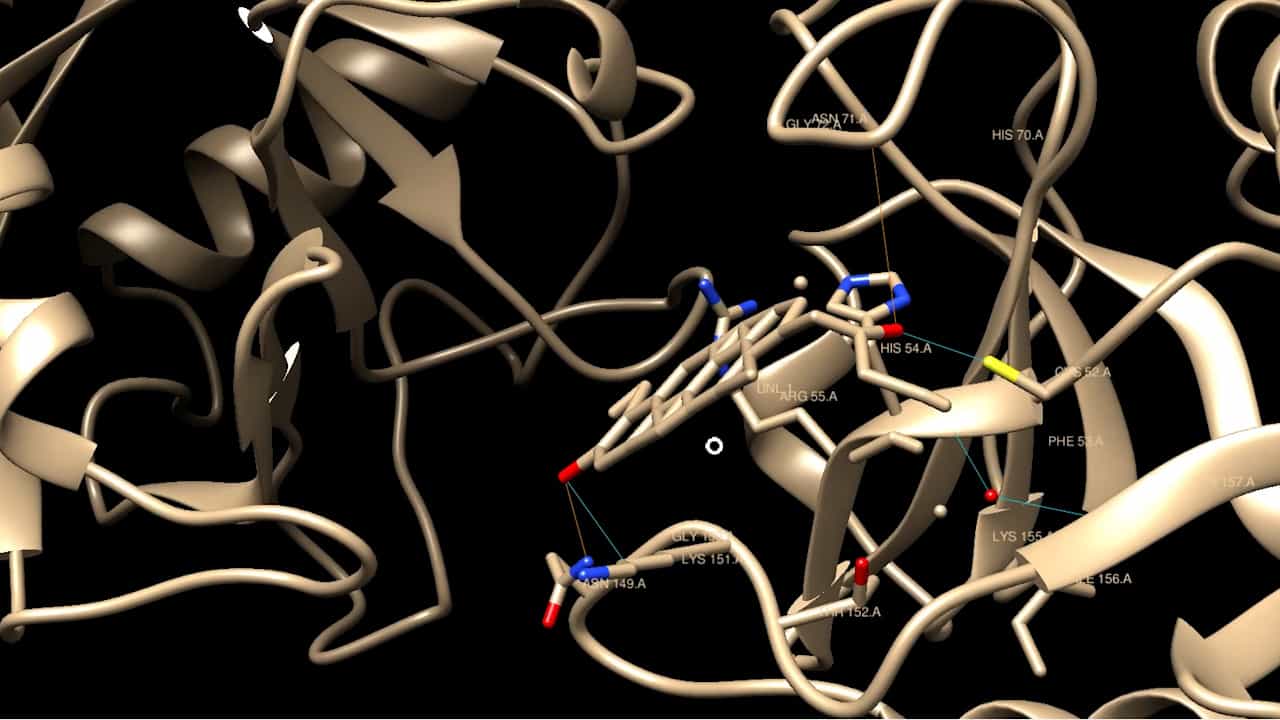
Computationally guided discovery of novel inhibitors targeting Acinetobacter-derived cephalosporinase (AmpC) through
Mohanraj Gopikrishnan1, George Priya Doss C1
1Department of Integrative Biology, School of Biosciences and Technology, Vellore Institute of Technology, Vellore, Tamil Nadu, 632 014, India.

Abstract:
Acinetobacter baumannii is a critical opportunistic pathogen responsible for a global health crisis, driven primarily by the rising prevalence of multidrug-resistant (MDR) strains. A central mechanism of β-lactam resistance in A. baumannii involves the overexpression of the class C β-lactamase AmpC, which degrades a wide spectrum of β-lactams, thereby limiting the effectiveness of antibiotics. The precise structural factors that dictate AmpC inhibition, however, have been less extensively studied and present opportunities to develop effective inhibitors against it. We explored natural inhibitors targeting the AmpC β-lactamase with an inclusive structure-based approach for drug discovery. Using an integrated computational methodology comprising molecular docking, pharmacokinetic analyses, and long-time MD simulations, a dataset of 374 phytocompounds was docked with AutoDock Vina. Among them, the phytocompounds 1521_0134 and N105_0095 with estimated binding affinities of -6.69 and -7.07 kcal/mol were identified as significant binding candidates. The detailed binding-mode analysis showed that both phytocompounds form strong, stable hydrogen bonds with key catalytic residues in the AmpC active pocket and have docking scores comparable to cefepime (binding energy of -7.48 kcal/mol), a well-known substrate for the AmpC β-lactamase. Additional SwissADME, pkCSM, and ProTox-3.0 analysis indicated promising drug-like properties for both phytocompounds with appropriate molecular weight, optimal lipophilicity, and good membrane permeability. We further performed 1000 ns all-atom MD simulations of AmpC in the presence of the top two phytocompounds to elucidate the dynamics and structural stability following ligand binding. Trajectory analyses demonstrated that ligand binding enhanced the structural stability of AmpC, as evidenced by reduced conformational fluctuations and restricted protein mobility. Principal component analysis (PCA) and free-energy landscape (FEL) analyses further indicated that the AmpC-ligand complexes were restricted within the stable conformational space throughout the simulations. Notably, the AmpC_1521_0134 complex presented two distinct free-energy basins, demonstrating structurally stable, thermodynamically controlled conformational ensembles. Moreover, the dynamic cross-correlation matrix (DCCM) analysis showed a decrease in correlated motion between remote residues, indicating the structural rigidity of the enzyme-ligand complexes. Collectively, our results provided a structural and energetic rationale for the interaction of natural phytocompounds with AmpC β-lactamase, with phytocompound 1521_0134 as a lead candidate that merits further development for the rational design of novel lactamase inhibitors to overcome multidrug-resistant A. baumannii.