
Structure-based drug repurposing for CDK6: Molecular docking and molecular dynamics characterization of FDA-approved
Syarifuddin Husain1, Ruzianisra Mohamed2, Khairul Bariyyah Abd Halim3
1Bioinformatics Unit, Faculty of Pharmacy, Universiti Teknologi MARA, Bandar Puncak Alam, Selangor, 42300, Malaysia.

Abstract:
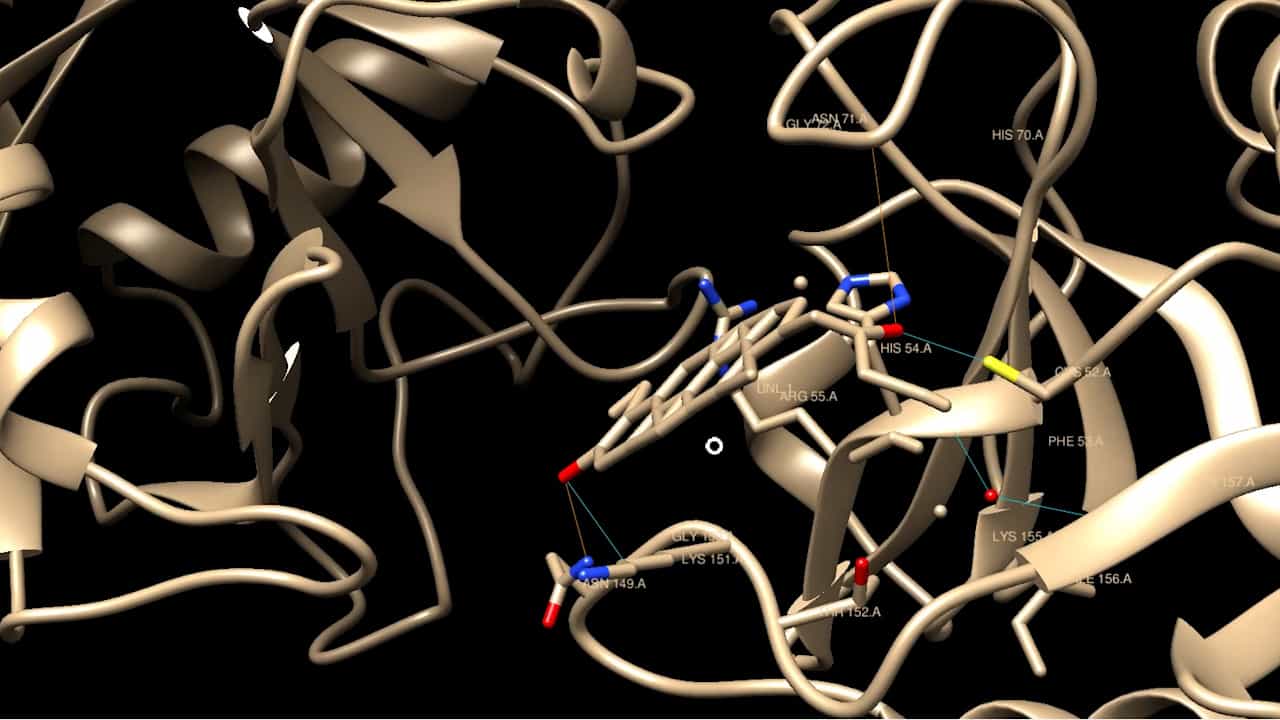
Cyclin-dependent kinase 6 (CDK6) contributes to G1-to-S cell-cycle progression and represents an established therapeutic target in hormone receptor-positive breast cancer. However, adverse effects and acquired resistance associated with current CDK4/6 inhibitors support continued investigation of alternative chemical scaffolds. In this study, a total of 319 FDA-approved non-oncological drugs were screened by molecular docking against the ATP-binding site of a predicted human CDK6-Cyclin D1 complex. The three highest-ranked compounds, dutasteride, danazol, and azelastine, were subsequently subjected to 100-ns molecular dynamics simulations, principal component analysis, MM-PBSA calculations, per-residue energy decomposition, and in silico pharmacokinetic profiling, with abemaciclib included as a reference inhibitor. All three repurposed candidates remained accommodated within the CDK6 binding pocket during their individual trajectories, although they exhibited different degrees of positional and conformational rearrangement and distinct energetic profiles. MM-PBSA ΔGtotal values were -20.47 ± 6.87 kcal/mol for dutasteride, -24.57 ± 5.09 kcal/mol for danazol, and -39.48 ± 6.10 kcal/mol for azelastine, compared with -41.79 ± 5.43 kcal/mol for abemaciclib. Per-residue decomposition showed that the candidates engaged overlapping regions of the CDK6 binding pocket while exhibiting distinct residue-level energetic contributions. Pharmacokinetic predictions further indicated differences in lipophilicity, gastrointestinal absorption, blood-brain barrier permeability, and P-glycoprotein substrate status among the selected compounds. Collectively, these findings provide structural, conformational, energetic, and pharmacokinetic characterization of dutasteride, danazol, and azelastine as candidate CDK6-binding scaffolds. Direct biochemical binding, kinase-inhibition, and cellular target-engagement studies are required to determine whether these predicted interactions have biological relevance.
Related Concept Videos
 01:26
Drug Discovery: Overview
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 02:34
Inhibition of Cdk Activity
 01:26
Targets for Drug Action: Overview
Receptors are either membrane-spanning or intracellular proteins, which upon binding a ligand, get activated and transmit the signal downstream to elicit a response. Drugs bind receptors, either mimicking the action of endogenous ligands or blocking the receptor activity to bring about a modified response. Nearly 35% of approved drugs target the G...
 02:15
M-Cdk Drives Transition Into Mitosis
Cyclin-dependent kinases, or Cdks, work in concert with cyclins to control cell cycle transitions. M-Cdk, a complex of Cdk1 bound to M cyclin, is a well-known example of this coordinated control that drives the transition from the G2 to the M phase.
M cyclin...
 01:15
Impact of Pharmacokinetic–Pharmacodynamic Models: Regulatory Decisions