Large-scale sequencing and the new animal phylogeny
Related Experiment Video
Updated: Aug 6, 2026
Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
Hervé Philippe1, Maximilian J Telford
1Canadian Institute for Advanced Research, Centre Robert-Cedergren, Département de Biochimie, Université de Montréal, Succursale Centre-Ville, Montréal, QC, Canada, H3C 3J7. herve.philippe@umontreal.ca

Abstract:
Although comparisons of gene sequences have revolutionised our understanding of the animal phylogenetic tree, it has become clear that, to avoid errors in tree reconstruction, a large number of genes from many species must be considered: too few genes and stochastic errors predominate, too few taxa and systematic errors appear. We argue here that, to gather many sequences from many taxa, the best use of resources is to sequence a small number of expressed sequence tags (1000-5000 per species) from as many taxa as possible. This approach counters both sources of error, gives the best hope of a well-resolved phylogeny of the animals and will act as a central resource for a carefully targeted genome sequencing programme.
Related Concept Videos
 02:54
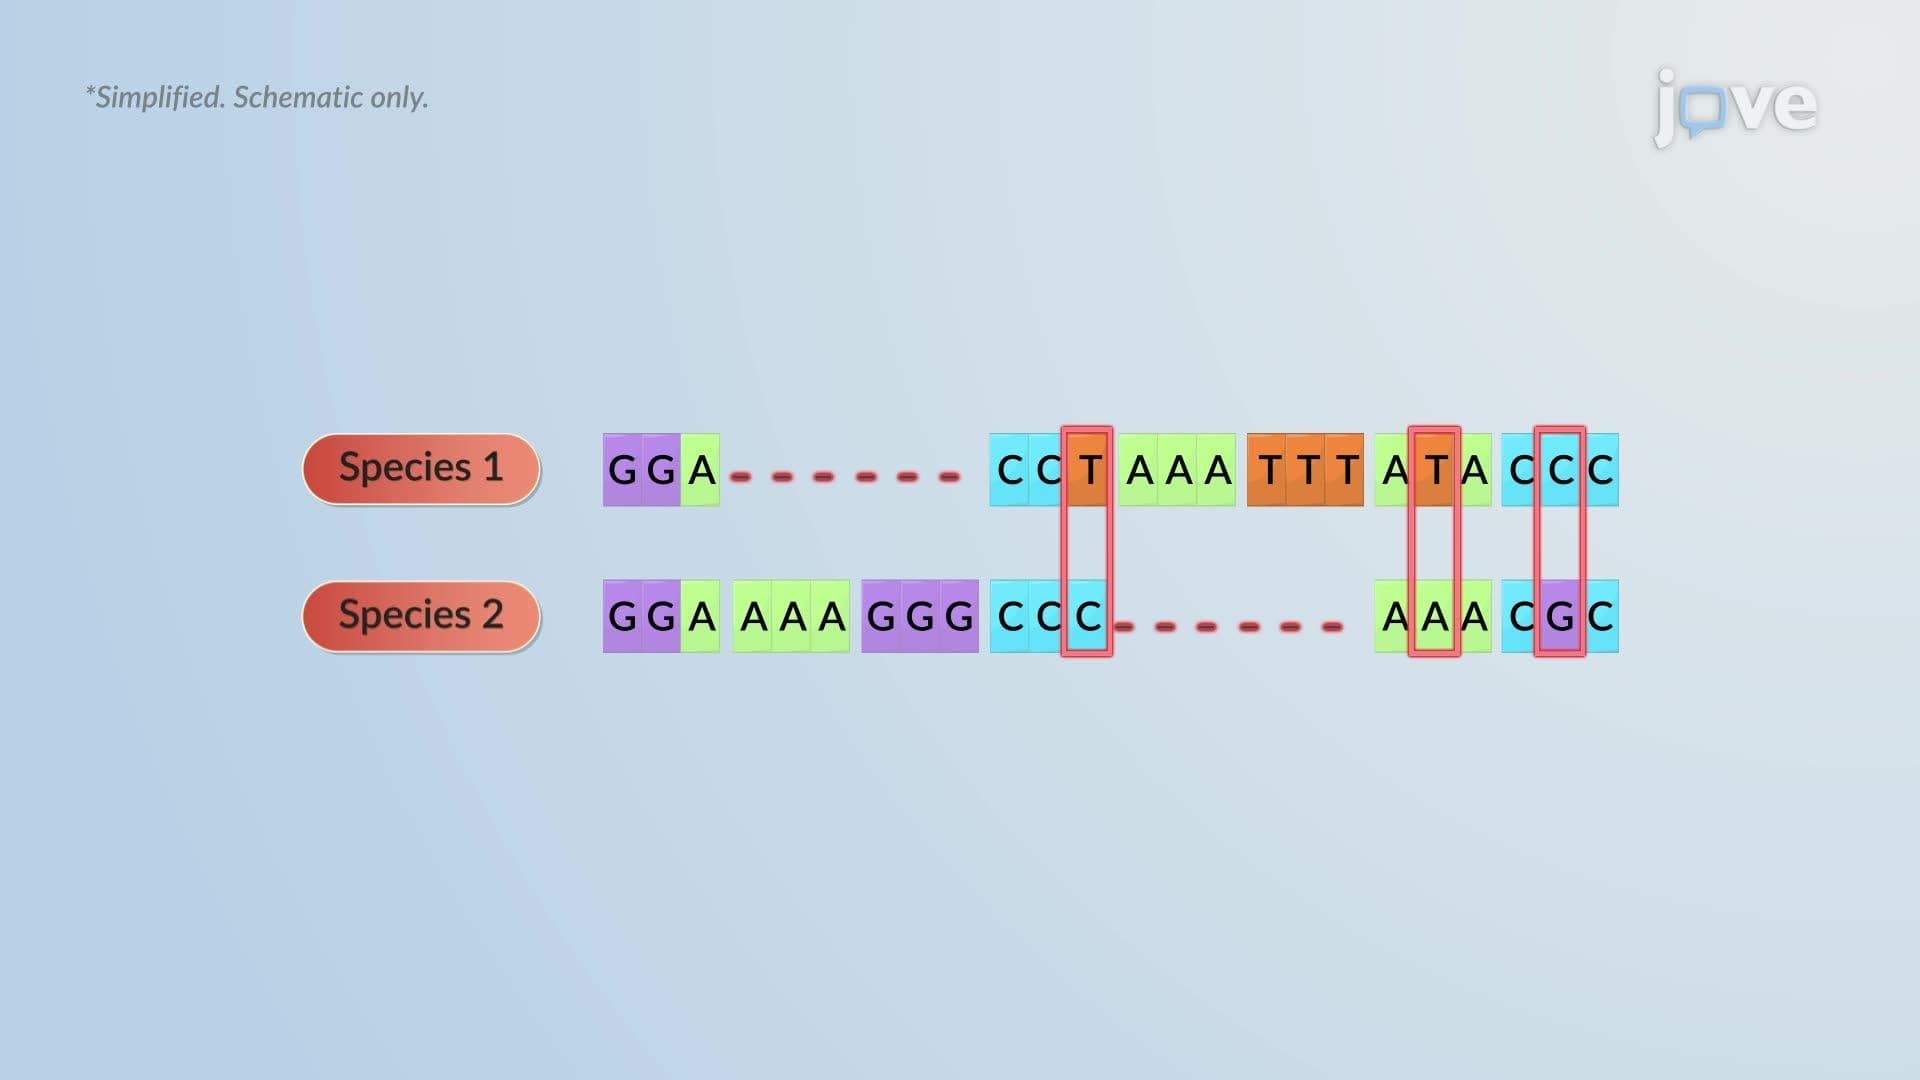
Evolutionary Relationships through Genome Comparisons
 01:28
Microbial Phylogeny
 02:40
The Tree of Life - Bacteria, Archaea, Eukaryotes
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...
 01:23
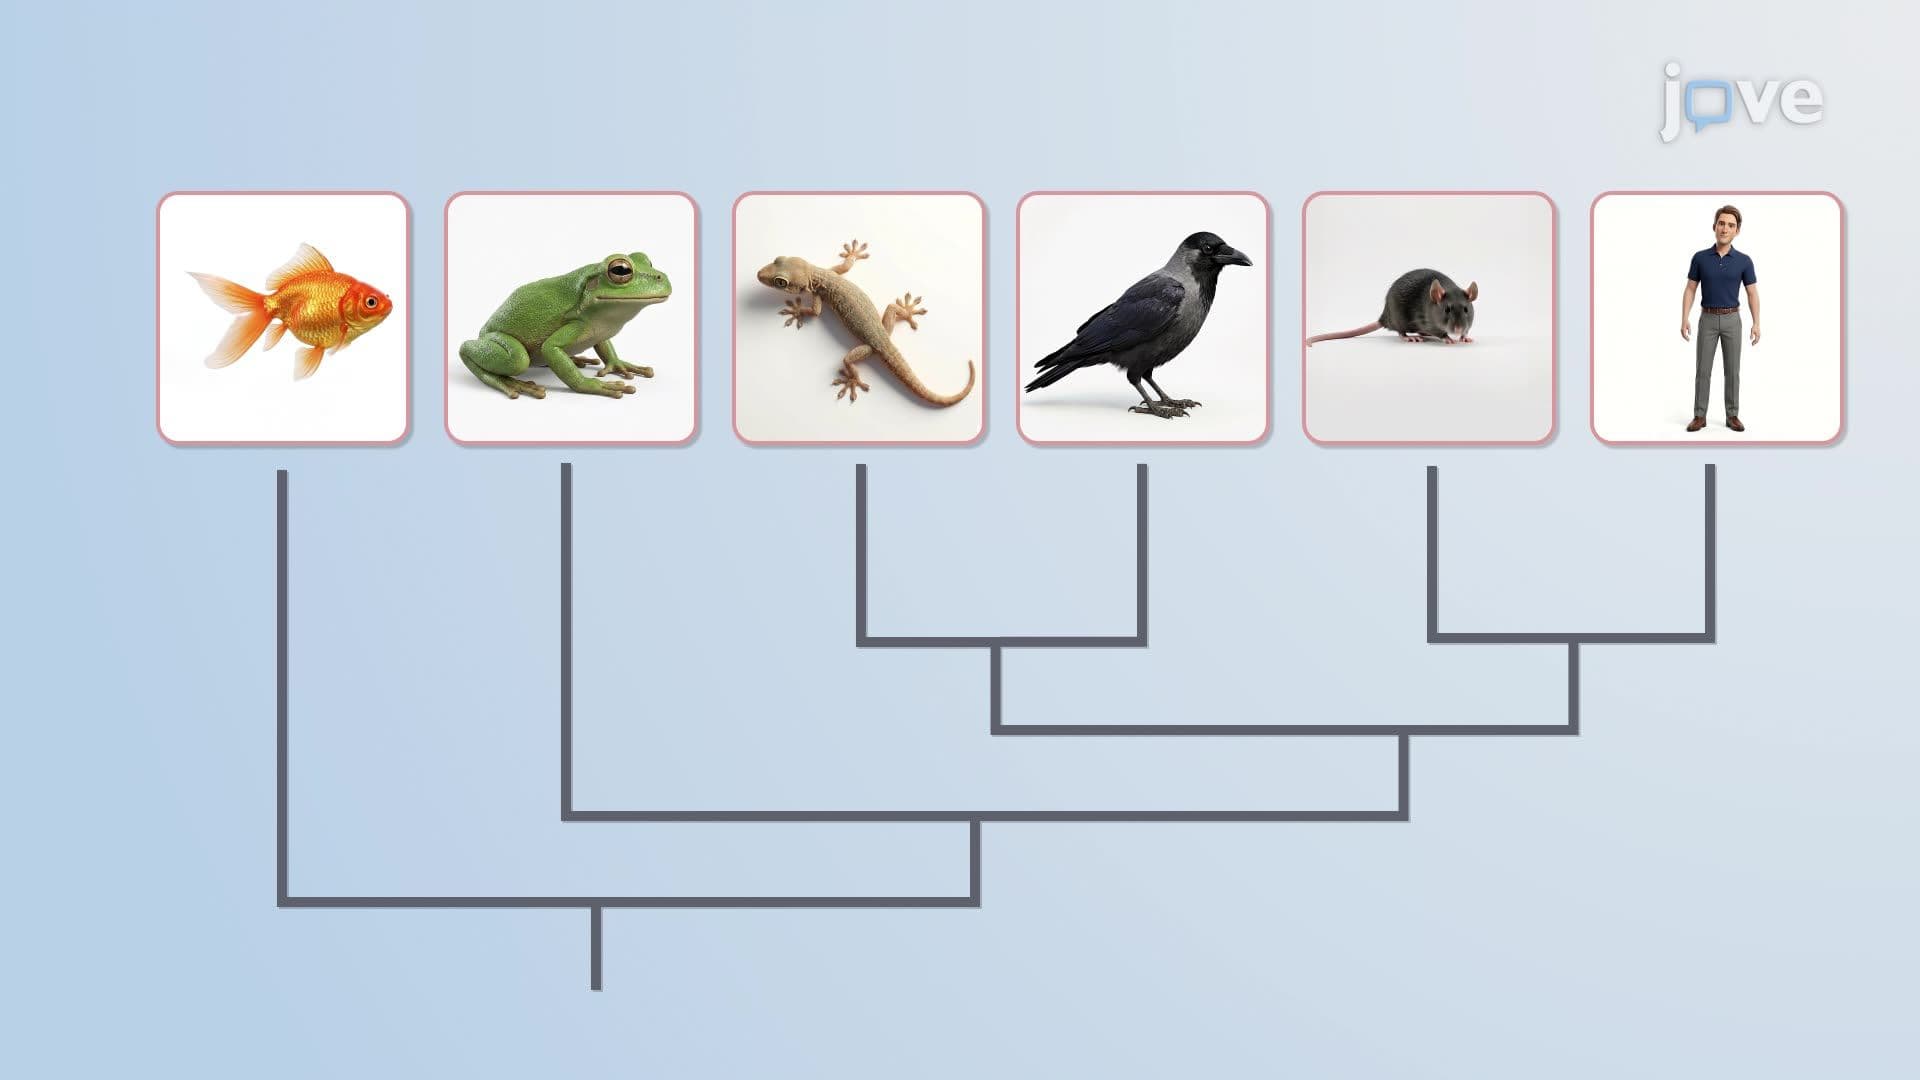
Phylogeny
 02:31
Synteny and Evolution
Around 80 million years ago, the human and mice lineages diverged from the common ancestor. During the course of evolution, the ancestral chromosome underwent...