How inaccuracies in protein structure models affect estimates of protein-ligand interactions: computational analysis of HIV-I protease inhibitor binding
Related Experiment Video
Updated: Jul 20, 2026

Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Holmfridur B Thorsteinsdottir1, Torsten Schwede, Vincent Zoete
1Biozentrum, University of Basel, Klingelbergstrasse 50,4056 Basel, Switzerland.

Abstract:
The influence of possible inaccuracies that can arise during homology modeling of protein structures used for ligand binding studies were investigated with the molecular mechanics generalized Born surface area (MM-GBSA) method. For this, a family of well-characterized HIV-I protease-inhibitor complexes was used. Validation of MM-GBSA led to a correlation coefficient ranging from 0.72 to 0.93 between calculated and experimental binding free energies DeltaG. All calculated DeltaG values were based on molecular dynamics simulations with explicit solvent. Errors introduced into the protein structure through misplacement of side-chains during rotamer modeling led to a correlation coefficient between DeltaG(calc) and DeltaG(exp) of 0.75 compared with 0.90 for the correctly placed side chains. This is in contrast to homology models for members of the retroviral protease family with template structures ranging in sequence identity between 32% and 51%. For these protein models, the correlation coefficients vary between 0.84 and 0.87, which is considerably closer to the original protein (0.90). It is concluded that HIV-I low sequence identity with the template structure still allows creating sufficiently reliable homology models to be used for ligand-binding studies, although placement of the rotamers is a critical step during the modeling.
Related Concept Videos
 02:40
Ligand Binding Sites
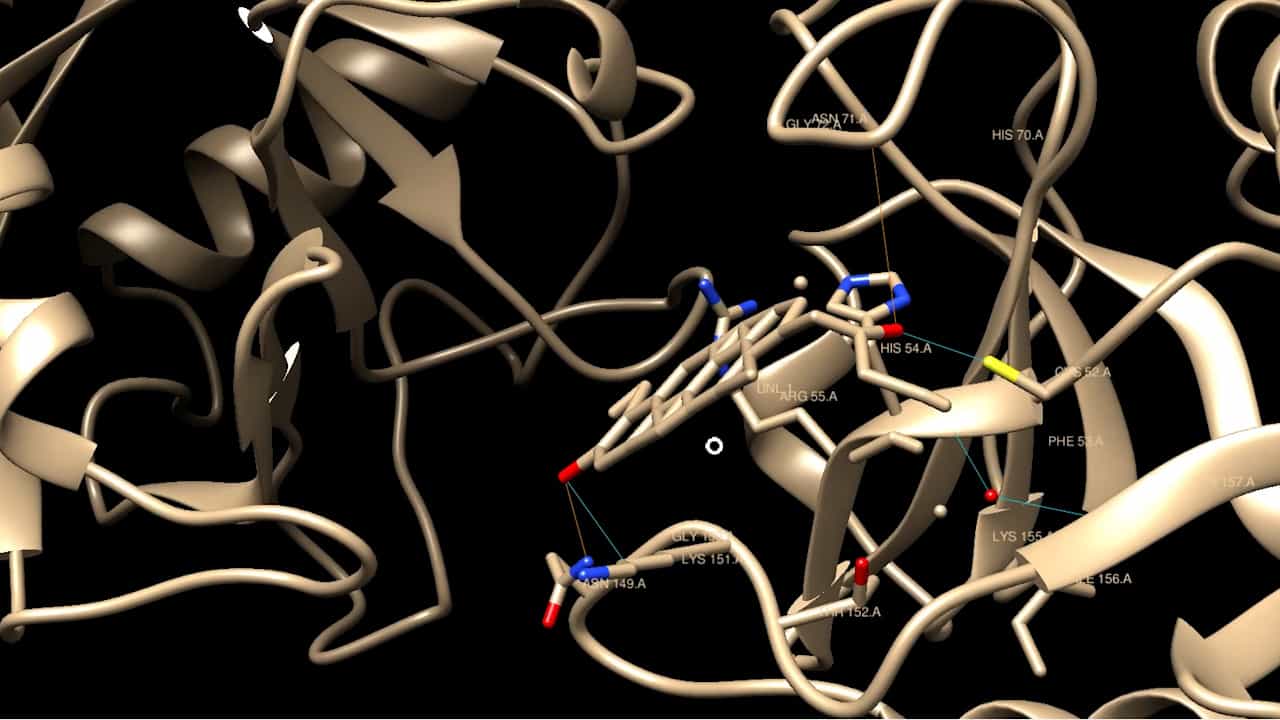
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:04
Protein-protein Interfaces
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence.
 01:19
Inhibitors of Virion Maturation and Assembly