Related Experiment Video
Updated: Oct 25, 2026
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Full-Dimensional Reactive Potential Energy Surfaces of OCS+ for S+ Dissociation: Ground and Excited States
Cangtao Yin1, Stefan Willitsch1, Markus Meuwly1
1Department of Chemistry, University of Basel, Klingelbergstrasse 80, CH-4056Basel, Switzerland.

Abstract:
Full-dimensional reactive potential energy surfaces (PESs) for the OCS+ cation are constructed to describe S+ loss in the electronic ground state and seven low-lying electronically excited states. High-level ab initio reference energies were computed at the MRCI+Q/aug-cc-pVTZ level and were used to generate PESs employing reproducing kernel Hilbert space representations (RKHS). The PESs accurately reproduce the measured dissociation limits to CO(X1Σ+) + S+ in different electronic states. The topology of the PESs reveals multiple linear and T-shaped minima, pronounced angular anisotropy, and state-crossing manifolds. Exploratory quasi-classical trajectory simulations on all PESs confirm numerical stability and energy conservation, illustrating the suitability of the surfaces for dynamical applications. The present work represents the most comprehensive characterization to date of the lowest PESs of OCS+ and provides a reliable foundation for future studies of the photodissociation of OCS+ and the chemi-ionization dynamics of OCS.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:22
Measurement of Ultrafast Vibrational Coherences in Polyatomic Radical Cations with Strong-Field Adiabatic Ionization
Published on: August 6, 2018
Related Concept Videos
 02:13
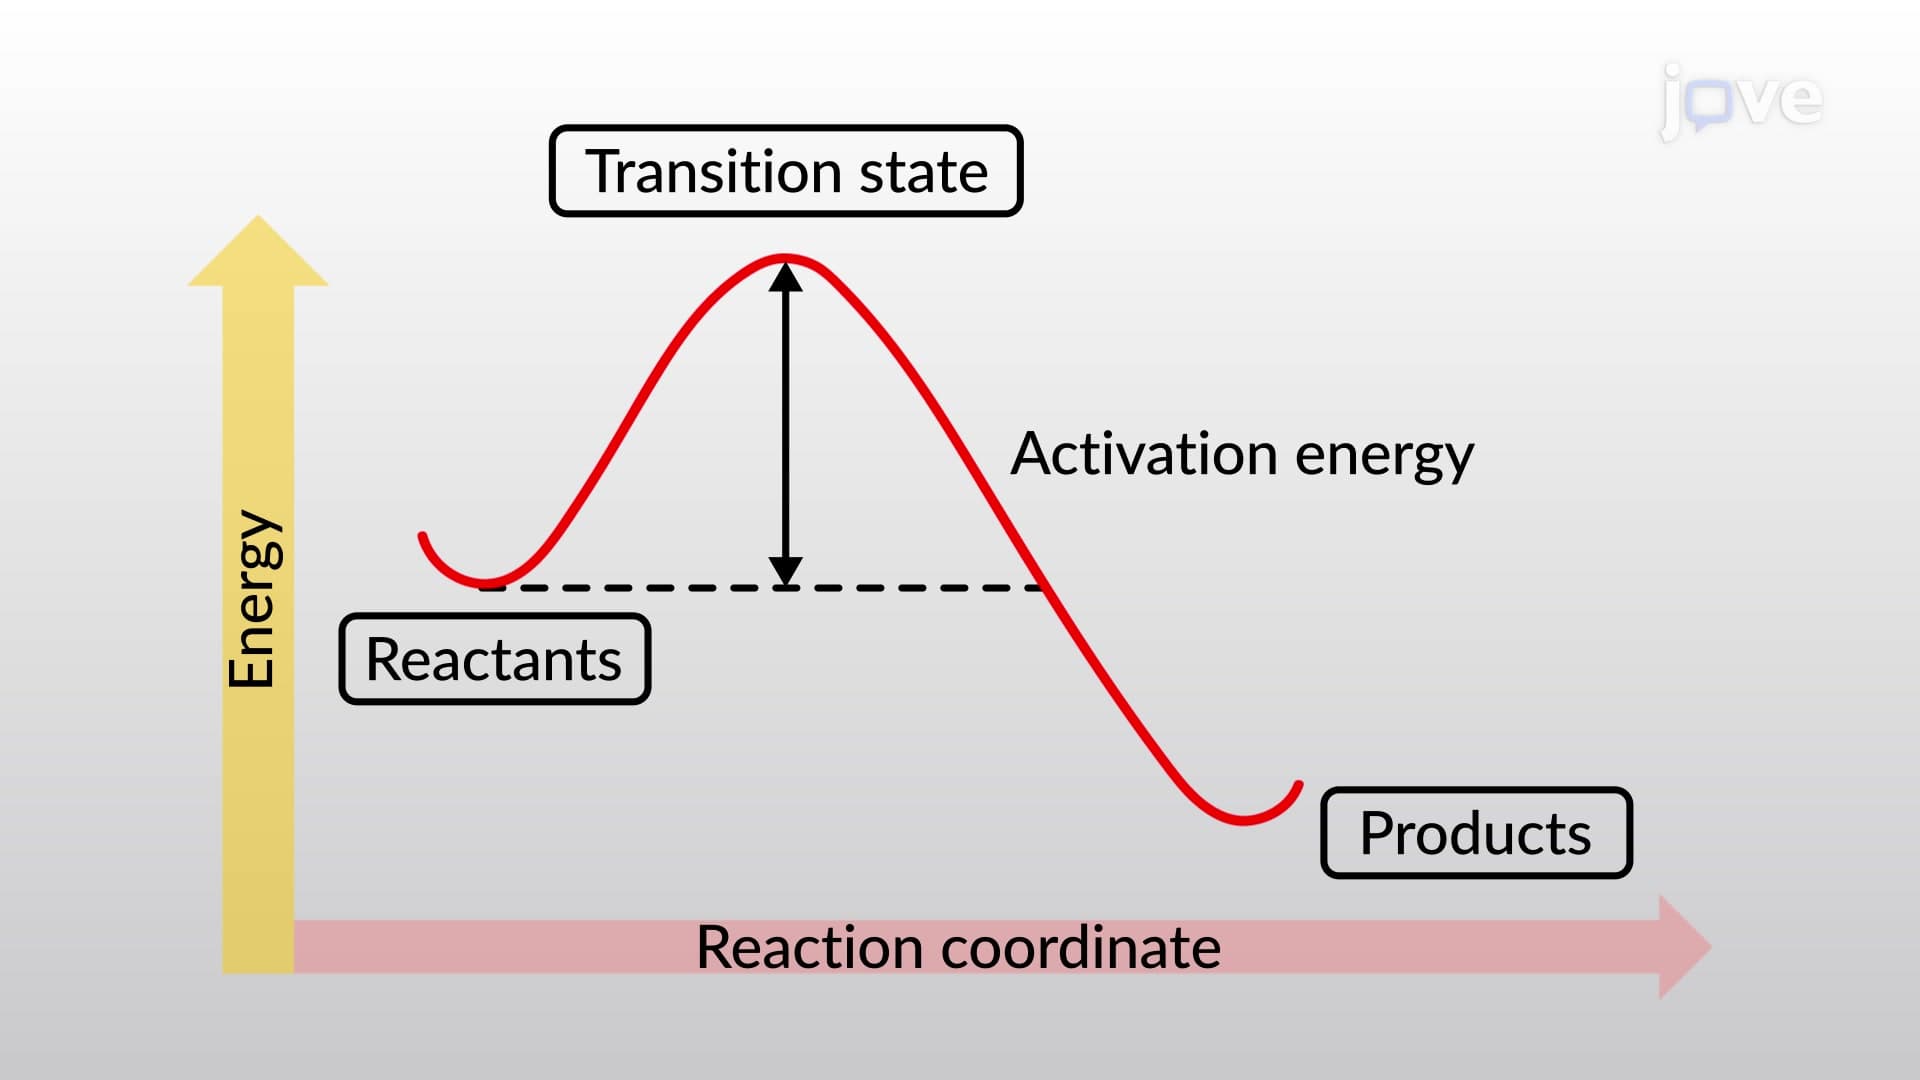
Bond Dissociation Energy and Activation Energy
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:13
Energy Diagrams, Transition States, and Intermediates
 03:51
Molecular Orbital Theory II
 02:35
Molecular Orbital Theory I
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...