Related Experiment Video
Updated: Oct 25, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Investigation of Medium-Sized Methane Clusters with Quantum Chemical First-Principles and Ab Initio Calculations
Xiaoyan Cao1, Michael Dolg1,2
1Laboratory of Theoretical and Computational Photochemistry, College of Chemistry, Beijing Normal University, Xinjiekouwaidajie 19, Beijing100875, China.

Abstract:
The structures and relative stabilities of medium-sized methane clusters (CH4)n (n = 10-40) have been studied using first-principles density functional theory calculations applying the Becke and Lee, Yang, Parr hybrid functional with a posteriori pairwise corrections for dispersion interactions due to Grimme, B3LYP+D3+BJ, and basis sets of doubly polarized valence triple-ζ quality. For all structures, vibrational frequencies were evaluated. Coupled-cluster single-point calculations including single, double, and perturbative triple substitutions with an explicit dependence on the interelectronic distance, CCSD(T)-F12, were performed using the incremental scheme accounting for up to third-order interactions between the monomers, applying correlation-consistent basis sets of polarized double- and triple-ζ quality especially optimized for F12 calculations. The structures obtained with the B3LYP+D3+BJ approach were found to have a higher stability than those of three sets published previously in the literature, which are based on pairwise additive potentials or density-functional calculations corrected for dispersion interactions by atom-centered potentials. By means of extrapolation, the heat of vaporization was estimated as 6.87 ± 0.16 kJ/mol and 6.97 ± 0.19 kJ/mol at the B3LYP+D3+BJ and CCSD(T)-F12 levels, respectively (exp. 8.19 kJ/mol). The rotation of CH4 monomers was studied at the B3LYP+D3+BJ and CCSD(T)-F12 level and was found to have barriers of only about 1.2-1.5 and 3.2-3.7 kJ/mol for monomers at the surface and in the center of a (CH4)30 cluster, respectively. The incremental CCSD(T)-F12 calculations exhibit a scaling better than O(n2) for clusters up to (CH4)40 comprising 5000 contracted basis functions, whereas simulations for larger clusters exhibit a linear size dependence.
Related Concept Videos
 01:23
Mass Spectrum
 01:21
Chemical Ionization (CI) Mass Spectrometry
 01:18
Mass Spectrometry: Long-Chain Alkane Fragmentation
 02:18
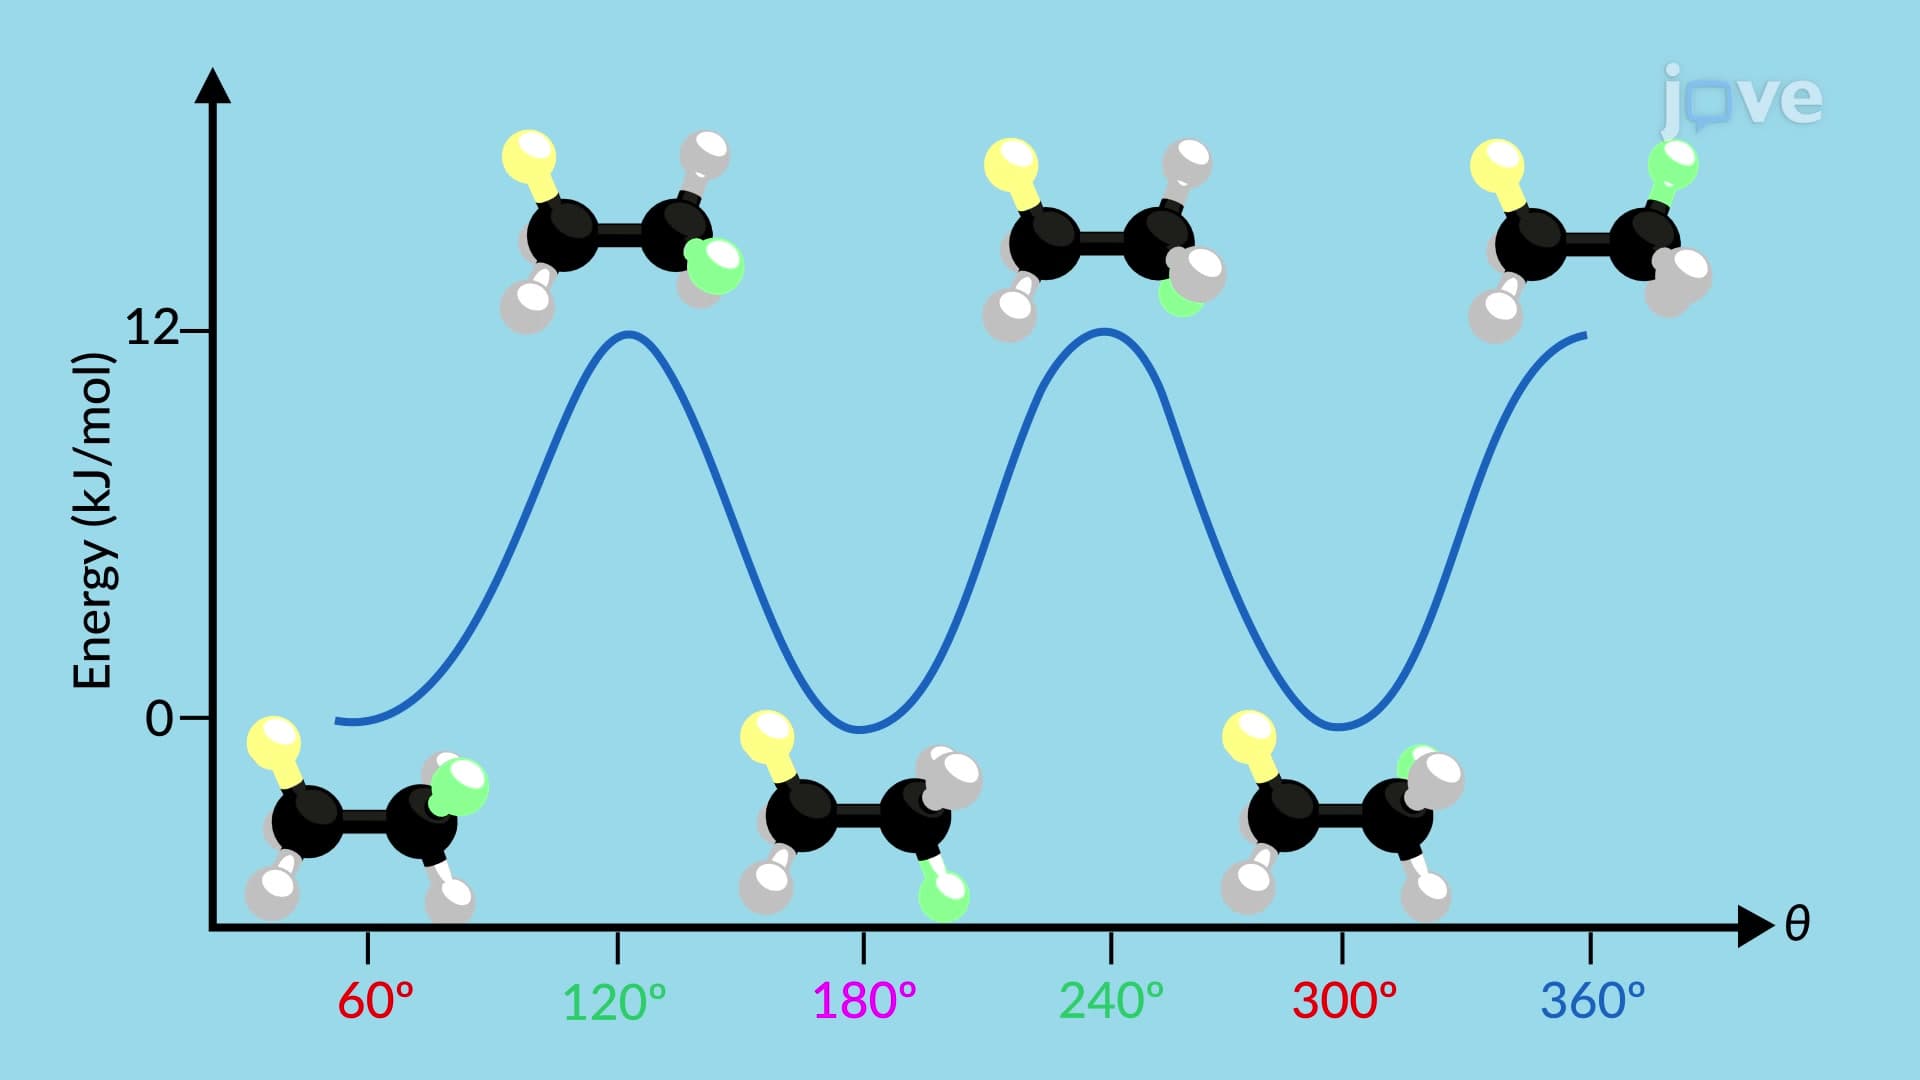
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered ethane, the...
 01:24
Mass Spectrum: Interpretation