Related Experiment Video
Updated: Jul 17, 2026
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Density functional theory study of the inner hydrogen atom transfer in metal-free porphyrins: meso-substitutional
Yuexing Zhang1, Ping Yao, Xue Cai
1Department of Chemistry, Shandong University, Jinan 250100, PR China.

Abstract:
Density functional theory (DFT) calculations were carried out to study the influence of both electron-withdrawing fluorine and electron-donating amidogen meso-substituents on the inner hydrogen transfer in metal-free porphyrins. Twenty-four stable structures and 21 transition states among the stable structures are fully optimized at the B3LYP/6-31G (d) level, and vibration analyses are carried out to verify the optimized structures. It is found that the acidity of the transferred hydrogen atom, the basicity of the nitrogen atoms of the adjacent pyrrole ring, the distance from the transferred hydrogen atom to the nitrogen atom of the adjacent pyrrole ring, and the electronic nature of meso-substituents in transfer paths, all have significant influences on the potential energy barrier of the inner hydrogen transfer in porphyrins. The different transfer paths of each substituted porphyrin are compared, all the transfer barriers are compared to unsubstituted metal free porphyrin, and the infrared and electronic absorption spectra of some important complexes are described. By placing substituents appropriately, the cis-trans transition energy barrier can be greatly decreased, and the cis-porphyrin conformer which can be detected spectroscopically may be significantly stabilized. The present work would shed light on tuning the transition barrier by selecting the most appropriate paths and detecting cis-porphyrins experimentally.
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Valence Bond Theory
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
 03:51
Molecular Orbital Theory II
 01:09
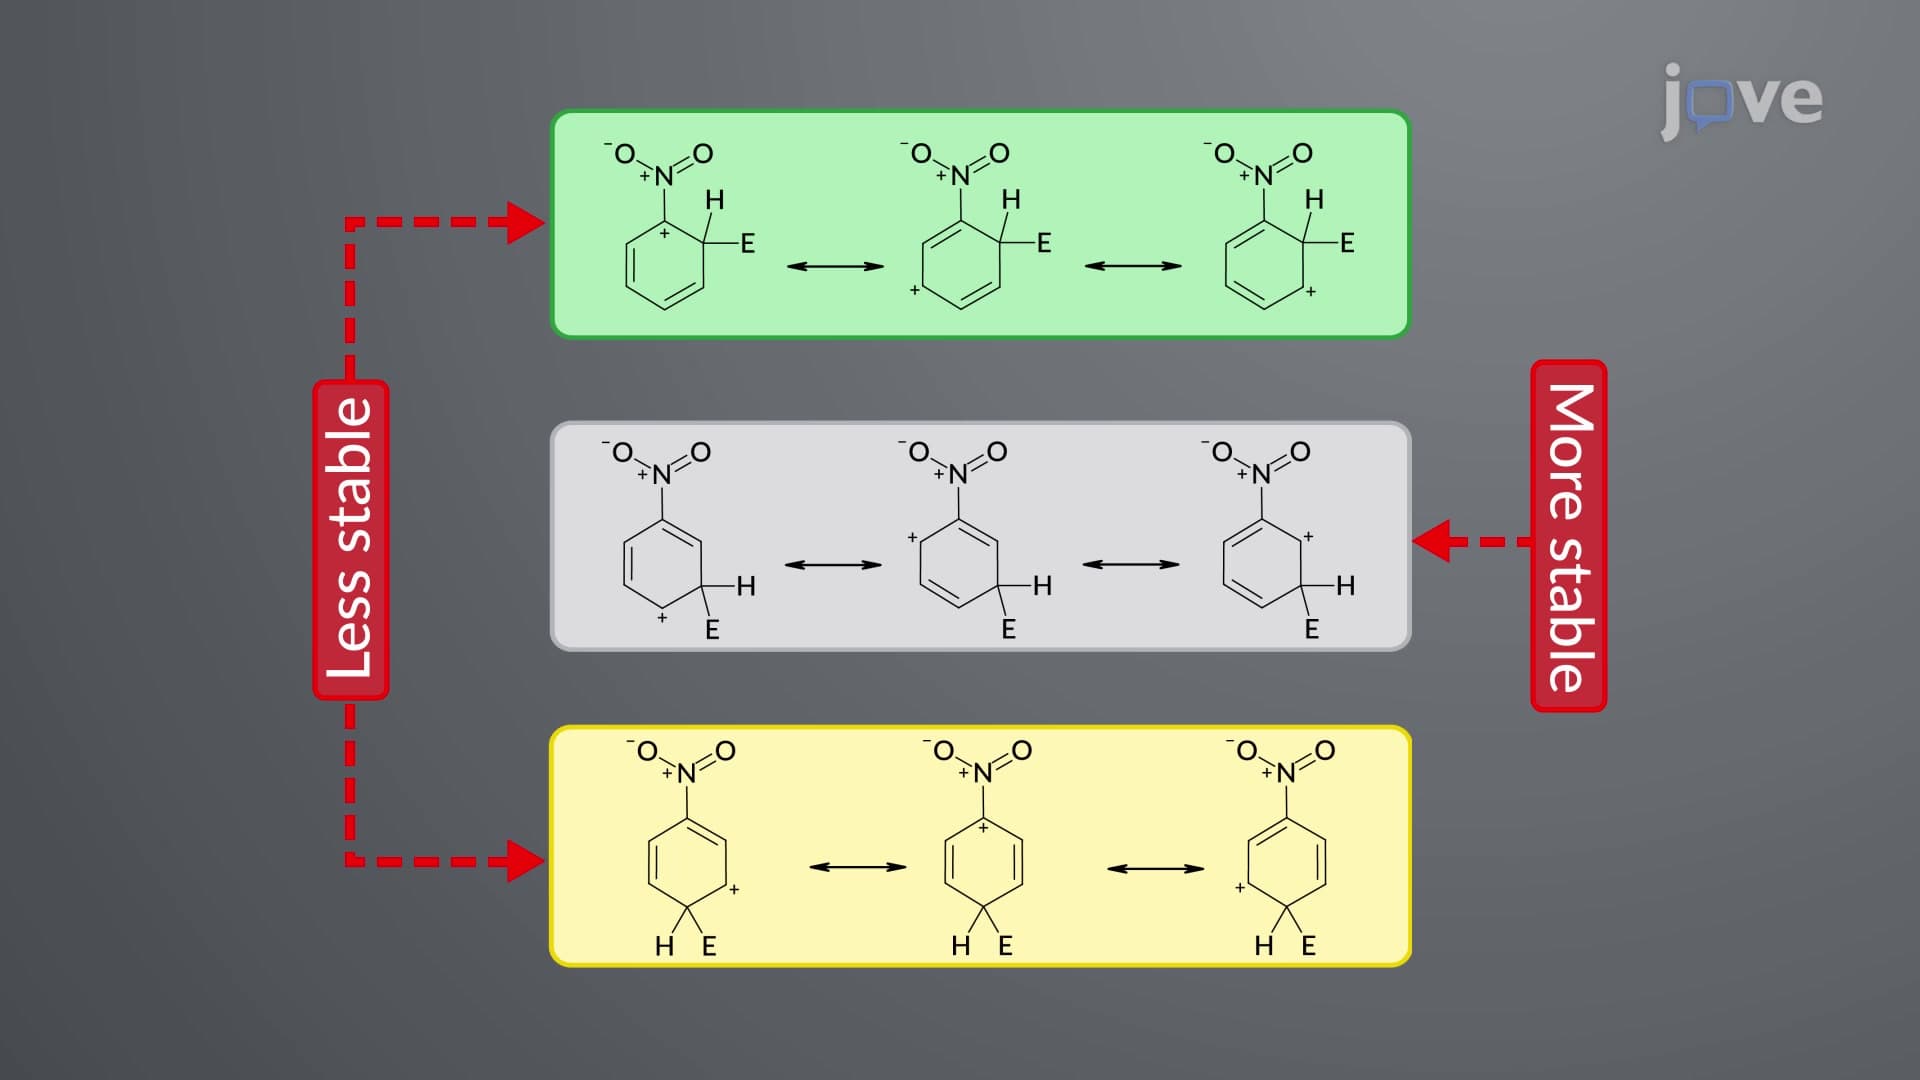
Directing Effect of Substituents: meta-Directing Groups