Related Experiment Video
Updated: Oct 16, 2026
Construction and Systematical Symmetric Studies of a Series of Supramolecular Clusters with Binary or Ternary Ammonium Triphenylacetates
Published on: February 15, 2016
Unveiling phase-dependent structure-property relationships in cyclohexaphosphane (P6H6) molecular blocks: A DFT and
Mohammad Qasemnazhand1, Farhad Khoeini2, Ali Ben Ahmed3
1Department of Physics Education, Farhangian University, P.O. Box 14665-889, Tehran, Iran.

Abstract:
Phosphorene-based nanomaterials have attracted considerable interest because of Physicochemical properties. However, the influence of local molecular geometry on the properties of finite phosphorene-related molecular motifs remains insufficiently explored. In this work, density functional theory (DFT) and time-dependent density functional theory (TD-DFT) calculations are performed for four isolated cyclohexaphosphane structures used as finite molecular motifs associated with the α, β, γ, and δ phosphorene configurations. This molecular-block representation is intended to compare local structure-property trends and does not imply quantitative equivalence with periodic phosphorene sheets. Optimized geometries, vibrational spectra, frontier molecular orbitals, density of states, electrostatic surface potential (ESP), and optical properties are analyzed. The calculations reveal clear phase-dependent electronic characteristics: the α- and β-motifs exhibit larger gaps of 5.34 and 4.89 eV, respectively, whereas the γ- and δ-motifs show smaller gaps of 4.14 and 4.11 eV, together with higher chemical softness, electrophilicity, and Electronic Interaction Index values. The optical calculations also show distinct responses, with absorption maxima at 357 and 339 nm for γ and δ, respectively, both in the near-UV region. The emission-related quantities and Stokes shifts obtained from the adopted triplet-relaxation protocol further indicate phase-dependent excited-state relaxation. ESP maps reveal heterogeneous electrostatic distributions that may be useful for guiding future explicit interaction studies. Overall, the calculations establish phase-dependent molecular structure-property relationships and provide a theoretical basis for subsequent studies of molecular sensing, adsorption, and optoelectronic behavior. Biomedical applications such as targeted delivery, toxicity assessment, and photothermal therapy are considered only as prospective directions requiring explicit intermolecular, environmental, and experimental validation.
More Related Videos
 07:20
Discovery and Synthesis Optimization of Isoreticular Al(III) Phosphonate-Based Metal-Organic Framework Compounds Using High-Throughput Methods
Published on: October 6, 2023
 06:55
Synthesis of Cyclic Polymers and Characterization of Their Diffusive Motion in the Melt State at the Single Molecule Level
Published on: September 26, 2016
Related Concept Videos
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group with both...
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal tetrahedral value,...
 01:18
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous overlap of p...
 01:18
Frost Circles for Different Conjugated Systems
 02:37
Disubstituted Cyclohexanes: cis-trans Isomerism
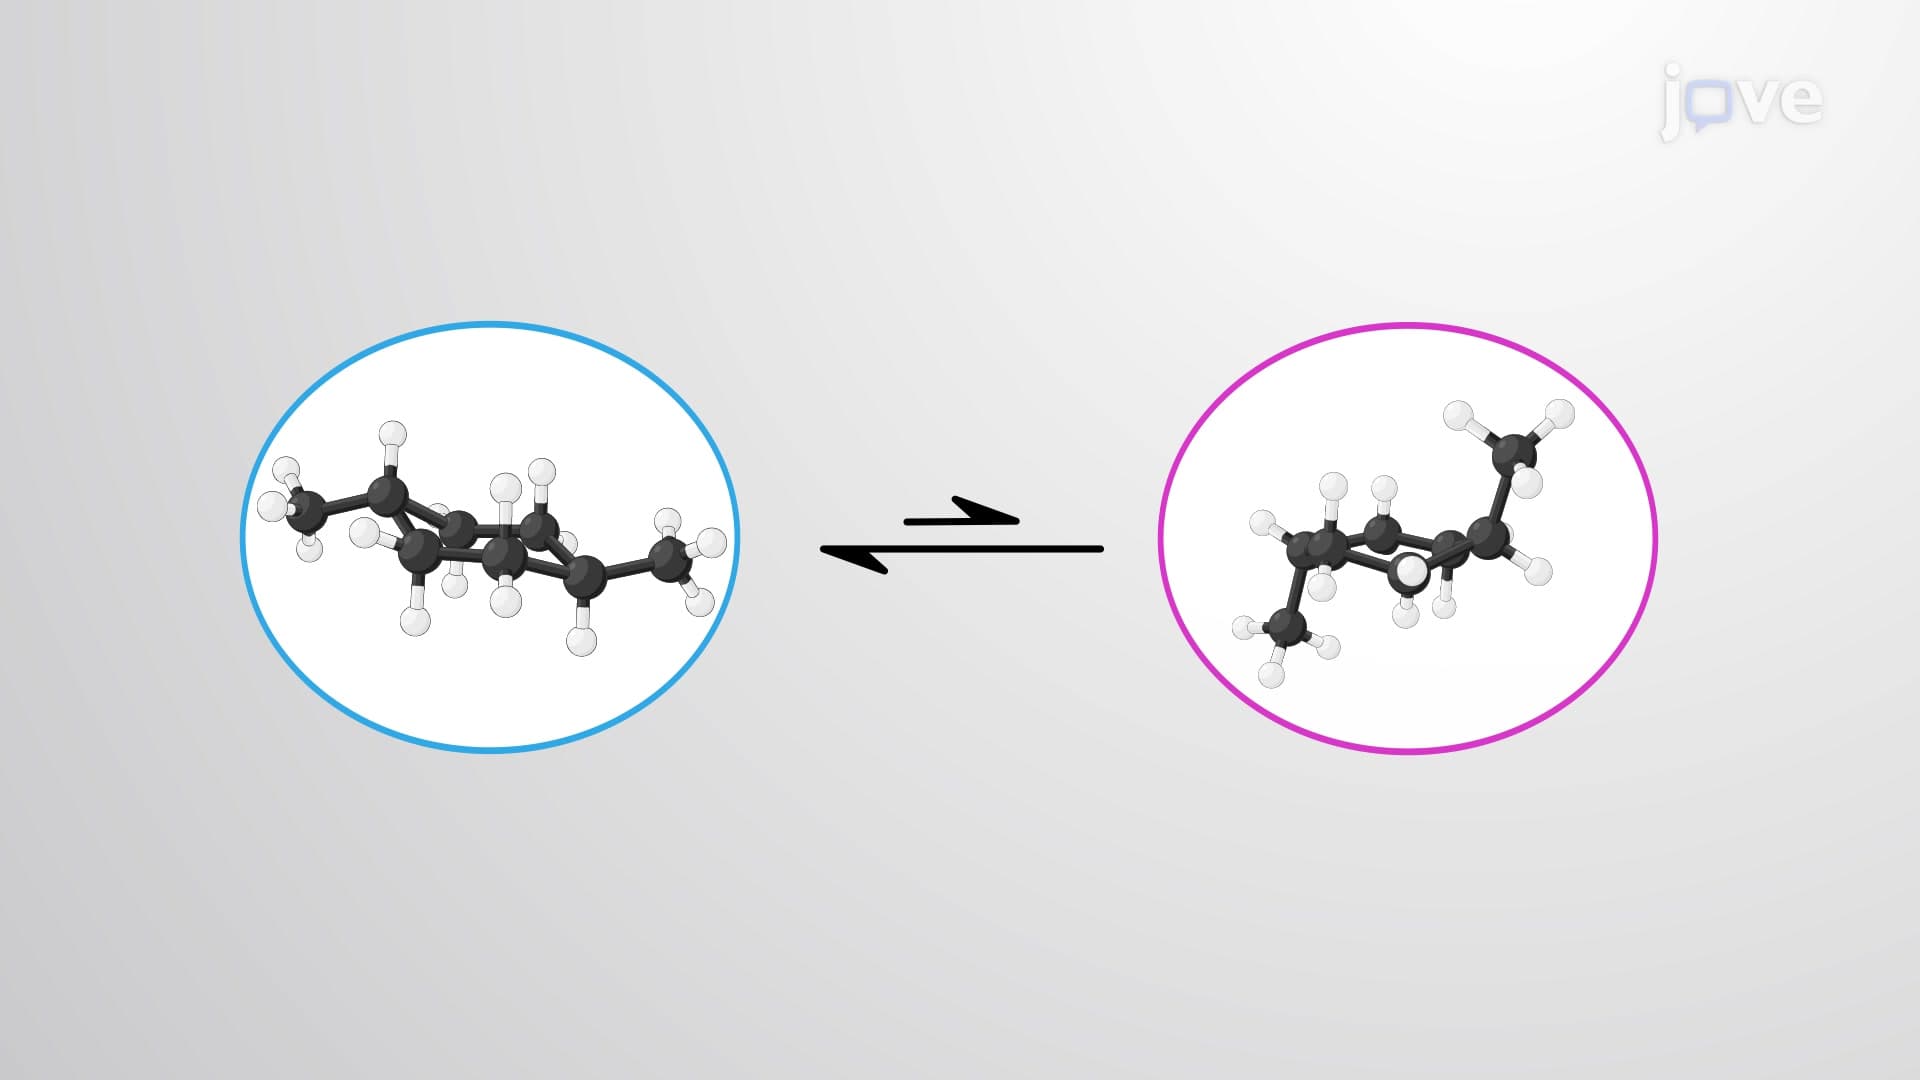
In cyclohexane, the substituents can occupy different positions generating distinct isomers.