Related Experiment Video
Updated: Jul 16, 2026

Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
Characterization of the potency, selectivity, and pharmacokinetic profile for six adenosine A2A receptor antagonists
Ming Yang1, Daniel Soohoo, Sandriyana Soelaiman
1Department of Pharmacology, CV Therapeutics, Inc., Palo Alto, CA 94304, USA. ming.yang@cvt.com

Abstract:
Antagonists of adenosine A2A receptors (A2A -antagonists) with different chemical structures have been developed by several pharmaceutical companies for the potential treatment of Parkinson's disease. Pharmacological characterization of these antagonists was incomplete, and different assay conditions were used in different labs. Therefore, we characterized the potencies, selectivities, and pharmacokinetic profiles of six prototypical A2A -antagonists. Displacements of [3H]MSX-2 and of [3H]CGS21680 binding to the human cloned and rat A2A receptors were performed. The rank order of potency of antagonists to displace [(3)H]MSX-2 binding to the human A2A was SCH58261 > or = Biogen-34 > or = Ver-6623 > or = MSX-2 > KW-6002 > > DMPX. For the rat striatal A2A, the order of potency was Biogen-34 > or = SCH58261 > or = Ver-6623 > or = MSX-2 > or = KW-6002 > > DMPX. SCH58261 was the most potent antagonist of the human A2A with a K(i) value of 4 nM, whereas Biogen-34 was the most potent antagonist of the rat A2A with a K(i) value of 1.2 nM. Similar results were obtained from cAMP assays. Selectivities of A2A-antagonists were determined using radioligands [3H]DPCPX, [3H]ZM241385, and [125I]-AB-MECA for A1, A2B, and A3 receptors, respectively. KW-6002 and Biogen-34 exhibited the highest selectivity for A2A vs A1 (human and rat), respectively. The pharmacokinetic profiles of antagonists were evaluated in vivo in rats. DMPX and KW-6002 had the greatest oral bioavailability. In contrast, SCH58261, MSX-2, and Ver-6623 had low or poor oral bioavailability. In summary, SCH58261, Biogen-34, MSX-2, and Ver-6623 had high affinities for both human and rat A2A receptors, with reasonable selectivity for A2A over A1 and A2B receptors. They are suitable as A2A -antagonists for in vitro pharmacological studies. Among the six A2A-antagonists, KW-6002 is the best for use in in vivo animal studies, particularly for a CNS target, based on its bioavailability, half life, and brain penetration.
Related Concept Videos
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:17
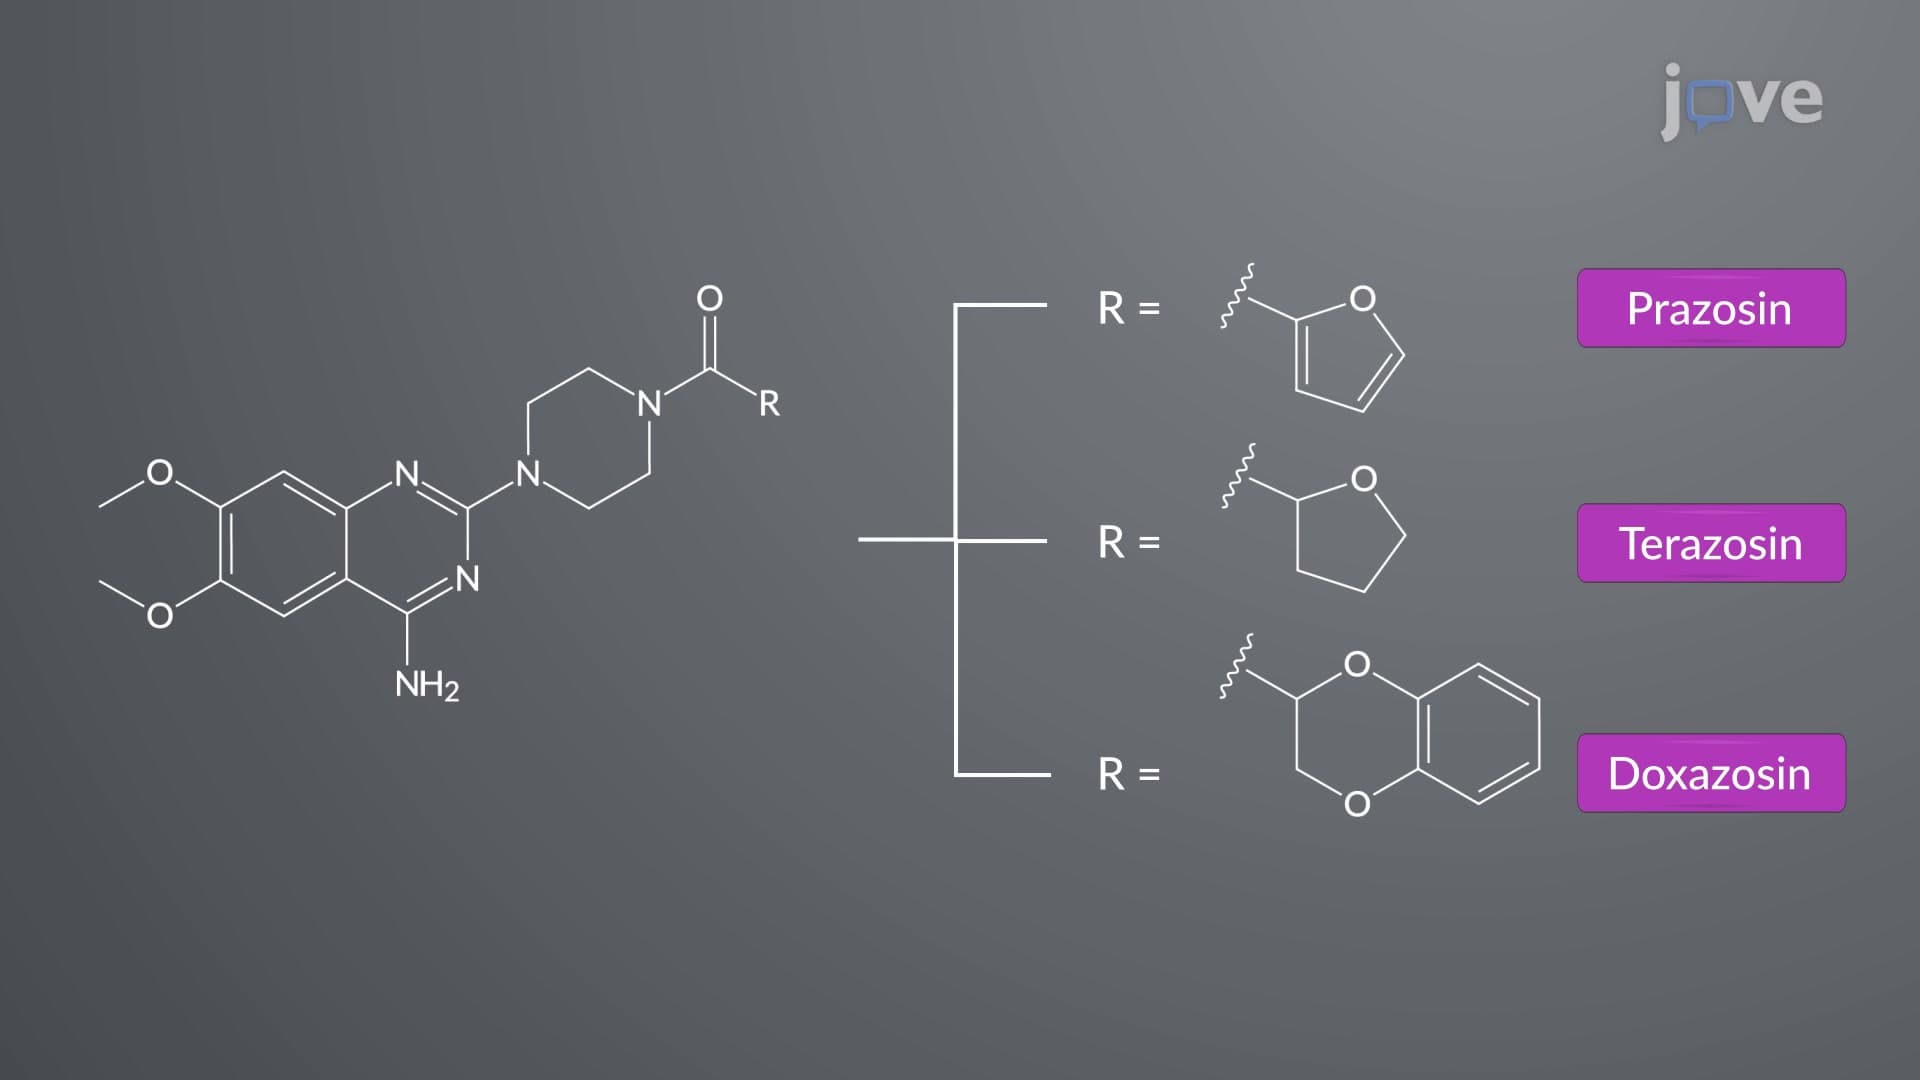
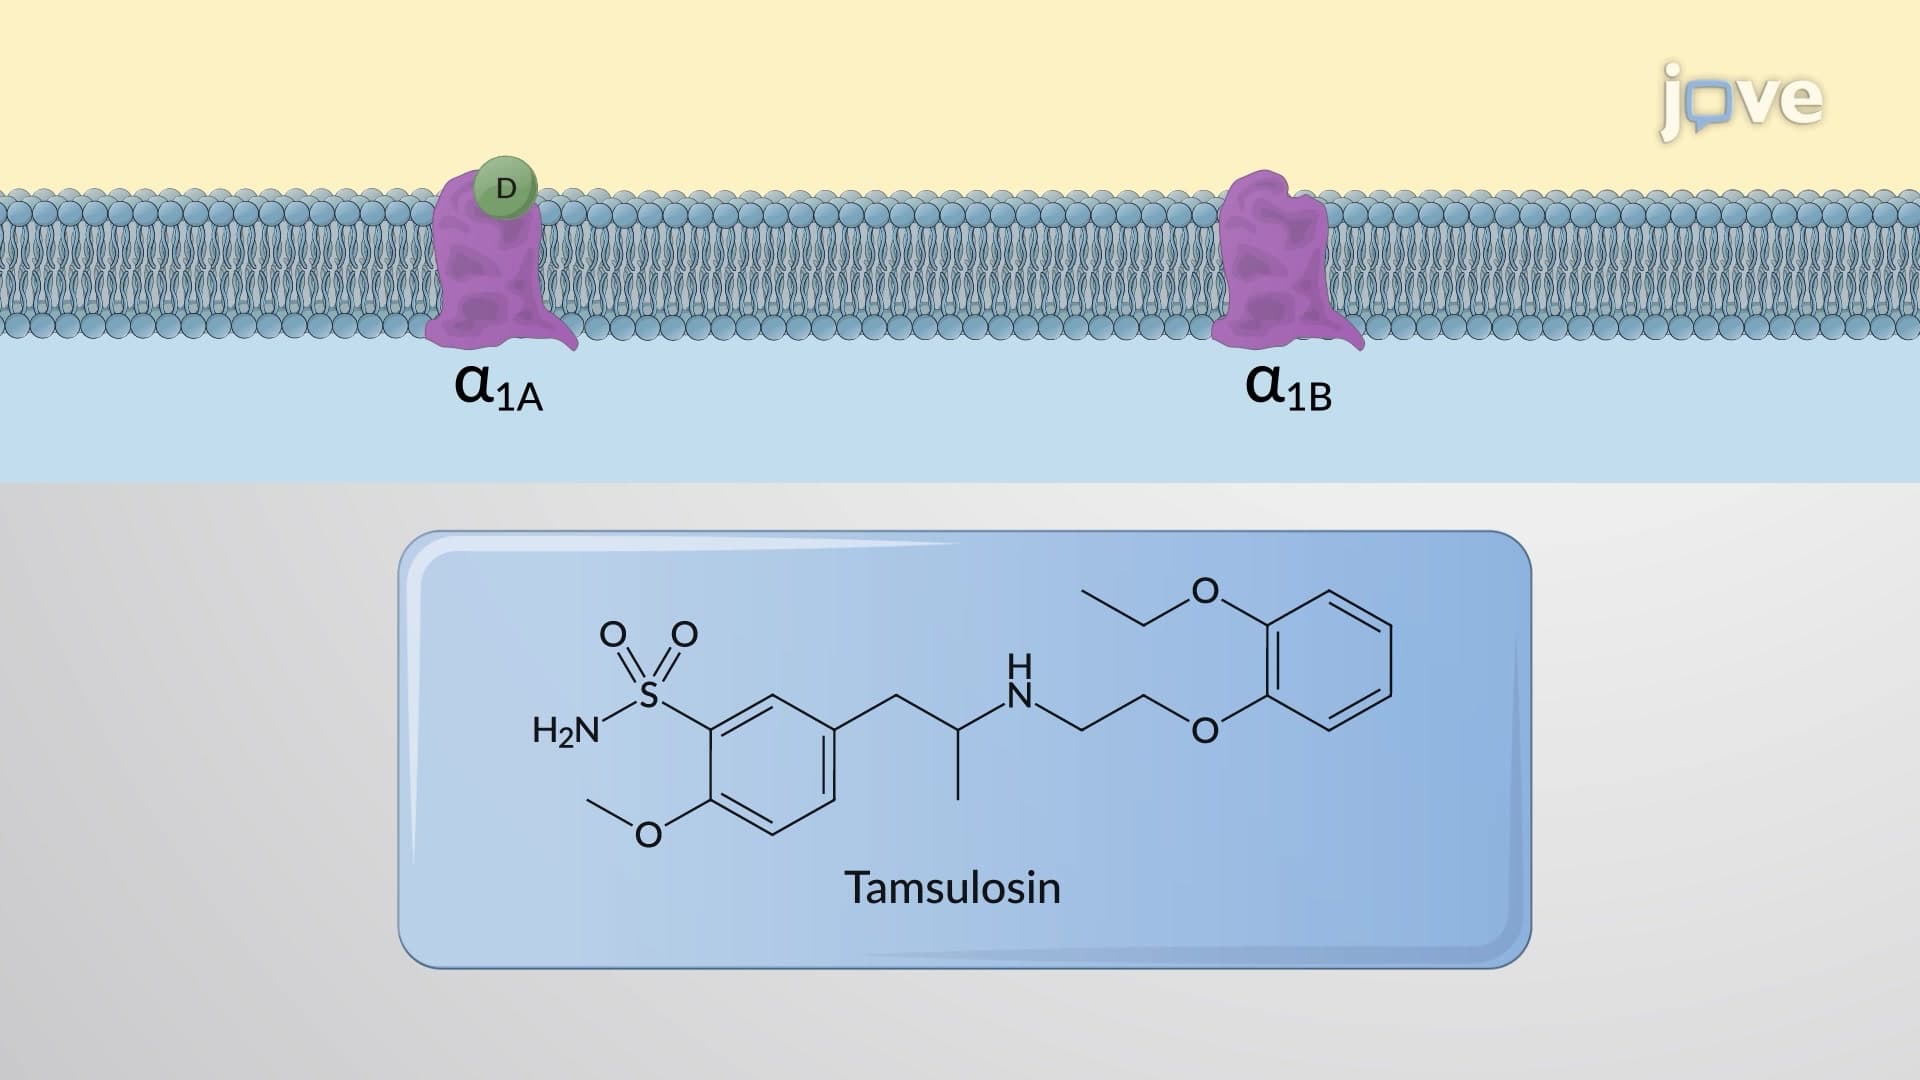
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:31
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:25
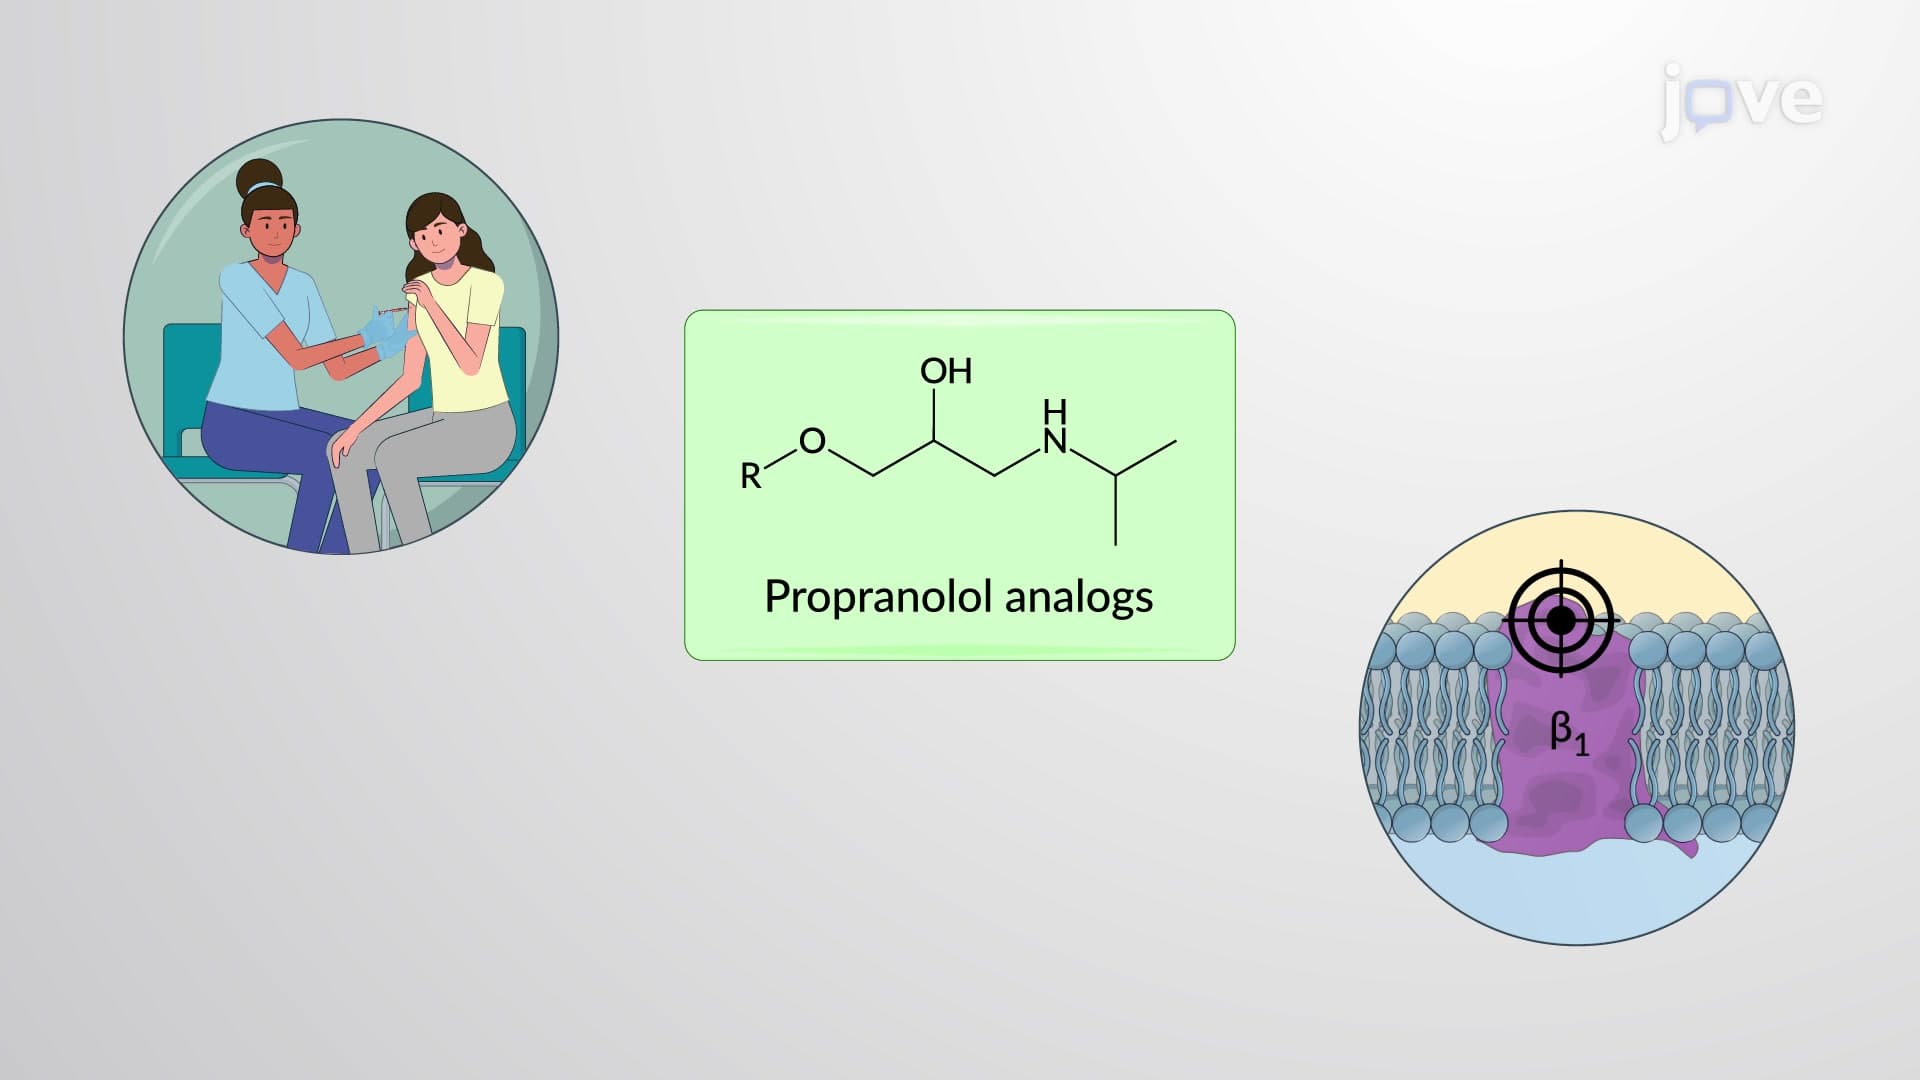
Adrenergic Antagonists: Chemistry and Classification of β-Receptor Blockers
 01:22
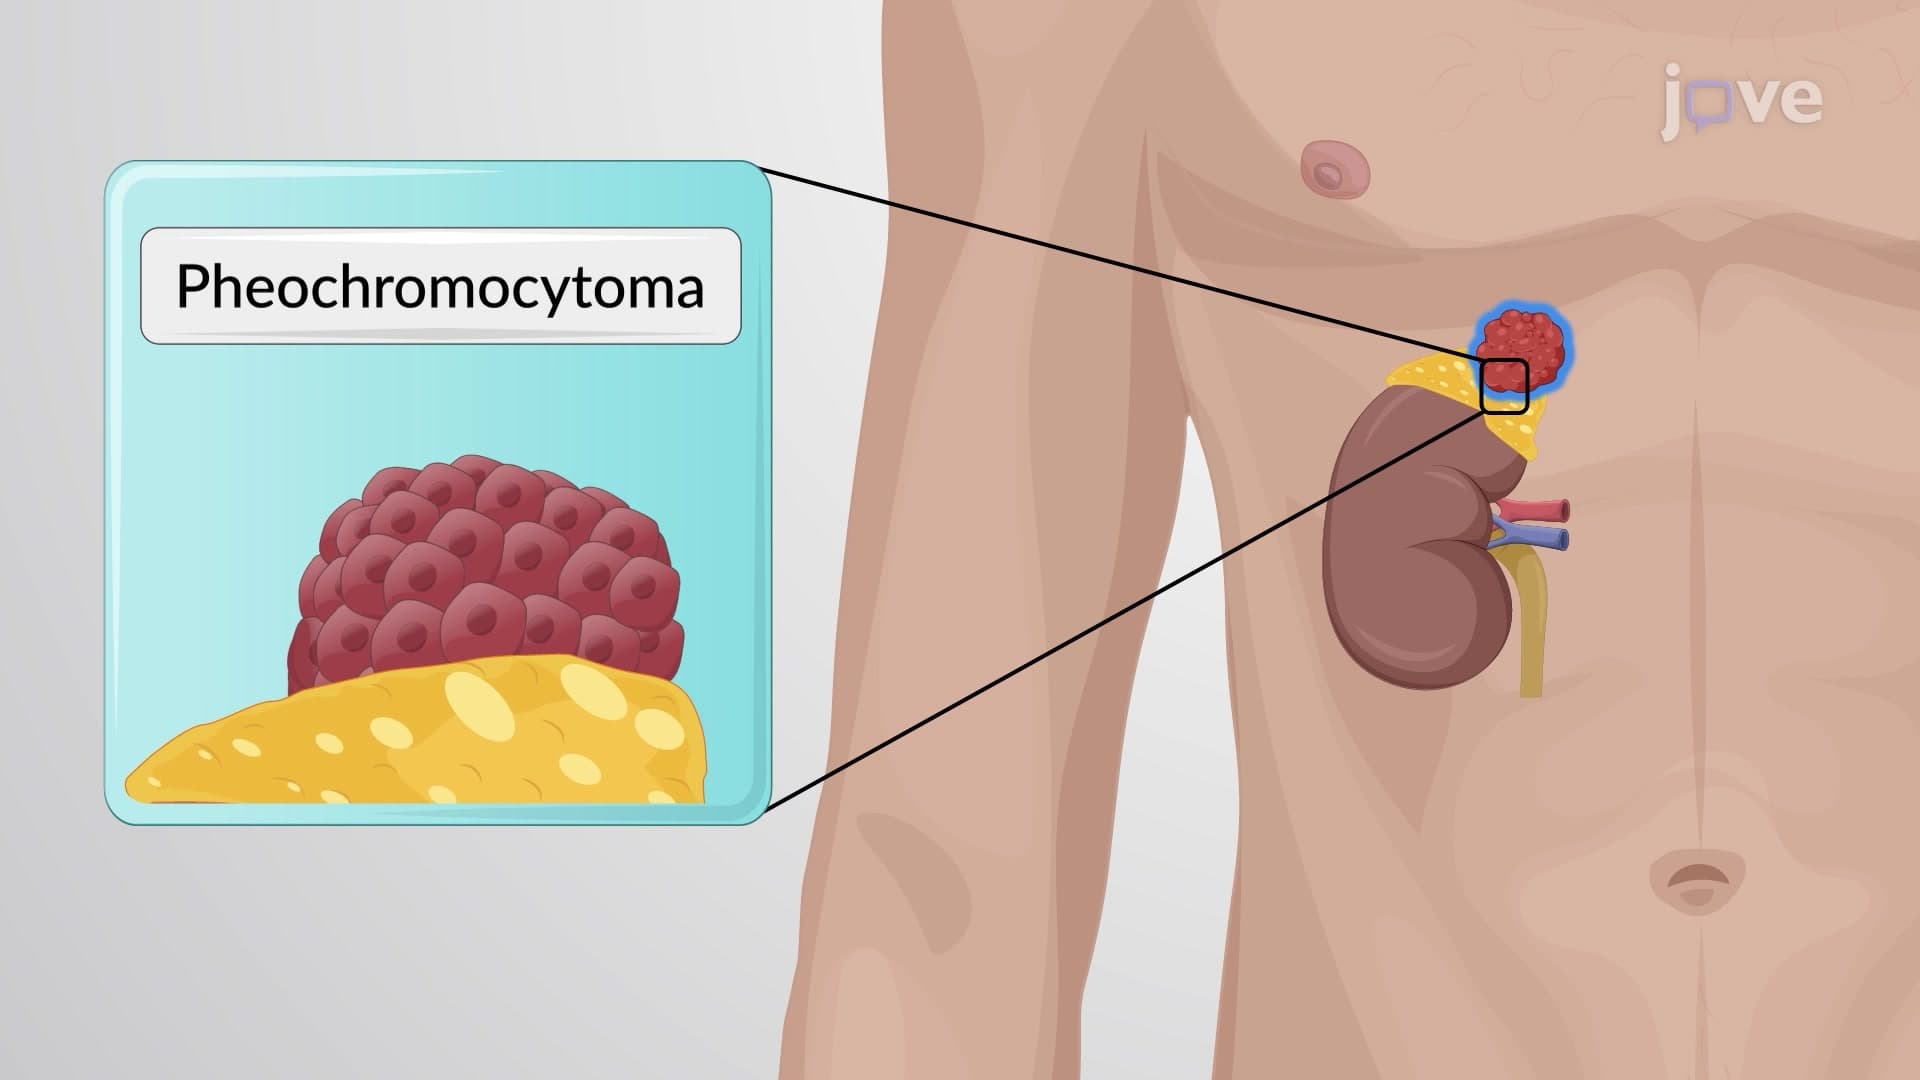
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally, α1-blockers effectively address urinary obstruction...
 01:25
Dose-Response Relationship: Selectivity and Specificity